UNITED STATES
SECURITIES AND EXCHANGE COMMISSION
Washington, D.C. 20549
FORM
(Mark One)
For the fiscal year ended
OR
For the transition period from to
Commission file number
(Exact name of registrant as specified in its charter)
(State or Other Jurisdiction of Incorporation or Organization) | (I.R.S. Employer Identification No.) | |
(Address of Principal Executive Offices) | (Zip Code) |
Registrant’s telephone number, including area code (
Securities registered pursuant to Section 12(b) of the Act:
Title of each class | Trading Symbol(s) | Name of each exchange on which registered | ||
Indicate by check mark if the registrant is a well-known seasoned issuer, as defined in Rule 405 of the Securities Act. Yes ☐
Indicate by check mark if the registrant is not required to file reports pursuant to Section 13 or Section 15(d) of the Act. Yes ☐
Indicate by check mark whether the registrant (1) has filed all reports required to be filed by Section 13 or 15(d) of the Securities Exchange Act of 1934 during the preceding 12 months (or for such shorter period that the registrant was required to file such reports), and (2) has been subject to such filing requirements for the past 90 days.
Indicate by check mark whether the registrant has submitted electronically, if any, every Interactive Data File required to be submitted pursuant to Rule 405 of Regulation S-T during the preceding 12 months (or for such shorter period that the registrant was required to submit and post such files).
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, a smaller reporting company, or an emerging growth company. See the definitions of “large accelerated filer,” “accelerated filer”, “smaller reporting company” and “emerging growth company” in Rule 12b-2 of the Exchange Act.
Large accelerated filer ☐ | Accelerated filer ☐ | Smaller reporting company | Emerging growth company |
If an emerging growth company, indicate by check mark if the registrant has elected not to use the extended transition period for complying with any new or revised financial accounting standards provided pursuant to Section 13(a) of the Exchange Act. ☐
Indicate by check mark whether the registrant has filed a report on and attestation to its management's assessment of the effectiveness of its internal control over financial reporting under Section 404(b) of the Sarbanes-Oxley Act (15 U.S.C 7262(b)) by the registered public accounting firm that prepared or issued its audit report. Yes
If securities are registered pursuant to Section 12(b) of the Act, indicate by check mark whether the financial statements of the registrant included in the filing reflect the correction of an error to previously issued financial statements.
Indicate by check mark whether any of those error corrections are restatements that required a recovery analysis of incentive‐based compensation received by any of the registrant’s executive officers during the relevant recovery period pursuant to §240.10D‐1(b).☐
Indicate by check mark whether the registrant is a shell company (as defined in Rule 12b-2 of the Exchange Act). Yes
The aggregate market value (approximate) of the registrant’s common equity held by non-affiliates based on the closing price of a share of the registrant’s common share for The Nasdaq Stock Market on June 30, 2025 (the last business day of the registrant’s most recently completed second fiscal quarter) was $
As of March 20, 2026, the total number of shares outstanding of the registrant’s Common Shares was
DOCUMENTS INCORPORATED BY REFERENCE:
Portions of the registrant’s definitive proxy statement for the registrant’s 2025 annual meeting of stockholders, to be filed within 120 days after the close of the registrant’s fiscal year, are incorporated by reference into Part III of this Annual Report.
TABLE OF CONTENTS
2
“Milestone Pharmaceuticals” and the Milestone logo appearing in this Annual Report on Form 10-K are unregistered trademarks of Milestone Pharmaceuticals Inc. All other trademarks, trade names and service marks appearing in this Annual Report on Form 10-K are the property of their respective owners. Solely for convenience, the trademarks and trade names in this Annual Report on Form 10-K may be referred to without the ® and ™ symbols, but such references should not be construed as any indicator that their respective owners will not assert their rights thereto.
This Annual Report on Form 10-K contains references to United States dollars and Canadian dollars. All dollar amounts referenced, unless otherwise indicated, are expressed in United States dollars. References to “$” are to United States dollars and references to “C$” are to Canadian dollars.
SPECIAL NOTE REGARDING FORWARD-LOOKING STATEMENTS
This Annual Report on Form 10-K contains forward-looking statements about us and our industry that involve substantial risks and uncertainties. All statements other than statements of historical facts contained in this Annual Report on Form 10-K, including statements regarding our strategy, future financial condition, future operations, projected costs, prospects, plans, objectives of management and expected market growth, are forward-looking statements. In some cases, you can identify forward-looking statements by terminology such as “aim,” “anticipate,” “assume,” “believe,” “contemplate,” “continue,” “could,” “design,” “due,” “estimate,” “expect,” “goal,” “intend,” “may,” “objective,” “plan,” “predict,” “positioned,” “potential,” “seek,” “should,” “target,” “will,” “would,” and other similar expressions that are predictions of or indicate future events and future trends, or the negative of these terms or other comparable terminology.
We have based these forward-looking statements largely on our current expectations and projections about future events and financial trends that we believe may affect our financial condition, results of operations, business strategy, and financial needs. These forward-looking statements are subject to a number of known and unknown risks, uncertainties, and assumptions, including risks described in the section titled “Risk Factors” and elsewhere in this Annual Report on Form 10-K, regarding, among other things:
| • | our expectations regarding the commercialization of CARDAMYST™ (etripamil) nasal spray for the conversion of acute symptomatic episodes of paroxysmal supraventricular tachycardia, or “PSVT,” to sinus rhythm in adults as well as our plans to develop and commercialize our product candidates; |
| • | the initiation, timing, progress, and results of our current and future clinical trials of etripamil, including our Phase 3 clinical trial of etripamil for the treatment of atrial fibrillation and rapid ventricular rate, and of our research and development programs; |
| • | our ability to develop and, if approved by regulatory authorities, commercialize etripamil in China, Hong Kong, Macau, and Taiwan through our license agreement with Corxel Pharmaceuticals, or “Corxel,” formerly Ji Xing Pharmaceuticals Limited, JIXING; |
| • | our estimates regarding expenses, future revenue, capital requirements, and needs for additional financing; |
| • | our ability to establish collaborations or obtain additional funding; |
| • | our ability to obtain regulatory approval of our current products for new indications and/or of our future product candidates; |
| • | our expectations regarding the potential market size and the rate and degree of market acceptance of etripamil and any future product candidates; |
| • | our ability to fund our working capital requirements and expectations regarding the sufficiency of our capital resources; |
3
| • | the implementation of our business model and strategic plans for our business, etripamil, and any future product candidates; |
| • | our intellectual property position and the duration of our patent rights; |
| • | developments or disputes concerning our intellectual property or other proprietary rights; |
| • | our expectations regarding government and third-party payor coverage and reimbursement; |
| • | our ability to compete in the markets we serve; |
| • | the impact of government laws and regulations; |
| • | developments relating to our competitors and our industry; |
| • | the effects of international trade policies, including tariffs, sanctions, and trade barriers; and |
| • | other factors that may impact our financial results. |
The foregoing list of risks is not exhaustive. Other sections of this Annual Report on Form 10-K may include additional factors that could harm our business and financial performance. Moreover, we operate in a very competitive and rapidly changing environment. New risk factors emerge from time to time, and it is not possible for our management to predict all risk factors, nor can we assess the impact of all factors on our business or the extent to which any factor, or combination of factors, may cause actual results to differ materially from those contained in, or implied by, any forward-looking statements.
In light of the significant uncertainties in these forward-looking statements, you should not rely upon forward-looking statements as predictions of future events. Although we believe that we have a reasonable basis for each forward-looking statement contained in this Annual Report on Form 10-K, we cannot guarantee that the future results, levels of activity, performance, or events and circumstances reflected in the forward-looking statements will be achieved or occur at all. You should refer to the section titled "Risk Factors" for a discussion of important factors that may cause our actual results to differ materially from those expressed or implied by our forward-looking statements. Furthermore, if our forward-looking statements prove to be inaccurate, the inaccuracy may be material. Except as required by law, we undertake no obligation to publicly update any forward-looking statements, whether as a result of new information, future events or otherwise. The Private Securities Litigation Reform Act of 1995 and Section 27A of the Securities Act of 1933, as amended, or the Securities Act, do not protect any forward-looking statements that we make in connection with this Annual Report on Form 10-K.
4
Summary of Risk Factors
Our business is subject to numerous risks and uncertainties, including those described in Item 1A “Risk Factors”. These risks include, but are not limited to the following:
| • | We have incurred significant operating losses since inception and anticipate that we will continue to incur substantial operating losses for the foreseeable future until revenue from CARDAMYST is sufficient to fund our operations, if ever, and may never achieve or maintain profitability. |
| • | We will require substantial additional funding to finance our operations. If we are unable to raise capital when needed, we could be forced to delay, reduce, or terminate our development of etripamil or other operations, including the continued commercialization of CARDAMYST. |
| • | Greater than expected returns of CARDAMYST may exceed our reserve for returns, which would adversely affect our revenue and operating results. |
| • | Raising additional capital may cause dilution to our shareholders, restrict our operations, or require us to relinquish rights to our product candidates. |
| • | We currently have one approved product, CARDAMYST™ (etripamil) nasal spray, a prescription medication for the conversion of acute symptomatic episodes of PSVT. We are currently pursuing clinical development for subsequent etripamil indications. If we are not able to obtain required regulatory approvals for subsequent etripamil indications or any future product candidates, our ability to generate revenue will be adversely affected. |
| • | We may not be successful in our efforts to expand our pipeline of product candidates beyond etripamil. |
| • | The development of additional product candidates is risky and uncertain. |
| • | Success in preclinical studies or earlier clinical trials may not be indicative of results in future clinical trials, and we cannot assure you that any ongoing, planned, or future clinical trials will lead to results sufficient for the necessary regulatory approvals. |
| • | We may encounter substantial delays or difficulties in our clinical trials. |
| • | Enrollment and retention of patients in clinical trials is an expensive and time-consuming process and could be delayed, made more difficult, or rendered impossible by multiple factors outside our control. |
| • | If we are unable to successfully implement and maintain sales, marketing and distribution capabilities for CARDAMYST for the treatment of PSVT or any product candidate that may receive regulatory approval, we may not be successful in commercializing CARDAMYST for the treatment of PSVT or our product candidates if and when they are approved. |
| • | CARDAMYST, along with any subsequent etripamil indications or any other product candidates, if approved, may fail to achieve market acceptance by physicians, patients, third-party payors or others in the medical community necessary for commercial success. |
| • | The success of etripamil will be dependent on its use in accordance with labeled instructions for use. |
| • | If the market opportunities for CARDAMYST, any subsequent indications for etripamil, and any future product candidates are smaller than we estimate, our business may suffer. |
| • | Coverage and adequate reimbursement may not be available for etripamil or any future product candidates, which could make it difficult for us to gain market acceptance. |
| • | Even if we maintain regulatory approval for etripamil or any future product candidates from the Food and Drug Administration, or “FDA,” we may never obtain approval of etripamil or any future product candidates outside of the United States, which would limit our market opportunities and could harm our business. |
| • | Even if we maintain approval for CARDAMYST, or we obtain regulatory approval for subsequent etripamil indications or any future product candidates, they, and etripamil for the treatment of PSVT, will remain subject to ongoing regulatory oversight. |
| • | We will rely on third parties to produce clinical and commercial supplies of etripamil and any future product candidates, and if those third parties perform in an unsatisfactory manner, it may harm our business. |
| • | We rely on third parties to conduct, supervise, and monitor our preclinical studies and clinical trials, and if those third parties perform in an unsatisfactory manner, it may harm our business. |
| • | Etripamil is intended to be used with a nasal spray device, which may result in additional regulatory and supply risks. |
5
| • | If we are unable to obtain and maintain patent protection for etripamil or any future product candidates, or if the scope of the patent protection obtained is not sufficiently broad, our competitors could develop and commercialize drugs similar or identical to ours, and our ability to successfully commercialize our product candidates may be impaired. |
| • | Our future success depends on our ability to retain key executives and to attract, retain, and motivate qualified personnel. |
| • | The market price of our common shares has been and may continue to be volatile and fluctuate substantially, and you could lose all or part of your investment. |
| • | Our common shares are thinly traded, and our shareholders may be unable to sell their shares quickly or at market price. |
| • | Concentration of ownership of our common shares among our existing executive officers, directors and principal shareholders may prevent new investors from influencing significant corporate decisions. |
6
PART I
ITEM 1. BUSINESS
Company Overview
We are a biopharmaceutical company focused on the development and commercialization of innovative cardiovascular medicines. Our approved product CARDAMYSTTM (etripamil) nasal spray is available in the United States and is the first and only FDA approved self-administered treatment for use by patients anywhere, anytime an attack of PSVT occurs. We continue to seek, either directly or through collaboration with our partners, marketing approval from regulatory agencies responsible for regions outside the United States. We are also developing etripamil for the indication of atrial fibrillation with rapid ventricular rate, or “AFib-RVR.”
We are currently focusing our efforts and financial resources on (i) the commercialization of CARDAMYSTTM (etripamil) nasal spray for the treatment of PSVT, (ii) the development of etripamil for the treatment of AFib-RVR and (iii) corporate development activities that have the potential to increase company value through strategic collaborations.
CARDAMYSTTM (etripamil) nasal spray for the Treatment of PSVT
On December 12, 2025, we announced that the FDA approved our first commercial product, CARDAMYST (etripamil) nasal spray, a prescription medication for the conversion of acute symptomatic episodes of PSVT, to sinus rhythm in adults. We are focused on the commercialization of CARDAMYST, which became available in retail pharmacies in the first quarter of 2026. We manufacture CARDAMYST with a global supply chain that produces the active pharmaceutical ingredient, etripamil, and then formulates and assembles it into the delivery device, a nasal spray, before completing packaging of the saleable product which is then distributed via a third party logistics provider to wholesalers for delivery to retail pharmacies once prescriptions are received. To create demand, our sales team of approximately 60 representatives expects to engage healthcare providers who we believe will treat around 500,000 patients with a diagnosis of PSVT in 2026.
We designed the molecule of etripamil, and we are commercializing and further developing the drug, a novel, potent, and rapid-onset calcium channel blocker, as a nasal spray to be administered by the patient to terminate episodes of transient cardiovascular conditions as they occur. Rapid pharmacological action is both appropriate and sufficient to resolve an episode of supraventricular tachycardia, or “SVT.” We are also developing etripamil to provide acute ventricular rate control for patients with symptomatic episodes of atrial fibrillation with rapid ventricular rate and are exploring other therapeutic applications in which a patient-administered, rapid-onset, non-dihydropyridine calcium channel blocking agent could provide patient benefit.
In our work to develop potential therapies, we sought to create new chemical entities as analogs of known molecular classes with clinically validated mechanisms of action. Our goal was to preserve the beneficial pharmacology of existing molecules while altering their pharmacokinetic profile with focused medicinal chemistry to produce drugs that are fast acting. As a result, we created a series of novel non-dihydropyridine, L-type calcium channel blockers containing chemical ester moieties that preserved the desired pharmacology on the heart but that could be rapidly metabolized in the blood by serum esterases. Etripamil resulted from this effort as a new chemical entity with a fast pharmacodynamic effect in humans, relative to oral calcium channel blockers.
We believe that the following attributes of etripamil make it a better treatment candidate for certain episodic cardiovascular conditions than current standards of care:
• | Action: Etripamil is designed to be rapidly metabolized by blood-borne esterases, with the goal of reducing either the duration of side effects that may occur with acute treatment or the long-term side effects that may occur with chronic drug therapy. |
7
| • | Absorption: Etripamil is designed to be absorbed into the bloodstream in less than 10 minutes through the inner lining of the nose, yielding a rapid-onset pharmacologic pattern consistent with parenterally administered drugs. |
| • | Administration: Etripamil is designed, and has been investigated, to empower patients to self-administer treatment outside of a medical setting and as prompted by a patient’s symptoms. |
PSVT
PSVT is a serious, markedly symptomatic, and recurring cardiac arrhythmia, caused by altered electrical conduction within the heart involving the atrioventricular, or “AV,” node and over an abnormal electrical circuit in most cases. The common form of PSVT, AV nodal reentrant tachycardia, or “AVNRT,” has an abnormal limb of electrical circuitry within the AV node that is the substrate of the tachycardia and forms the basis for a reentrant arrhythmia, resulting in excessively rapid beating of both the atria and ventricles.
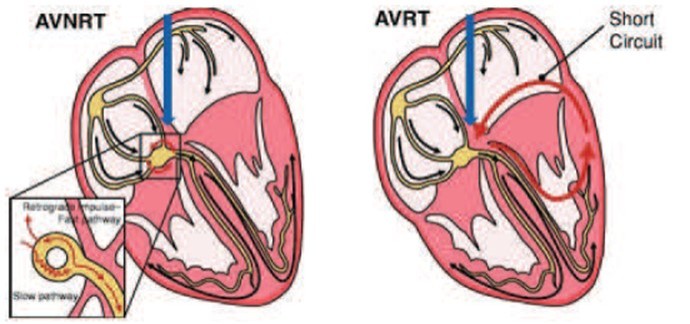
In the next most common form of PSVT, atrioventricular reciprocating tachycardia, or “AVRT,” there is an abnormal limb of electrical tissue, or bypass tract, that directly connects the atria and the ventricles. In AVRT, the bypass tract allows the signal to travel between the atria and ventricles as a “short circuit.” AVRT also involves the AV node, in addition to the bypass tract. Thus, for both AVNRT and AVRT – two PSVT types that require the AV node as a part of their abnormal circuit (i.e., AV-nodal dependent PSVTs) – a drug which targets the AV node can represent a potential treatment. Non-dihydropyridine calcium channel blockers are a class of drugs that target the AV node, reflecting why etripamil has been an excellent therapeutic candidate for AVNRT and AVRT disruption and termination.
Current Treatment Options for PSVT
Treatment for PSVT depends on the frequency, duration, and severity of the episodes, as well as patient preference for pharmacologic or invasive treatment. Current options for patients with PSVT to acutely terminate an episode of arrhythmia include intravenous, or “IV,” medication (e.g., IV adenosine, IV calcium channel blockers), administered in an emergency department, which transiently alter electrical conduction over the AV node leading to termination of AV-dependent arrhythmias and restoration of sinus rhythm; or an external shock delivered in the emergency department. Of note, IV adenosine temporarily causes complete heart block and patients have reported experiencing chest tightness, flushing, and a sense of impending doom. Two other modalities, though of low effectiveness, are vagal maneuvers or oral medications administered at the onset of an episode (e.g., calcium channel blockers, beta blockers). Long-term, preventative strategies include chronic drug therapy to reduce the frequency of episodes, though patients exhibit break-through episodes of PSVT and emergency department visits for the same, despite chronic medications and cardiac ablation to potentially cure the disease. Patients may also elect not to treat their PSVT and simply endure episodes of SVT when they occur.
8
Of note, chronic medication can lead to side effects such as sexual dysfunction or fatigue in the case of beta blockers and constipation in the case of verapamil. Some patients discontinue chronic oral medication due to intolerable side effects. Based on our market research, we estimate that approximately two thirds of patients with PSVT have been prescribed chronic medications such as beta blockers or calcium channel blockers to prevent episodes of SVT or to treat other concomitant conditions such as hypertension.
Ablations, a potentially curative treatment for PSVT, is an invasive procedure, which works by directly cauterizing or freezing the short circuit that is the substrate for the abnormal rhythm. This is achieved in an electrophysiology lab via catheters that enter the patient’s heart through the blood vessels and that deliver burning or freezing to ablate the abnormal electrical tissue. Ablation single-procedure success rates for PSVT are reported to be 91% to 96%. However, we estimate that less than 10% of eligible patients with PSVT per year choose this option, which we believe is due primarily to anxiety related to the procedure. Although ablations are generally considered to be safe by the treating community, as with any invasive procedure there are potential complications, which include bleeding, blood clots, pericardial tamponade, and transient or permanent heart block, with the latter requiring permanent pacemaker implantation.
PSVT Market Overview
PSVT is a condition that causes a patient’s heart to suddenly start beating faster than normal. It can be life-altering as PSVT is highly symptomatic, characterized by unpredictable attacks of a racing heart, often exceeding 150 beats per minute. Symptoms of PSVT arise suddenly and may include palpitations, sweating, chest pressure or pain, shortness of breath, sudden onset of fatigue, lightheadedness or dizziness, fainting, and anxiety, causing many patients to interrupt their daily activities at the time of symptom-onset. The impact and morbidity from an episode of PSVT can be especially detrimental in patients with underlying cardiovascular or medical conditions, such as heart failure, obstructive coronary disease, or dehydration. The uncertainty of when such an attack of PSVT will strike or how long it will persist is often anxiety-provoking, reducing patients’ quality of life and preventing participation in many desired activities. Drugs approved for the treatment of attacks of PSVT include adenosine, verapamil, and diltiazem, with all being administered intravenously under medical supervision, usually in the emergency department. Other oral drugs are sometimes used to treat attacks in a concept called “pill in the pocket.” However, those drugs have never been proven effective or safe and are not approved for this use. Doctors are often frustrated by the limited effective treatment options with the only approved options involving prolonged, unpleasant, and costly trips to the emergency department or, for some patients, an invasive ablation procedure. PSVT can be traumatic for patients, frustrating for healthcare providers, and costly for payors. With no pharmaceutical innovation in the treatment of PSVT for more than 30 years and a movement in the healthcare system to enable patient-centered care, we believe there is an opportunity to help patients living with PSVT to take greater control over their PSVT.
We believe that PSVT is a large and under-recognized market which we estimate affects more than two million Americans. From this diagnosed population, we define the immediate target addressable market for CARDAMYST as approximately 50% of patients with PSVT who have sufficient disease burden that they are compelled to seek care for their condition and are primarily managed by approximately 40,000 healthcare providers composed primarily of clinical cardiologists, interventional cardiologists and electrophysiologists. The remaining patients with PSVT can become addressable over time, as they are inconsistently managed (cycling in and out of the healthcare system). Furthermore, PSVT is expected to increase in diagnosed prevalence in coming years as wearable electrocardiogram, or “ECG,” technology (e.g., smartphone, watches) becomes both more adept at diagnosing PSVT and more widely used by patients and clinical practitioners, in turn shortening the time to diagnosis.
Following the release of data from the RAPID clinical study, in market research, cardiologists reported a willingness to prescribe CARDAMYST to approximately 50% of the patients with PSVT in their care, which suggests approximately 500,000 to 800,000 patients can potentially be treated with CARDAMYST in peak years. Additionally, we believe that this cardiology-identified group of patients may use CARDAMYST to treat a median of three to five episodes per year, based on the projected number of self-reported longer and more intense episodes experienced by patients, as well as the patient utilization experience in our Phase 3 clinical trials. This implies a peak demand potential in the United States for CARDAMYST of 2.5 million to 4 million episodes treated per year.
9
Current treatment for PSVT also consumes significant healthcare resources. Research published in the American Journal of Cardiology in 2020 shows that total healthcare expenditures in the year following a diagnosis of PSVT ranged from $20,000 to $30,000 per patient which were significantly higher than the expenditures observed for patients without PSVT. These significant increases included increased emergency department visits and hospitalization costs. Of note, catheter ablations following diagnosis represented only 23% of this increased spend, meaning most costs were unrelated to ablations. Recent data from the Healthcare Cost and Utilization Project, or “HCUP,” database indicate that in 2019 there were approximately 140,000 emergency department, or “ED,” visits for PSVT when coded (for billing) in the primary diagnostic position, and a total of approximately 525,000 ED visits when PSVT was coded in any diagnostic position. Of these, approximately 25% of ED admissions for PSVT resulted in a hospital admission. HCUP estimates a total of approximately 40,000 to approximately 120,000 inpatient admissions for PSVT in 2019 (based again if a PSVT billing code was found in the primary v. any diagnostic position). Despite the effectiveness of catheter ablation, claims data suggests that only approximately 15% of patients with PSVT are ablated over a three-year period, leading to a total of approximately 100,000 catheter ablations annually. In total, up to $15.0 billion is spent annually in the United States on the management of PSVT.
PSVT Continued Development Highlights
In November 2025, we submitted our marketing authorization application, or “MAA,” to the European Medicines Agency, or “EMA,” for review for approval to market etripamil nasal spray, under the trade name TACHYMISTTM, in European Union. The MAA utilizes much of the clinical, manufacturing and quality data package that was submitted in the NDA which lead to the US FDA approval.
In January 2025, our licensing partner, Corxel (formerly Ji Xing Pharmaceuticals Limited, JIXING), announced that the Center for Drug Evaluation, or “CDE,” of the National Medical Products Administration, or “NMPA,” of the People’s Republic of China has accepted the New Drug Application, or “NDA,” for etripamil nasal spray for the treatment of PSVT. The NDA to the NMPA included data from the successful Phase 3 JX02002 clinical trial of etripamil nasal spray in patients with PSVT in China in addition to data included in the NDA that Milestone submitted to the FDA.
The 500-patient Phase 3 trial (JX02002) met its primary endpoint, with a Kaplan Meier analysis shows a statistically significantly greater proportion of patients who self-administered etripamil converted from PSVT to sinus rhythm within 30 minutes compared to placebo (40.5% vs. 15.9%, respectively; hazard ratio [HR] = 3.00; 95% CI 1.58-5.71; p<0.001). Statistically significant (p<0.05) results were also shown for the secondary efficacy endpoints for percent of patients’ PSVT converted to sinus rhythm by 10, 15, 45 and 60 minutes after self-administration of study drug.
Corxel further reported that, overall, treatment emergent adverse events were comparable between treatment groups, and there were no reported serious adverse events related to etripamil. The safety and tolerability data from the JX02002 trial were consistent with previous clinical studies. This important study further expands the etripamil global development program to more than 2,000 unique patients treated with etripamil.
Etripamil Nasal Spray for the Treatment of AFib-RVR
Similar to our approach for PSVT, we believe that etripamil has the potential to help people experiencing a symptomatic episode of AFib-RVR to self-treat and to conveniently, reliably, and quickly reduce their elevated heart rate, with the goal of reducing the need for emergency department utilization. We completed a successful Phase 2 study, named “ReVeRA” or the “ReVeRA study”, in patients presenting urgently with episodes of AFib-RVR to the emergency department. We have published the ReVeRA study and results, which demonstrated that patients receiving etripamil nasal spray experienced rapid and statistically superior ventricular rate reduction and improved symptom-relief compared to placebo, with safety and tolerability findings generally consistent with those observed in our PSVT program. We believe these data support the continued development of etripamil, self-administered in the medically unmonitored setting, for the treatment of AFib-RVR.
Incorporating FDA’s guidance, we have developed a Phase 3 registrational program to evaluate self-administered etripamil as a potential treatment for patients with AFib-RVR. We intend to pursue a supplemental new drug application, or “sNDA,”
10
regulatory approval pathway for a potential second indication for etripamil in Afib-RVR. As such, we will leverage the PSVT indication and PSVT program data, along with a single pivotal Phase 3 study in patients with AFib-RVR.
Atrial Fibrillation
AFib is a common arrhythmia with an irregular and often rapid heart rate that can increase the risk of stroke, embolism, heart failure, and other cardiac morbidities. AFib is often highly symptomatic, particularly when episodes have rapid heart rates. Symptoms include heart palpitations, shortness of breath, fatigue, and weakness, and underlying cardiac disorders can be worsened. Episodes of atrial fibrillation can come and go, or patients may have AFib that does not resolve. Although the heart arrhythmia in AFib itself usually is not life-threatening, it is a serious medical condition that sometimes requires emergency treatment. Because AFib is associated with elevated risk of embolism and stroke, anticoagulant medications, also called blood thinners, are commonly prescribed to manage this risk. Uncertainty around symptom timing and episode length may impact a patient’s quality of life. During AFib, the heart’s two upper chambers, the atria, beat chaotically and irregularly—out of coordination with the two lower chambers, the ventricles, of the heart, as shown in the figure below.
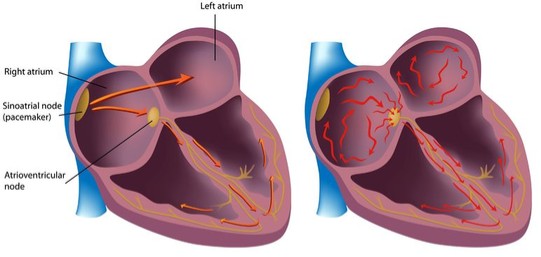
Categorization of AFib is customarily defined as: paroxysmal, which involves episodes of AFib that resolve spontaneously within seven days of symptom onset; persistent, which involves episodes that fail to terminate within seven days of symptom onset and require treatment to convert back to sinus rhythm; long-standing persistent, which involves episodes of atrial fibrillation that last longer than one year despite continued attempts to restore sinus rhythm; and permanent, which involves a joint decision by the treating provider and patient to no longer pursue cardioversion and leave the patient in AFib, focusing on rate control and symptom management. Prevalence of categories includes approximately 40% of patients with AFib as paroxysmal stage, 30% as persistent and long-standing persistent stage, and 30% as permanent. Concomitant structural heart irregularities including valvular dysfunction and the presence of active symptoms may also help to characterize patients and influence treatment decisions.
Current Treatment Options for Atrial Fibrillation with Rapid Ventricular Rate
A common complication of AFib is an RVR which can be defined as a heart rate of ≥110 beats per minute. Irregular and inefficient cardiac pumping function caused by a RVR accounts for hemodynamic instability and many of the arrhythmia’s symptoms.
There are currently two major approaches to managing AFib: rate control to lower a RVR or upward excursions in rate, and rhythm control to both restore and maintain normal sinus rhythm and to prevent recurrent AFib. Either of these treatment approaches, often performed by pharmacological means, may be given chronically or acutely, depending on patient preference and episode frequency and severity, though each has limitations. The decision to pursue rate or rhythm control for episodes of AFib is dependent on factors including episode severity, episode frequency, patient preference, and safety and tolerability of treatments. Several rhythm control strategies exist, including electrical cardioversion, catheter-based cardiac ablation, and antiarrhythmic drug therapy. For rate control, the rapid heart rate of AFib is can be treated with chronically administered AV nodal acting drugs (e.g., calcium channel blockers, beta blockers, digoxin) to control RVR and symptoms from RVR, and to improve cardiac function/hemodynamic stability. Oral rate control drugs used acutely do not provide immediate ventricular rate control due to a 30-to-120-minute delayed onset of action. Breakthrough
11
episodes of symptomatic atrial fibrillation often require urgent medical treatment with IV calcium channel blockers and IV beta blockers under medical supervision in an emergency department to quickly reduce heart rate before transitioning a patient back to oral therapy.
Of note, AHA and ACC guidelines do not explicitly acknowledge the as needed (PRN) approach to acutely administering oral rate control agents, however, market research conducted by us indicates a significant share of patients are managed this way. This market research, from 2018, estimated that approximately 40% of patients use acutely administered rate control medication to manage symptoms of atrial fibrillation. Additionally, our physician market research, from 2021, suggests that both clinical/interventional cardiologists and electrophysiologists prescribe PRN rate control for some of their patients with AFib.
Clinical Development Plan for Atrial Fibrillation with Rapid Ventricular Rate
In November 2023, we presented positive Phase 2 data from the ReVeRA study as a Featured Science Presentation at the American Heart Association Scientific Meetings (Philadelphia, PA) and as simultaneously published in Circulation: Arrhythmia and Electrophysiology. The Phase 2, double-blind, placebo-controlled, study, Reduction of Ventricular Rate in Patients with Atrial Fibrillation, or “ReVeRA”, was conducted in patients presenting urgently (e.g., to an emergency department) with AFib-RVR. ReVeRA’s objectives included evaluation of the potential effectiveness of etripamil NS to reduce RVR and to improve symptoms in patients with AFib and evaluation of the drug’s safety. The trial enrolled 87 patients and dosed 56 patients with blinded study drug (the latter having a ventricular rate of ≥110 beats per minute, or “bpm”) prior to receiving study drug.
Data showed that delivery of etripamil nasal spray significantly and rapidly reduced ventricular rate, with a time course consistent with the drug’s pharmacologic profile.
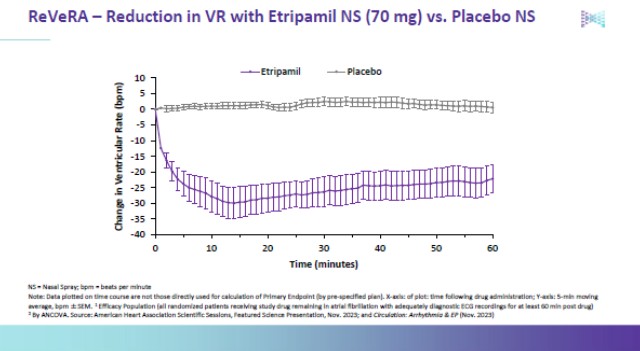
Etripamil achieved the primary endpoint with high statistical significance with patients experiencing a ventricular rate reduction of 29.91 bpm (95% confidence interval: -40.31, -19.52; p<0.0001) in the etripamil arm compared to placebo. The maximum absolute reduction in rate in the etripamil arm was 34.97 bpm. The median time to maximum reduction in ventricular rate, or “VR,” was 13 minutes in patients taking etripamil, and the duration of effect (reduction in VR from
12
baseline and from placebo) was at least 150 min. The median duration of maintaining a VR <100 bpm was 45.5 min in the first 60 min following drug in the etripamil arm.
ReVeRA Primary Endpoint – Maximum Reduction in Ventricular Rate from Baseline
Primary Endpoint: Maximum Reduction in VR from Baseline | Placebo NS, | Etripamil NS, 70 mg, |
Mean (95% CI), bpm | -5.06 (-7.44, -2.67) | -34.97 (-45.13, -24.87) |
Difference in means (95% CI), bpm | ─ | -29.91 (-40.31, -19.52) |
p-value2 | ─ | <0.0001 |
1 Efficacy population (all randomized patients receiving study drug remaining in atrial fibrillation with adequately diagnostic ECG recordings for at least 60 min post drug)
2 From ANCOVA; calculations adjust for variance in baseline
Bpm = beats per minute, ECG = electrocardiogram, CI = confidence interval, NS = nasal spray, VR = ventricular rate
--------------------------------
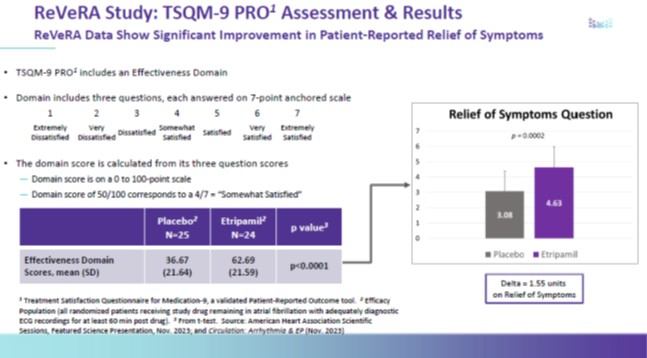
Etripamil treatment was associated with significant improvement in symptom relief and in treatment satisfaction as measured by the TSQM-9 patient reported outcome, or “PRO,” instrument. Patients treated with etripamil, compared to those treated with placebo, demonstrated significant improvements in satisfaction ratings in the TSQM-9 Effectiveness Domain (p<0.0001) and on the Relief of Symptoms Question from the Effectiveness Domain (p = 0.0002, with a treatment effect of 1.55 units). Moreover, patients in the placebo arm were given additional AFib-RVR treatments during the window of 60 minutes to 24 hours after study drug to an approximately two-fold greater degree than patients in the etripamil arm.
13
In ReVeRA, safety and tolerability data were generally consistent with that observed in our PSVT program. Treatment-emergent serious adverse events, or “TESAEs,” were rare, with two occurring in one patient in the etripamil arm (3.7%) and four occurring in two patients in the placebo arm (6.9%). The TESAEs in the etripamil arm (transient severe bradycardia and syncope, assessed as due to hyper-vagotonia) occurred in a patient with a history of vagal events, and fully resolved by placing the patient supine and was without sequelae. The majority of common AEs were localized to the drug-administration site, and there was a low incidence of serious adverse events. The most common (≥ 5%) adverse events were mild or moderate in intensity and included nasal discomfort, rhinorrhea, increased lacrimation, throat irritation and dizziness.
These positive Phase 2 data have supported interactions with the FDA (as summarized in “AFib-RVR Clinical Development Highlights”) that have established a path to potential approval for an indication for etripamil in patients with AFib-RVR. The Phase 3 study is expected to use an optional repeat-dose, self-administration approach of etripamil nasal spray, outside of the medically supervised setting, that is similar to that used in the PSVT development program.
AFib-RVR Clinical Development Highlights
During 2024, we met with the FDA on the ReVeRA study, during which the FDA confirmed its guidance from our Pre-IND meeting (2023) regarding the availability of a sNDA pathway for the marketing approval for etripamil for the indication of AFib-RVR. The sNDA pathway potentially permits a single pivotal efficacy study to be sufficient for filing for marketing approval if etripamil is already approved for PSVT. FDA further concurred with respect to key proposed study elements including powering, inclusion criteria, patient population, and statistical analyses, and offered clarification with respect to the endpoints to guide the design of the Phase 3 study. In our mid-2023 Pre-IND meeting, the FDA provided guidance that our primary endpoint can be the reduction of ventricular rate, and the primary analysis would be performed on the intent to treat, or “ITT,” population. In addition, the study would have to show statistical significance (p<0.05) on the key secondary endpoint of symptom relief as a patient benefit, also in the ITT population. The secondary endpoint could use a PRO measure, and various PROs were discussed with the FDA. We have finalized the Phase 3 study protocol following FDA’s review and obtained concurrence with the FDA to proceed.
The Phase 3 study will be conducted in a medically unmonitored setting (e.g., at-home) in a manner very similar to the conduct of our Phase 3 development program for PSVT. The Phase 3 AFib-RVR study will enroll patients with a history of symptomatic AFib episodes, and will use a self-administered, repeat-dose regimen of 70 mg per dose (the dose and dosing approach that was studied in the RAPID trial in patients with PSVT). The Phase 3 study’s target population will be patients with verified history of AFib-RVR, and the ITT population will be all patients self-administering the study drug for perceived AFib-RVR. The primary endpoint is the mean change from baseline ventricular rate to nadir ventricular rate for patients treated with etripamil versus placebo, as was studied in the ReVeRA trial in AFib-RVR. The key secondary endpoint will be based on a PRO of symptomatic improvement, discussed with the FDA, which is similar to the PRO questions utilized in our PSVT and AFib-RVR programs. The study has been powered and sized based upon approximately 150 events from 150 unique patients with a history of symptomatic episodes of AFib-RVR.
AFib-RVR Market Overview
Atrial Fibrillation, or “AFib,” is a common cardiac arrhythmia with an irregular and often rapid heart rate that is often markedly symptomatic and, without proper treatment, can increase the risk of stroke, heart failure, and other cardiovascular complications. A common complication of AFib is a rapid heart rate, also referred to as AFib-RVR, which is frequently defined as a heart rate ≥ 110 beats per minute. The occurrence of a rapid ventricular rate, or “RVR,” in patients with atrial fibrillation increases the likelihood of marked symptoms including heart palpitations, shortness of breath and weakness. There are two commonly used pharmacological approaches to chronically manage AFib, rhythm control and rate control. Regardless of the chronic approach, break-through episodes of rapid heart rate occur frequently; and when faced with a sudden episode of AFib-RVR, acute rate control is needed, with most treatments being AV-nodal targeted drugs such as a beta blocker or calcium channel blocker. These treatments can be given intravenously; however, this requires a burdensome trip to an emergency department which may lead to a hospital admission. Acute treatment can be attempted by administration of an oral rate control drug; however, such drugs do not adequately provide immediate or dependable enough ventricular rate control due to a 30- to 90-minute delayed onset of action, and, as a result, many patients need faster
14
and more certain rate-reduction and symptom-resolution and so seek acute-medical care by going to the emergency department for treatment utilizing intravenous rate control and/or electrical cardioversion of their atrial fibrillation. Furthermore, the chronic administration of oral rate-control drugs does not broadly prevent episodes of AFib-RVR. Similar to PSVT, patients may feel a loss of control by needing to visit the emergency department for overcoming their AFib-RVR episode and the unpredictable nature of these episodes, which can occur anytime and anywhere. Doctors have expressed frustration at the lack of options for patients to self-manage these acute rate attacks; and payor organizations would prefer to treat the AFib-RVR attacks in a more cost effective and time-efficient manner.
An estimated 10 million Americans suffer from AFib. The prevalence of AFib is expected to grow to greater than 12 million by 2030. A subset of patients with AFib experiences episodes of abnormally high heart rate, most often accompanied by palpitations, shortness of breath, dizziness, and weakness. While these episodes, known as AFib-RVR, may be treated by oral calcium channel blockers and/or beta blockers, patients frequently seek acute care in the ED to address symptoms. In 2019, nearly 1.1 million patients were admitted to the ED due to AFib symptoms. Initial data suggests that approximately 60% of all AFib ED visits were attributable to AFib-RVR, as symptoms driving patients to seek care generally become more pronounced at higher heart rates. Treatment for such symptoms typically includes medically supervised intravenous administration of calcium channel blockers or beta blockers, or electrical cardioversion. With little available data for AFib-RVR, we believe, based on our initial market research, that 30% to 40% of patients with AFib experience one or more symptomatic episodes of RVR per year that require treatment, suggesting a current target addressable market of approximately three to four million patients for etripamil in patients with AFib-RVR.
We believe that etripamil has the potential to be developed such that it can be used by patients to rapidly reduce their heart rate to provide a supplemental option to either the acute oral rate or rhythm control strategy their physician would use. When presented with a target product profile reflecting this potential use case, cardiologists and electrophysiologists, in a 2021 market research study perceived utility in the product profile and indicated that they would prescribe to approximately two-thirds of their patients that experience episodes of AFib-RVR. They further indicated that a rapidly-acting intranasal calcium channel blocker could serve as a “bridge” to the longer onset times of acute oral agents. According to physicians, it can take hours for patients to feel an alleviation of symptoms using acute oral rate or rhythm control. During this time, patients may experience concerning symptoms that often prompt them to seek emergency care. We believe that the combination of convenient delivery, potency, rapid onset and short duration of action of etripamil has the potential to move the current treatment setting for some acute episodes of AFib out of the burdensome and costly emergency department.
Current AFib management consumes significant healthcare resources in the United States. The American Heart Association published a report in 2016 summarizing the current and projected cost burden of cardiovascular diseases in the United States. This report suggests atrial fibrillation resulted in $25 billion in direct medical costs in 2016 (approximately 7% of all cardiovascular diseases) and another $7 billion in indirect costs (i.e., up to $32 billion in total costs). Additionally, the forecasted growth in atrial fibrillation prevalence is anticipated to result in healthcare expenditures of $46 billion in direct costs and $10 billion in indirect costs in the United States by 2030.
Our Strategy
Our goal is to identify, develop, and commercialize innovative cardiovascular medicines, including etripamil for the treatment of PSVT, AFib-RVR, and other cardiovascular indications, and potentially including additional clinical stage compounds for other cardiovascular conditions. The key elements of our business strategy to achieve this goal include the following:
| • | Maximize the value of our launch of CARDAMYST in the United States. We have launched CARDAMYST by building out our sales team of approximately 60 sales representatives who will be targeting cardiology and high value primary care healthcare providers who we believe will treat up to an estimated 500,000 patients with a diagnosis of PSVT during 2026. We have implemented retail distribution and target most of our launch year marketing mix on increasing healthcare professional, or “HCP,” awareness and the number of trials of CARDAMYST. |
15
| • | Expand the scope of cardiovascular indications for etripamil beyond PSVT. We are investigating, through a completed Phase 2 study and an upcoming Phase 3 study, the use of etripamil for the treatment of patients with AFib-RVR. We believe that etripamil could benefit patients with AFib-RVR based on the approved use of IV calcium channel blockers in this indication. We also intend to explore additional cardiovascular opportunities for the use of etripamil. |
| • | Successfully complete development and obtain regulatory approval of etripamil in other major markets. We are currently seeking regulatory approval in China through our partner, Corxel. We intend to seek regulatory approvals in other major markets through additional strategic partnerships. |
| • | Leverage our expertise and experience to expand our pipeline of product candidates. We seek to maximize our commercial opportunities by acquiring or in-licensing product candidates for indications with significant unmet need with a focus on novel treatments for cardiovascular or other conditions. Our leadership team has extensive experience in developing and commercializing successful drugs and has developed a deep understanding of the PSVT and Atrial Fibrillation markets. We intend to leverage the collective talent within our organization and our network to guide our development plans and pipeline expansion. |
Etripamil in Other Therapeutic Applications
Our goal in expanding our pipeline around etripamil is to apply the same paradigm-changing aspiration that we have for supraventricular tachycardias, like PSVT and AFib-RVR, to other cardiac and potentially non-cardiac conditions where we believe that a rapid-onset dihydropyridine L-type calcium channel blocker could potentially deliver significant clinical and quality of life benefits for patients. We believe that the insights that led to the development of etripamil for the treatment of PSVT are relevant in other indications where AV-nodal blocking agents with blood vessel widening activity have demonstrated clinical utility. Both calcium channel blockers and beta blockers are commonly used to manage supraventricular tachycardias like PSVT or AFib-RVR, and other conditions.
Sales and Marketing
We are commercializing CARDAMYST in the United States by targeting high volume healthcare providers, primarily cardiologists, with a focused specialty sales force supported by digital/omnichannel investments and robust reimbursement and patient support. Specifically, the launch of CARDAMYST is being led by a focused national field force calling on approximately 8,000 clinical/interventional cardiologists, 1,500 electrophysiologists, and 500 primary care physicians who treat large populations of patients with PSVT, supported by strategic investments in medical affairs, digital/targeted non-personal promotion, and robust reimbursement and patient support programs. Although we will initially focus on obtaining broad commercial insurance coverage from large Pharmacy Benefit Managers, or “PBMs,” and targeted national and regional commercial payors, as we expand reimbursement for CARDAMYST to include government payors and targeted integrated delivery networks we anticipate that this investment in omnichannel promotion will grow, targeting both prescribers and patients. As we expand, we believe our field force could potentially grow to call on up to 25,000 clinical cardiologists, interventional cardiologists, electrophysiologists, and high-volume primary care physicians/advanced practice providers who have a history of prescribing cardiovascular therapies. We believe an organization of this size could potentially allow us to reach prescribers that collectively care for most patients diagnosed with PSVT in the United States. Given the importance of increasing awareness and educating patients with PSVT, as we establish favorable broad payor reimbursement, we also anticipate deploying focused direct-to-patient marketing campaigns for CARDAMYST.
We anticipate that our sales force could also support the commercialization of additional product candidates treating cardiovascular diseases through partnerships and/or co-promotion deals. In addition, we may pursue and believe that we can maximize the value of etripamil by entering into collaboration agreements for certain territories outside the United States, including the European Union, while retaining commercialization rights in the United States.
16
Manufacturing of Commercial Supply
We do not have any manufacturing facilities. We rely on, and expect to continue to rely on, third party contract manufacturing organizations, or “CMOs,” for all of our required raw materials, nasal spray device, active pharmaceutical ingredient, or “API,” and finished product for our commercial product, clinical trials and for our preclinical research. We require all of our CMOs to conduct manufacturing activities in compliance with current good manufacturing practice, or “cGMP,” requirements. We have assembled a team of experienced employees and consultants to provide the necessary technical, quality, and regulatory oversight over our CMOs and have implemented a comprehensive plan for audits of our CMOs. Currently, we have contracts and quality agreements with our CMOs for the manufacturing of etripamil drug substance and drug product. We believe we currently have enough manufactured supply of etripamil to support our commercial launch. We also may elect to pursue additional CMOs for manufacturing supplies of regulatory starting materials in the future and for the filling of the nasal spray device, labeling, packaging, storage, and distribution of investigational and commercial drug products. We plan to continue to rely on third party manufacturers for any future trials and commercialization of etripamil. We anticipate that these CMOs will have capacity to support commercial scale production.
Competition
Drug development is highly competitive and subject to rapid and significant technological advancements. Our ability to compete will significantly depend upon our ability to complete necessary clinical trials, regulatory approval processes, and effectively market any drug that we may successfully develop. Our current and potential future competitors include pharmaceutical and biotechnology companies, academic institutions, and government agencies. The primary competitive factors that will affect the commercial success of etripamil or any other product candidate for which we may receive marketing approval, include differentiation of any competitor’s product regarding efficacy, safety, tolerability, dosing convenience, price, coverage, and reimbursement. A number of our potential competitors have substantially greater financial, technical, and human resources than we do and significantly greater experience in the discovery and development of product candidates, as well as in obtaining regulatory approvals and commercializing those product candidates in the United States and in foreign countries. It is also possible that a competitor may develop a cure or more effective treatment method for the diseases we are targeting, which could render our current or future product candidates non-competitive or obsolete, or reduce the demand for our product candidates before we can recover our development and commercialization expenses.
We are not aware of any approved drug or any drug candidate in clinical development for a patient with PSVT to self-administer treatment to terminate PSVT episodes. In the acute setting, IV treatments of generic drugs such as adenosine, verapamil, and diltiazem, are routinely given. Additionally, some practitioners prescribe oral medications, such as calcium channel blockers, beta blockers and antiarrhythmics to be taken at the onset of an episode. However, these interventions are not acutely effective and are not approved by the FDA or other regulatory agencies for this use.
For AFib, there are a number of marketed generic antiarrhythmic drugs that are used for chronic and/or acute rate control, such as metoprolol, propranolol, esmolol, pindolol, atenolol, nadolol, verapamil, and diltiazem. For acute rate control, the intravenous forms of these drugs are the ones most often used. We are aware of several drugs or new formulations of existing drugs under development or recently under development for atrial fibrillation, including InRhythm (flecainide), a sodium channel blocker in Phase 3 from InCarda Therapeutics, Inc., and an IV and potentially oral small-molecule SK channel inhibitor being developed by Acesion Pharma for acute conversion and chronic maintenance of sinus rhythm, respectively, in patients with AFib. Acesion’s lead asset, AAP30663, is the IV-formulated short acting AFib conversion therapy for hospital use that has successfully completed a Phase 2 trial. Rapiblyk (landiolol) is an intravenous beta blocker approved for the short-term reduction of ventricular rate with supraventricular tachycardia including atrial fibrillation and atrial flutter.
Intellectual Property
We have filed numerous patent applications pertaining to etripamil and possible future product candidates, formulations containing etripamil, methods of making such formulations, and clinical use. We strive to protect and enhance the
17
proprietary technology, invention, and improvements that are commercially important to the development of our business by seeking, maintaining, and defending our intellectual property. We also rely on know-how, continuing technological innovation and potential in-licensing opportunities to develop, strengthen, and maintain our position in the field of cardiac arrhythmias, such as PSVT, and immediate rate control in atrial fibrillation, as well as other medical conditions affecting the cardiovascular system. Additionally, we intend to rely on regulatory protection afforded through data exclusivity and market exclusivity, as well as patent term extensions, where available.
As of December 31, 2025, our patent portfolio as it pertains to etripamil included:
| • | a patent family containing six U.S. patents, projected to expire in 2028, a pending U.S. patent application, which, if granted, is projected to expire in 2028, as well as corresponding patents in Australia, Brazil, Canada, China, Europe, Hong Kong, India, Japan, Mexico, New Zealand, and South Korea, directed to etripamil, pharmaceutical compositions including etripamil, and uses of etripamil such as to treat cardiac arrhythmias, including PSVT and atrial fibrillation; and |
| • | a patent family containing one U.S. patent, projected to expire in 2036, a pending U.S. patent application, which, if granted, is projected to expire in 2036, as well as corresponding patents in Australia, Brazil, Canada, China, Europe, Hong Kong, Israel, Japan, Macau, Mexico, Russia, South Africa, South Korea and Ukraine and corresponding patent applications in China, Europe, Hong Kong, and New Zealand, directed to formulations including etripamil, methods of making such formulations, and uses of such formulations to treat cardiac arrhythmias, such as PSVT and atrial fibrillation. |
| • | a patent family containing one U.S. patent, projected to expire in 2042, a pending U.S. application which, if granted, is projected to expire in 2042, and pending applications in Canada and Europe, which, if granted, are projected to expire in 2041 in Canada and 2042 in Europe, directed to uses of formulations including etripamil to treat cardiac arrhythmias, such as PSVT and atrial fibrillation, or migraines. |
The terms of individual patents may vary based on the countries in which they are obtained. Generally, patents issued for applications filed in the United States are effective for 20 years from the earliest effective non-provisional filing date in the absence, for example, of a terminal disclaimer shortening the term of the patent or patent term adjustment increasing the term of the patent. In addition, in certain instances, a patent term can be extended to recapture a portion of the term effectively lost as a result of FDA regulatory review periods. The restoration period cannot be longer than five years and the total term, including the restoration period, must not exceed 14 years following FDA approval. The duration of patents outside the United States varies in accordance with provisions of applicable local law, but typically is also 20 years from the earliest non-provisional filing date.
In addition to patents and patent applications that we own, we rely on know-how to develop and maintain our competitive position. We seek to protect our proprietary technology and processes, and obtain and maintain ownership of certain technologies, in part, through confidentiality agreements and invention assignment agreements with our employees, consultants, scientific advisors, contractors, and commercial partners.
Our future commercial success depends, in part, on our ability to obtain and maintain patent and other proprietary protection for commercially important technology, inventions and know-how related to our business; defend and enforce our patents; and operate without infringing valid enforceable patents and proprietary rights of third parties. Our ability to stop third parties from making, using, selling, offering to sell, or importing our products may depend on the extent to which we have rights under valid and enforceable patents that cover these activities. With respect to our owned intellectual property, we cannot be sure that patents will issue from any of the pending patent applications which we own or from any patent applications that we may file in the future, nor can we be sure that any patents that may be issued in the future to us will be commercially useful in protecting etripamil or any future product candidates and methods of using or manufacturing the same. Moreover, we may be unable to obtain patent protection for certain aspects of etripamil or future product candidates generally, as well as with respect to certain indications. See the section entitled “Risk Factors—Risks Related to Our Intellectual Property” for a more comprehensive description of risks related to our intellectual property.
18
Government Regulation and Product Approval
Government authorities in the United States at the federal, state, and local levels, and in other countries, extensively regulate, among other things, the research, development, testing, manufacture, packaging, storage, recordkeeping, labeling, advertising, promotion, distribution, marketing, import, and export of pharmaceutical products, such as those we are developing. The processes for obtaining regulatory approvals in the United States and in foreign countries, along with subsequent compliance with applicable statutes and regulations, require the expenditure of substantial time and financial resources.
United States Government Regulation
In the United States, the FDA regulates drugs under the Federal Food, Drug, and Cosmetic Act, or “FDCA,” and its implementing regulations. The process of obtaining regulatory approvals and the subsequent compliance with appropriate federal, state, local and foreign statutes and regulations require the expenditure of substantial time and financial resources. Failure to comply with the applicable United States requirements at any time during the drug development process, approval process or after approval, may subject an applicant to a variety of administrative or judicial sanctions, such as the FDA’s refusal to approve an NDA, withdrawal of an approval, imposition of a clinical hold, issuance of warning or untitled letters, product recalls, product seizures, total or partial suspension of production or distribution, injunctions, fines, refusals of government contracts, restitution, disgorgement, or civil or criminal penalties.
The process required by the FDA before a drug may be marketed in the United States generally involves:
• | completion of preclinical laboratory tests, animal studies and formulation studies in compliance with the FDA’s good laboratory practice, or “GLP,” regulations; |
• | submission to the FDA of an IND, which must become effective before human clinical trials may begin; |
• | approval by an independent institutional review board, or “IRB,” at each clinical site before each trial may be initiated; |
• | performance of adequate and well controlled clinical trials, in accordance with good clinical practice, or “GCP,” requirements to establish the safety and efficacy of the proposed drug for each indication; |
• | submission to the FDA of an NDA; |
• | satisfactory completion of an FDA advisory committee review, if applicable; |
• | satisfactory completion of an FDA inspection of the manufacturing facility or facilities at which the product is produced to assess compliance with cGMP requirements, and to assure that the facilities, methods and controls are adequate to preserve the drug’s identity, strength, quality, and purity; |
• | satisfactory completion of an FDA inspection of selected clinical sites to assure compliance with GCPs and the integrity of the clinical data; |
• | payment of user fees; and |
• | FDA review and approval of the NDA. |
Preclinical Studies
Preclinical studies include laboratory evaluation of product chemistry, toxicity, and formulation, as well as animal studies to assess potential safety and efficacy. An IND sponsor must submit the results of the nonclinical tests, together with
19
manufacturing information, analytical data, and any available clinical data or literature, among other things, to the FDA as part of an IND. Some nonclinical testing may continue even after the IND is submitted. An IND automatically becomes effective 30 days after receipt by the FDA, unless before that time the FDA raises concerns or questions related to one or more proposed clinical trials and places the clinical trial on a clinical hold.
In such a case, the IND sponsor and the FDA must resolve any outstanding concerns before the clinical trial can begin. As a result, submission of an IND may not result in the FDA allowing clinical trials to commence.
Clinical Trials
Clinical trials involve the administration of the investigational new drug to human subjects under the supervision of qualified investigators in accordance with GCP requirements, which include the requirement that all research subjects provide their informed consent in writing for their participation in any clinical trial. Clinical trials are conducted under protocols detailing, among other things, the objectives of the trial, the parameters to be used in monitoring safety and the effectiveness criteria to be evaluated. A protocol for each clinical trial and any subsequent protocol amendments must be submitted to the FDA as part of the IND. In addition, an IRB at each institution participating in the clinical trial must review and approve the plan for any clinical trial before it commences at that institution, and the IRB must continue to oversee the clinical trial while it is being conducted. Information about certain clinical trials must be submitted within specific timeframes to the National Institutes of Health, or “NIH,” for public dissemination on their ClinicalTrials.gov website.
Human clinical trials are typically conducted in three sequential phases, which may overlap or be combined. In Phase 1, the drug is initially introduced into healthy human subjects or patients with the target disease or condition and tested for safety, dosage tolerance, absorption, metabolism, distribution, excretion and, if possible, to gain an initial indication of its effectiveness. In Phase 2, the drug typically is administered to a limited patient population to identify possible adverse effects and safety risks, to preliminarily evaluate the efficacy of the product for specific targeted diseases and to determine dosage tolerance and optimal dosage. In Phase 3, the drug is administered to an expanded patient population, generally at geographically dispersed clinical trial sites, in well controlled clinical trials to generate enough data to statistically evaluate the safety and efficacy of the product for approval, to establish the overall risk benefit profile of the product and to provide adequate information for the labeling of the product.
Progress reports detailing the results of the clinical trials must be submitted, at least annually, to the FDA, and more frequently if serious adverse events occur. Phase 1, Phase 2, and Phase 3 clinical trials may not be completed successfully within any specified period, or at all. Furthermore, the FDA or the sponsor may suspend or terminate a clinical trial at any time on various grounds, including a finding that the research subjects are being exposed to an unacceptable health risk. Similarly, an IRB can suspend or terminate approval of a clinical trial at its institution if the clinical trial is not being conducted in accordance with the IRB’s requirements, or if the drug has been associated with unexpected serious harm to patients.
Marketing Approval
Assuming successful completion of the required clinical testing, the results of the preclinical and clinical studies, together with detailed information relating to the product’s chemistry, manufacture, controls, and proposed labeling, among other things, are submitted to the FDA as part of an NDA requesting approval to market the product for one or more indications. In most cases, the submission of an NDA is subject to a substantial application user fee. Under the PDUFA guidelines that are currently in effect, the FDA has a goal of ten months from the date of “filing” of a standard NDA for a new molecular entity to review and act on the submission. This review typically takes twelve months from the date the NDA is submitted to the FDA because the FDA has approximately two months to make a “filing” decision.
In addition, under the Pediatric Research Equity Act, certain NDAs or supplements to an NDA must contain data that are adequate to assess the safety and effectiveness of the drug for the claimed indications in all relevant pediatric subpopulations, and to support dosing and administration for each pediatric subpopulation for which the product is safe and effective. The FDA may, on its own initiative or at the request of the applicant, grant deferrals for submission of some
20
or all pediatric data until after approval of the product for use in adults, or full or partial waivers from the pediatric data requirements. Unless otherwise required by regulation, the pediatric data requirements do not apply to products with orphan designation.
The FDA also may require submission of a risk evaluation and mitigation strategy, or “REMS,” plan to ensure that the benefits of the drug outweigh its risks. The REMS plan could include medication guides, physician communication plans, assessment plans, and/or elements to assure safe use, such as restricted distribution methods, patient registries, or other risk minimization tools.
The FDA conducts a preliminary review of all NDAs within the first 60 days after submission, before accepting them for filing, to determine whether they are sufficiently complete to permit substantive review. The FDA may request additional information rather than accept an NDA for filing. In this event, the application must be resubmitted with the additional information. The resubmitted application is also subject to review before the FDA accepts it for filing. Once the submission is accepted for filing, the FDA begins an in-depth substantive review. The FDA reviews an NDA to determine, among other things, whether the drug is safe and effective and whether the facility in which it is manufactured, processed, packaged or held meets standards designed to assure the product’s continued safety, quality and purity.
The FDA may refer an application for a novel drug to an advisory committee. An advisory committee is a panel of independent experts, including clinicians and other scientific experts, that reviews, evaluates, and provides a recommendation as to whether the application should be approved and under what conditions. The FDA is not bound by the recommendations of an advisory committee, but it considers such recommendations carefully when making decisions.
Before approving an NDA, the FDA typically will inspect the facility or facilities where the product is manufactured. The FDA will not approve an application unless it determines that the manufacturing processes and facilities are in compliance with cGMP requirements and adequate to assure consistent production of the product within required specifications. Additionally, before approving an NDA, the FDA will typically inspect one or more clinical trial sites to assure compliance with GCP requirements.
After evaluating the NDA and all related information, including the advisory committee recommendation, if any, and inspection reports regarding the manufacturing facilities and clinical trial sites, the FDA may issue an approval letter, or, in some cases, a complete response letter. A complete response letter generally contains a statement of specific conditions that must be met in order to secure final approval of the NDA and may require additional clinical or preclinical testing in order for FDA to reconsider the application. Even with submission of this additional information, the FDA ultimately may decide that the application does not satisfy the regulatory criteria for approval. If and when those conditions have been met to the FDA’s satisfaction, the FDA will typically issue an approval letter. An approval letter authorizes commercial marketing of the drug with specific prescribing information for specific indications.
Even if the FDA approves a product, it may limit the approved indications for use of the product, require that contraindications, warnings, or precautions be included in the product labeling, require that post-approval studies, including Phase 4 clinical trials, be conducted to further assess a drug’s safety after approval, require testing, and surveillance programs to monitor the product after commercialization, or impose other conditions, including distribution and use restrictions or other risk management mechanisms under a REMS, which can materially affect the potential market and profitability of the product. The FDA may prevent or limit further marketing of a product based on the results of post-marketing studies or surveillance programs. After approval, some types of changes to the approved product, such as adding new indications, manufacturing changes, and additional labeling claims, are subject to further testing requirements and FDA review and approval.
Post-Approval Requirements
Drugs manufactured or distributed pursuant to FDA approvals are subject to pervasive and continuing regulation by the FDA, including, among other things, requirements relating to recordkeeping, periodic reporting, product sampling and distribution, advertising, and promotion and reporting of adverse experiences with the product. After approval, most changes to the approved product, such as adding new indications, manufacturing changes or other labeling claims, are
21
subject to further testing requirements and prior FDA review and approval. There also are continuing annual user fee requirements for any marketed products and the establishments at which such products are manufactured, as well as application fees for supplemental applications with clinical data. The FDA may prevent or limit further marketing of a product based on the results of post-marketing studies or surveillance programs.
In addition, drug manufacturers and other entities involved in the manufacture and distribution of approved drugs are required to register their establishments with the FDA and state agencies, and are subject to periodic unannounced inspections by the FDA and these state agencies for compliance with cGMP requirements. Changes to the manufacturing process are strictly regulated and often require prior FDA approval before being implemented. FDA regulations also require investigation and correction of any deviations from cGMP and impose reporting and documentation requirements upon the sponsor and any third-party manufacturers that the sponsor may decide to use. Accordingly, manufacturers must continue to expend time, money, and effort in the area of production and quality control to maintain cGMP compliance.
Once an approval is granted, the FDA may withdraw the approval if compliance with regulatory requirements and standards is not maintained or if problems occur after the product reaches the market.
Later discovery of previously unknown problems with a product, including adverse events of unanticipated severity or frequency, or with manufacturing processes, or failure to comply with regulatory requirements, may result in mandatory revisions to the approved labeling to add new safety information; imposition of post-market studies or clinical trials to assess new safety risks, or imposition of distribution or other restrictions under a REMS program. Other potential consequences include, among other things:
• | restrictions on the marketing or manufacturing of the product, complete withdrawal of the product from the market, or product recalls; |
• | fines, warning letters or holds on post-approval clinical trials; |
• | refusal of the FDA to approve pending NDAs or supplements to approved NDAs, or suspension or revocation of product approvals; |
• | product seizure or detention, or refusal to permit the import or export of products; or |
• | injunctions or the imposition of civil or criminal penalties. |
The FDA strictly regulates marketing, labeling, advertising and promotion of products that are placed on the market. Drugs may be promoted only for the approved indications and in accordance with the provisions of the approved label, although physicians, based on their independent medical judgement, may prescribe approved drugs for unapproved (or “off-label”) indications. The FDA and other agencies actively enforce the laws and regulations prohibiting their promotion of off-label uses, and a company that is found to have improperly promoted off-label uses may be subject to significant civil, criminal and administrative liability.
In addition, the distribution of prescription pharmaceutical products is subject to the Drug Supply Security Chain Act, which regulates the distribution of drugs and drug samples at the federal level, and sets minimum standards for the registration and regulation of drug distributors by the states.
Federal and State Fraud and Abuse, Data Privacy and Security, and Transparency Laws and Regulations
In addition to FDA restrictions on marketing of pharmaceutical products, federal and state healthcare laws and regulations restrict business practices in the biopharmaceutical industry. These laws regulate, among other things, our current and future business operations, including our clinical research activities, sales, marketing, and education programs. These laws may also constrain the business or financial arrangements and relationships with healthcare providers and other parties through which we market, sell, and distribute our products for which we obtain marketing approval. These laws include
22
anti-kickback and false claims laws and regulations, data privacy and security laws and regulations, and transparency laws and regulations, including, without limitation, those laws described below.
The federal Anti-Kickback Statute prohibits any person or entity from, among other things, knowingly and willfully offering, paying, soliciting, or receiving remuneration to induce or in return for purchasing, leasing, ordering, or arranging for or recommending the purchase, lease, or order of any item or service reimbursable under Medicare, Medicaid, or other federal healthcare programs. The term “remuneration” has been broadly interpreted to include anything of value. The federal Anti-Kickback Statute has been interpreted to apply to arrangements between pharmaceutical manufacturers, on the one hand, and prescribers, purchasers, and formulary managers on the other. There are a number of statutory exceptions and regulatory safe harbors protecting some common activities from prosecution. Practices that involve remuneration that may be alleged to be intended to induce prescribing, purchases or recommendations may be subject to scrutiny if they do not qualify for an exception or safe harbor. Several courts have interpreted the statute’s intent requirement to mean that if any one purpose of an arrangement involving remuneration is to induce referrals of federal healthcare covered business, the statute has been violated.
A person or entity does not need to have actual knowledge of this statute or specific intent to violate it in order to have committed a violation. In addition, the government may assert that a claim including items or services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the federal civil False Claims Act or the civil monetary penalties laws.
Federal civil and criminal false claims laws, including the federal civil False Claims Act, which can be enforced by individuals through civil whistleblower and qui tam actions and civil monetary penalties laws, prohibit any person or entity from, among other things, knowingly presenting, or causing to be presented, a false claim for payment to the federal government or knowingly making, using or causing to be made or used a false record or statement material to a false or fraudulent claim to the federal government. A claim includes “any request or demand” for money or property presented to the U.S. government. Several pharmaceutical and other healthcare companies have been prosecuted under these laws for allegedly providing free product to customers with the expectation that the customers would bill federal programs for the product. Other companies have been prosecuted for causing false claims to be submitted because of the companies’ marketing of products for unapproved, and thus non-reimbursable, uses.
The federal Health Insurance Portability and Accountability Act of 1996, or “HIPAA,” created additional federal criminal statutes that prohibit, among other things, knowingly and willfully executing a scheme to defraud any healthcare benefit program, including private third-party payors and knowingly and willfully falsifying, concealing or covering up a material fact or making any materially false, fictitious, or fraudulent statement in connection with the delivery of or payment for healthcare benefits, items or services. Also, many states have similar fraud and abuse statutes or regulations that apply to items and services reimbursed under Medicaid and other state programs, or, in several states, apply regardless of the payor.
In addition, we are subject to data privacy and security regulation by both the federal government and the states in which we conduct our business. HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act, or “HITECH,” and their respective implementing regulations, impose specified requirements on certain types of individuals and entities relating to the privacy, security, and transmission of individually identifiable health information. Among other things, HITECH makes HIPAA’s security standards directly applicable to “business associates,” defined as independent contractors or agents of covered entities, which include certain healthcare providers, healthcare clearinghouse and health plans, that create, receive, maintain, or transmit individually identifiable health information in connection with providing a service for or on behalf of a covered entity, and their covered subcontractors. HITECH also increased the civil and criminal penalties that may be imposed against covered entities, business associates, and possibly other persons, and gave state attorneys general new authority to file civil actions for damages or injunctions in federal courts to enforce HIPAA and seek attorney’s fees and costs associated with pursuing federal civil actions. In addition, state laws govern the privacy and security of health information in certain circumstances, many of which are not preempted by HIPAA, differ from each other in significant ways and may not have the same effect, thus complicating compliance efforts.
The federal Physician Payments Sunshine Act requires certain manufacturers of drugs, devices, biologics and medical supplies for which payment is available under Medicare, Medicaid or the Children’s Health Insurance Program, with
23
specific exceptions, to report annually to the Centers for Medicare & Medicaid Services, or “CMS,” information related to payments or other transfers of value made to physicians (defined to include doctors, dentists, optometrists, podiatrists and chiropractors), other healthcare professionals (such as physician assistants and nurse practitioners), and teaching hospitals, and applicable manufacturers and applicable group purchasing organizations to report annually to CMS ownership and investment interests held by physicians and their immediate family members.
We may also be subject to state laws that require pharmaceutical companies to comply with the pharmaceutical industry’s voluntary compliance guidelines and the relevant compliance guidance promulgated by the federal government, state laws that require drug manufacturers to report information related to payments and other transfers of value to physicians and other healthcare providers or marketing expenditures, state laws that require drug manufacturers to report information on the pricing of certain drugs, and state and local laws that require certain regulatory licenses to manufacture or distribute our products commercially and/or the registration of pharmaceutical sales representatives.
Because of the breadth of these laws, it is possible that some of our business activities could be subject to challenge under one or more of such laws. There are a number of statutory exceptions and regulatory safe harbors protecting some common activities from prosecution. If our operations are found to be in violation of any of the federal and state laws described above or any other governmental regulations that apply to us, we may be subject to significant criminal, civil and administrative penalties including damages, fines, imprisonment, additional reporting requirements and oversight if we become subject to a corporate integrity agreement or similar agreement to resolve allegations of non-compliance with these laws, contractual damages, reputational harm, diminished profits and future earnings, disgorgement, exclusion from participation in government healthcare programs and the curtailment or restructuring of our operations, any of which could adversely affect our ability to operate our business and our results of operations. To the extent that any of our products are sold in a foreign country, we may be subject to similar foreign laws and regulations, which may include, for instance, applicable post-marketing requirements, including safety surveillance, anti-fraud and abuse laws, implementation of corporate compliance programs, reporting of payments or transfers of value to healthcare professionals, and additional data privacy and security requirements.
Coverage and Reimbursement
The future commercial success of our, or any of our collaborators’, product candidates, if approved, will depend in part on the extent to which third-party payors, such as governmental payor programs at the federal and state levels, including Medicare and Medicaid, private health insurers and other third-party payors, provide coverage of and establish adequate reimbursement levels for our product candidates. Third-party payors generally decide which products they will pay for and establish reimbursement levels for those products. In particular, in the United States, no uniform policy for coverage and reimbursement exists. Private health insurers and other third-party payors often provide coverage and reimbursement for products based on the level at which the government, through the Medicare program, provides coverage and reimbursement for such products, but also have their own methods and approval process apart from Medicare determinations. Therefore, coverage and reimbursement can differ significantly from payor to payor.
In the United States, the European Union, or “EU,” and other potentially significant markets for our product candidates, government authorities and third-party payors are increasingly attempting to limit or regulate the price of products, particularly for new and innovative products, which often has resulted in average selling prices lower than they would otherwise be. Further, the increased emphasis on managed healthcare in the United States and on country and regional pricing and reimbursement controls in the EU will put additional pressure on product pricing, reimbursement and usage. These pressures can arise from rules and practices of managed care groups, judicial decisions and laws and regulations related to Medicare, Medicaid and healthcare reform, pharmaceutical coverage and reimbursement policies and pricing in general. Further, the U.S. Department of Health and Human Services, or “HHS,” imposes rebates on many Medicare Part B and Medicare Part D products to penalize price increases that outpace inflation on an annual basis. HHS has also been empowered to negotiate the price of certain single-source drugs that have been on the market for at least seven (7) years covered under Medicare as part of the Medicare Drug Price Negotiation Program. Each year up to twenty (20) products will be selected by HHS for the Medicare Drug Price Negotiation Program. Products subject to the Medicare Drug Price Negotiation Program are expected to experience a significant reduction in reimbursement from the Medicare program on a per unit basis.
24
Third-party payors are increasingly imposing additional requirements and restrictions on coverage and limiting reimbursement levels for products. For example, federal and state governments reimburse products at varying rates generally below average wholesale price. These restrictions and limitations influence the purchase of products. Third-party payors may limit coverage to specific products on an approved list, or formulary, which might not include all of the FDA-approved products for a particular indication. Third-party payors are increasingly challenging the price and examining the medical necessity and cost-effectiveness of products, in addition to their safety and efficacy. We may need to conduct expensive pharmacoeconomic studies in order to demonstrate the medical necessity and cost-effectiveness of our product candidates, in addition to the costs required to obtain the FDA approvals. Our product candidates may not be considered medically necessary or cost-effective. A payor’s decision to provide coverage for a product does not imply that an adequate reimbursement rate will be approved. Adequate third-party payor reimbursement may not be available to enable us to realize an appropriate return on our investment in product development. Legislative proposals to reform healthcare or reduce costs under government insurance programs may result in lower reimbursement for our product candidates, if approved, or exclusion of our product candidates from coverage and reimbursement. The cost containment measures that third-party payors and providers are instituting and any healthcare reform could significantly reduce our revenues from the sale of any approved product candidates. Even if favorable coverage and reimbursement status is attained for one or more products for which we receive regulatory approval, less favorable coverage policies and reimbursement rates may be implemented in the future.
Healthcare Reform
The United States and some foreign jurisdictions are considering enacting or have enacted a number of additional legislative and regulatory proposals to change the healthcare system in ways that could affect our ability to sell our product candidates profitably, if approved. Among policy makers and payors in the United States and elsewhere, there is significant interest in promoting changes in healthcare systems with the stated goals of containing healthcare costs, improving quality and expanding access. In the United States, the pharmaceutical industry has been a particular focus of these efforts, which include major legislative initiatives to reduce the cost of care through changes in the healthcare system, including limits on the pricing, coverage, and reimbursement of pharmaceutical and biopharmaceutical products, especially under government-funded health care programs, and increased governmental control of drug pricing.
There have been several U.S. government initiatives over the past few years to fund and incentivize certain comparative effectiveness research, including creation of the Patient-Centered Outcomes Research Institute under the Patient Protection and Affordable Care Act of 2010, as amended by the Health Care and Education Reconciliation Act of 2010, or collectively the “PPACA.” It is also possible that comparative effectiveness research demonstrating benefits in a competitor’s product could adversely affect the sales of our product candidates.
There have been amendments to, and executive, judicial and Congressional challenges to certain aspects of the Patient Protection and Affordable Care Act, or the “PPACA.” For example, on July 4, 2025, the One Big Beautiful Bill Act, or the “OBBBA,” was signed into law, which narrowed access to PPACA marketplace exchange enrollment and declined to extend the PPACA enhanced advanced premium tax credits that expired at the end of 2025, which, among other provisions in the law, are anticipated to reduce the number of Americans with health insurance. The OBBBA also is expected to reduce Medicaid spending and enrollment by implementing work requirements for some beneficiaries, capping state-directed payments, reducing federal funding, and limiting provider taxes used to fund the program. Congress is considering proposed legislation intended to further reduce healthcare costs with alternatives to replace the expired PPACA subsidies.
In addition, other legislative changes have been proposed and adopted since the PPACA was enacted. These changes include aggregate reductions to Medicare payments to providers of 2% per fiscal year, which began in 2013 and will remain in effect until 2032 unless additional Congressional action is taken.
The current administration is pursuing policies to reduce regulations and expenditures across government agencies including at HHS, the FDA, CMS and related agencies. These actions, presently directed by executive orders or memoranda from the Office of Management and Budget, may propose policy changes that create additional uncertainty for our business. For example, the current administration has announced agreements with several pharmaceutical companies that require the drug manufacturers to offer, through a direct to consumer platform, or “TrumpRx”, U.S. patients
25
and Medicaid programs prescription drug Most-Favored Nation pricing equal to or lower than those paid in other developed nations, with additional mandates for direct-to-patient discounts and repatriation of foreign revenues. Other recent actions, for example, include (1) directing agencies to reduce agency workforce and cut programs; (2) directing HHS and other agencies to lower prescription drug costs through a variety of initiatives; (3) imposing tariffs on imported pharmaceutical products; and (4) as part of the Make America Healthy Again, or “MAHA”, Commission’s Strategy Report released in September 2025, working across government agencies to increase enforcement on direct-to-consumer pharmaceutical advertising. Additionally, the current administration recently called on Congress to enact "The Great Healthcare Plan," to codify and expand Most-Favored Nation pricing, lower government subsidies to private insurance companies, increase healthcare price transparency, expand pharmaceutical drugs available for over-the-counter purchase, and enact restrictions on pharmacy benefit manager, or “PBM”, payment methodologies, among other things. These actions and policies may significantly reduce U.S. drug prices, potentially impacting manufacturers’ global pricing strategies and profitability, while increasing their operational costs and compliance risks. In June 2024, the U.S. Supreme Court’s Loper Bright decision greatly reduced judicial deference to regulatory agencies, which could increase successful legal challenges to federal regulations affecting our operations. Congress may introduce and ultimately pass health care related legislation that could impact the drug approval process and make changes to the Medicare Drug Price Negotiation Program.
Individual states in the United States have also become increasingly active in passing legislation and implementing regulations designed to control pharmaceutical and biological product pricing, including price or patient reimbursement constraints, discounts, restrictions on certain product access and marketing cost disclosure and transparency measures, and, in some cases, designed to encourage importation from other countries and bulk purchasing. Legally mandated price controls on payment amounts by third-party payors or other restrictions could materially and adversely affect our business, financial condition, results of operations and prospects. In addition, regional healthcare authorities and individual hospitals are increasingly using bidding procedures to determine what pharmaceutical products and which suppliers will be included in their prescription drug and other healthcare programs. This could reduce the ultimate demand for our product candidates or put pressure on our product pricing.
Foreign Regulation
In order to market any product outside of the United States, we would need to comply with numerous and varying regulatory requirements of other countries regarding safety and efficacy and governing, among other things, clinical trials, marketing authorization, commercial sales and distribution of our product candidates. For example, in the EU, we must obtain authorization of a clinical trial application, or “CTA,” in each member state in which we intend to conduct a clinical trial. Whether or not we obtain FDA approval for a drug, we would need to obtain the necessary approvals by the comparable regulatory authorities of foreign countries before we can commence clinical trials or marketing of the drug in those countries. The approval process varies from country to country and can involve additional product testing and additional administrative review periods. The time required to obtain approval in other countries might differ from and be longer than that required to obtain FDA approval. Regulatory approval in one country does not ensure regulatory approval in another, but a failure or delay in obtaining regulatory approval in one country may negatively impact the regulatory process in others.
Employees and Human Capital
Patients inspire all we do. Our employees are passionate about creating a solution for patients who suffer from PSVT and other related illnesses as we work together on our mission to develop innovative cardiovascular medicines. We have built a culture of high performance based on our core values:
| • | Patients First: Everything we do is with the patient in mind. We listen to and partner with patients and place their well-being at the core of all our initiatives. |
Our patients inspire us.
| • | Teamwork: Our employees support, challenge and care for each other. |
Employees engage with one another through their teams, but also through our weekly gatherings, outings and friendly competitions and challenges.
26
Collaboration is key.
| • | Entrepreneurial Mindset: We place a high value on grit, courage and resolve. Our organizational energy has the sense of a startup. |
Employees are encouraged to think like an owner.
| • | Every Idea Matters: Sometimes the best ideas evolve from where it is least expected. |
All ideas are welcome.
| • | Humility, Empathy and Integrity: We act individually and as a team with these three attributes in mind in all we do. |
We care to do what is right.
Our human capital objectives include, as applicable, identifying, recruiting, retaining, incentivizing and integrating our existing and additional employees. The principal purposes of our equity incentive plans are to attract, retain and motivate selected employees, consultants and directors through the granting of stock-based compensation awards.
As of December 31, 2025, we had 38 full-time employees, 13 of whom were primarily engaged in research and development activities. Four of these employees have an M.D. or Ph.D. degree. None of our employees is represented by a labor union and we consider our employee relations to be excellent.
Facilities
Our headquarters is currently located in Montréal (Québec), Canada and consists of 7,700 square feet of leased office space under a lease that expires in November 2027. We also have a U.S. subsidiary in Charlotte, North Carolina that occupies 13,050 square feet of leased office space under a lease that expires in September 2027.
Legal Proceedings
From time to time, we may become involved in legal proceedings arising in the ordinary course of our business. We are not currently a party to any material legal proceedings, and we are not aware of any pending or threatened legal proceedings against us that we believe could have an adverse effect on our business, operating results or financial condition.
Corporate Information
Milestone Pharmaceuticals Inc. was incorporated under the laws of the province of Québec in July 2003. Our principal executive offices are located at 1111 Dr. Frederik-Philips Blvd., Suite 420, Montréal, Québec, Canada H4M 2X6, and our telephone number is (514) 336-0444. Our US offices are located at 6210 Ardrey Kell Rd, Suite 650, Charlotte, NC 28277 and our telephone number is (704) 848-5316.
Available Information
We maintain an internet website at www.milestonepharma.com and make available free of charge through our website our Annual Reports on Form 10-K, Quarterly Reports on Form 10-Q, Current Reports on Form 8-K and amendments to those reports filed or furnished pursuant to Sections 13(a) and 15(d) of the Exchange Act of 1934, or the “Exchange Act”. We make these reports available through our website as soon as reasonably practicable after we electronically file such reports with, or furnish such reports to, the Securities and Exchange Commission, or “SEC”. You can review our electronically filed reports and other information that we file with the SEC on the SEC’s website at http://www.sec.gov. We also make available, free of charge on our website, the reports filed with the SEC by our executive officers, directors and 10% stockholders pursuant to Section 16 under the Exchange Act as soon as reasonably practicable after copies of those filings are provided to us by those persons. In addition, we regularly use our website to post information regarding our business, product development programs and governance, and we encourage investors to use our website, particularly the information in the section entitled “Investors,” as a source of information about us.
27
The information on our website is not incorporated by reference into this Annual Report on Form 10-K and should not be considered to be a part of this Annual Report on Form 10-K. Our website address is included in this Annual Report on Form 10-K as an inactive technical reference only.
Investors and others should note that we announce material information to our investors using one or more of the following: SEC filings, press releases and our corporate website, including without limitation the “Investors” and “Events and Presentations” sections of our website. We use these channels, as well as social media channels such as LinkedIn, in order to achieve broad, non-exclusionary distribution of information to the public and for complying with our disclosure obligations under Regulation FD. It is possible that the information we post on our corporate website or other social media could be deemed to be material information. Therefore, we encourage investors, the media, and others interested in our Company to review the information we post on the “Investors” and “Events and Presentations” sections of our corporate website and on our social media channels. The contents of our corporate website and social media channels are not, however, a part of this Annual Report.
28
ITEM 1A. RISK FACTORS
An investment in our common shares involves a high degree of risk. You should carefully consider the following information about these risks, together with the other information appearing elsewhere in this Annual Report on Form 10-K, consolidated financial statements and related notes thereto and “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” before deciding to invest in our common shares. The occurrence of any of the following risks could have a material adverse effect on our business, financial condition, results of operations and future growth prospects or cause our actual results to differ materially from those contained in forward-looking statements we have made in this report and those we may make from time to time. In these circumstances, the market price of our common shares could decline, and you may lose all or part of your investment. We cannot assure you that any of the events discussed below will not occur.
Risks Related to Our Financial Position and Capital Needs
We have incurred significant operating losses since inception and anticipate that we will continue to incur substantial operating losses for the foreseeable future until revenue from CARDAMYST is sufficient to fund our operations, if ever, and may never achieve or maintain profitability.
Since inception in 2003, we have incurred significant operating losses. Our net loss was $63.1 million and $41.5 million for the years ended December 31, 2025 and 2024, respectively. As of December 31, 2025, we had an accumulated deficit of $430.6 million. We expect to continue to incur significant expenses and increase operating losses for the foreseeable future. Since inception, we have devoted substantially all of our efforts to research and preclinical and clinical development of etripamil, as well as to expanding our management team and infrastructure. We expect that our existing cash and cash equivalents and short-term investments will be sufficient to fund our operations for at least the next 12 months from the date of issuance of this 10-K for the year ending December 31, 2025 and that there are no events or conditions that may cast substantial doubt on our ability to continue as a going concern for at least the next 12 months from the date of this filing.
The net losses we incur may fluctuate significantly from quarter to quarter and year to year. We anticipate that our expenses will increase if, and as, we:
• | continue our ongoing and planned development of etripamil, including future Phase 3 clinical trials for the treatment of AFib-RVR and potential Phase 4 clinical trials for treatment of PSVT; |
• | seek, either directly or through collaboration with our partners, marketing approval for etripamil nasal spray for the treatment of PSVT from regulatory agencies responsible for regions outside the United States; |
• | seek marketing approvals for etripamil for the treatment of AFib-RVR and other cardiovascular indications; |
• | increase our sales, marketing, manufacturing and distribution capabilities, either directly or indirectly with third parties; |
• | build a portfolio of product candidates through development, or the acquisition or in-license of drugs, product candidates or technologies; |
• | initiate preclinical studies and clinical trials for etripamil for any additional indications we may pursue, including the clinical trials for the treatment of atrial fibrillation and rapid ventricular rate as well as other areas of unmet medical need, and for any additional product candidates that we may pursue in the future; |
• | maintain, protect and expand our intellectual property portfolio; |
• | hire additional clinical, regulatory and scientific personnel; |
29
• | add operational, financial and management information systems and personnel, including personnel to support our product development and potential expansion of our future commercialization efforts; and |
• | incur additional legal, insurance related, accounting and other expenses associated with operating as a public company. |
To become and remain profitable, we must succeed in commercializing CARDAMYST for the treatment of PSVT and developing and eventually commercializing product candidates that generate significant revenue. This will require us to be successful in a range of challenging activities, including commercialization of CARDAMYST for the treatment of PSVT, completing any further clinical trials of etripamil and any future product candidates that way may pursue, obtaining regulatory approval, and manufacturing, marketing and selling etripamil and any future products for which we may obtain regulatory approval, as well as discovering or acquiring and then developing additional product candidates. We are only in the preliminary stages of some of these activities. We may never succeed in these activities and, even if we do, may never generate revenues that are significant enough to achieve profitability.-scale manufacturing, marketing and selling etripamil and any future products for which we may obtain regulatory approval, as well as discovering or acquiring and then developing additional product candidates. We are only in the preliminary stages of some of these activities. We may never succeed in these activities and, even if we do, may never generate revenues that are significant enough to achieve profitability.
Our revenue will be dependent, in part, upon the size of the markets in the territories for which we have gained or may gain regulatory approval, the accepted price for the product, the ability to obtain coverage and reimbursement, and whether we own the commercial rights for that territory. If the number of our addressable patients is not as significant as we estimate, the indication approved by regulatory authorities is narrower than we expect, or the treatment population is narrowed by competition, physician choice or treatment guidelines, we may not generate significant revenue from sales of such products.
Because of the numerous risks and uncertainties associated with drug development, we are unable to accurately predict the timing or amount of expenses or when, or if, we will be able to achieve profitability.
Our expenses could increase beyond our expectations if we are required by the FDA, the European Medicines Agency or other regulatory authorities to perform studies in addition to those we currently expect, or if there are any delays in the initiation and completion of our clinical trials or the development of etripamil or any future product candidates.
Even if we do achieve profitability, we may not be able to sustain or increase profitability on a quarterly or annual basis. Our failure to become and remain profitable would decrease the value of our Company and could impair our ability to raise capital, maintain our research and development efforts, expand our business or continue our operations. A decline in the value of our common shares could also cause you to lose all or part of your investment.
Our limited operating history and limited history of commercializing products may make it difficult for you to evaluate the success of our business to date and to assess our future viability.
We are a commercial pharmaceutical company founded in 2003, and our operations to date have been largely focused on raising capital, organizing, staffing our Company and undertaking preclinical studies and conducting clinical trials for etripamil. As an organization, we have not yet demonstrated the ability to conduct sales and marketing activities necessary for successful commercialization. Consequently, any predictions about our future success or viability may not be as accurate as they could be if we had a longer operating history or a history of scaling product or arranging for a third party to do so on our behalf, or conducting sales and marketing activities necessary for successful commercialization. Consequently, any predictions about our future success or viability may not be as accurate as they could be if we had a longer operating history or a history of successful clinical development and commercialization of products.
We may encounter unforeseen expenses, difficulties, complications, delays and other known or unknown factors in achieving our business objectives. We are currently transitioning from a company with a research and development focus to a company primarily focused on commercial activities. We may not be successful in such a transition.
30
Additionally, we expect our financial condition and operating results to continue to fluctuate from quarter to quarter and year to year due to a variety of factors, many of which are beyond our control. Accordingly, you should not rely upon the results of any quarterly or annual periods as indications of future operating performance.
We will require substantial additional funding to finance our operations. If we are unable to raise capital when needed, we could be forced to delay, reduce or terminate our development of etripamil or other operations, including the continued commercialization of CARDAMYST.
Based on our commercialization and research and development plans, we expect that our existing cash and cash equivalents and short-term investments will be sufficient to fund our operations for at least the next 12 months from the date of issuance of this 10-K for the year ended December 31, 2025 and that there are no events or conditions that we are aware of that cast substantial doubt on our ability to continue as a going concern for at least the next 12 months from the date of this filing. However, we will need to obtain substantial additional funding in connection with our continuing operations and planned activities, including the continued commercialization of CARDAMYST. Our future capital requirements will depend on many factors, including:
• | the timing, progress of and results of our ongoing and planned clinical trials of etripamil in AFib-RVR and in other indications; |
• | the success of our commercialization efforts; |
• | the scope, progress, results and costs of preclinical development, laboratory testing and clinical trials of etripamil for additional indications or any future product candidates that we may pursue; |
• | our ability to establish collaborations on favorable terms, if at all; |
• | the ability of vendors who we rely on to accurately forecast expenses and deliver on expectations; |
• | the costs, timing and outcome of regulatory review of etripamil in subsequent indications and any future product candidates; |
• | the costs and timing of commercialization activities, including product manufacturing, marketing, sales and distribution, for etripamil and any future product candidates for which we receive marketing approval; |
• | the costs and timing of preparing, filing and prosecuting patent applications, maintaining and enforcing our intellectual property rights and defending any intellectual property-related claims; |
• | the extent to which we acquire or in-license other product candidates and technologies; and license other product candidates and technologies; and |
• | the costs of operating as a public company. |
31
Identifying potential product candidates and conducting preclinical testing and clinical trials is a time-consuming, expensive, and uncertain process that takes years to complete, and we may never generate the necessary data or results required to obtain regulatory approval of additional product candidates and achieve product sales related thereto. For example, our NODE-301 trial of etripamil for PSVT did not meet its primary endpoint. In addition, CARDAMYST, additional indications of etripamil and any future product candidates may not achieve commercial success. In the near term, our commercial revenues will be derived from sales of CARDAMYST, which may not be sufficient to fund our operations. Accordingly, we will need to continue to rely on additional financing to achieve our business objectives. Adequate additional financing may not be available to us on acceptable terms, or at all. We may seek additional capital due to favorable market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. If banks and financial institutions enter receivership or become insolvent in the future in response to financial conditions affecting the banking system and financial markets, our ability to access our existing cash, cash equivalents and investments may be threatened and could have a material adverse effect on our business and financial condition. Weakness and volatility in capital markets and the economy, in general or as a result of bank failures or macroeconomic conditions such as rising inflation, could limit our access to capital markets and increase our costs of borrowing. If we are unable to raise capital when needed or on attractive terms, we could be forced to delay, reduce, or altogether terminate our research and development programs or future commercialization efforts.
Greater than expected returns of CARDAMYST may exceed our reserve for returns, which would adversely affect our revenue and operating results.
The pharmaceutical wholesalers and distributors to which we sell CARDAMYST are permitted to return purchased product under certain circumstances. We estimate expected returns based on our review of similar products in the industry and record discrete reserves if products held by distributors, forecasted sales and expiration of product warrant a reserve. Any significant increase in returns that exceeds our reserve could adversely affect our revenue and operating results.
Unfavorable global economic conditions could adversely affect our business, financial condition or results of operations.
Our results of operations could be adversely affected by general conditions in the global economy. Unfavorable conditions in the economy both in the United States and abroad, including conditions resulting from changes in gross domestic product growth in the United States or abroad, financial and credit market fluctuations, inflation, fluctuating interest rates, international tariff policies, trade wars and other concerns regarding international trade relations, political turmoil, natural catastrophes, outbreaks of contagious diseases, geopolitical tensions, warfare and terrorist attacks, could cause a decrease in business investments, disrupt the timing and cadence of key industry events, and negatively affect the growth of our business and our results of operations. For example, the COVID-19 pandemic adversely affected workforces, economies and financial markets globally, leading to a reduction in the ability of, or the inability of, partners, suppliers, vendors or other parties to meet their contractual obligations, and for a period of time, a reduction in customer spending on technology, and such conditions may reoccur in the future. Geopolitical conflict, including wars, and the related political and economic responses thereto, such as sanctions, may also exacerbate these issues. A severe or prolonged economic downturn could result in a variety of risks to our business, including weakened demand for CARDAMYST, our product candidates and our ability to raise additional capital when needed on acceptable terms, if at all. A weak or declining economy could also strain our suppliers, possibly resulting in supply disruption, or cause delays in payments for our services by third-party payors or our collaborators. Any of the foregoing could harm our business and we cannot anticipate all of the ways in which the current economic climate and financial market conditions could adversely impact our business, financial condition, results of operations and prospects.
Raising additional capital may cause dilution to our shareholders, restrict our operations or require us to relinquish rights to our product candidates.
Until such time, if ever, as we can generate substantial product revenue, we expect to finance our cash needs through public or private equity or debt financings, third-party funding, marketing and distribution arrangements, as well as other collaborations, strategic alliances, and licensing arrangements, or any combination of these approaches. We do not have any committed external source of funds. To the extent that we raise additional capital through the sale of equity or convertible debt securities, such as the 2029 Convertible Notes (as defined herein), your ownership interest may be diluted,
32
and the terms of these securities may include liquidation or other preferences that adversely affect your rights as a shareholder. Debt and equity financings, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as redeeming our shares, making investments, incurring additional debt, making capital expenditures, declaring dividends or placing limitations on our ability to acquire, sell or license intellectual property rights.
If we raise additional capital through future collaborations, strategic alliances or third-party licensing arrangements, we may have to relinquish valuable rights to our intellectual property, future revenue streams, research programs or product candidates, or grant licenses on terms that may not be favorable to us. If we are unable to raise additional capital when needed, we may be required to delay, limit, reduce or terminate our drug development or future commercialization efforts, or grant rights to develop and market product candidates that we would otherwise develop and market ourselves.
Our ability to use our non-capital loss carry-forwards to offset future taxable income may be subject to certain limitations.
In general, where control of a corporation has been acquired by a person or group of persons, subsection 111(5) of the Income Tax Act (Canada), or the “Canadian Tax Act,” and equivalent provincial income tax legislation restrict the corporation’s ability to carry forward non-capital losses from preceding taxation years. We have not performed a detailed analysis to determine whether an acquisition of control for the purposes of subsection 111(5) of the Canadian Tax Act has occurred after each of our previous issuances of common shares or preferred shares. In addition, if we undergo an acquisition of control, our ability to utilize non-capital losses could be limited by subsection 111(5) of the Canadian Tax Act. As of December 31, 2025, we had Canadian federal and provincial non-capital loss carry forwards of $230.7 million and $226.6 million, respectively, which expire beginning in 2026 through 2045. In addition, we also have scientific research and experimental development expenditures of $32.7 million and $38.8 million, respectively, for Canadian federal and provincial income tax purposes, which have not been deducted. These expenditures are available to reduce future taxable income and have an unlimited carry-forward period. Research and development tax credits and expenditures are subject to verification by the tax authorities, and, accordingly, these amounts may vary. Future changes in our share ownership, some of which are outside of our control, could result in an acquisition of control for the purposes of subsection 111(5) of the Canadian Tax Act. Furthermore, our ability to utilize non-capital losses (or U.S. equivalents) of companies that we may acquire in the future may be subject to limitations. As a result, even if we attain profitability, we may be unable to use a material portion of our non-capital losses and other tax attributes, which could negatively impact our future cash flow.
Our subsidiary’s ability to use its U.S. net operating loss carryforwards and certain other tax attributes for U.S. income tax purposes may be limited.
As of December 31, 2025, we had U.S. federal net operating loss carry-forwards, or “NOLs,” of $103.2 million as a result of expenses incurred by Milestone Pharmaceuticals USA, Inc., our wholly owned subsidiary. Under current U.S. federal tax law, NOLs incurred in taxable years ending beginning after December 31, 2017 may be carried forward indefinitely. However, the deductibility of such NOLs is limited to 80% of taxable income. It is uncertain if and to what extent various states will conform to the federal law. In addition, under Sections 382 and 383 of the Internal Revenue Code of 1986, as amended, and corresponding provisions of state law, if a corporation undergoes an “ownership change,” which is generally defined as a greater than 50% change (by value) in its equity ownership over a three year period, the corporation’s ability to use its pre change NOL carryforwards and other pre change tax attributes (such as research tax credits) to offset its post change income may be limited. It is possible that we have experienced one or more ownership changes in the past. In addition, we may also experience ownership changes in the future as a result of subsequent shifts in our share ownership, some of which may be outside of our control. As a result, if we earn net taxable income, our ability to use our pre ownership change NOL carryforwards to offset U.S. federal taxable income may be subject to limitations, which could potentially result in increased future tax liability to us. In addition, at the state level, there may be periods during which the use of NOLs is suspended or otherwise limited, which could accelerate or permanently increase state taxes owed.
33
Risks Related to the Development of Our Product Candidates
We currently have one approved product, CARDAMYST™ (etripamil) nasal spray, a prescription medication for the conversion of acute symptomatic episodes of PSVT. We are currently pursuing clinical development for subsequent etripamil indications. If we are not able to obtain required regulatory approvals for subsequent etripamil indications or any future product candidates, our ability to generate revenue will be adversely affected.
Etripamil is currently our only product. We have not obtained regulatory approval for additional indications of etripamil or any other product candidate, and it is possible that neither new indications of etripamil nor any product candidates we may seek to develop in the future will ever obtain regulatory approval. Neither we nor any future collaborator is permitted to market any drug product candidates in the United States or other countries until we receive regulatory approval from the FDA or applicable foreign regulatory agency. The time required to obtain approval or other marketing authorizations by the FDA and comparable foreign regulatory authorities is unpredictable but typically takes many years following the commencement of clinical trials and depends upon numerous factors, including the substantial discretion of the regulatory authorities. In addition, approval policies, regulations, or the type and amount of clinical data necessary to gain approval may change during the course of a product candidate’s clinical development and may vary among jurisdictions.
Prior to obtaining approval to commercialize etripamil for additional indications and any other drug product candidate in the United States or elsewhere, we must demonstrate with substantial evidence from well controlled clinical trials, and to the satisfaction of the FDA or comparable foreign regulatory authorities, that such product candidates are safe and effective for their intended uses. Results from nonclinical studies and clinical trials can be interpreted in different ways. Even if we believe the nonclinical or clinical data for our product candidates are promising, such data may not be sufficient to support approval by the FDA and other regulatory authorities. The FDA may also require us to conduct additional nonclinical studies, including human factor studies, or clinical trials for our product candidates either prior to or post-approval, or it may object to elements of our clinical development program. For example, the approval of CARDAMYST is subject to a postmarketing requirement to conduct a safety, pharmacokinetics and pharmacodynamics trial in pediatric patients with PSVT.
Of the large number of products in development, only a small percentage successfully complete the FDA or comparable foreign regulatory authorities’ approval processes and are commercialized. The lengthy approval or marketing authorization process as well as the unpredictability of future clinical trial results may result in our failing to obtain regulatory approval or marketing authorization to market etripamil or any future product candidates, which would significantly harm our business, financial condition, results of operations and prospects.
We have invested a significant portion of our time and financial resources in the development of etripamil. Our business would be enhanced if we are able to successfully develop etripamil and obtain regulatory approval for subsequent indications and other potential product candidates.
Even if we eventually complete clinical testing and receive approval of an NDA, or foreign marketing application for additional indications of etripamil and any future product candidates, the FDA or the comparable foreign regulatory authorities may grant approval or other marketing authorization contingent on the performance of costly additional clinical trials, including post-market clinical trials. The FDA or the comparable foreign regulatory authorities also may approve or authorize for marketing a product candidate for a more limited indication or patient population that we originally request, and the FDA or comparable foreign regulatory authorities may not approve or authorize the labeling that we believe is necessary or desirable for the successful commercialization of a product candidate. Any delay in obtaining, or inability to obtain, applicable regulatory approval or other marketing authorization would delay or prevent commercialization of that product candidate and would materially adversely impact our business and prospects.
Although we have received approval of CARDAMYST in the United States, the FDA may change their policies, adopt additional regulations or revise existing regulations or take other actions, which may prevent or delay approval of our future products under development on a timely basis. Such policy or regulatory changes could impose additional requirements upon us that could delay our ability to obtain approvals, increase the costs of compliance, or restrict our ability to maintain any marketing authorizations we may have obtained.
34
We may not be successful in our efforts to expand our pipeline of product candidates beyond etripamil for PSVT.
We intend to build a pipeline of product candidates beyond etripamil for PSVT and progress these product candidates through clinical development. For example, on November 11, 2023, we announced positive Phase 2 clinical trial data on etripamil for the treatment of AFib-RVR and we intend to conduct Phase 3 development. We may not be able to successfully expand the scope of cardiovascular indications for etripamil beyond PSVT, or leverage our expertise and experience with etripamil in PSVT to other product candidates. We may not be able to in-license, acquire or develop future product candidates that are safe and effective. Even if we are successful in continuing to expand etripamil to other indications and further build our pipeline, the potential product candidates that we identify may not be suitable for clinical development, including as a result of safety, tolerability, efficacy or other characteristics that indicate that they are unlikely to be drugs that will receive marketing approval, achieve market acceptance or obtain reimbursements from third-party payors. If we do not successfully execute on our strategy of expanding our product pipeline, it could significantly harm our financial position and adversely affect the trading price of our common shares.
The development of additional product candidates is risky and uncertain.
Efforts to identify, acquire or in-license, and then develop product candidates require substantial technical, financial and human resources, whether or not any product candidates are ultimately identified. Our efforts may initially show promise in identifying potential product candidates, yet fail to yield product candidates for clinical development, approved products or commercial revenues for many reasons, including the following:
• | the methodology used may not be successful in identifying potential product candidates; |
• | competitors may develop alternatives that render any product candidates we develop obsolete; |
• | any product candidates we develop may nevertheless be covered by third parties’ patents or other exclusive rights; |
• | a product candidate may be shown to have harmful side effects or other characteristics that indicate it is unlikely to be effective or otherwise does not meet applicable regulatory criteria; |
• | a product candidate may not be capable of being produced in commercial quantities at an acceptable cost, or at all; and |
• | a product candidate may not be accepted as safe and effective by physicians, patients, the medical community, or third-party payors. |
We have limited financial and management resources and, as a result, we may forego or delay pursuit of opportunities with other product candidates or for other indications that later prove to have greater market potential. Our resource allocation decisions may cause us to fail to capitalize on viable commercial drugs or profitable market opportunities. If we do not accurately evaluate the commercial potential or target market for a particular product candidate, we may relinquish valuable rights to that product candidate through collaboration, licensing, or other royalty arrangements in circumstances under which it would have been more advantageous for us to retain sole development and commercialization rights to such product candidate. If we are unsuccessful in identifying and developing additional product candidates or are unable to do so, our business may be harmed.
Success in preclinical studies or earlier clinical trials may not be indicative of results in future clinical trials, and we cannot assure you that any ongoing, planned, or future clinical trials will lead to results sufficient for the necessary regulatory approvals.
Success in preclinical testing and earlier clinical trials does not ensure that later clinical trials will generate the same results or otherwise provide adequate data to demonstrate the efficacy and safety of a product candidate. Preclinical tests and Phase 1 and Phase 2 clinical trials are primarily designed to test safety, to study pharmacokinetics and pharmacodynamics
35
and to understand the side effects of product candidates at various doses and schedules. Success in preclinical studies and earlier clinical trials does not ensure that later efficacy trials will be successful, nor does it predict final results. For example, our Phase 2 clinical trial of etripamil for PSVT was conducted in an electrophysiology lab, a controlled setting, in which episodes of SVT were induced and etripamil was administered by healthcare providers. Our Phase 3 clinical trials were conducted in an at-home setting with patients self-administering etripamil and monitoring their cardiac activity as episodes of SVT occur. Additionally, in our Phase 2 clinical trial, four sprays of study drug were dispensed to patients using four separate FDA approved single spray devices. In our Phase 3 clinical trials, patients self-administered two to four sprays of study drug from an FDA approved device that is capable of delivering two separate sprays. While our RAPID Phase 3 trial did meet its primary endpoint, our NODE-301 clinical trial did not meet its primary endpoint. Additional indications of etripamil and any future product candidates may fail to show the desired safety and efficacy in clinical development despite positive results in preclinical studies or having successfully advanced through earlier clinical trials.
In addition, the design of a clinical trial can determine whether its results will support approval of a product, and flaws in the design of a clinical trial may not become apparent until the clinical trial is well advanced. Clinical trial design flaws are more likely in therapy areas, such as PSVT, where there are limited previous trials from which to learn and model clinical trials. As an organization, we have limited experience designing clinical trials for future indications of etripamil or other product candidates and may be unable to design and execute a clinical trial to support regulatory approval. Many companies in the pharmaceutical and biotechnology industries have suffered significant setbacks in late-stage clinical trials even after achieving promising results in preclinical testing and earlier clinical trials. Data obtained from preclinical and clinical activities are subject to varying interpretations, which may delay, limit, or prevent regulatory approval. In addition, we may experience regulatory delays or rejections as a result of many factors, including changes in regulatory policy during the period of our product candidate development. Any such delays could negatively impact our business, financial condition, results of operations, and prospects.
We may encounter substantial delays or difficulties in our clinical trials.
We may not commercialize, market, promote or sell any product candidate without obtaining marketing approval from the FDA or comparable foreign regulatory authorities, and we may never receive such approvals in subsequent etripamil indications or any other potential unapproved product candidates. It is impossible to predict when or if any of our future product candidates will prove effective or safe in humans and will receive regulatory approval. Before obtaining marketing approval from regulatory authorities for the sale of our product candidates, we must complete preclinical development and then conduct extensive clinical trials to demonstrate the safety and efficacy of our product candidates in humans. Clinical testing is expensive, difficult to design and implement, can take many years to complete, and is uncertain as to outcome. A failure of one or more clinical trials can occur at any stage of testing. Moreover, preclinical and clinical data are often susceptible to varying interpretations and analyses, and many companies that have believed their product candidates performed satisfactorily in preclinical studies and clinical trials have nonetheless failed to obtain marketing approval of their products.
We may experience numerous unforeseen events prior to, during, or as a result of, clinical trials that could delay or prevent our ability to receive marketing approval or commercialize etripamil and any future product candidates, including:
• | delays in reaching a consensus with regulatory authorities on design or implementation of our clinical trials; |
• | regulators or institutional review boards, or “IRBs,” may not authorize us or our investigators to commence a clinical trial or conduct a clinical trial at a prospective trial site; |
• | delays in reaching agreement on acceptable terms with prospective clinical research organizations, or “CROs,” and clinical trial sites; |
• | the number of patients required for clinical trials of our product candidates may be larger than we anticipate, enrollment in these clinical trials may be slower than we anticipate, patients may drop out of these clinical |
36
trials at a higher rate than we anticipate or fail to return for post-treatment follow up or we may fail to recruit suitable patients to participate in a trial;
• | clinical trials of our product candidates may produce negative or inconclusive results; |
• | imposition of a clinical hold by regulatory authorities as a result of a serious adverse event, concerns with a class of product candidates or after an inspection of our clinical trial operations, trial sites or manufacturing facilities; |
• | occurrence of serious adverse events associated with the product candidate that are viewed to outweigh its potential benefits; |
• | changes in regulatory requirements and guidance that require amending or submitting new clinical protocols; |
• | interruptions resulting from public health emergencies, or other geopolitical tensions; |
| • | interruptions to government agencies, such as the FDA, including as a result of levels of government funding, the ability to fill key leadership roles; or |
• | we may decide, or regulators may require us, to conduct additional clinical trials or abandon product development programs. |
Any inability to successfully complete preclinical and clinical development could result in additional costs to us or impair our ability to generate revenue from future drug sales or other sources. In addition, if we make manufacturing or formulation changes to our product candidates, we may need to conduct additional testing to bridge our modified product candidate to earlier versions. Clinical trial delays could also shorten any periods during which we may have the exclusive right to commercialize our product candidates, if approved, or allow our competitors to bring competing drugs to market before we do, which could impair our ability to successfully commercialize our product candidates and may harm our business, financial condition, results of operations and prospects.
Additionally, if the results of our clinical trials are inconclusive or if there are safety concerns or serious adverse events associated with our product candidates, we may:
• | be delayed in obtaining marketing approval, if at all; |
• | obtain approval for indications or patient populations that are not as broad as intended or desired; |
• | obtain approval with labeling that includes significant use or distribution restrictions or safety warnings; |
• | be subject to additional post-marketing testing requirements; |
• | be required to perform additional clinical trials to support approval or be subject to additional post-marketing testing requirements; |
• | have regulatory authorities withdraw or suspend their approval of the drug or impose restrictions on its distribution in the form of a modified risk evaluation and mitigation strategy, or “REMS”; |
• | be subject to the addition of labeling statements, such as warnings or contraindications; |
• | be sued; or |
• | experience damage to our reputation. |
37
Our product development costs will also increase if we experience delays in testing or obtaining marketing approvals. We do not know whether any of our preclinical studies or clinical trials will begin as planned, need to be restructured or be completed on schedule, if at all.
Further, we, the FDA or an IRB may suspend our clinical trials at any time if it appears that we or our collaborators are failing to conduct a trial in accordance with regulatory requirements, including the FDA’s current GCP regulations, that we are exposing participants to unacceptable health risks, or if the FDA finds deficiencies in our investigational new drug applications, or “INDs,” or the conduct of these trials. Therefore, we cannot predict with any certainty the schedule for commencement and completion of future clinical trials. If we experience delays in the commencement or completion of our clinical trials, or if we terminate a clinical trial prior to completion, the commercial prospects of our product candidates could be negatively impacted, and our ability to generate revenues from our product candidates may be delayed.
Clinical trials are very expensive, time-consuming, and difficult to design and implement.
Our product candidates will require clinical testing before we are prepared to submit an NDA, or comparable application to foreign regulatory authorities, for regulatory approval. We cannot predict with any certainty if or when we might submit an application for regulatory approval for any of our product candidates or whether any such application will be approved by the FDA or foreign regulatory authority. Human clinical trials are very expensive and difficult to design and implement, in part because they are subject to rigorous regulatory requirements. For instance, the FDA or foreign regulatory authority may not agree with our proposed endpoints for any future clinical trial of our product candidates, which may delay the commencement of our clinical trials. In addition, we may not succeed in developing and validating disease relevant clinical endpoints based on insights regarding biological pathways for the diseases we are studying. The clinical trial process is also time-consuming. We estimate that the successful completion of clinical trials for etripamil and any future product candidates will take several years to complete. Furthermore, failure can occur at any stage, and we could encounter problems that cause us to abandon or repeat clinical trials.
Enrollment and retention of patients in clinical trials is an expensive and time-consuming process and could be delayed, made more difficult or rendered impossible by multiple factors outside our control.
Identifying and qualifying patients to participate in our clinical trials is critical to our success. If the actual number of patients with PSVT, AFib-RVR, or any other indications that we may pursue for etripamil or future product candidates, is smaller than we anticipate, we may encounter difficulties in enrolling patients in our clinical trials, thereby delaying or preventing development and approval of etripamil and any future product candidates. Even once enrolled, we may be unable to retain a sufficient number of patients to complete any of our trials. Patient enrollment and retention in clinical trials depends on many factors, including the size of the patient population, the nature of the trial protocol, the existing body of safety and efficacy data, the number and nature of competing treatments and ongoing clinical trials of competing therapies for the same indication, the proximity of patients to clinical sites, the experience and capabilities of the clinical sites to recruit the correct patients, and the eligibility criteria for the trial. Patient enrollment may also be affected by public health crises, such as patients experiencing difficulty accessing clinical trial sites and complying with clinical trial protocols.
Furthermore, our efforts to build relationships with patient communities may not succeed, which could result in delays in patient enrollment in our clinical trials. In addition, any negative results we may report in clinical trials of etripamil, and any future product candidate may make it difficult or impossible to recruit and retain patients in other clinical trials of that same product candidate. For example, we reported a failed primary endpoint from our NODE-301 trial in March 2020. Delays or failures in planned patient enrollment or retention may result in increased costs, program delays, or both, which could have a harmful effect on our ability to develop etripamil or any future product candidates or could render further development impossible. In addition, we expect to rely on CROs and clinical trial sites to ensure proper and timely conduct of our future clinical trials and, while we intend to enter into agreements governing their services, we will be limited in our ability to compel their actual performance. Similarly, our formulation of etripamil is designed to be self-administered as a nasal spray. While we expect enrolled patients to adhere to the protocol, our ability to ensure patient compliance is limited.
38
Our product candidates may cause undesirable side effects or have other properties that could delay or prevent their regulatory approval, limit the commercial potential, or result in significant negative consequences following any potential marketing approval.
During the conduct of clinical trials, patients report changes in their health, including illnesses, injuries and discomforts, to their doctor. Often, it is not possible to determine whether or not the product candidate being studied caused these conditions. Regulatory authorities may draw different conclusions or require additional testing to confirm these determinations, if they occur. For example, in our Phase 2 clinical trial for PSVT, three serious adverse events, or “SAEs,” were considered possibly related to etripamil, including an episode of second-degree AV block that subsequently and spontaneously resolved. Calcium channel blockers have known side effects, such as slowing AV conduction, slowing the heart rate below normal levels and hypotension, or low blood pressure. While we designed etripamil to have a short pharmacodynamic effect to lower these risks, if etripamil is not quickly metabolized as designed, these known side effects may become more pronounced in patients who use etripamil.
In addition, it is possible that as we test etripamil or any future product candidates in larger, longer, and more extensive clinical trials, or as use of etripamil or any future product candidates becomes more widespread if they receive regulatory approval, illnesses, injuries, discomforts, and other adverse events that were observed in earlier trials, as well as conditions that did not occur or went undetected in previous trials, will be reported by subjects or patients. Many times, side effects are only detectable after investigational drugs are tested in large-scale pivotal trials or, in some cases, after they are made available to patients on a commercial scale after approval. If additional clinical experience indicates that etripamil or any future product candidates have side effects or causes serious or life-threatening side effects, the development of the product candidate may fail or be delayed, or, if the product candidate has received regulatory approval, such approval may be revoked, which would harm our business, prospects, operating results and financial condition.
Interim, “top-line” and preliminary data from our clinical trials that we announce or publish from time to time may change as more patient data become available and are subject to audit and verification procedures that could result in material changes in the final data.
From time to time, we may publish interim, “topline” or preliminary data from our clinical trials. Interim data from clinical trials that we may complete are subject to the risk that one or more of the clinical outcomes may materially change as patient enrollment continues and more patient data becomes available. Preliminary or “topline” data also remain subject to audit and verification procedures that may result in the final data being materially different from the preliminary data we previously published. As a result, interim and preliminary data should be viewed with caution until the final data are available. Differences between preliminary or interim data and final data could significantly harm our business prospects and may cause the trading price of our common shares to fluctuate significantly.
We may explore additional strategic collaborations that may never materialize, or we may be required to relinquish important rights to and control over the development of our product candidates to any future collaborators.
We intend to continue to periodically explore a variety of possible strategic collaborations in an effort to gain access to additional product candidates or resources. We are likely to face significant competition in seeking appropriate strategic collaborators, and strategic collaborations can be complicated and time consuming to negotiate and document. We may not be able to negotiate strategic collaborations on acceptable terms, or at all. We are unable to predict when, if ever, we will enter into any strategic collaborations because of the numerous risks and uncertainties associated with establishing them.
Future collaborations could subject us to a number of risks, including:
• | we may be required to undertake the expenditure of substantial operational, financial, and management resources; |
• | we may be required to issue equity securities that would dilute our shareholders’ percentage ownership of our Company; |
39
• | we may be required to assume substantial actual or contingent liabilities; |
• | we may not be able to control the amount and timing of resources that our strategic collaborators devote to the development or commercialization of our product candidates; |
• | strategic collaborators may select indications or design clinical trials in a way that may be less successful than if we were doing so; |
• | strategic collaborators may delay clinical trials, provide insufficient funding, terminate a clinical trial, or abandon a product candidate, repeat, or conduct new clinical trials or require a new version of a product candidate for clinical testing; |
• | strategic collaborators may not pursue further development of products resulting from the strategic collaboration arrangement or may elect to discontinue research and development programs; |
• | strategic collaborators may not commit adequate resources to the marketing and distribution of our product candidates, limiting our potential revenues from these products; |
• | disputes may arise between us and our strategic collaborators that result in the delay or termination of the research or development of our product candidates or that result in costly litigation or arbitration that diverts management’s attention and consumes resources; |
• | strategic collaborators may experience financial difficulties; |
• | strategic collaborators may not properly maintain or defend our intellectual property rights or may use our proprietary information in a manner that could jeopardize or invalidate our proprietary information or expose us to potential litigation; |
• | business combinations or significant changes in a strategic collaborator’s business strategy may adversely affect a strategic collaborator’s willingness or ability to complete its obligations under any arrangement; |
• | strategic collaborators could decide to move forward with a competing product candidate developed either independently or in collaboration with others, including our competitors; and |
• | strategic collaborators could terminate the arrangement or allow it to expire, which would delay the development and may increase the cost of developing our product candidates. |
Risks Related to the Commercialization of Our Product and Product Candidates
If we are unable to successfully implement and maintain sales, marketing and distribution capabilities for CARDAMYST for the treatment of PSVT or any product candidate that may receive regulatory approval, we may not be successful in commercializing CARDAMYST for the treatment of PSVT or our product candidates if and when they are approved .
We are in the early stages of commercializing CARDAMYST for the treatment of PSVT. To achieve commercial success for CARDAMYST for the treatment of PSVT, any additional indications for etripamil and any other product candidate for which we may obtain marketing approval, we will need to continue to implement and maintain an effective sales and marketing organization. We have built a focused sales and marketing organization to launch CARDAMYST for the treatment of PSVT in the United States. However, we may choose to expand upon our sales force in the future for PSVT, additional etripamil indications, or other product candidates. There are inherent risks to implementing and maintaining a standalone commercial organization, which is also time-consuming and requires significant financial resources.
Factors that create risk and may inhibit our efforts to commercialize our products on our own include:
40
| • | our inability to recruit, train and retain adequate numbers of effective sales and marketing personnel; |
| • | the inability of sales personnel to obtain access to physicians or educate adequate numbers of physicians on the benefits of prescribing any future products; |
| • | inability to obtain favorable insurance coverage of any approved product; |
| • | the lack of complementary products to be offered by sales personnel, which may put us at a competitive disadvantage relative to companies with more extensive product lines; and |
| • | unforeseen costs and expenses associated with creating an independent sales and marketing organization. |
If we are unable to successfully implement and maintain our own sales, marketing and distribution capabilities and are forced to enter into arrangements with, and rely on, third parties to perform these services, our revenue and our profitability, if any, are likely to be lower than if we had developed such capabilities ourselves. In addition, we may not be successful in entering into arrangements with third parties to sell, market and distribute our product candidates or may be unable to do so on terms that are favorable to us. We likely will have little control over such third parties, and any of them may fail to devote the necessary resources and attention to sell and market our products effectively. If we do not maintain sales, marketing and distribution capabilities successfully, either on our own or in collaboration with third parties, we will not be successful in commercializing our product candidates
CARDAMYST, along with any subsequent etripamil indications or any other product candidates, if approved, may fail to achieve market acceptance by physicians, patients, third-party payors or others in the medical community necessary for commercial success
CARDAMYST, any subsequent etripamil indications and any other product candidates that receive marketing approval may fail to gain market acceptance by physicians, patients, third-party payors and others in the medical community. If such CARDAMYST, any subsequent etripamil indications or any future product candidates do not achieve an adequate level of acceptance, we may not generate significant drug revenue and may not become profitable. The degree of market acceptance of CARDAMYST, any subsequent etripamil indications or any future product candidates, if approved for commercial sale, will depend on a number of factors, including but not limited to third-party payors and others in the medical community. If such CARDAMYST, any subsequent etripamil indications or any future product candidates do not achieve an adequate level of acceptance, we may not generate significant drug revenue and may not become profitable. The degree of market acceptance of CARDAMYST, any subsequent etripamil indications or any future product candidates, if approved for commercial sale, will depend on a number of factors, including but not limited to:
• | the convenience and ease of administration compared to alternative treatments and therapies; |
• | the willingness of the target patient population to try new therapies and of physicians to prescribe these therapies; |
• | the efficacy and potential advantages compared to alternative treatments and therapies; |
• | the effectiveness of sales and marketing efforts; |
• | the prevalence and severity of any side effects; |
• | the strength of our relationships with patient communities; |
• | the cost of treatment in relation to alternative treatments and therapies, including any similar generic treatments; |
• | our ability to offer such drug for sale at competitive prices; |
41
• | the strength of marketing and distribution support; and |
• | the availability of third-party coverage and adequate reimbursement; any restrictions on the use of the drug together with other medications; and the awareness and support from key opinion leaders in cardiology. |
Our efforts to educate physicians, patients, third-party payors and others in the medical community on the benefits of CARDAMYST, any subsequent etripamil indications or any future product candidates may require significant resources and may never be successful. Because we expect sales of CARDAMYST, any subsequent etripamil indications or any future product candidates, if approved, to generate substantially all of our revenues for the foreseeable future, the failure of these product candidates to find market acceptance would harm our business.
The success of etripamil for the treatment of PSVT will be dependent on its proper use.
While we have designed etripamil to be self-administered, we cannot control the successful use of the product. While we have conducted, and intend in the future to conduct, human factors studies to determine how to optimize the instructions for use, the results in our clinical trials may not be replicated by users in the future. If we are not successful in promoting the proper use of etripamil, we may not be able to achieve market acceptance or effectively commercialize the drug. In addition, even in the event of proper use of etripamil, individual devices may fail.
If the market opportunities for CARDAMYST, any subsequent indications for etripamil and any future product candidates are smaller than we estimate, our business may suffer.
Our eligible patient population may differ significantly from the actual market addressable by our products and product candidates. Our projections of both the number of people who have these conditions, as well as the subset of people with these diseases who have the potential to benefit from treatment with our product candidates, are based on our beliefs and estimates. These estimates have been derived from a variety of sources, including scientific literature, insurance claims databases, or market research, and may prove to be incorrect. Further, new studies may change the estimated incidence or prevalence of these diseases. The number of patients may turn out to be lower than expected. Likewise, the potentially addressable patient population for each of our product candidates may be limited or may not be amenable to treatment with our product candidates, and new patients may become increasingly difficult to identify or access. If the market opportunities for our product candidates are smaller than we estimate, our business and results of operations could be adversely affected.
We may face substantial competition, which may result in others developing or commercializing drugs before or more successfully than us.
The development and commercialization of new drugs is highly competitive. We may face competition from major pharmaceutical companies, specialty pharmaceutical companies, and biotechnology companies worldwide. Potential competitors also include academic institutions, government agencies, and other public and private research organizations that conduct research, seek patent protection, and establish collaborative arrangements for research, development, manufacturing, and commercialization.
More established companies may have a competitive advantage over us due to their greater size, resources, and institutional experience. In particular, these companies have greater experience and expertise in securing reimbursement, negotiating government contracts, establishing relationships with key opinion leaders, conducting testing and clinical trials, obtaining and maintaining regulatory approvals, and distribution relationships. These companies also have significantly greater research and marketing capabilities than we do. If we are not able to compete effectively against existing and potential competitors, our business and financial condition may be harmed.
As a result of these factors, our competitors may obtain regulatory approval of their drugs before we are able to, which may limit our ability to develop or commercialize future product candidates. Our competitors may also develop therapies that are safer, more effective, more widely accepted or less expensive than ours, and may also be more successful than we
42
in manufacturing and marketing their drugs. These advantages could render our product candidates obsolete or noncompetitive before we can recover the costs of such product candidates’ development and commercialization.
Mergers and acquisitions in the pharmaceutical and biotechnology industries may result in even more resources being concentrated among a smaller number of our competitors. Smaller and early-stage companies may also prove to be significant competitors, particularly through collaborative arrangements with large and established companies. These third parties compete with us in recruiting and retaining qualified scientific, management, and commercial personnel, establishing clinical trial sites, and subject registration for clinical trials, as well as in acquiring technologies complementary to, or necessary for, our programs.
If we commercialize etripamil or any future product candidates outside of the United States, a variety of risks associated with international operations could harm our business.
We intend to seek approval to market etripamil outside of the United States and may do so for future product candidates. If we market approved products outside of the United States, we expect that we will be subject to additional risks in commercialization including:
• | different regulatory requirements for approval of therapies in foreign countries; |
• | reduced protection for intellectual property rights; |
• | unexpected changes in tariffs, trade barriers and regulatory requirements; |
• | economic weakness, including inflation, or political instability in particular foreign economies and markets; |
• | compliance with tax, employment, immigration, and labor laws for employees living or traveling abroad; |
• | foreign currency fluctuations, which could result in increased operating expenses and reduced revenues, and other obligations incident to doing business in another country; |
• | foreign reimbursement, pricing, and insurance regimes; |
• | workforce uncertainty in countries where labor unrest is more common than in the United States; |
• | production shortages resulting from any events affecting raw material supply or manufacturing capabilities abroad; and |
• | business interruptions resulting from pandemics and public health crisis (such as the COVID-19 pandemic), geopolitical actions, including war , and terrorism or natural disasters including earthquakes, typhoons, floods and fires. |
We have no prior experience in these areas. In addition, there are complex regulatory, tax, labor, and other legal requirements imposed by many of the individual countries in and outside of Europe with which we will need to comply. Many biopharmaceutical companies have found the process of marketing their own products in foreign countries to be very challenging.
The pharmaceutical industry in China is highly regulated, and such regulations are subject to change, which may negatively affect the commercialization of the Company’s medicines and drug candidates in that country.
On May 15, 2021, the Company entered into a license and collaboration agreement, or the License Agreement, with Corxel, which is an entity affiliated with RTW Investments, LP, or “RTW,” an existing shareholder of the Company. Under the License Agreement, the Company granted Corxel exclusive development and commercialization rights to any pharmaceutical product that uses a device to deliver etripamil by nasal spray for all prophylactic and therapeutic uses in
43
humans in the following territories: People’s Republic of China, including mainland China, Hong Kong, Macau, and Taiwan. The pharmaceutical industry in China is subject to comprehensive government regulation and supervision, encompassing the approval, registration, manufacturing, packaging, licensing, and marketing of new medicines. In recent years, the regulatory framework in China for pharmaceutical companies has undergone significant changes, and the Company expects that such changes will continue in the foreseeable future. Any such change may cause delays in or prevent the successful research, development, manufacturing, or commercialization of etripamil in the greater China region and may reduce the current benefits the Company believes are available to it from licensing such products to be developed, manufactured, and sold in the greater China region. In addition, any failure by the Company or its partners to maintain compliance with applicable laws and regulations or obtain and maintain required licenses and permits may result in the suspension or termination of its business activities in China or create other legal risks and the continuing compliance of such changing regulation may require the Company to incur substantial legal and regulatory expenses.
Coverage and adequate reimbursement may not be available for CARDAMSYT or any future product candidates, which could make it difficult for us to gain market acceptance.
Market acceptance and sales of any product candidates that we commercialize, will depend in part on the extent to which reimbursement for these drugs and related treatments will be available from third-party payors, including government health administration authorities, managed care organizations and other private health insurers. Third-party payors decide which therapies are reimbursed and establish reimbursement levels. While no uniform policy for coverage and reimbursement exists in the United States, third-party payors often rely upon Medicare coverage policy and payment limitations in setting their own coverage and reimbursement policies. However, decisions regarding the extent of coverage and amount of reimbursement to be provided for any product candidates that we develop will be made on a payor-by-payor basis. Therefore, one payor’s determination to provide coverage for a drug does not assure that other payors will also provide coverage, and adequate reimbursement, for the drug. Additionally, a third-party payor’s decision to provide coverage for a therapy does not imply that an adequate reimbursement rate will be approved. Each payor determines whether or not it will provide coverage for a therapy, what amount it will pay the manufacturer for the therapy, and on what tier of its formulary it will be placed. The position on a payor’s list of covered drugs, or formulary, generally determines the copayment that a patient will need to make to obtain the therapy and can strongly influence the adoption of such therapy by patients and physicians. Patients who are prescribed treatments for their conditions and providers prescribing such services generally rely on third-party payors to reimburse all or part of the associated healthcare costs. Patients are unlikely to use our products unless coverage is provided and reimbursement is adequate to cover a significant portion of the cost of our products. Further, the increased emphasis on cost containment in the United States and abroad will put additional pressure on product pricing, reimbursement and usage. For example, HHS imposes rebates on many Medicare Part B and Medicare Part D products to penalize price increases that outpace inflation on an annual basis. HHS has also been empowered to negotiate the price of certain single-source drugs that have been on the market for at least seven (7) years covered under Medicare as part of the Medicare Drug Price Negotiation Program. Each year up to twenty (20) products will be selected by HHS for the Medicare Drug Price Negotiation Program. Products subject to the Medicare Drug Price Negotiation Program are expected to experience a significant reduction in reimbursement from the Medicare program on a per unit basis.
Third-party payors have attempted to control costs by limiting coverage and the amount of reimbursement for particular medications. We cannot be sure that coverage and reimbursement will be available for any drug that we commercialize and, if reimbursement is available, what the level of reimbursement will be. Inadequate coverage and reimbursement may impact the demand for, or the price of, any drug for which we obtain marketing approval. If coverage and adequate reimbursement are not available, or are available only to limited levels, we may not be able to successfully commercialize etripamil or any future product candidates that we develop.
In addition, we expect that the increased emphasis on managed care and cost containment measures in the United States by third-party payors and government authorities will continue and will place pressure on pharmaceutical pricing and coverage. Coverage policies and third-party reimbursement rates may change at any time. Therefore, even if favorable coverage and reimbursement status is attained for one or more drug products for which we receive regulatory approval, less favorable coverage policies and reimbursement rates may be implemented in the future. If we are unable to obtain and maintain sufficient third-party coverage and adequate reimbursement for our drug products, the commercial success
44
of our drug products may be greatly hindered, and our financial condition and results of operations may be materially and adversely affected.
Product liability lawsuits against us could cause us to incur substantial liabilities and could limit commercialization of any product candidates that we may develop.
We face an inherent risk of product liability exposure related to commercializing etripamil or testing and commercializing any future product candidates in clinical trials. If we cannot successfully defend ourselves against claims that any such product candidates caused injuries, we could incur substantial liabilities. Regardless of merit or eventual outcome, liability claims may result in:
• | decreased demand for any product candidate that we may develop; |
• | loss of revenue; |
• | substantial monetary awards to trial participants or patients; |
• | significant time and costs to defend the related litigation; |
• | withdrawal of clinical trial participants; |
• | increased insurance costs; |
• | the inability to commercialize any product candidate that we may develop; and |
• | injury to our reputation and significant negative media attention. |
Although we maintain clinical trial liability insurance coverage with maximum coverage of $10 million per incident and an aggregate loss limit of $10 million such insurance may not be adequate to cover all liabilities that we may incur with a medical product during the clinical trials. We anticipate that we will need to increase our insurance coverage each time we commence a clinical trial and maintain a product liability insurance if we successfully commercialize any product candidate. Insurance coverage is increasingly expensive. We may not be able to maintain insurance coverage at a reasonable cost or in an amount adequate to satisfy any liability that may arise.
Risks Related to Regulatory Compliance
Even if we maintain approval for CARDAMYST, or we obtain and maintain approval for subsequent etripamil indications or any future product candidates from the FDA, we may never obtain approval of etripamil or any future product candidates outside of the United States, which would limit our market opportunities and could harm our business.
Approval of a product candidate in the United States by the FDA does not ensure approval of such product candidate by regulatory authorities in other countries or jurisdictions, and approval by one foreign regulatory authority does not ensure approval by regulatory authorities in other foreign countries or by the FDA. Sales of etripamil or any future product candidates outside of the United States will be subject to foreign regulatory requirements governing clinical trials and marketing approval. Even if the FDA grants marketing approval for a product candidate, comparable foreign regulatory authorities also must approve the manufacturing and marketing of the product candidate in those countries. Approval procedures vary among jurisdictions and can involve requirements and administrative review periods different from, and more onerous than, those in the United States, including additional preclinical studies or clinical trials. In many countries outside the United States, a product candidate must be approved for reimbursement before it can be approved for sale in that country. In some cases, the price that we intend to charge for any product candidates, if approved, is also subject to approval. Obtaining approval for etripamil or any future product candidates in the European Union from the European Commission following the opinion of the European Medicines Agency, if we choose to submit a marketing authorization
45
application there, would be a lengthy and expensive process. Even if a product candidate is approved, the FDA or the European Commission, as the case may be, may limit the indications for which the drug may be marketed, require extensive warnings on the drug labeling or require expensive and time-consuming additional clinical trials or reporting as conditions of approval. Obtaining foreign regulatory approvals and compliance with foreign regulatory requirements could result in significant delays, difficulties and costs for us and could delay or prevent the introduction of etripamil or any future product candidates in certain countries.
Further, clinical trials conducted in one country may not be accepted by regulatory authorities in other countries. Also, regulatory approval for our product candidates may be withdrawn. If we fail to comply with the regulatory requirements, our target market will be reduced and our ability to realize the full market potential of etripamil or any future product candidates will be harmed and our business, financial condition, results of operations and prospects could be harmed.
CARDAMYST, and any subsequent etripamil indications or any future product candidates, if approved, will remain subject to ongoing regulatory oversight.
CARDAMYST, and any subsequent etripamil indications or any future product candidates, if approved, will be subject to ongoing regulatory requirements for manufacturing, labeling, packaging, storage, advertising, promotion, sampling, record-keeping, and submission of safety and other post-market information. Any regulatory approvals that we receive for etripamil or any future product candidates may also be subject to a REMS, limitations on the approved indicated uses for which the drug may be marketed or to the conditions of approval, or contain requirements for potentially costly post-marketing testing, including Phase 4 trials, and surveillance to monitor the quality, safety, and efficacy of the drug. Such regulatory requirements may differ from country to country depending on where we have received regulatory approval.
In addition, drug manufacturers and their facilities are subject to payment of user fees and continual review and periodic inspections by the FDA and other regulatory authorities for compliance with cGMP requirements and adherence to commitments made in the NDA or foreign marketing application. If we, or a regulatory authority, discover previously unknown problems with a drug, such as adverse events of unanticipated severity or frequency, or problems with the facility where the drug is manufactured or if a regulatory authority disagrees with the promotion, marketing, or labeling of that drug, a regulatory authority may impose restrictions relative to that drug, the manufacturing facility or us, including requesting a recall or requiring withdrawal of the drug from the market or suspension of manufacturing.
If we fail to comply with applicable regulatory requirements for CARDAMYST or any subsequent etripamil indications, a regulatory authority may:
• | issue an untitled letter or warning letter asserting that we are in violation of the law; |
• | seek an injunction or impose administrative, civil or criminal penalties or monetary fines; |
• | suspend or withdraw regulatory approval; |
• | suspend any ongoing clinical trials; |
• | refuse to approve a pending NDA or comparable foreign marketing application or any supplements thereto submitted by us or our partners; |
• | restrict the marketing or manufacturing of the drug; |
• | seize or detain the drug or otherwise require the withdrawal of the drug from the market; |
• | refuse to permit the import or export of product candidates; or |
• | refuse to allow us to enter into supply contracts, including government contracts. |
46
Moreover, the FDA strictly regulates the promotional claims that may be made about drug products. In particular, a product may not be promoted for uses that are not approved by the FDA as reflected in the product’s approved labeling. Physicians, on the other hand, may prescribe products for off-label uses. Although the FDA and other regulatory agencies do not regulate a physician’s choice of drug treatment made in the physician’s independent medical judgment, they do regulate promotional communications from companies or their sales force with respect to off-label uses of products for which marketing clearance has not been issued. However, biopharmaceutical companies may share truthful and not misleading information that is otherwise consistent with the labeling. The FDA and other agencies actively enforce the laws and regulations prohibiting the promotion of off-label uses, and a company that is found to have improperly promoted off-label uses may be subject to significant civil, criminal, and administrative penalties.
Any government investigation of alleged violations of law could require us to expend significant time and resources in response and could generate negative publicity. The occurrence of any event or penalty described above may inhibit our ability to commercialize etripamil or any future product candidates and harm our business, financial condition, results of operations and prospects.
The FDA’s and other regulatory authorities’ policies may change, and additional government regulations may be enacted that could prevent, limit, or delay regulatory approval of our product candidates. If we are slow or unable to adapt to changes in existing requirements or the adoption of new requirements or policies, or if we are not able to maintain regulatory compliance, we may lose any marketing approval that we may have obtained and we may not achieve or sustain profitability, which would adversely affect our business, prospects, financial condition and results of operations.
In addition, we cannot predict the likelihood, nature or extent of government regulation that may arise from future legislation or administrative or executive action, either in the United States or abroad.
Our relationships with customers, physicians, and third-party payors are subject, directly or indirectly, to federal and state healthcare fraud and abuse laws, false claims laws, health information privacy and security laws, and other healthcare laws and regulations. If we are unable to comply, or have not fully complied, with such laws, we could face substantial penalties.
Healthcare providers, including physicians and third-party payors in the United States and elsewhere will play a primary role in the recommendation and prescription of any product candidates for which we obtain marketing approval. Our current and future arrangements with healthcare professionals, principal investigators, consultants, customers, and third-party payors subject us to various federal and state fraud and abuse laws, data privacy and security laws, transparency laws, and other healthcare laws that may constrain the business or financial arrangements and relationships through which we research, sell, market, and distribute our products, if we obtain marketing approval.
The federal Anti-Kickback Statute prohibits the offer, receipt, or payment of remuneration in exchange for or to induce the referral of patients or the use of products or services that would be paid for in whole or part by Medicare, Medicaid, or other federal healthcare programs. Remuneration has been broadly defined to include anything of value, including cash, improper discounts, and free or reduced-price items and services. Additionally, the intent standard under the federal Anti-Kickback Statute was amended by PPACA, to a stricter standard such that a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation. Further, PPACA codified case law that a claim including items or services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the False Claims Act.
The federal false claims, including the False Claims Act, and civil monetary penalties laws, which prohibit individuals or entities from, among other things, knowingly presenting, or causing to be presented, false or fraudulent claims for payment of federal funds, and knowingly making, or causing to be made, a false record or statement material to a false or fraudulent claim to avoid, decrease or conceal an obligation to pay money to the federal government.
The federal Health Information Insurance Portability and Accountability Act of 1996, or “HIPAA,” prohibits, among other things, knowingly and willfully executing, or attempting to execute, a scheme to defraud any healthcare benefit program, or knowingly and willfully falsifying, concealing, or covering up a material fact or making any materially false statement,
47
in connection with the delivery of, or payment for, healthcare benefits, items, or services. Similar to the U.S. federal Anti-Kickback Statute, a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation.
HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act, or “HITECH,” also imposes, among other things, certain standards and obligations on covered entities including certain healthcare providers, health plans, and healthcare clearinghouses, and their respective business associates that create, receive, maintain, or transmit individually identifiable health information for or on behalf of a covered entity as well as their covered subcontractors relating to the privacy, security, transmission, and breach reporting of individually identifiable health information.
The federal Physician Payments Sunshine Act, and its implementing regulations, require certain manufacturers of drugs, devices, biologics, and medical supplies that are reimbursable under Medicare, Medicaid, or the Children’s Health Insurance Program to report annually to Centers for Medicare & Medicaid Services information related to certain payments and other transfers of value to physicians (defined to include doctors, dentists, optometrists, podiatrists, and chiropractors), other healthcare professionals (such as physician assistants and nurse practitioners), and teaching hospitals, as well as ownership and investment interests held by the physicians described above and their immediate family members.
We will also be subject to healthcare regulation and enforcement by the U.S. federal government and the states and any other countries in which we conduct our business, including our research, and the sales, marketing, and distribution of our product candidates and products once they have obtained marketing authorization.
Additionally, some state and local laws require certain regulatory licenses to manufacture or distribute our products commercially and/or the registration of pharmaceutical sales representatives in the jurisdiction. Further, some state laws require pharmaceutical companies to comply with the pharmaceutical industry’s voluntary compliance guidelines and the relevant compliance guidance promulgated by the federal government in addition to requiring manufacturers to report information related to payments to physicians and other health care providers or marketing expenditures. State and foreign laws also govern the privacy and security of health information in some circumstances, many of which differ from each other in significant ways and often are not preempted by HIPAA, thus complicating compliance efforts.
Ensuring that our business arrangements with third parties comply with applicable healthcare laws and regulations will likely be costly. It is possible that governmental authorities will conclude that our business practices may not comply with current or future statutes, regulations, or case law involving applicable fraud and abuse or other healthcare laws and regulations. If our operations are found to be in violation of any of these laws or any other governmental regulations that may apply to us, we may be subject to significant civil, criminal, and administrative penalties, damages, fines, disgorgement, imprisonment, exclusion from participating in government-funded healthcare programs, such as Medicare and Medicaid, additional reporting requirements and oversight if we become subject to a corporate integrity agreement or similar agreement to resolve allegations of non-compliance with these laws, contractual damages, reputational harm, and the curtailment or restructuring of our operations.
If the physicians or other providers or entities with whom we expect to do business are found not to be in compliance with applicable laws, they may be subject to significant criminal, civil, or administrative sanctions, including exclusions from government-funded healthcare programs. Even if resolved in our favor, litigation or other legal proceedings relating to healthcare laws and regulations may cause us to incur significant expenses and could distract our technical and management personnel from their normal responsibilities. In addition, there could be public announcements of the results of hearings, motions, or other interim proceedings or developments. If securities analysts or investors perceive these results to be negative, it could have a substantial adverse effect on the price of our common shares. Such litigation or proceedings could substantially increase our operating losses and reduce the resources available for development, manufacturing, sales, marketing, or distribution activities. Uncertainties resulting from the initiation and continuation of litigation or other proceedings relating to applicable healthcare laws and regulations could have a material adverse effect on our ability to compete in the marketplace.
48
Healthcare legislative reform measures may have a negative impact on our business and results of operations.
In the United States and some foreign jurisdictions, there have been, and continue to be, several legislative and regulatory changes and proposed changes regarding the healthcare system that could prevent or delay marketing approval of product candidates, restrict or regulate post approval activities, and affect our ability to profitably sell any product candidates for which we obtain marketing approval. In particular, there have been and continue to be a number of initiatives at the U.S. federal and state levels that seek to reduce healthcare costs and improve the quality of healthcare. For example, in March 2010 the PPACA, was passed, which substantially changed the way healthcare is financed by both governmental and private payors in the United States. There have been amendments, as well as executive, judicial and Congressional challenges, to certain aspects of the PPACA. For example, on July 4, 2025, the One Big Beautiful Bill Act, or the OBBBA, was signed into law, which narrowed access to PPACA marketplace exchange enrollment and declined to extend the PPACA enhanced advanced premium tax credits that expired at the end of 2025, which, among other provisions in the law, are anticipated to reduce the number of Americans with health insurance. The OBBBA is also expected to reduce Medicaid spending and enrollment by implementing work requirements for some beneficiaries, capping state-directed payments, reducing federal funding, and limiting provider taxes used to fund the program. Congress is considering proposed legislation intended to further reduce healthcare costs with alternatives to replace the expired PPACA subsidies.
In addition, other legislative changes have been proposed and adopted since the PPACA was enacted. These changes include aggregate reductions to Medicare payments to providers of 2% per fiscal year, which began in 2013 and will remain in effect until 2032 unless additional Congressional action is taken.
The current administration is pursuing policies to reduce regulations and expenditures across government agencies including at HHS, the FDA, the Centers for Medicare & Medicaid Services, or “CMS,” and related agencies. These actions, presently directed by executive orders or memoranda from the Office of Management and Budget, may propose policy changes that create additional uncertainty for our business. For example, the current administration has announced agreements with several pharmaceutical companies that require the drug manufacturers to offer, through a direct to consumer platform, or “TrumpRx” U.S. patients and Medicaid programs prescription drug Most-Favored Nation pricing equal to or lower than those paid in other developed nations, with additional mandates for direct-to-patient discounts and repatriation of foreign revenues. Other recent actions, for example, include (1) directing agencies to reduce agency workforce and cut programs; (2) directing HHS and other agencies to lower prescription drug costs through a variety of initiatives; (3) imposing tariffs on imported pharmaceutical products; and (4) as part of the Make America Healthy Again, or “MAHA,” Commission’s Strategy Report released in September 2025, working across government agencies to increase enforcement on direct-to-consumer pharmaceutical advertising. Additionally, the current administration recently called on Congress to enact "The Great Healthcare Plan," to codify and expand Most-Favored Nation pricing, lower government subsidies to private insurance companies, increase healthcare price transparency, expand pharmaceutical drugs available for over-the-counter purchase, and enact restrictions on pharmacy benefit manager, or “PBM,” payment methodologies, among other things. These actions and policies may significantly reduce U.S. drug prices, potentially impacting manufacturers’ global pricing strategies and profitability, while increasing their operational costs and compliance risks. In June 2024, the U.S. Supreme Court’s Loper Bright decision greatly reduced judicial deference to regulatory agencies, which could increase successful legal challenges to federal regulations affecting our operations. Congress may introduce and ultimately pass health care related legislation that could impact the drug approval process and make changes to the Medicare Drug Price Negotiation Program.
Our business involves the use of hazardous materials and we and our third-party manufacturers and suppliers must comply with environmental laws and regulations, which can be expensive and restrict how we do business.
Our research and development activities and our third-party manufacturers’ and suppliers’ activities involve the controlled storage, use and disposal of hazardous materials owned by us, including the components of etripamil and any future product candidates and other hazardous compounds. We and our manufacturers and suppliers are subject to laws and regulations governing the use, manufacture, storage, handling and disposal of these hazardous materials. In some cases, these hazardous materials and various wastes resulting from their use are stored at our and our manufacturers’ facilities pending their use and disposal. We cannot eliminate the risk of contamination, which could cause an interruption of our commercialization efforts, research and development efforts and business operations, environmental damage resulting in costly clean-up and liabilities under applicable laws and regulations governing the use, storage, handling and disposal of these materials and specified waste products. Although we believe that the safety procedures utilized by our third-party
49
manufacturers for handling and disposing of these materials generally comply with the standards prescribed by these laws and regulations, we cannot guarantee that this is the case or eliminate the risk of accidental contamination or injury from these materials. In such an event, we may be held liable for any resulting damages and such liability could exceed our resources and state or federal or other applicable authorities may curtail our use of certain materials and/or interrupt our business operations. Furthermore, environmental laws and regulations are complex, change frequently and have tended to become more stringent. We cannot predict the impact of such changes and cannot be certain of our future compliance. We do not currently carry hazardous waste insurance coverage.
Risks Related to Our Dependence on Third Parties
We will rely on third parties to produce clinical and commercial supplies of etripamil and any future product candidates.
We do not own or operate facilities for drug manufacturing, storage and distribution, or testing. We are dependent on third parties to manufacture the clinical supplies of etripamil and any future product candidates. The facilities used by our contract manufacturers to manufacture etripamil and any future product candidates must be approved by the FDA pursuant to inspections. We do not control the manufacturing process of, and are completely dependent on, our contract manufacturing partners for compliance with regulatory requirements including current Good Manufacturing Practices or, “cGMPs,” for manufacture of active drug substances, nasal spray devices, and finished product candidates. If our contract manufacturers cannot successfully manufacture material that conforms to our specifications and the strict regulatory requirements of the FDA or others, we will not be able to secure and/or maintain regulatory approval for our product candidates. In addition, we have no control over the ability of our contract manufacturers to maintain adequate quality control, quality assurance and qualified personnel. We intend to use multiple contract manufacturers for clinical and commercial supply of our drug product and drug substance. As such, we will need to demonstrate to the FDA that the drug product and drug substance from these contract manufacturers are comparable, which may include conducting additional equivalence studies. If the FDA or a comparable foreign regulatory authority does not approve these facilities for the manufacture of our product candidates or if it withdraws any such approval in the future, we may need to find alternative manufacturing facilities, which would significantly impact our ability to develop, obtain regulatory approval for, or market our approved products and product candidates, if and when approved. Any significant delay in the supply of a product candidate, or the raw material components thereof, for an ongoing clinical trial due to the need to replace a third-party manufacturer could considerably delay completion of our clinical trials, product testing, and potential regulatory approval of our product candidates
Further, we also will rely on third-party manufacturers to supply us with sufficient quantities of etripamil and any future product candidates, if approved, for commercialization. We do not yet have a commercial supply agreement for the nasal spray device. If we are not able to meet market demand for any approved product, it would negatively affect our ability to generate revenue, harm our reputation, and could have a material and adverse effect on our business and financial condition. Increasing the scale of production inherently creates increased risk of manufacturing errors, and we may not be able to adequately inspect every device that is produced, and it is possible that individual devices may fail to perform as designed. Manufacturing errors could negatively impact market acceptance of any of our product candidates that receive approval, result in negative press coverage, or increase our liability.
Further, our reliance on third-party manufacturers entails risks to which we may not be subject if we manufactured product candidates ourselves, including:
• | inability to meet our product specifications and quality requirements consistently; |
• | delay or inability to procure or expand sufficient manufacturing capacity; |
• | issues related to scale-up of manufacturing; |
• | costs and validation of new equipment and facilities required for scale-up; |
50
• | our third-party manufacturers may not be able to execute our manufacturing procedures and other logistical support requirements appropriately; |
• | our third-party manufacturers may fail to comply with cGMP-compliance and other inspections by the FDA or other comparable regulatory authorities; |
• | our inability to negotiate manufacturing agreements with third parties under commercially reasonable terms, if at all; |
• | breach, termination, or nonrenewal of manufacturing agreements with third parties in a manner or at a time that is costly or damaging to us; |
• | reliance on a single source for the nasal spray device; |
• | our third-party manufacturers may not devote sufficient resources to our product candidates; |
• | we may not own, or may have to share, the intellectual property rights to any improvements made by our third-party manufacturers in the manufacturing process for our product candidates; |
• | operations of our third-party manufacturers or suppliers could be disrupted by conditions unrelated to our business or operations, including public health emergencies, natural disasters, such as earthquakes, fires or floods, the bankruptcy of the manufacturer or supplier, carrier disruptions or increased costs that are beyond our control, and global macro uncertainty related to geopolitical conflicts; and |
• | the possibility that, to the extent we need to complete technology transfers to a backup CMO or alternative CMO, we may be subject to additional costs associated with the technology transfer or dependent on the cooperation of our current CMOs. |
Any of these events could lead to clinical trial delays, failure to obtain regulatory approval, or affect our ability to successfully commercialize etripamil or any future product candidates once approved. Some of these events could be the basis for FDA action, including injunction, request for recall, seizure, or total or partial suspension of production.
We rely on third parties to conduct, supervise and monitor our preclinical studies and clinical trials, and if those third parties perform in an unsatisfactory manner, it may harm our business.
We have engaged CROs to conduct our Phase 2 clinical trial of etripamil for the treatment of AFib-RVR, and we expect to engage a CRO for future clinical trials of etripamil and any future product candidates. We do not currently have the ability to independently conduct any clinical trials. We rely on CROs and clinical trial sites to ensure the proper and timely conduct of our preclinical studies and clinical trials, and we expect to have limited influence over their actual performance. We rely upon CROs to monitor and manage data for our clinical programs, as well as the execution of future nonclinical studies. We expect to control only certain aspects of our CROs’ activities. Nevertheless, we will be responsible for ensuring that each of our preclinical studies and clinical trials is conducted in accordance with the applicable protocol, legal, regulatory and scientific standards, and our reliance on the CROs does not relieve us of our regulatory responsibilities.
We and our CROs are required to comply with good laboratory practices, or “GLPs,” and GCPs, which are regulations and guidelines enforced by the FDA and comparable foreign regulatory authorities in the form of International Conference on Harmonization guidelines for any of our product candidates that are in preclinical and clinical development. Regulatory authorities enforce GCPs through periodic inspections of trial sponsors, principal investigators, and clinical trial sites. Although we rely on CROs to conduct GCP-compliant clinical trials, we remain responsible for ensuring that each of our GLP preclinical studies and clinical trials is conducted in accordance with its investigational plan, protocol, and applicable laws and regulations. Our reliance on the CROs does not relieve us of our regulatory responsibilities. If we or our CROs fail to comply with GCPs, the clinical data generated in our clinical trials may be deemed unreliable, and the FDA or comparable foreign regulatory authorities may require us to perform additional clinical trials before approving our
51
marketing applications. Accordingly, if our CROs fail to comply with these regulations or fail to recruit a sufficient number of subjects, we may be required to repeat clinical trials, which would delay the regulatory approval process.
Our reliance on third parties to conduct clinical trials will result in less direct control over the management of data developed through clinical trials than would be the case if we were relying entirely upon our own staff. Any failure by third parties to prevent unauthorized access, use, or disclosure of data, including personal data regarding our patients or employees, could harm our reputation, cause us not to comply with federal and/or state breach notification laws and foreign law equivalents, and otherwise subject us to liability under laws and regulations that protect the privacy and security of personal data.
Communicating with CROs and other third parties can be challenging, potentially leading to mistakes as well as difficulties in coordinating activities. Such parties may:
• | have staffing difficulties; |
• | fail to comply with contractual obligations; |
• | experience regulatory compliance issues; |
• | experience business disruptions from public health emergencies; or |
• | undergo changes in priorities or become financially distressed. |
These factors may materially adversely affect the willingness or ability of third parties to conduct our clinical trials and may subject us to unexpected cost increases that are beyond our control. If our CROs do not successfully carry out their contractual duties or obligations, fail to meet expected deadlines, fail to comply with regulatory requirements, or if the quality or accuracy of the clinical data they obtain is compromised due to the failure to adhere to our clinical protocols or regulatory requirements or for any other reasons, our clinical trials may be extended, delayed, or terminated, and we may not be able to obtain regulatory approval for, or successfully commercialize, any product candidate that we develop. As a result, our financial results and the commercial prospects for any product candidate that we develop would be harmed, our costs could increase, and our ability to generate revenue could be delayed. While we will have agreements governing their activities, our CROs will not be our employees, and we will not control whether or not they devote sufficient time and resources to our future clinical and nonclinical programs. These CROs may also have relationships with other commercial entities, including our competitors, for whom they may also be conducting clinical trials, or other drug development activities that could harm our business. We face the risk of potential unauthorized disclosure or misappropriation of our intellectual property by CROs, which may reduce our trade secret protection and allow our potential competitors to access and exploit our proprietary technology.
If our relationship with any of these CROs terminates, we may not be able to enter into arrangements with alternative CROs or do so on commercially reasonable terms. Switching or adding additional CROs involves substantial cost and requires management time and focus. In addition, there is a natural transition period when a new CRO commences work. As a result, delays occur, which can negatively affect our ability to meet our desired clinical development timelines. Though we intend to manage carefully our relationships with our CROs, there can be no assurance that we will not encounter challenges or delays in the future or that these delays or challenges will not have a negative impact on our business, financial condition, and prospects.
In addition, principal investigators for our clinical trials may serve as scientific advisors or consultants to us from time to time and receive compensation in connection with such services. Under certain circumstances, we may be required to report some of these relationships to the FDA. The FDA may conclude that a financial relationship between us and a principal investigator has created a conflict of interest or otherwise affected interpretation of the trial. The FDA may therefore question the integrity of the data generated at the applicable clinical trial site and the utility of the clinical trial itself may be jeopardized. This could result in a delay in approval or rejection of our marketing applications by the FDA and may ultimately lead to the denial of marketing approval of etripamil and any future product candidates.
52
Etripamil is intended to be used with a nasal-spray device, which may result in additional regulatory and supply risks.
Etripamil is administered through a nasal-spray device that we obtain from a single-source supplier, and that supplier is relying on multiple component suppliers, some of whom are single-source suppliers. There are a limited number of device suppliers that address our particular design requirements. In addition, increasing the scale of production inherently creates increased risk of manufacturing errors, and we may not be able to adequately inspect every device that is produced, and it is possible that individual devices may fail to perform as designed. While we intend to explore alternative nasal spray devices for the delivery of etripamil that are produced by other suppliers to have backup sources for future commercial needs, we may not identify other nasal device suppliers that meet our requirements, and such alternative devices may not be as effective at the delivery of etripamil as our current supplier’s device. We do not currently have a formal supply agreement with our current sole nasal spray device supplier, and we are obtaining such devices as needed. Even if we reach an agreement for commercial supply, if we do not have additional nasal spray device suppliers, our sole supplier may be unable to meet our demands. Unpredictability of supply could have a material adverse effect on our commercialization plans for etripamil and could have a material adverse effect on our business and financial condition.
Delays in or failure of the studies conducted by us, or failure of our Company, our collaborators, if any, or the third-party providers or suppliers to obtain or maintain regulatory approval could result in increased development costs, delays in or failure to obtain regulatory approval, and associated delays in etripamil reaching the market. Further, failure to successfully develop or supply the device, or to gain or maintain its approval, could adversely affect sales of etripamil.
Risks Related to Our Intellectual Property
If we are unable to obtain and maintain patent protection for etripamil or any future product candidates, or if the scope of the patent protection obtained is not sufficiently broad, our competitors could develop and commercialize drugs similar or identical to ours, and our ability to commercialize successfully our product candidates may be impaired.
Our success depends in large part on our ability to obtain and maintain patent protection in the United States and other countries with respect to etripamil and any future product candidates. We seek to protect our proprietary position by filing patent applications in the United States and abroad related to our product candidates. The patent application and prosecution process is expensive and time-consuming. We may not be able to file and prosecute all necessary or desirable patent applications at a reasonable cost or in a timely manner. We may also fail to identify patentable aspects of our research and development before it is too late to obtain patent protection. Therefore, these and any of our patents and applications may not be prosecuted and enforced in a manner consistent with the best interests of our business. It is possible that defects of form in the preparation or filing of our patents or patent applications may exist, or may arise in the future, such as with respect to proper priority claims, inventorship, claim scope, or patent term adjustments. If any future licensors or licensees are not fully cooperative or disagree with us as to the prosecution, maintenance or enforcement of any patent rights, such patent rights could be compromised and we might not be able to prevent third parties from making, using and selling competing products. If there are material defects in the form or preparation of our patents or patent applications, such patents or applications may be invalid and unenforceable. Moreover, our competitors may independently develop equivalent knowledge, methods, and know-how. Any of these outcomes could impair our ability to prevent competition from third parties.
The patent position of biotechnology and pharmaceutical companies generally is highly uncertain. Changes in either the patent laws or interpretation of the patent laws in the United States and other countries may diminish the value of our patents or narrow the scope of our patent protection. In addition, the laws of foreign countries may not protect our rights to the same extent as the laws of the United States. No consistent policy regarding the breadth of claims allowed in biotechnology and pharmaceutical patents has emerged to date in the United States or in many foreign jurisdictions. In addition, the determination of patent rights with respect to pharmaceutical compounds and technologies commonly involves complex legal and factual questions, which has in recent years been the subject of much litigation. As a result, the issuance, scope, validity, enforceability, and commercial value of our patent rights are highly uncertain. Furthermore, recent changes in patent laws in the United States, including the America Invents Act of 2011, may affect the scope, strength, and enforceability of our patent rights or the nature of proceedings that may be brought by us related to our patent rights.
53
We may not be aware of all third-party intellectual property rights potentially relating to etripamil or any future product candidates. Publications of discoveries in the scientific literature often lag behind the actual discoveries, and patent applications in the United States and other jurisdictions are typically not published until 18 months after filing, or in some cases not at all. For example, U.S. applications filed before November 28, 2000, and certain U.S. applications filed after that date that will not be filed outside the United States remain confidential until a patent issues. Therefore, we cannot be certain that we were the first to make the inventions claimed in our patents or pending patent applications, or that we were the first to file for patent protection of such inventions. Similarly, should we own any patents or patent applications in the future, we may not be certain that we were the first to file for patent protection for the inventions claimed in such patents or patent applications. As a result, the issuance, scope, validity, and commercial value of our patent rights cannot be predicted with any certainty. Moreover, we may be subject to a third-party preissuance submission of prior art to the U.S. Patent and Trademark Office, or “USPTO,” or become involved in opposition, derivation, reexamination, inter parties review, post grant review, or interference proceedings, in the United States or elsewhere, challenging our patent rights or the patent rights of others. An adverse determination in any such submission, proceeding, or litigation could reduce the scope of, or invalidate, our patent rights, allow third parties to commercialize our technology or product candidates and compete directly with us, without payment to us, or result in our inability to manufacture or commercialize products without infringing third-party patent rights.
Our pending and future patent applications may not result in patents being issued that protect our technology or product candidates, in whole or in part, or which effectively prevent others from commercializing competitive technologies and products. Even if our patent applications are issued as patents, they may not be issued in a form that will provide us with any meaningful protection against competing products or processes sufficient to achieve our business objectives, prevent competitors from competing with us or otherwise provide us with any competitive advantage. Our competitors may be able to circumvent our owned or licensed patents by developing similar or alternative technologies or products in a non-infringing manner. Our competitors may seek to market generic versions of any approved products by submitting abbreviated new drug applications to the FDA in which they claim that patents owned or licensed by us are invalid, unenforceable and/or not infringed. Alternatively, our competitors may seek approval to market their own products similar to or otherwise competitive with our products. In these circumstances, we may need to defend and/or assert our patents, including by filing lawsuits alleging patent infringement. In any of these types of proceedings, a court or other agency with jurisdiction may find our patents invalid and/or unenforceable.
The issuance of a patent is not conclusive as to its inventorship, scope, validity, or enforceability, and our owned and licensed patents may be challenged in the courts or patent offices in the United States and abroad. Such challenges may result in loss of exclusivity or freedom to operate or in patent claims being narrowed, invalidated, or held unenforceable, in whole or in part, which could limit our ability to stop others from using or commercializing similar or identical technology and products, or limit the duration of the patent protection of our technology and products. In addition, given the amount of time required for the development, testing, and regulatory review of new product candidates, patents protecting such candidates might expire before or shortly after such candidates are commercialized.
Obtaining and maintaining our patent protection depends on compliance with various procedural, document submission, fee payment, and other requirements imposed by government patent agencies, and our patent protection could be reduced or eliminated for non-compliance with these requirements.
The USPTO and various foreign governmental patent agencies require compliance with a number of procedural, documentary, fee payment, and other similar provisions during the patent application process. In addition, periodic maintenance fees, renewal fees, annuity fees, and various other government fees on patents and/or applications will have to be paid to the USPTO and various government patent agencies outside of the United States over the lifetime of our owned patents and/or applications and any patent rights we may own or license in the future. We rely on our outside counsel to pay these fees due to non-U.S. patent agencies. The USPTO and various non-U.S. government patent agencies require compliance with several procedural, documentary, fee payment, and other similar provisions during the patent application process. We employ reputable law firms and other professionals to help us comply.
Non-compliance events that could result in abandonment or lapse of a patent or patent application include, but are not limited to, failure to respond to official actions within prescribed time limits, non-payment of fees, and failure to properly legalize and submit formal documents. If we fail to maintain the patents and patent applications covering our products or
54
technologies, we may not be able to stop a competitor from marketing products that are the same as or similar to etripamil or any future product candidates, which would have a material adverse effect on our business. In many cases, an inadvertent lapse can be cured by payment of a late fee or by other means in accordance with the applicable rules. There are situations, however, in which non-compliance can result in abandonment or lapse of the patent or patent application, resulting in partial or complete loss of patent rights in the relevant jurisdiction. In such an event, potential competitors might be able to enter the market, and this circumstance could harm our business.
Patent terms may be inadequate to protect our competitive position on our product candidates for an adequate amount of time.
Given the amount of time required for the development, testing, and regulatory review of new product candidates, such as etripamil, patents protecting such candidates might expire before or shortly after such candidates are commercialized. We expect to seek extensions of patent terms in the United States and, if available, in other countries where we are prosecuting patents. In the United States, the Drug Price Competition and Patent Term Restoration Act of 1984 permits a patent term extension of up to five years beyond the normal expiration of the patent, which is limited to the approved indication (or any additional indications approved during the period of extension). However, the applicable authorities, including the FDA and the USPTO in the United States, and any equivalent regulatory authority in other countries, may not agree with our assessment of whether such extensions are available, and may refuse to grant extensions to our patents, or may grant more limited extensions than we request. If this occurs, our competitors may be able to take advantage of our investment in development and clinical trials by referencing our clinical and preclinical data and launch their drug earlier than might otherwise be the case.
Intellectual property rights do not necessarily address all potential threats to our business.
The degree of future protection afforded by our intellectual property rights is uncertain because intellectual property rights have limitations and may not adequately protect our business. The following examples are illustrative:
• | others may be able to make compounds or formulations that are similar to etripamil or formulations of etripamil or our future product candidates but that are not covered by the claims of any patents, should they issue, that we own or control; |
• | we or any strategic partners might not have been the first to make the inventions covered by the issued patents or pending patent applications that we own or control; |
• | we might not have been the first to file patent applications covering certain of our inventions; |
• | others may independently develop similar or alternative technologies or duplicate any of our technologies without infringing our intellectual property rights; |
• | it is possible that our pending patent applications will not lead to issued patents; |
• | issued patents that we own or control may not provide us with any competitive advantages, or may be held invalid or unenforceable as a result of legal challenges; |
• | our competitors might conduct research and development activities in the United States and other countries that provide a safe harbor from patent infringement claims for certain research and development activities, as well as in countries where we do not have patent rights and then use the information learned from such activities to develop competitive drugs for sale in our major commercial markets; |
• | we may not develop additional proprietary technologies that are patentable; and |
• | the patents of others may have an adverse effect on our business. |
55
Should any of these events occur, they could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We may become involved in lawsuits to protect or enforce our patents or other intellectual property rights, which could be expensive, time consuming and unsuccessful.
Competitors may infringe our issued patents, future trademarks, copyrights or other intellectual property. To counter infringement or unauthorized use, we may be required to file infringement claims, which can be expensive and time-consuming and divert the time and attention of our management and scientific personnel. Any claims we assert against perceived infringers could provoke these parties to assert counterclaims against us alleging that we infringe their patents, trademarks, copyrights, or other intellectual property. In addition, in a patent infringement proceeding, there is a risk that a court will decide that a patent of ours is invalid or unenforceable, in whole or in part, and that we do not have the right to stop the other party from using the invention at issue. There is also a risk that, even if the validity of such patents is upheld, the court will construe the patent’s claims narrowly or decide that we do not have the right to stop the other party from using the invention at issue on the grounds that our patents do not cover the invention. An adverse outcome in a litigation or proceeding involving our patents could limit our ability to assert our patents against those parties or other competitors and may curtail or preclude our ability to exclude third parties from making and selling similar or competitive products. Similarly, if we assert trademark infringement claims, a court may determine that the marks we have asserted are invalid or unenforceable, or that the party against whom we have asserted trademark infringement has superior rights to the marks in question. In this case, we could ultimately be forced to cease use of such trademarks.
In any infringement litigation, any award of monetary damages we receive may not be commercially valuable. Furthermore, because of the substantial amount of discovery required in connection with intellectual property litigation, there is a risk that some of our confidential information could be compromised by disclosure during litigation. In addition, there could be public announcements of the results of hearings, motions, or other interim proceedings or developments and if securities analysts or investors perceive these results to be negative, it could have a substantial adverse effect on the price of our common shares. Moreover, there can be no assurance that we will have sufficient financial or other resources to file and pursue such infringement claims, which typically last for years before they are concluded. Some of our competitors may be able to sustain the costs of such litigation or proceedings more effectively than we can because of their greater financial resources and more mature and developed intellectual property portfolios. Even if we ultimately prevail in such claims, the monetary cost of such litigation and the diversion of the attention of our management and scientific personnel could outweigh any benefit we receive as a result of the proceedings. Accordingly, despite our efforts, we may not be able to prevent third parties from infringing, misappropriating, or successfully challenging our intellectual property rights. Uncertainties resulting from the initiation and continuation of patent litigation or other proceedings could have a negative impact on our ability to compete in the marketplace.
Third parties may initiate legal proceedings alleging that we are infringing their intellectual property rights, the outcome of which would be uncertain and could have a negative impact on the success of our business.
Our commercial success depends, in part, upon our ability and the ability of future collaborators, if any, to develop, manufacture, market, and sell etripamil and any future product candidates and use our proprietary technologies without infringing the proprietary rights and intellectual property of third parties. The biotechnology and pharmaceutical industries are characterized by extensive and complex litigation regarding patents and other intellectual property rights. We may in the future become party to, or be threatened with, adversarial proceedings or litigation regarding intellectual property rights with respect to etripamil and any future product candidates and technology, including interference proceedings, post grant review, and inter parties review before the USPTO. Third parties may assert infringement claims against us based on existing patents or patents that may be granted in the future, regardless of their merit. There is a risk that third parties may choose to engage in litigation with us to enforce or to otherwise assert their patent rights against us. Even if we believe such claims are without merit, a court of competent jurisdiction could hold that these third-party patents are valid, enforceable, and infringed, which could have a negative impact on our ability to commercialize our current and any future product candidates. In order to successfully challenge the validity of any such U.S. patent in federal court, we would need to overcome a presumption of validity. As this burden is a high one requiring us to present clear and convincing evidence as to the invalidity of any such U.S. patent claim, there is no assurance that a court of competent jurisdiction would invalidate the claims of any such U.S. patent. If we are found to infringe a third party’s valid and enforceable intellectual
56
property rights, we could be required to obtain a license from such third party to continue developing, manufacturing and marketing our product candidate(s) and technology. However, we may not be able to obtain any required license on commercially reasonable terms or at all. Even if we were able to obtain a license, it could be nonexclusive, thereby giving our competitors and other third parties access to the same technologies licensed to us, and it could require us to make substantial licensing and royalty payments. We could be forced, including by court order, to cease developing, manufacturing and commercializing the infringing technology or product candidate. In addition, we could be found liable for monetary damages, including treble damages and attorneys’ fees, if we are found to have willfully infringed a patent or other intellectual property right. A finding of infringement could prevent us from manufacturing and commercializing etripamil or any future product candidates or force us to cease some or all of our business operations, which could materially harm our business. Claims that we have misappropriated the confidential information or trade secrets of third parties could have a similar negative impact on our business, financial condition, results of operations and prospects. See the section herein titled “Legal Proceedings” for additional information.
We may need to license intellectual property from third parties, and such licenses may not be available or may not be available on commercially reasonable terms.
A third party may hold intellectual property rights, including patent rights, which are important or necessary to the development of etripamil or any future product candidates. It may be necessary for us to use the patented or proprietary technology of third parties to commercialize our product candidates, in which case we would be required to obtain a license from these third parties. Such a license may not be available on commercially reasonable terms, or at all, and we could be forced to accept unfavorable contractual terms. If we are unable to obtain such licenses on commercially reasonable terms, our business could be harmed.
We may be subject to claims asserting that our employees, consultants, or advisors have wrongfully used or disclosed alleged trade secrets of their current or former employers or claims asserting ownership of what we regard as our own intellectual property.
Many of our employees, consultants, and advisors are currently, or were previously, employed at universities or other biotechnology or pharmaceutical companies, including our competitors or potential competitors. Although we try to ensure that our employees, consultants and advisors do not use the proprietary information or know-how of others in their work for us, we may be subject to claims that these individuals or we have used or disclosed intellectual property, including trade secrets or other proprietary information, of any such individual’s current or former employer. Litigation may be necessary to defend against these claims. If we fail in defending any such claims, in addition to paying monetary damages, we may lose valuable intellectual property rights or personnel. Even if we are successful in defending against such claims, litigation could result in substantial costs and be a distraction to management.
In addition, we may in the future be subject to claims by our former employees or consultants asserting an ownership right in our patents or patent applications, as a result of the work they performed on our behalf. Although it is our policy to require our employees and contractors who may be involved in the development of intellectual property to execute agreements assigning such intellectual property to us, we may be unsuccessful in executing such an agreement with each party who, in fact, conceives or develops intellectual property that we regard as our own, and we cannot be certain that our agreements with such parties will be upheld in the face of a potential challenge or that they will not be breached, for which we may not have an adequate remedy. The assignment of intellectual property rights may not be self-executing, or the assignment agreements may be breached, and we may be forced to bring claims against third parties, or defend claims that they may bring against us, to determine the ownership of what we regard as our intellectual property.
Changes in U.S. patent law or the patent law of other countries or jurisdictions could diminish the value of patents in general, thereby impairing our ability to protect etripamil and any future product candidates.
The United States has recently enacted and implemented wide ranging patent reform legislation. The U.S. Supreme Court has ruled on several patent cases in recent years, either narrowing the scope of patent protection available in certain circumstances or weakening the rights of patent owners in certain situations. In addition to increasing uncertainty with regard to our ability to obtain patents in the future, this combination of events has created uncertainty with respect to the
57
value of patents, once obtained. Depending on actions by the U.S. Congress, the federal courts, and the USPTO, the laws and regulations governing patents could change in unpredictable ways that could weaken our ability to obtain new patents or to enforce patents that we have licensed or that we might obtain in the future. Similarly, changes in patent law and regulations in other countries or jurisdictions, changes in the governmental bodies that enforce them or changes in how the relevant governmental authority enforces patent laws or regulations may weaken our ability to obtain new patents or to enforce patents that we have licensed or that we may obtain in the future. For example, the complexity and uncertainty of European patent laws have also increased in recent years. In Europe, a unitary patent system was launched on June 1, 2023. Under the unitary patent system, European applications have the option, upon grant of a patent, of becoming a Unitary Patent which is subject to the jurisdiction of the Unitary Patent Court, or “UPC.” As the UPC is a relatively new court system, there is little precedent for the court, increasing the uncertainty of any litigation. Patents granted before the implementation of the UPC had the option of opting out of the jurisdiction of the UPC and remaining as national patents in the UPC countries. Patents that remain under the jurisdiction of the UPC are potentially vulnerable to a single UPC-based revocation challenge that, if successful, could invalidate the patent in all countries who are signatories to the UPC. We cannot predict with certainty the long-term effects of any potential changes.
We may not be able to protect our intellectual property rights throughout the world, which could negatively impact our business.
Filing, prosecuting, and defending patents on product candidates in all countries throughout the world would be prohibitively expensive, and our intellectual property rights in some countries outside the United States could be less extensive than those in the United States. In some cases, we may not be able to obtain patent protection for certain technology outside the United States. In addition, the laws of some foreign countries do not protect intellectual property rights to the same extent as federal and state laws in the United States, even in jurisdictions where we do pursue patent protection. Consequently, we may not be able to prevent third parties from practicing our inventions in all countries outside the United States, even in jurisdictions where we do pursue patent protection or from selling or importing products made using our inventions in and into the United States or other jurisdictions.
Many companies have encountered significant problems in protecting and defending intellectual property rights in foreign jurisdictions. The legal systems of certain countries, particularly certain developing countries, do not favor the enforcement of patents and other intellectual property protection, which could make it difficult for us to stop the infringement of our patents, if pursued and obtained, or marketing of competing products in violation of our proprietary rights generally. Proceedings to enforce our patent rights in foreign jurisdictions could result in substantial costs and divert our efforts and attention from other aspects of our business, could put our patents at risk of being invalidated or interpreted narrowly and our patent applications at risk of not issuing, and could provoke third parties to assert claims against us. We may not prevail in any lawsuits that we initiate, and the damages or other remedies awarded, if any, may not be commercially meaningful. Accordingly, our efforts to enforce our intellectual property rights around the world may be inadequate to obtain a significant commercial advantage from the intellectual property that we develop or license.
The ongoing conflict in Ukraine and related sanctions could significantly devalue our Russian and Ukrainian patents. Recent Russian decrees may significantly limit our ability to enforce Russian patents. We cannot predict when or how this situation will change.
Reliance on third parties requires us to share our proprietary information, which increases the possibility that such information will be misappropriated or disclosed.
Because we rely on third parties to develop and manufacture etripamil and any future product candidates, or if we collaborate with additional third parties for the development or commercialization of etripamil or any future product candidates, we must, at times, share proprietary information with them. We seek to protect our proprietary technology in part by entering into confidentiality agreements and, if applicable, material transfer agreements, consulting agreements, or other similar agreements with our advisors, employees, third-party contractors, and consultants prior to beginning research or disclosing proprietary information. These agreements typically limit the rights of the third parties to use or disclose our confidential information. Despite the contractual provisions employed when working with third parties, the need to share confidential information increases the risk that such information becomes known by our competitors, is inadvertently
58
incorporated into the technology of others, or is disclosed or used in violation of these agreements. Given that our proprietary position is based, in part, on our know-how, a competitor’s discovery of our know-how or other unauthorized use or disclosure could have an adverse effect on our business and results of operations.
In addition, these agreements typically restrict the ability of our advisors, employees, third-party contractors and consultants to publish data potentially relating to our know-how. Despite our efforts to protect our know-how, we may not be able to prevent the unauthorized disclosure or use of our technical know-how by the parties to these agreements. Moreover, we cannot guarantee that we have entered into such agreements with each party that may have or have had access to our confidential information or proprietary technology and processes. Monitoring unauthorized uses and disclosures is difficult, and we do not know whether the steps we have taken to protect our proprietary technologies will be effective. If any of the collaborators, scientific advisors, employees, contractors, and consultants who are parties to these agreements breaches or violates the terms of any of these agreements, we may not have adequate remedies for any such breach or violation. Moreover, if confidential information that is licensed or disclosed to us by our partners, collaborators, or others is inadvertently disclosed or subject to a breach or violation, we may be exposed to liability to the owner of that confidential information. Enforcing a claim that a third-party illegally obtained and is using our proprietary information, like patent litigation, is expensive and time consuming, and the outcome is unpredictable. In addition, courts outside the United States are sometimes less willing to protect proprietary information.
Any trademarks we may obtain may be infringed or successfully challenged, resulting in harm to our business.
We expect to rely on trademarks as one means to distinguish any of our product candidates that are approved for marketing from the products of our competitors. In the event that our trademarks are successfully challenged, we could be forced to rebrand our products, which could result in loss of brand recognition and could require us to devote resources to advertising and marketing new brands. Our competitors may infringe our trademarks, and we may not have adequate resources to enforce our trademarks.
In addition, any proprietary name we propose to use with any future product candidate in the United States must be approved by the FDA, regardless of whether we have registered it, or applied to register it, as a trademark. The FDA typically conducts a review of proposed product names, including an evaluation of the potential for confusion with other product names. If the FDA objects to any of our proposed proprietary product names, we may be required to expend significant additional resources in an effort to identify a suitable proprietary product name that would qualify under applicable trademark laws, not infringe the existing rights of third parties and be acceptable to the FDA.
If we are unable to protect the confidentiality of our proprietary information, our business and competitive position would be harmed.
In addition to seeking and/or maintaining patent and trademark protection for etripamil and any future product candidate, we also rely on unpatented know-how, technology and other proprietary information, to maintain our competitive position. We seek to protect our proprietary information, in part, by entering into non-disclosure and confidentiality agreements with parties who have access to them, such as our employees, corporate collaborators, outside scientific collaborators, contract manufacturers, consultants, advisors, and other third parties. We also enter into confidentiality and invention or patent assignment agreements with our employees and consultants. Despite these efforts, any of these parties may breach the agreements and disclose our proprietary information. Monitoring unauthorized uses and disclosures of our intellectual property is difficult, and we do not know whether the steps we have taken to protect our intellectual property will be effective. In addition, we may not be able to obtain adequate remedies for any such breaches. Enforcing a claim that a party illegally disclosed or misappropriated proprietary information is difficult, expensive, and time consuming, and the outcome is unpredictable. In addition, some courts inside and outside the United States are less willing or unwilling to protect trade secrets.
Moreover, our competitors may independently develop knowledge, methods, and know-how equivalent to our proprietary information. Competitors could purchase our products and replicate some or all of the competitive advantages we derive from our development efforts for technologies on which we do not have patent protection. If any of our proprietary information were to be lawfully obtained or independently developed by a competitor, we would have no right to prevent
59
them, or those to whom they communicate, from using that technology or information to compete with us. If any of our proprietary information were to be disclosed to or independently developed by a competitor, our competitive position would be harmed.
We also seek to preserve the integrity and confidentiality of our data and other confidential information by maintaining physical security of our premises and physical and electronic security of our information technology systems. While we have confidence in these individuals, organizations, and systems, agreements or security measures may be breached and detecting the disclosure or misappropriation of confidential information and enforcing a claim that a party illegally disclosed or misappropriated confidential information is difficult, expensive and time-consuming, and the outcome is unpredictable. Further, we may not be able to obtain adequate remedies for any breach. In addition, our confidential information may otherwise become known or be independently discovered by competitors, in which case we would have no right to prevent them, or those to whom they communicate, from using that technology or information to compete with us.
Risks Related to Our Business Operations, Employee Matters and Managing Growth
Our future success depends on our ability to retain key executives and to attract, retain and motivate qualified personnel.
We are highly dependent on our President and Chief Executive Officer, Joseph Oliveto, our Chief Medical Officer, David Bharucha, our Chief Commercial Officer, Lorenz Muller and our Chief Financial Officer, Amit Hasija. Each of these officers may currently terminate their employment with us at any time. The loss of the services of any of these persons could impede the achievement of our research, development, and commercialization objectives. We do not currently maintain “key person” life insurance on the lives of our executives or any of our employees other than on our President and Chief Executive Officer, Joseph Oliveto.
Recruiting and retaining qualified scientific and clinical personnel and, if we progress the development of any of our product candidates, commercialization, manufacturing, and sales and marketing personnel will be critical to our success. The loss of the services of our executive officers or other key employees could impede the achievement of our research, development, and commercialization objectives and seriously harm our ability to successfully implement our business strategy. Furthermore, replacing executive officers and key employees may be difficult and may take an extended period of time because of the limited number of individuals in our industry with the breadth of skills and experience required to successfully develop, gain regulatory approval of, and commercialize our product candidates. Competition to hire from this limited pool is intense, and we may be unable to hire, train, retain, or motivate these key personnel on acceptable terms given the competition among numerous pharmaceutical and biotechnology companies for similar personnel. We also experience competition for the hiring of scientific and clinical personnel from universities and research institutions. In addition, we rely on consultants and advisors, including scientific and clinical advisors, to assist us in formulating our research and development and commercialization strategy. Our consultants and advisors may have commitments under consulting or advisory contracts with other entities that may limit their availability to us. If we are unable to continue to attract and retain high-quality personnel, our ability to pursue our growth strategy will be limited.
We may expand our organization in the future, and we may experience difficulties in managing this growth, which could disrupt our operations.
As we continue our efforts to commercialize CARDAMYST for the treatment of PSVT, pursue the development of subsequent etripamil indications and as we look to expand our pipeline, we may, in the future, experience significant growth in the number of our employees and the scope of our operations, particularly in the areas of research, drug development, regulatory affairs, sales, marketing and distribution. To manage any future growth, we will be required to continue to implement and improve our managerial, operational and financial systems, expand our facilities and continue to recruit and train additional qualified personnel. Due to our limited financial resources and the limited experience of our management team in managing a company with growth, we may not be able to effectively manage any expansion of our operations or recruit and train additional qualified personnel. Any expansion of our operations may lead to significant
60
costs and may divert our management and business development resources. Any inability to manage growth could delay the execution of our business plans or disrupt our operations.
Our internal computer systems, or those of our collaborators, contractors, consultants, or other third parties with whom we work, may fail or suffer security breaches, which could result in a material disruption of our product development programs and can lead to adverse consequences, including but not limited to regulatory investigations or actions; litigation; fines and penalties; reputational harm; loss of revenue or profits; loss of customers or sales; disruption of our business operations.
In the ordinary course of our business, we and the third parties with whom we work, process proprietary, confidential, and sensitive data, including personal data (such as health-related data), intellectual property and trade secrets, or collectively, “sensitive information”. Our internal computer systems and those of our current and any future collaborators, contractors, consultants, or other third parties with whom we work, are vulnerable to a variety of evolving threats, including but not limited to damage from malicious code (such as computer viruses and worms), unauthorized access, natural disasters, terrorism, war, social-engineering attacks (including through deep fakes, which may be increasingly more difficult to identify as fake, and phishing attacks), malware (including as a result of advanced persistent threat intrusions), denial-of-service attacks, credential stuffing attacks, credential harvesting, personnel misconduct or error, ransomware attacks, supply-chain attacks, software bugs, server malfunctions, software or hardware failures, loss of data or other information technology assets, adware, attacks enhanced or facilitated by artificial intelligence, telecommunication and electrical failures, and other similar threats. In particular, severe ransomware attacks are becoming increasingly prevalent and can lead to significant interruptions in our operations, ability to provide our products or services, loss of sensitive data and income, reputational harm, and diversion of funds. Extortion payments may alleviate the negative impact of a ransomware attack, but we may be unwilling or unable to make such payments due to, for example, applicable laws or regulations prohibiting such payments. Cyber-attacks are increasing in their frequency, sophistication, and intensity, and have become increasingly difficult to detect. Cyber-attacks, malicious internet-based activity, online and offline fraud, and other similar activities can affect service reliability and threaten the confidentiality, integrity, and availability our sensitive information and information technology systems, and those of the third parties with whom we work. Cyber-attacks also could include phishing attempts or e-mail fraud to cause payments or information to be transmitted to an unintended recipient.
Some actors now engage and are expected to continue to engage in cyber-attacks, including without limitation nation-state actors for geopolitical reasons and in conjunction with military conflicts and defense activities. During times of war and other major conflicts, we, the third parties with whom we work, may be vulnerable to a heightened risk of these attacks, including retaliatory cyber-attacks, which could materially disrupt our systems and operations, supply chain, and ability to produce, sell, and distribute our goods and services. A significant system failure, accident, or security could cause interruptions in our operations, and could result in a disruption of our development programs and our business operations, whether due to a loss of our trade secrets or other proprietary information or other similar disruptions. For example, the loss of clinical trial data from completed or future clinical trials by us or our CROs could result in delays in our regulatory approval efforts and significantly increase our costs to recover or reproduce the data. Additionally, any such event that leads to unauthorized access, use or disclosure of personal data, including personal data regarding our patients or employees, could harm our reputation, cause us not to comply with federal and/or state breach notification laws and foreign law equivalents and otherwise subject us to liability under laws and regulations that protect the privacy and security of personal data. Security breaches and other inappropriate access can be difficult to detect, and any delay in identifying them may lead to increased harm of the type described above. Applicable data privacy and security obligations may require us, or we may voluntarily choose, to notify relevant stakeholders, including affected individuals, customers, regulators, and investors, of security incidents, or to take other actions, such as providing credit monitoring and identity theft protection services. Such disclosures and related actions can be costly, and the disclosure or failure to comply with such requirements could lead to adverse consequences.
While we have implemented security measures designed to protect against security incidents, there can be no assurance that these measures will be effective. We take steps to detect, mitigate, and remediate vulnerabilities in our information systems (such as our hardware and/or software, including that of third parties with whom we work), but we may not, in the future, be able to detect and remediate all such vulnerabilities including on a timely basis. Further, we have experienced and may in the future experience delays in developing and deploying remedial measures and patches designed to address identified vulnerabilities. Vulnerabilities could be exploited and result in a security incident. To the extent that any
61
disruption or security breach were to result in a loss of, or damage to, our data or applications, or inappropriate disclosure of sensitive information, we could incur liability, our competitive position could be harmed, and the further development and commercialization of our product candidates could be delayed. We may expend significant resources or modify our business activities (including our clinical trial activities) to try to protect against security incidents. Certain data privacy and security obligations have required us to implement and maintain specific security measures or industry-standard or reasonable security measures to protect our information technology systems and sensitive information. It may be difficult and/or costly to detect, investigate, mitigate, contain, and remediate a security incident. Our efforts to do so may not be successful. Actions taken by us or the third parties with whom we work to detect, investigate, mitigate, contain, and remediate a security incident could result in outages, data losses, and disruptions of our business. Threat actors may also gain access to other networks and systems after a compromise of our networks and systems. For example, threat actors may use an initial compromise of one part of our environment to gain access to other parts of our environment, or leverage a compromise of our networks or systems to gain access to the networks or systems of third parties with whom we work, such as through phishing or supply chain attacks.
Future or past business transactions (such as acquisitions or integrations) could expose us to additional cybersecurity risks and vulnerabilities, as our systems could be negatively affected by vulnerabilities present in acquired or integrated entities’ systems and technologies. Furthermore, we may discover security issues that were not found during due diligence of such acquired or integrated entities, and it may be difficult to integrate companies into our information technology environment and security program.
We rely on third parties to operate critical business systems to process sensitive information in a variety of contexts, including, without limitation, cloud-based infrastructure, data center facilities, encryption and authentication technology, employee email, content delivery to customers, and other functions. Our ability to monitor these third parties’ information security practices is limited, and these third parties may not have adequate information security measures in place. If the third parties with whom we work experience a security incident or other interruption, we could experience adverse consequences. While we may be entitled to damages if the third parties with whom we work fail to satisfy their privacy or security-related obligations to us, any award may be insufficient to cover our damages, or we may be unable to recover such an award. In addition, supply-chain attacks have increased in frequency and severity, and we cannot guarantee that third parties’ infrastructure in our supply chain or that of the third parties with whom we work have not been compromised.
Any of the previously identified or similar threats have in the past and may in the future cause a security incident or other interruption that have in the past and may in the future result in unauthorized, unlawful, or accidental acquisition, modification, destruction, loss, alteration, encryption, disclosure of, or access to our sensitive information or our information technology systems, or those of the third parties with whom we work. A security incident or other interruption could disrupt our ability (and that of the third parties with whom we work) to provide our services.
If we (or the third parties with whom we work) experience a security incident or are perceived to have experienced a security incident, we may experience adverse consequences, such as government enforcement actions (for example, investigations, fines, penalties, audits, and inspections); additional reporting requirements and/or oversight; restrictions on processing sensitive information (including personal data); litigation (including class-action claims); indemnification obligations; negative publicity; reputational harm; monetary fund diversions; diversion of management attention; interruptions in our operations (including availability of data); financial loss; and other similar harms. Security incidents and attendant consequences may prevent or cause customers to stop using our services, deter new customers from using our services, and negatively impact our ability to grow and operate our business. Our contracts may not contain limitations of liability, and even where they do, there can be no assurance that limitations of liability in our contracts are sufficient to protect us from liabilities, damages, or claims related to our data privacy and security obligations.
In addition to experiencing a security incident, third parties may gather, collect, or infer sensitive information about us from public sources, data brokers, or other means that reveals competitively sensitive details about our organization and could be used to undermine our competitive advantage or market position. Additionally, sensitive information of the Company could be leaked, disclosed, or revealed as a result of or in connection with our employees’, personnel’s, or vendors’ use of generative artificial intelligence, or “AI,” technologies. Any sensitive information (including confidential, competitive, proprietary, or personal data) that we input into a third-party generative AI or platform could be leaked or disclosed to others, including if sensitive information is used to train the third parties’ AI technologies.
62
Additionally, where an AI technology ingests personal data and makes connections using such data, those technologies may reveal other personal or sensitive information generated by the model. Moreover, AI technologies may create flawed, incomplete, or inaccurate outputs, some of which may appear correct. This may happen if the inputs that the model relied on were inaccurate, incomplete or flawed (including if a bad actor “poisons” the AI technology with bad inputs or logic), or if the logic of the AI technology is flawed (a so-called “hallucination”). We may use AI technology outputs to make certain decisions. Due to these potential inaccuracies or flaws, the model could be biased and could lead us to make decisions that could bias certain individuals (or classes of individuals), and adversely impact their rights, employment, and ability to obtain certain pricing, products, services, or benefits.
We (and the third parties with whom we work) are subject to stringent and evolving U.S. and foreign laws, regulations, and rules, contractual obligations, industry standards, policies, and other obligations related to data privacy and security. Our (or the third parties with whom we work) actual or perceived failure to comply with such obligations could lead to regulatory investigations or actions; litigation (including class-action claims) and mass arbitration demands; fines and penalties; disruptions of our business operations; reputational harm; loss of revenue or profits; and other adverse business consequences.
In the ordinary course of business, we collect, receive, store, process, generate, use, transfer, disclose, make accessible, protect, secure, dispose of, transmit, and share (collectively, “process”) personal data and other sensitive information, including proprietary and confidential business data, trade secrets, intellectual property, data we collect about trial participants in connection with clinical trials and sensitive third-party data. Our data processing activities subject us to numerous data privacy and security obligations, such as various laws, regulations, guidance, industry standards, external and internal privacy and security policies, contractual requirements, and other obligations relating to data privacy and security. In the United States, federal, state, and local governments have enacted numerous data privacy and security laws, including data breach notification laws, personal data privacy laws, consumer protection laws (e.g., Section 5 of the Federal Trade Commission Act), and other similar laws (e.g., wiretapping laws). Numerous U.S. states have enacted comprehensive privacy laws that impose certain obligations on covered businesses, including providing specific disclosures in privacy notices and affording residents with certain rights concerning their personal data. The exercise of these rights may impact our business and ability to provide our products and services. Certain states also impose stricter requirements for processing certain personal data, including sensitive information, such as conducting data privacy impact assessments. These state laws allow for statutory fines for noncompliance. For example, the California Consumer Privacy Act of 2018 applies to personal data of consumers, business representatives, and employees who are California residents, and requires businesses to provide specific disclosures in privacy notices and honor requests of such individuals to exercise certain privacy rights. The CCPA provides for and allows private litigants affected by certain data breaches to recover significant statutory damages. Similar laws are being considered in several other states, as well as at the federal and local levels, and we expect more states to pass similar laws in the future. While these states, like the CCPA, also exempt some data processed in the context of clinical trials, these developments further complicate compliance efforts, and increase legal risk and compliance costs for us, the third parties upon whom we rely. We are also subject to new laws governing the privacy of consumer health data. For example, Washington’s My Health My Data Act, or “MHMD,” broadly defines consumer health data, places restrictions on processing consumer health data (including imposing stringent requirements for consents), provides consumers certain rights with respect to their health data, and creates a private right of action to allow individuals to sue for violations of the law. Other states have passed, are considering, and may adopt similar laws.
Outside the United States, an increasing number of laws, regulations, and industry standards govern data privacy and security. For example, the European Union’s General Data Protection Regulation, or “EU GDPR,” the United Kingdom’s GDPR, or “UK GDPR,”, and China’s Personal Information Protection Law, or “PIPL,” impose strict requirements for processing personal data. Additionally, in Canada, the Personal Information Protection and Electronic Documents Act, or “PIPEDA,” and various related provincial laws, as well as Canada’s Anti-Spam Legislation, or “CASL,” may apply to our operations.
We anticipate seeking regulatory approval for, and commercializing, etripamil for the treatment of PSVT in Europe. We may also elect to do so for future product candidates. We are conducting clinical trial activities in Europe, which subject us to European data protection laws, including the EU GDPR and the UK GDPR. The GDPR establishes requirements applicable to the processing of personal data (i.e., data which identifies an individual or from which an individual is
63
identifiable). The GDPR creates significant and complex compliance burdens for companies such as: limiting permitted processing of personal data to only that which is necessary for specified, explicit and legitimate purposes; requiring the establishment a legal basis for processing personal data; expressly confirming that ‘pseudonymized’ or key coded data constitutes personal data to which the GDPR applies; creating obligations for controllers and processors to appoint data protection officers in certain circumstances; increasing transparency obligations to data subjects for controllers (including presentation of certain information in a concise, intelligible and easily accessible form about how their personal data is used and their rights vis-à-vis that data and its use); introducing the obligation to carry out so-called data protection impact assessments in certain circumstances; establishing limitations on collection and retention of personal data through ‘data minimization’ and ‘storage limitation’ principles; establishing obligations to implement ‘privacy by design’; introducing obligations to honor increased rights for data subjects (such as rights for individuals to be ‘forgotten,’ rights to data portability, rights to object etc. in certain circumstances); formalizing a heightened and codified standard of data subject consent; establishing obligations to implement certain technical and organizational safeguards to protect the security and confidentiality of personal data; introducing obligations to agree to certain specific contractual terms and to take certain measures when engaging third-party processors and joint controllers; introducing the obligation to provide notice of certain significant personal data breaches to the relevant supervisory authority(ies) and affected individuals; and mandating the appointment of representatives in the United Kingdom, or the “UK,” and/or European Union in certain circumstances. The processing of “special category personal data”, such as health information, may also impose heightened compliance burdens under the GDPR. The GDPR has robust regulatory enforcement and penalties for noncompliance, including fines of up to €20 million Euros under the EU GDPR, 17.5 million pounds sterling under the UK GDPR, or, in each case 4% of global annual revenue of any noncompliant company for the preceding financial year, whichever is higher or private litigation related to processing of personal data brought by classes of data subjects or consumer protection organizations authorized at law to represent their interests. In addition to administrative fines, a wide variety of other potential enforcement powers are available to competent supervisory authorities in respect of potential and suspected violations of the GDPR, including extensive audit and inspection rights, and powers to order temporary or permanent bans on all or some processing of personal data carried out by noncompliant actors. The GDPR also confers a private right of action on data subjects and consumer associations to lodge complaints with supervisory authorities, seek judicial remedies, and obtain compensation for damages resulting from violations of the GDPR. There may be circumstances under which a failure to comply with GDPR, or the exercise of individual rights under the GDPR, would limit our ability to utilize clinical trial data collected on certain subjects. The GDPR will likely impose additional responsibility and liability in relation to our processing of personal data. This may be onerous and materially adversely affect our business, financial condition, results of operations, and prospects.
In the ordinary course of business, we transfer personal data from Europe and other jurisdictions to the United States or other countries. Europe and other jurisdictions have enacted laws requiring data to be localized or limiting the transfer of personal data to other countries. In particular, the European Economic Areas, or “EEA,” and the UK have significantly restricted the transfer of personal data to the United States and other countries whose privacy laws it generally believes are inadequate. One of the primary safeguards allowing U.S. companies to import personal data from Europe is the European Commission’s Standard Contractual Clauses and we have relied on Standard Contractual Clauses to comply with the GDPR’s restrictions on transfer of personal data out of Europe. However, in July 2020 the Court of Justice of the European Union, or “CJEU,” in a case known colloquially as “Schrems II” raised questions about whether the Standard Contractual Clauses can lawfully be used for personal data transfers from Europe to the United States or other third countries that are not the subject of an adequacy decision of the European Commission. At present, there are few viable alternatives to the Standard Contractual Clauses including the UK’s International Data Transfer Agreement / Addendum, and the EU-U.S. Data Privacy Framework and the UK extension thereto (which allows for transfers to relevant U.S.-based organizations who self-certify compliance and participate in the Framework). However, these mechanisms are subject to legal challenges, and there is no assurance that we can satisfy or rely on these measures to lawfully transfer personal data to the United States. As such, if we are unable to implement a valid solution to transfer personal data from the EEA, the UK or other jurisdictions to the United States, or if the requirements for a legally-compliant transfer are too onerous, we could face significant adverse consequences, including the interruption or degradation of our operations, increased exposure to regulatory actions, substantial fines and penalties, injunctions against our processing or transferring of personal data necessary to operate our business, the need to relocate part of or all of our business or data processing activities to other jurisdictions (such as Europe) at significant expense, and limit our ability to collaborate with partners as well as other service providers, contractors and other companies subject to European data protection laws. Restrictions on our ability to import personal data from Europe could therefore impact our clinical trial activities in Europe and limit our
64
ability to collaborate with CROs and other third parties subject to European data protection laws. Additionally, other countries outside of Europe have enacted or are considering enacting similarly stringent data localization and cross-border data transfer restrictions and laws, which could increase the cost and complexity of delivering our services and operating our business. The type of challenges we face in Europe will likely also arise in other jurisdictions that adopt laws similar in construction to the GDPR or regulatory frameworks of equivalent complexity.
Additionally, the U.S. Department of Justice issued a rule entitled Preventing Access to U.S. Sensitive Personal Data and Government-Related Data by Countries of Concern or Covered Persons, which places additional restrictions on certain data transactions involving countries of concern (e.g., China, Russia, Iran) and covered individuals (i.e., individuals and entities located in or controlled by individuals or entities located in those jurisdictions) that may impact certain business activities such as vendor engagements, employment of certain individuals and investor agreements. Violations of the rule could lead to significant civil and criminal fines and penalties. The rule applies regardless of whether data is anonymized, key-coded, pseudonymized, de-identified or encrypted.
In addition to data privacy and security laws, we are contractually subject to industry standards adopted by industry groups, and we are and may become in the future, subject to such obligations. We are also bound by contractual obligations related to data privacy and security, and our efforts to comply with such obligations may not be successful.
We publish privacy policies, marketing materials, whitepapers, and other statements, such as statements related to compliance with certain certifications or self-regulatory principles, concerning data privacy, security and AI. Regulators in the United States are increasingly scrutinizing these statements, and if these policies, materials or statements are found to be deficient, lacking in transparency, deceptive, unfair, misleading or misrepresentative of our practices, we may be subject to investigation, enforcement actions by regulators, or other adverse consequences.
Obligations related to data privacy and security (and consumers’ data privacy expectations) are quickly changing, becoming increasingly stringent and creating uncertainty. Additionally, these obligations may be subject to differing applications and interpretations, which may be inconsistent or conflict among jurisdictions. Preparing for and complying with these obligations requires us to devote significant resources, which may necessitate changes to our services, information technologies, systems, and practices and to those of any third parties that process personal data on our behalf. In addition, these obligations may require us to change our business model. We may at times fail (or be perceived to have failed) in our efforts to comply with our data privacy and security obligations. Moreover, despite our efforts, our personnel or third parties with whom we work may fail to comply with such obligations, which could negatively impact our business operations. If we or the third parties with whom we work fail, or are perceived to have failed, to address or comply with applicable data privacy and security obligations, we could face significant consequences, including but not limited to: government enforcement actions (e.g., investigations, fines, penalties, audits, inspections, and similar); litigation (including class-action claims) and mass arbitration demands; additional reporting requirements and/or oversight; bans or restrictions on processing personal data; orders to destroy or not use personal data; and imprisonment of company officials. In particular, plaintiffs have become increasingly more active in bringing privacy-related claims against companies, including class claims and mass arbitration demands. Some of these claims allow for the recovery of statutory damages on a per violation basis, and, if viable, carry the potential for monumental statutory damages, depending on the volume of data and the number of violations. Any of these events could have a material adverse effect on our reputation, business, or financial condition, including but not limited to: loss of customers; interruptions or stoppages in our business operations including, as relevant, clinical trials; inability to process personal data or to operate in certain jurisdictions; limited ability to develop or commercialize our products; expenditure of time and resources to defend any claim or inquiry; adverse publicity; or substantial changes to our business model or operations. We use generative AI to assist us in making certain decisions, which is regulated by certain privacy laws. Due to inaccuracies or flaws in the inputs, outputs, or logic of the AI technologies, the model could be biased and could lead us to make decisions that could bias certain individuals (or classes of individuals), and adversely impact their rights, employment, and ability to obtain certain pricing, products, services, or benefits. The development and use of AI technologies present various privacy and security risks that may impact our business. AI technologies are subject to privacy and data security laws, as well as increasing regulation and scrutiny.
Our employees and personnel may use AI technologies to perform their work, and the disclosure and use of personal data in generative AI technologies is subject to various privacy laws and other privacy obligations. Governments have passed
65
and are likely to pass additional laws and regulations which regulate generative AI. Our use of this technology could result in additional compliance costs, regulatory investigations and actions, and lawsuits. If we are unable to use generative AI, it could make our business less efficient and result in competitive disadvantages.
Our employees, principal investigators, consultants, and commercial partners may engage in misconduct or other improper activities, including non-compliance with regulatory standards and requirements and insider trading.
We are exposed to the risk of fraud or other misconduct by our employees, principal investigators, consultants, and commercial partners. Misconduct by these parties could include intentional failures to comply with FDA regulations or the regulations applicable in other jurisdictions, provide accurate information to the FDA and other regulatory authorities, comply with healthcare fraud and abuse laws and regulations in the United States and abroad, report financial information or data accurately, or disclose unauthorized activities to us. In particular, sales, marketing, and business arrangements in the healthcare industry are subject to extensive laws and regulations intended to prevent fraud, misconduct, kickbacks, self-dealing, and other abusive practices. These laws and regulations restrict or prohibit a wide range of pricing, discounting, marketing and promotion, sales commission, customer incentive programs, and other business arrangements. Such misconduct also could involve the improper use of information obtained in the course of clinical trials or interactions with the FDA or other regulatory authorities, which could result in regulatory sanctions and cause serious harm to our reputation. It is not always possible to identify and deter employee misconduct, and the precautions we take to detect and prevent this activity may not be effective in controlling unknown or unmanaged risks or losses or in protecting us from government investigations or other actions or lawsuits stemming from a failure to comply with these laws or regulations. If any such actions are instituted against us and we are not successful in defending ourselves or asserting our rights, those actions could result in significant civil, criminal, and administrative penalties, damages, fines, disgorgement, imprisonment, exclusion from participating in government-funded healthcare programs, such as Medicare and Medicaid, additional reporting requirements and oversight if we become subject to a corporate integrity agreement or similar agreement to resolve allegations of non-compliance with these laws, contractual damages, reputational harm, and the curtailment or restructuring of our operations, any of which could have a negative impact on our business, financial condition, results of operations, and prospects.
Disruptions at the FDA, the SEC and other government agencies and regulatory authorities caused by funding shortages or global health concerns could hinder their ability to hire and retain key leadership and other personnel, prevent new products and services from being developed or commercialized in a timely manner or otherwise prevent those agencies from performing normal business functions on which the operation of our business may rely, which could negatively impact our business.
The ability of the FDA and comparable foreign regulatory authorities to review and approve new products can be affected by a variety of factors, including government budget and funding levels, ability to hire and retain key personnel and accept the payment of user fees, and statutory, regulatory, and policy changes. Average review times at the agency have fluctuated in recent years as a result. In addition, government funding of the SEC, and other government agencies on which our operations may rely, including those that fund research and development activities, is subject to the political process, which is inherently fluid and unpredictable.
Disruptions at the FDA may slow the time necessary for new products to be reviewed and/or approved, which would adversely affect our business. For example, over the last several years, including most recently in October 2025, the U.S. government has shut down several times and certain regulatory agencies, such as the FDA and the SEC, have had to furlough critical FDA, SEC and other government employees and stop critical activities. If a prolonged government shutdown occurs, it could significantly impact the ability of the FDA to timely review and process our regulatory submissions, which could have a material adverse effect on our business.
The ability of the FDA and other government agencies to properly administer their functions is highly dependent on the levels of government funding and the ability to fill key leadership appointments, among various factors. Changes in FDA staffing could result in delays in the FDA’s responsiveness or in its ability to review submissions or applications, issue regulations or guidance, or implement or enforce regulatory requirements in a timely fashion or at all. If any legislation, executive orders, or lapses in agency funding impose constraints on the FDA’s ability to engage in oversight and
66
implementation activities in the normal course, our business may be negatively impacted. In addition, government funding of other government agencies that fund research and development activities is subject to the political process, which is inherently fluid and unpredictable.
Our current or future acquisitions or strategic collaborations could increase our capital requirements, dilute our shareholders, cause us to incur debt, or assume contingent liabilities and subject us to other risks.
We actively search for and continually evaluate various acquisition and strategic collaboration opportunities, including licensing or acquiring complementary drugs, intellectual property rights, technologies or businesses, as deemed appropriate to carry out our business plan. Our collaborations, including any future acquisitions or strategic partnerships, may entail numerous risks, including:
• | increased operating expenses and cash requirements; |
• | the assumption of additional indebtedness or contingent liabilities; |
• | assimilation of operations, intellectual property, and drugs of an acquired company, including difficulties associated with integrating new personnel; |
• | the diversion of our management’s attention from our existing drug programs and initiatives in pursuing such a strategic partnership, merger, or acquisition; |
• | retention of key employees, the loss of key personnel, and uncertainties in our ability to maintain key business relationships; |
• | risks and uncertainties associated with the other party to such a transaction, including the prospects of that party and their existing drugs or product candidates and regulatory approvals; and |
• | our inability to generate revenue from acquired technology and/or drugs sufficient to meet our objectives in undertaking the acquisition or even to offset the associated acquisition and maintenance costs. |
In addition, in connection with our current or future acquisitions or strategic partnerships, we may issue dilutive securities, assume or incur debt obligations, incur large one-time expenses, and acquire intangible assets that could result in significant future amortization expense. Moreover, we may not be able to locate suitable future acquisition opportunities, and this inability could impair our ability to grow or obtain access to technology or drugs that may be important to the development of our business.
Risks Related to Ownership of Our Common Shares
The market price of our common shares has been and may continue to be volatile and fluctuate substantially, and you could lose all or part of your investment.
The market price of our common shares has been and may continue to be highly volatile and could be subject to wide fluctuations in price in response to various factors, many of which are beyond our control. From January 1, 2025 through March 20, 2026, the price of our common shares has ranged from $0.65 per share to $2.95 per share.
The stock market in general and the market for biopharmaceutical and pharmaceutical companies in particular, has experienced extreme volatility that has often been unrelated to the operating performance of particular companies. As a result of this volatility, you may not be able to sell your common shares at or above the price paid for the shares. In addition to the factors discussed in this “Risk Factors” section and elsewhere in this Annual Report on Form 10-K, the market price for our common shares may be influenced by the following:
67
• | our ability to meet external revenue and profitability expectations for CARDAMYSTTM for the treatment of PSVT; |
• | the commencement, enrollment, or results of our planned or future clinical trials of etripamil and any future product candidates or those of our competitors; |
• | the success of competitive drugs or therapies; |
• | regulatory or legal developments in the United States and other countries; |
• | the success of competitive products or technologies; |
• | developments or disputes concerning patent applications, issued patents or other proprietary rights; |
• | the recruitment or departure of key personnel; |
• | the level of expenses related to etripamil and any future product candidates or clinical development programs; |
• | the results of our efforts to discover, develop, acquire or in-license additional product candidates; |
• | actual or anticipated changes in estimates as to financial results, development timelines or recommendations by securities analysts; |
• | our inability to obtain or delays in obtaining adequate drug supply for any approved drug or inability to do so at acceptable prices; |
• | disputes or other developments relating to proprietary rights, including patents, litigation matters, and our ability to obtain patent protection for our technologies; |
• | significant lawsuits, including patent or shareholder litigation; |
• | variations in our financial results or those of companies that are perceived to be similar to us; |
• | changes in the structure of healthcare payment systems, including coverage and adequate reimbursement for any approved drug; |
• | market conditions in the pharmaceutical and biotechnology sectors; |
• | general economic, political, and market conditions, including deteriorating market conditions due to investor concerns regarding inflation and global geopolitical conflicts, changes in trade and tariff policies, and overall fluctuations in the financial markets in the United States and abroad; and |
• | investors’ general perception of us and our business. |
These and other market and industry factors may cause the market price and demand for our common shares to fluctuate substantially, regardless of our actual operating performance, which may limit or prevent investors from selling their shares at or above the price paid for the shares and may otherwise negatively affect the liquidity of our common shares. In addition, the stock market in general, and biopharmaceutical companies in particular, have experienced extreme price and volume fluctuations that have often been unrelated or disproportionate to the operating performance of these companies.
Some companies that have experienced volatility in the trading price of their shares have been the subject of securities class action litigation. Any lawsuit to which we are a party, with or without merit, may result in an unfavorable judgment. We also may decide to settle lawsuits on unfavorable terms. Any such negative outcome could result in payments of
68
substantial damages or fines, damage to our reputation, or adverse changes to our business practices. Defending against litigation is costly and time-consuming, and could divert our management’s attention and our resources. Furthermore, during the course of litigation, there could be negative public announcements of the results of hearings, motions, or other interim proceedings or developments, which could have a negative effect on the market price of our common shares.
We have a substantial number of outstanding warrants. The exercise of our outstanding warrants will dilute existing stockholders and could adversely affect the trading price of our common stock.
While the holders of our outstanding pre-funded warrants and warrants are subject to beneficial ownership limitations, as of December 31, 2025, we had (i) outstanding pre-funded warrants to purchase, without regard to any beneficial ownership limitations, up to 16,412,925 shares of our common stock at an exercise price of $0.001 per share and (ii) outstanding warrants to purchase, without regard to any beneficial ownership limitations, up to 55,707,997 shares of common stock at a weighted average exercise price of $1.73 per share. The exercise of our outstanding warrants could result in significant dilution to existing stockholders, cause the trading price of our common stock to decline and impair our ability to raise capital through the sale of additional equity securities. Moreover, the expectation of such exercises could encourage the short selling of our common stock, which could place further downward pressure on the trading price of our common stock.
Unstable market and economic conditions, including as a result of recent bank closures, public health crises or geopolitical tensions may have serious adverse consequences on our business, financial condition, and share price.
The global economy, including credit and financial markets, has experienced extreme volatility and disruptions, including severely diminished liquidity and credit availability, declines in consumer confidence, declines in economic growth, increases in unemployment rates, increases in inflation rates, and uncertainty about economic stability. For example, the macroeconomic uncertainty and volatile business environment have resulted in ongoing inflation, volatility in the capital markets, significantly reduced liquidity and credit availability, elevated interest rates, decreases in consumer demand and confidence, declines in economic growth, increases in unemployment rates, and uncertainty about economic stability. There is inherent risk, based on the complex relationships among the United States and the countries in which we conduct our business, that political, diplomatic and national security factors could lead to global trade restrictions and changes in trade policies and regulations that may adversely affect our business and operations. Our general business strategy may be materially or adversely impacted if these unpredictable and unstable market conditions continue. Our general business strategy may be materially or adversely impacted if these unpredictable and unstable market conditions continue. Additionally, the global geopolitical conflicts have created extreme volatility in the global capital markets and are expected to have further global economic consequences, including potential disruptions of the global supply chain, manufacturing, and energy markets. Additionally, the introduction of or changes in tariffs or trade barriers, such as the enactment of tariffs on goods imported into the United States, could also increase our expenses. For example, in February 2026, the United States Supreme Court (SCOTUS) invalidated certain tariffs imposed by the U.S. government under emergency statutory authority in 2025. Shortly thereafter, President Trump signed an executive order implementing a new 10% global tariff pursuant to an alternative statutory authority, which may be raised up to 15%. It remains unclear whether and to what extent duties previously collected under the invalidated tariffs will be refunded, whether refunds will be subject to administrative or judicial processes, or whether offsets or alternative measures may be imposed. This evolving legal and policy landscape has contributed to continued volatility in the trade environment. Any such volatility and disruptions may have adverse consequences on us or the third parties on whom we rely. If the equity and credit markets deteriorate, including as a result of inflation expectations, recent bank closures, the changing interest rate environment, political unrest or war, it may make any necessary debt or equity financing more difficult to obtain in a timely manner or on favorable terms, more costly or more dilutive. Increased inflation rates can adversely affect us by increasing our costs, including labor and employee benefit costs. Any significant increases in inflation and related increases in interest rates could have a material adverse effect on our business, results of operations and financial condition.
69
Our common shares are thinly traded, and our shareholders may be unable to sell their shares quickly or at market price.
Although we have had periods of high-volume daily trading in our common shares, generally our shares are thinly traded. As a consequence of this lack of liquidity, the trading of relatively small quantities of shares by our shareholders may disproportionately influence the price of those shares in either direction. The price for our shares could, for example, decline significantly in the event that a large number of our common shares are sold on the market without commensurate demand, as compared to a seasoned issuer that could better absorb those sales without adverse impact on its share price.
Concentration of ownership of our common shares among our existing executive officers, directors, and principal shareholders may prevent new investors from influencing significant corporate decisions.
Based upon our common shares outstanding as of December 31, 2025, our executive officers, directors, and shareholders who owned more than 5% of our outstanding common shares, in the aggregate, beneficially owned shares representing 12.6% of our outstanding common shares. If our executive officers, directors, and shareholders who owned more than 5% of our outstanding common shares acted together, they may be able to significantly influence all matters requiring shareholder approval, including the election and removal of directors and approval of any merger, consolidation, or sale of all or substantially all of our assets. The concentration of voting power and transfer restrictions could delay or prevent an acquisition of our Company on terms that other shareholders may desire or result in the management of our Company in ways with which other shareholders disagree.
If research analysts do not publish research or reports, or publish unfavorable research or reports, about us, our business or our market, our share price and trading volume could decline.
The trading market for our common shares will be influenced by the research and reports that industry or financial analysts publish about us or our business. Equity research analysts may discontinue research coverage of our common shares, and such lack of research coverage may adversely affect the market price of our common shares. We do not have any control over the analysts or the content and opinions included in their reports. The price of our shares could decline if one or more equity research analysts downgrade our shares or issue other unfavorable commentary or research about us. If one or more equity research analysts cease coverage of us or fails to publish reports on us regularly, demand for our shares could decrease, which in turn could cause the trading price or trading volume of our common shares to decline.
Because we do not anticipate paying any cash dividends on our share capital in the foreseeable future, capital appreciation, if any, will be your sole source of gain.
You should not rely on an investment in our common shares to provide dividend income. We have never declared or paid cash dividends on our share capital. We currently intend to retain all of our future earnings, if any, to finance the growth and development of our business. In addition, the terms of any future debt agreements or preferred equity may preclude us from paying dividends. As a result, capital appreciation, if any, of our common shares will be your sole source of gain for the foreseeable future. Investors seeking cash dividends should not purchase our common shares.
We have broad discretion in the use of our cash and cash equivalents and may use them in ways in which you do not agree or in ways that do not increase the value of your investment.
Our management has broad discretion in the application of our cash and cash equivalents and could spend these funds in ways that do not improve our results of operations or enhance the value of our common shares. The failure by our management to apply these funds effectively could result in financial losses that could have a negative impact on our business, causing the price of our common shares to decline and delay the development of our product candidates. Pending their use, we may invest our cash and cash equivalents, in a manner that does not produce income or that loses value.
70
If we are a passive foreign investment company, there could be adverse U.S. federal income tax consequences to U.S. Holders (as defined below).
We have not yet made a determination with respect to our status as a passive foreign investment company, or “PFIC,” for our taxable year ending December 31, 2025. If we are a PFIC for the current taxable year, or any subsequent taxable years, we intend to annually furnish U.S. Holders, upon request, a “PFIC Annual Information Statement,” with the information required to allow U.S. Holders to make a “qualified electing fund” election, or “QEF Election,” for United States federal income tax purposes. No assurances regarding our PFIC status can be provided for any past, current, or future taxable years. The determination of whether we are a PFIC is a fact-intensive determination made on an annual basis and the applicable law is subject to varying interpretation. In particular, the characterization of our assets as active or passive may depend in part on our current and intended future business plans, which are subject to change. In addition, the total value of our assets for PFIC testing purposes may be determined in part by reference to the market price of our common shares from time to time, which may fluctuate considerably. As a result, our PFIC status may change from year to year. Further, our status as a PFIC depends on the composition of our income which will depend on the transactions we enter into in the future and our corporate structure. The composition of our income and assets is also affected by how, and how quickly, we spend the cash we raise in any offering.
If we are a PFIC, U.S. Holders (as defined below) may be subject to adverse U.S. federal income tax consequences, such as ineligibility for preferential tax rates for individuals on capital gains or on actual or deemed dividends, interest charges on certain taxes treated as deferred, and additional reporting requirements under U.S. federal income tax laws and regulations.
A “U.S. Holder” is a holder of our common shares who, for U.S. federal income tax purposes, is: (i) an individual who is a citizen or resident of the United States; (ii) a corporation, or another entity taxable as a corporation, created or organized in or under the laws of the United States, any state therein or the District of Columbia; (iii) an estate the income of which is subject to U.S. federal income taxation regardless of its source; or (iv) a trust if (1) a U.S. court is able to exercise primary supervision over the administration of the trust and one or more U.S. persons have authority to control all substantial decisions of the trust or (2) the trust has a valid election to be treated as a U.S. person under applicable U.S. Treasury Regulations.
71
Future changes to tax laws could materially adversely affect our Company and reduce net returns to our shareholders.
Our tax treatment is subject to the enactment of, or changes in, tax laws, regulations, and treaties, or the interpretation thereof, tax policy initiatives and reforms under consideration and the practices of tax authorities in jurisdictions in which we operate, including those related to the Organization for Economic Co-Operation and Development, or “OECD,” the Base Erosion and Profit Shifting, or “BEPS,” Project, the European Commission’s state aid investigations, and other initiatives. Such changes may include (but are not limited to) the taxation of operating income, investment income, dividends received, or (in the specific context of withholding tax) dividends paid. We are unable to predict what tax reform may be proposed or enacted in the future or what effect such changes would have on our business, but such changes, to the extent they are brought into tax legislation, regulations, policies, or practices, could affect our financial position and overall or effective tax rates in the future in countries where we have operations, reduce post-tax returns to our shareholders, and increase the complexity, burden, and cost of tax compliance.
For example, legislation commonly referred to as the OBBBA, enacted in 2025, the Inflation Reduction Act enacted in 2022, the Coronavirus Aid, Relief, and Economic Security Act enacted in 2020, and the Tax Cuts and Jobs Act enacted in 2017 made many significant changes to the U.S. Internal Revenue Code of 1986, as amended, or the “Code.” The Tax Cuts and Jobs Act required taxpayers to capitalize and amortize U.S.-based and non-U.S.-based research and experimental, or “R&E,” expenditures over five and fifteen years, respectively. The OBBBA restored the deductibility of domestic R&E expenditures in the year incurred for tax years beginning after December 31, 2024, but retained the capitalization and amortization requirement for foreign R&E expenditures. Future guidance from the Internal Revenue Service and other tax authorities with respect to any legislation may affect us, and certain aspects of such legislation could be repealed or modified in future legislation or sunset in future years. It is possible that changes in or interpretations under the OBBBA, the Tax Cuts and Jobs Act, or other tax legislation, or the enactment of new tax legislation, could increase our future tax liability, which could in turn adversely impact our business and future profitability.
We continue to evaluate the impact that these and other tax reforms may have on our business. The impact of these and other tax reforms is uncertain and one or more of these or similar measures could seriously harm our business. We urge you to consult with your legal and tax advisors with respect to these tax reforms and the potential tax consequences of investing in or holding our common shares.
Tax authorities may disagree with our positions and conclusions regarding certain tax positions, resulting in unanticipated costs, taxes, or non-realization of expected benefits.
A tax authority may disagree with tax positions that we have taken, which could result in increased tax liabilities. For example, the Canadian Revenue Agency, the IRS, or another tax authority could challenge our allocation of income by tax jurisdiction and the amounts paid between our affiliated companies pursuant to our intercompany arrangements and transfer pricing policies, including amounts paid with respect to our intellectual property development. Similarly, a tax authority could assert that we are subject to tax in a jurisdiction where we believe we have not established a taxable connection, often referred to as a “permanent establishment” under international tax treaties, and such an assertion, if successful, could increase our expected tax liability in one or more jurisdictions. A tax authority may take the position that material income tax liabilities, interest and penalties are payable by us, in which case, we expect that we might contest such assessment. Contesting such an assessment may be lengthy and costly and if we were unsuccessful in disputing the assessment, the result could increase our anticipated effective tax rate.
We are a “smaller reporting company” and, because we have opted to use the reduced reporting requirements available to us, certain investors may find investing in our securities less attractive.
We are a “smaller reporting company” under the SEC’s disclosure rules, meaning that we have either: (i) a public float of less than $250 million; or (ii) annual revenues of less than $100 million during the most recently completed fiscal year; and no public float; or a public float of less than $700 million.
As a smaller reporting company, we are permitted to comply with scaled-back disclosure obligations in our SEC filings compared to other issuers, including with respect to disclosure obligations regarding executive compensation in our periodic reports and proxy statements. We have elected to adopt the accommodations available to smaller reporting
72
companies. Until we cease to be a smaller reporting company, the scaled-back disclosure in our SEC filings will result in less information about our company being available than for other public companies. If investors consider our common shares less attractive as a result of our election to use the scaled-back disclosure permitted for smaller reporting companies, there may be a less active trading market for our common shares, and our share price may be more volatile.
We are also a non-accelerated filer under the Exchange Act, and we are not required to comply with the auditor attestation requirements of Section 404(b) of the Sarbanes-Oxley Act of 2002. Therefore, our internal controls over financial reporting will not receive auditor attestation included in annual reports of issuers that are subject to the auditor attestation requirements. In addition, we cannot predict if investors will find our common shares less attractive because we are not required to comply with the auditor attestation requirements. We cannot predict if investors will find our securities less attractive because we rely on these available exemptions. If some investors find our securities less attractive as a result, there may be a less active trading market for our securities, and the market price of our securities may be more volatile.
We are incurring, and expect to continue to incur, additional costs as a result of operating as a public company, and our management will be required to devote substantial time to new compliance initiatives.
As a public company, we are incurring, and expect to continue to incur, significant legal, accounting, and other expenses. In addition, the Sarbanes-Oxley Act of 2002, or the “Sarbanes-Oxley” Act, and rules subsequently implemented by the SEC and The Nasdaq Stock Market LLC have imposed various requirements on public companies, including establishment and maintenance of effective disclosure and financial controls and corporate governance practices. Our management and other personnel need to devote a substantial amount of time to these compliance initiatives. Moreover, these rules and regulations have increased, and will continue to increase, our legal and financial compliance costs and make some activities more time-consuming and costly.
As a non-accelerated filer and smaller reporting company, we are exempt from Section 404(b) of the Sarbanes-Oxley Act, which requires auditor attestation to the effectiveness of internal control over financial reporting. We are, however, subject to Section 404(a) of the Sarbanes-Oxley Act.
Additionally, pursuant to Section 404 of the Sarbanes-Oxley Act, we may be required to furnish an attestation on internal control over financial reporting issued by our independent registered public accounting firm in the future. To achieve compliance with Section 404(b), we will be engaged in additional internal processes to document and evaluate our internal control over financial reporting, which will be both costly and challenging. In this regard, we will need to continue to dedicate internal resources, potentially engage outside consultants, and adopt a detailed work plan to assess and document the adequacy of internal control over financial reporting, continue steps to improve control processes as appropriate, validate through testing that controls are functioning as documented, and implement a continuous reporting and improvement process for internal control over financial reporting. Despite our efforts, our independent registered public accounting firm may determine we have a material weakness or significant deficiency in our internal control over financial reporting once such firm begin its Section 404(b) audit in the future, there is a risk that neither we nor our independent registered public accounting firm will be able to conclude within the prescribed timeframe that our internal control over financial reporting is effective as required by Section 404(b). This could result in an adverse reaction in the financial markets due to a loss of confidence in the reliability of our consolidated financial statements.
Because we are a Canadian company, it may be difficult to serve legal process or enforce judgments against us.
We are a domestic filer in the United States; however, we are incorporated and have our corporate headquarters in Canada. In addition, while many of our directors and officers reside in the United States, several of them reside outside of the United States. Accordingly, service of process upon us may be difficult to obtain within the United States. Furthermore, because substantially all of our assets are located outside the United States, any judgment obtained in the United States against us, including one predicated on the civil liability provisions of the U.S. federal securities laws, may not be collectible within the United States. Therefore, it may not be possible to enforce those actions against us.
In addition, it may be difficult to assert U.S. securities law claims in original actions instituted in Canada. Canadian courts may refuse to hear a claim based on an alleged violation of U.S. securities laws against us or these persons on the grounds
73
that Canada is not the most appropriate forum in which to bring such a claim. Even if a Canadian court agrees to hear a claim, it may determine that Canadian law and not U.S. law is applicable to the claim. If U.S. law is found to be applicable, the content of applicable U.S. law must be proved as a fact, which can be a time-consuming and costly process. Certain matters of procedure will also be governed by Canadian law. Furthermore, it may not be possible to subject foreign persons or entities to the jurisdiction of the courts in Canada. Similarly, to the extent that our assets are located in Canada, investors may have difficulty collecting from us any judgments obtained in the U.S. courts and predicated on the civil liability provisions of U.S. securities provisions.
We are governed by the corporate laws of Québec, Canada which in some cases have a different effect on shareholders than the corporate laws of Delaware.
We are governed by the Business Corporations Act (Québec), or the “QBCA,” and other relevant laws, which may affect the rights of shareholders differently than those of a company governed by the laws of a U.S. jurisdiction, and may, together with our charter documents, have the effect of delaying, deferring or discouraging another party from acquiring control of us by means of a tender offer, a proxy contest or otherwise, or may affect the price an acquiring party would be willing to offer in such an instance. The material differences between the QBCA and Delaware General Corporation Law, or the “DGCL”, that may have the greatest such effect include but are not limited to the following: (i) for material corporate transactions (such as mergers and amalgamations, other extraordinary corporate transactions or amendments to our articles), the QBCA generally requires a two-thirds majority vote by shareholders, whereas the DGCL generally only requires a majority vote; and (ii) under the QBCA, a holder of 10% or more of our common shares can requisition a special meeting of shareholders, whereas such right does not exist under the DGCL.
Our bylaws and certain Canadian legislation contain provisions that may have the effect of delaying or preventing certain change in control transactions or shareholder proposals.
Certain provisions of our bylaws and certain Canadian legislation, together or separately, could discourage or delay certain change in control transactions or shareholder proposals.
Our bylaws contain provisions that establish certain advance notice procedures for nomination of candidates for election as directors at shareholders’ meetings. The BCA requires that any shareholder proposal that includes nominations for the election of directors must be signed by one or more holders of shares representing in the aggregate not less than 5% of the shares or 5% of the shares of a class or series of shares of the corporation entitled to vote at the meeting to which the proposal is to be presented.
The Investment Canada Act (Canada) requires that a non-Canadian must file an application for review with the Minister responsible for such statute and obtain approval of the Minister prior to acquiring control of a “Canadian business” within the meaning of such statute, where prescribed financial thresholds are exceeded. Furthermore, limitations on the ability to acquire and hold our common shares may be imposed by the Competition Act (Canada). This legislation permits the Commissioner of Competition, or Commissioner, to review any acquisition or establishment, directly or indirectly, including through the acquisition of shares, of control over or of a significant interest in our Company. Otherwise, there are no limitations either under the laws of Canada or Quebec, or in our articles on the rights of non-Canadians to hold or vote our common shares.
Any of these provisions may discourage a potential acquirer from proposing or completing a transaction that may have otherwise presented a premium to our shareholders.
ITEM 1B. UNRESOLVED STAFF COMMENTS.
None.
74
ITEM 1C. CYBERSECURITY.
We have implemented and maintain various information security processes designed to identify, assess and manage material risks from cybersecurity threats to our critical computer networks, communications systems, hardware and software, and our critical data, including intellectual property and confidential information that is proprietary, strategic or competitive in nature, or “Information Systems and Data.”.
Our Vice President of Information Technology, or “VP of IT,” with assistance from our third-party managed services team, legal, quality, finance and human resources, identifies, assesses and manages the Company’s cybersecurity threats and risks. The Company identifies and assesses risks from cybersecurity threats by monitoring and evaluating our threat environment using various methods including, for example using manual and automated tools, analyzing reports of threat actors, conducting scans of the threat environment, evaluating threats reported to us, and coordinating with law enforcement concerning threats.
Depending on the environment, we implement and maintain various technical, physical, and organizational measures, processes, standards, and policies designed to manage and mitigate material risks from cybersecurity threats to our Information Systems and Data, including, for example: an incident response plan; incident detection and response; disaster recovery and business continuity plans; implementation of security standards; network security controls; access controls; system monitoring; and employee training.
Our assessment and management of material risks from cybersecurity threats are
We use
For a description of the risks from cybersecurity threats that may materially affect the Company and how they may do so, see our risk factors under Part I. Item 1A. Risk Factors in this Annual Report on Form 10-K, including “Our internal computer systems, or those of our collaborators, contractors, consultants, or other third parties with whom we work, may fail or suffer security breaches, which could result in a significant disruption of our product development programs and can lead to adverse consequences, including but not limited to regulatory investigations or actions; litigation; fines and penalties; reputational harm; loss of revenue or profits; loss of customers or sales; disruption of our business operations.”
Governance
Our board of directors and audit committee addresses the Company’s cybersecurity risk management as part of its general oversight function.
75
Our cybersecurity risk assessment and management processes are implemented and maintained by certain Company management, including our Chief Financial Officer and our
The audit committee receives periodic reports concerning the Company’s significant cybersecurity threats and risk and the processes the Company has implemented to address them. The audit committee also receives various reports, summaries or presentations related to cybersecurity threats, risk and mitigation.
76
ITEM 2. PROPERTIES.
Our headquarters is currently located in Montréal (Québec), Canada and consists of 7,700 square feet of leased office space under a lease that expires in November 2027. We also have a U.S. subsidiary based in Charlotte, North Carolina. We believe that our facilities are adequate to meet our current needs, and that additional space can be obtained on commercially reasonable terms as needed.
ITEM 3. LEGAL PROCEEDINGS.
From time to time, we may become involved in legal proceedings arising in the ordinary course of our business. We are not currently a party to any material legal proceedings, and we are not aware of any pending or threatened legal proceeding against us that we believe could have an adverse effect on our business, operating results or financial condition.
ITEM 4. MINE SAFETY DISCLOSURES.
Not applicable.
77
PART II
ITEM 5. MARKET FOR REGISTRANT’S COMMON EQUITY, RELATED STOCKHOLDER MATTERS AND ISSUER PURCHASES OF EQUITY SECURITIES.
MARKET INFORMATION
Our common shares began trading on The Nasdaq Global Select Market on May 9, 2019. Our common shares trade under the symbol “MIST”. Prior to the commencement of trading on the Nasdaq Global Select Market on May 9, 2019, there was no public market for our common shares.
HOLDERS OF RECORD
As of December 31, 2025, there were 21 holders of record of our common shares, including Cede & Co., a nominee for The Depository Trust Company, or “DTC,” which holds our common shares on behalf of an indeterminate number of beneficial owners. All of the common shares held by brokerage firms, banks, and other financial institutions as nominees for beneficial owners are deposited into participant accounts at DTC, and are considered to be held of record by Cede & Co. as one shareholder. Because many of our shares are held by brokers and other institutions on behalf of shareholders, we are unable to estimate the total number of shareholders represented by these record holders.
DIVIDEND POLICY
We have never declared or paid any cash dividends on our capital stock and do not anticipate paying any cash dividends in the foreseeable future. Payment of cash dividends, if any, in the future will be at the discretion of our board of directors and will depend on then-existing conditions, including our financial condition, operating results, contractual restrictions, capital requirements, business prospects, and other factors our board of directors may deem relevant.
Recent Sales of Unregistered Securities
None.
Purchase of Equity Securities by the Issuer and Affiliated Purchasers
None.
Securities Authorized for Issuance Under Equity Compensation Plans
Information about securities authorized for issuance under our equity compensation plan is incorporated herein by reference to Item 12 of Part III of this Annual Report on Form 10-K.
ITEM 6. SELECTED FINANCIAL DATA.
Not Applicable.
78
ITEM 7. MANAGEMENT’S DISCUSSION AND ANALYSIS OF FINANCIAL CONDITION AND RESULTS OF OPERATIONS.
You should read the following discussion and analysis of our financial condition and results of operations together with our consolidated financial statements and related notes included elsewhere in this Annual Report on Form 10-K. This discussion contains forward-looking statements based upon current expectations that involve risks and uncertainties. Our actual results may differ materially from those anticipated in these forward-looking statements as a result of various factors, including those discussed in “Risk Factors” and in other parts of this Annual Report on Form 10-K.
Company Overview
We are a biopharmaceutical company focused on the development and commercialization of innovative cardiovascular medicines. Our approved product CARDAMYSTTM (etripamil) nasal spray is available in the United States and is the first and only U.S. Food and Drug Administration, or the “FDA,” approved self-administered treatment for use by patients anywhere, anytime an attack of paroxysmal supraventricular tachycardia, or “PSVT,” occurs. We continue to seek, either directly or through collaboration with our partners, marketing approval from regulatory agencies responsible for regions outside the United States. We are also developing etripamil for the indication of atrial fibrillation with rapid ventricular rate, or “AFib-RVR”.
We are currently focusing our efforts and financial resources on (i) the commercialization of CARDAMYSTTM (etripamil) nasal spray for the treatment of PSVT, (ii) the development of etripamil for the treatment of AFib-RVR and (iii) corporate development activities that have the potential to increase company value through strategic collaborations.
PSVT Market Overview
On December 12, 2025, we announced the FDA approved our first commercial product, CARDAMYST™ (etripamil) nasal spray, a prescription medication for the conversion of acute symptomatic episodes of PSVT to sinus rhythm in adults. We have currently begun focusing on the commercialization of CARDAMYST, which became available in retail pharmacies in the first quarter of 2026. PSVT is a condition that causes a patient’s heart to suddenly start beating faster than normal. It can be life-altering as PSVT is highly symptomatic, characterized by unpredictable attacks of a racing heart, often exceeding 150 beats per minute. Symptoms of PSVT arise suddenly and may include palpitations, sweating, chest pressure or pain, shortness of breath, sudden onset of fatigue, lightheadedness or dizziness, fainting, and anxiety, causing many patients to interrupt their daily activities at the time of symptom-onset. The impact and morbidity from an episode of PSVT can be especially detrimental in patients with underlying cardiovascular or medical conditions, such as heart failure, obstructive coronary disease, or dehydration. The uncertainty of when such an attack of PSVT will strike or how long it will persist is often anxiety-provoking, reducing patients’ quality of life and preventing participation in many desired activities. Drugs approved for the treatment of attacks of PSVT include adenosine, verapamil, and diltiazem, with all being administered intravenously under medical supervision, usually in the emergency department. Other oral drugs are sometimes used to treat attacks in a concept called “pill in the pocket.” However, those drugs have never been proven effective or safe and are not approved for this use. Doctors are often frustrated by the limited effective treatment options with the only approved options involving prolonged, unpleasant, and costly trips to the emergency department or, for some patients, an invasive ablation procedure. PSVT can be traumatic for patients, frustrating for healthcare providers, and costly for payors. With no pharmaceutical innovation in the treatment of PSVT for more than 30 years and a movement in the healthcare system to enable patient-centered care, we believe there is an opportunity to help patients living with PSVT to take greater control over their PSVT.
We believe that PSVT is a large and under-recognized market which we estimate affects more than two million Americans. From this diagnosed population, we define the immediate target addressable market for CARDAMYST as approximately 50% of patients with PSVT who have sufficient disease burden that they are compelled to seek care for their condition and are primarily managed by approximately 40,000 healthcare providers composed primarily of clinical cardiologists, interventional cardiologists and electrophysiologists. The remaining patients with PSVT can become addressable over time, as they are inconsistently managed (cycling in and out of the healthcare system). Furthermore, PSVT is expected to
79
increase in diagnosed prevalence in coming years as wearable electrocardiogram, or “ECG,” technology (e.g., smartphone, watches) becomes both more adept at diagnosing PSVT and more widely used by patients and clinical practitioners, in turn shortening the time to diagnosis.
Following the release of data from the RAPID clinical study, in market research, cardiologists reported a willingness to prescribe CARDAMYST to approximately 50% of the patients with PSVT in their care, which suggests approximately 500,000 to 800,000 patients can potentially be treated with CARDAMYST in peak years. Additionally, we believe that this cardiology-identified group of patients may use CARDAMYST to treat a median of three to five episodes per year, based on the projected number of self-reported longer and more intense episodes experienced by patients, as well as the patient utilization experience in our Phase 3 clinical trials. This implies a peak demand potential in the United States for CARDAMYST of 2.5 million to 4 million episodes treated per year.
Current treatment for PSVT also consumes significant healthcare resources. Research published in the American Journal of Cardiology in 2020 shows that total healthcare expenditures in the year following a diagnosis of PSVT ranged from $20,000 to $30,000 per patient which were significantly higher than the expenditures observed for patients without PSVT. These significant increases included increased emergency department visits and hospitalization costs. Of note, catheter ablations following diagnosis represented only 23% of this increased spend, meaning most costs were unrelated to ablations. Recent data from the Healthcare Cost and Utilization Project, or “HCUP,” database indicate that in 2019 there were approximately 140,000 emergency department, or “ED,” visits for PSVT when coded (for billing) in the primary diagnostic position, and a total of approximately 525,000 ED visits when PSVT was coded in any diagnostic position. Of these, approximately 25% of ED admissions for PSVT resulted in a hospital admission. HCUP estimates a total of approximately 40,000 to approximately 120,000 inpatient admissions for PSVT in 2019 (based again if a PSVT billing code was found in the primary versus any diagnostic position). Despite the effectiveness of catheter ablation, claims data suggests that only approximately 15% of patients with PSVT are ablated over a three-year period, leading to a total of approximately 100,000 catheter ablations annually. In total, up to $15.0 billion is spent annually in the United States on the management of PSVT.
PSVT Continued Development Highlights
In November 2025, we submitted our marketing authorization application, or “MAA,” to the European Medicines Agency, or “EMA,” for review for approval to market etripamil nasal spray, under the trade name TACHYMISTTM, in the European Union. The MAA utilizes much of the clinical, manufacturing and quality data package that was submitted in the NDA which lead to the US FDA approval.
In January 2025, our licensing partner, Corxel Pharmaceuticals, or “Corxel,” formerly Ji Xing Pharmaceuticals Limited, JIXING, announced that the Center for Drug Evaluation, or “CDE,” of the National Medical Products Administration, or “NMPA,” of the People’s Republic of China has accepted the New Drug Application, or “NDA,” for etripamil nasal spray for the treatment of PSVT. The NDA to the NMPA included data from the successful Phase 3 JX02002 clinical trial of etripamil nasal spray in patients with PSVT in China in addition to data included in the NDA that Milestone submitted to the FDA.
The 500-patient Phase 3 trial (JX02002) met its primary endpoint, with a Kaplan Meier analysis shows a statistically significantly greater proportion of patients who self-administered etripamil converted from PSVT to sinus rhythm within 30 minutes compared to placebo (40.5% vs. 15.9%, respectively; hazard ratio [HR] = 3.00; 95% CI 1.58-5.71; p<0.001). Statistically significant (p<0.05) results were also shown for the secondary efficacy endpoints for percent of patients’ PSVT converted to sinus rhythm by 10, 15, 45 and 60 minutes after self-administration of study drug.
Corxel further reported that, overall, treatment emergent adverse events were comparable between treatment groups, and there were no reported serious adverse events related to etripamil. The safety and tolerability data from the JX02002 trial were consistent with previous clinical studies. This important study further expands the etripamil global development program to more than 2,000 unique patients treated with etripamil.
80
Etripamil Nasal Spray for the Treatment of AFib-RVR
Similar to our approach for PSVT, we believe that etripamil has the potential to help people experiencing a symptomatic episode of AFib-RVR to self-treat and to conveniently, reliably, and quickly reduce their elevated heart rate, with the goal of reducing the need for emergency department utilization. We completed a successful Phase 2 study, named “ReVeRA” or the “ReVeRA study”, in patients presenting urgently with episodes of AFib-RVR to the emergency department. We have published the ReVeRA study and results, which demonstrated that patients receiving etripamil nasal spray experienced rapid and statistically superior ventricular rate reduction and improved symptom-relief compared to placebo, with safety and tolerability findings generally consistent with those observed in our PSVT program. We believe these data support the continued development of etripamil, self-administered in the medically unmonitored setting, for the treatment of AFib-RVR.
Incorporating FDA’s guidance, we have developed a Phase 3 registrational program to evaluate self-administered etripamil as a potential treatment for patients with AFib-RVR. We intend to pursue a supplemental new drug application, or “sNDA,” regulatory approval pathway for a potential second indication for etripamil in Afib-RVR. As such, we will leverage the PSVT indication and PSVT program data, along with a single pivotal Phase 3 study in patients with AFib-RVR.
AFib-RVR Market Overview
Atrial Fibrillation, or “AFib,” is a common cardiac arrhythmia with an irregular and often rapid heart rate that is often markedly symptomatic and, without proper treatment, can increase the risk of stroke, heart failure, and other cardiovascular complications. A common complication of AFib is a rapid heart rate, also referred to as AFib-RVR, which is frequently defined as a heart rate ≥ 110 beats per minute. The occurrence of a rapid ventricular rate, or “RVR,” in patients with atrial fibrillation increases the likelihood of marked symptoms including heart palpitations, shortness of breath and weakness. There are two commonly used pharmacological approaches to chronically manage AFib, rhythm control and rate control. Regardless of the chronic approach, break-through episodes of rapid heart rate occur frequently; and when faced with a sudden episode of AFib-RVR, acute rate control is needed, with most treatments being AV-nodal targeted drugs such as a beta blocker or calcium channel blocker. These treatments can be given intravenously; however, this requires a burdensome trip to an emergency department which may lead to a hospital admission. Acute treatment can be attempted with an oral rate-control drug; however, such agents often fail to provide immediate or dependable ventricular rate control because they have a 30- to 90-minute delayed onset of action. As a result, many patients require faster and more reliable rate reduction and symptom resolution, leading them to seek acute medical care in the emergency department for intravenous rate control and/or electrical cardioversion of their atrial fibrillation. Furthermore, the chronic administration of oral rate-control drugs does not broadly prevent episodes of AFib-RVR. Similar to PSVT, patients may feel a loss of control by needing to visit the emergency department for overcoming their AFib-RVR episode and the unpredictable nature of these episodes, which can occur anytime and anywhere. Doctors have expressed frustration at the lack of options for patients to self-manage these acute rate attacks; and payor organizations would prefer to treat the AFib-RVR attacks in a more cost effective and time-efficient manner.
An estimated 10 million Americans suffer from AFib. The prevalence of AFib is expected to grow to greater than 12 million by 2030. A subset of patients with AFib experiences episodes of abnormally high heart rate, most often accompanied by palpitations, shortness of breath, dizziness, and weakness. While these episodes, known as AFib-RVR, may be treated by oral calcium channel blockers and/or beta blockers, patients frequently seek acute care in the ED to address symptoms. In 2019, nearly 1.1 million patients were admitted to the emergency department due to AFib symptoms. Initial data suggests that approximately 60% of all AFib emergency department visits were attributable to AFib-RVR, as symptoms driving patients to seek care generally become more pronounced at higher heart rates. Treatment for such symptoms typically includes medically supervised intravenous administration of calcium channel blockers or beta blockers, or electrical cardioversion. With little available data for AFib-RVR, we believe, based on our initial market research, that 30% to 40% of patients with AFib experience one or more symptomatic episodes of RVR per year that require treatment, suggesting a current target addressable market of approximately three to four million patients for etripamil in patients with AFib-RVR.
81
We believe that etripamil has the potential to be developed such that it can be used by patients to rapidly reduce their heart rate to provide a supplemental option to either the acute oral rate or rhythm control strategy their physician would use. When presented with a target product profile reflecting this potential use case, cardiologists and electrophysiologists, in a 2021 market research study perceived utility in the product profile and indicated that they would prescribe to approximately two-thirds of their patients that experience episodes of AFib-RVR. They further indicated that a rapidly-acting intranasal calcium channel blocker could serve as a “bridge” to the longer onset times of acute oral agents. According to physicians, it can take hours for patients to feel an alleviation of symptoms using acute oral rate or rhythm control. During this time, patients may experience concerning symptoms that often prompt them to seek emergency care. We believe that the combination of convenient delivery, potency, rapid onset and short duration of action of etripamil has the potential to move the current treatment setting for some acute episodes of AFib out of the burdensome and costly emergency department.
Current AFib management consumes significant healthcare resources in the United States. The American Heart Association published a report in 2016 summarizing the current and projected cost burden of cardiovascular diseases in the United States. This report suggests atrial fibrillation resulted in $25 billion in direct medical costs in 2016 (approximately 7% of all cardiovascular diseases) and another $7 billion in indirect costs (i.e., up to $32 billion in total costs). Additionally, the forecasted growth in atrial fibrillation prevalence is anticipated to result in healthcare expenditures of $46 billion in direct costs and $10 billion in indirect costs in the United States by 2030.
AFib-RVR Clinical Development Highlights
In November 2023, we presented positive Phase 2 data from the ReVeRA study as a Featured Science Presentation at the American Heart Association Scientific Meetings (Philadelphia, PA) and as simultaneously published in Circulation: Arrhythmia and Electrophysiology. The randomized, double-blinded, placebo-controlled ReVeRA trial of etripamil nasal spray enrolled 87 patients and dosed 56 patients aged 18 years and older with AFib who urgently presented experiencing AFib and a ventricular rate of 110 or more beats per minute, or “bpm.” The trial was designed to assess the magnitude, rapidity, and duration of reduction and the patient satisfaction with treatment using an established patient reported outcome, or “PRO,” tool. Data showed that delivery of etripamil nasal spray (70 mg) significantly and rapidly reduced ventricular rate, in a pattern consistent with the drug’s pharmacologic profile. Etripamil achieved the primary endpoint with a high degree of statistical significance; patients experienced a ventricular rate reduction of 29.91 bpm (95% confidence interval: -40.31, -19.52; p<0.0001) in the etripamil arm relative to placebo. The absolute maximum reduction in rate in the etripamil arm was 34.97 bpm. Using the Treatment Satisfaction Questionnaire 9, or “TSQM-9,” PRO, compared to placebo, patients treated with etripamil demonstrated significant improvements in two satisfaction ratings: effectiveness (p<0.0001) and relief of symptoms (p=0.0002), with the degrees of improvement consistent with those customarily described as clinically meaningful. Treatment-emergent serious adverse events, or “TESAEs,” were rare and the most common (≥ 5%) adverse events were mild or moderate in intensity and included nasal discomfort, rhinorrhea, increased lacrimation, throat irritation and dizziness. Further trial details are below in this document.
During 2024, we met with the FDA on the ReVeRA study, during which the FDA confirmed its guidance from our Pre-IND meeting (2023) regarding the availability of a sNDA pathway for the marketing approval for etripamil for the indication of AFib-RVR. The sNDA pathway potentially permits a single pivotal efficacy study to be sufficient for filing for marketing approval if etripamil is already approved for PSVT. FDA further concurred with respect to key proposed study elements including powering, inclusion criteria, patient population, and statistical analyses, and offered clarification with respect to the endpoints to guide the design of the Phase 3 study. In our mid-2023 Pre-IND meeting, the FDA provided guidance that our primary endpoint can be the reduction of ventricular rate, and the primary analysis would be performed on the intent to treat, or “ITT,” population. In addition, the study would have to show statistical significance (p<0.05) on the key secondary endpoint of symptom relief as a patient benefit, also in the ITT population. The secondary endpoint could use a PRO measure, and various PROs were discussed with the FDA. We have finalized the Phase 3 study protocol following FDA’s review and obtained concurrence with the FDA to proceed.
82
The Phase 3 study will be conducted in a medically unmonitored setting (e.g., at-home) in a manner very similar to the conduct of our Phase 3 development program for PSVT. The Phase 3 AFib-RVR study will enroll patients with a history of symptomatic AFib episodes, and will use a self-administered, repeat-dose regimen of 70 mg per dose (the dose and dosing approach that was studied in the RAPID trial in patients with PSVT). The Phase 3 study’s target population will be patients with verified history of AFib-RVR, and the ITT population will be all patients self-administering the study drug for perceived AFib-RVR. The primary endpoint is the mean change from baseline ventricular rate to nadir ventricular rate for patients treated with etripamil versus placebo, as was studied in the ReVeRA trial in AFib-RVR. The key secondary endpoint will be based on a PRO of symptomatic improvement, discussed with the FDA, which is similar to the PRO questions utilized in our PSVT and AFib-RVR programs. The study has been powered and sized based upon approximately 150 events from 150 unique patients with a history of symptomatic episodes of AFib-RVR.
Operations Overview
Since the commencement of our operations in 2003, we have devoted substantially all of our resources to performing research and development activities in support of our product development efforts, hiring personnel, raising capital to support and expand such activities, providing general and administrative support for these operations and, more recently, preparing for commercialization. We have historically operated our business using a significant outsourcing model. As such, our team is currently composed of a relatively smaller core of employees who direct a significantly larger number of team members who are outsourced in the forms of vendors and consultants to enable execution of our operational plans. On December 12, 2025, we announced that the FDA approved our first commercial product, CARDAMYST™ (etripamil) nasal spray, a prescription medication for the conversion of acute symptomatic episodes of paroxysmal supraventricular tachycardia, or “PSVT,” to sinus rhythm in adults. As a result, we expect our future commercial expenses will increase as we invest in the infrastructure, personnel, and operational expenses required to commercialize CARDAMYST in the United States. We also expect inventory balances to increase as we capitalize costs related to the production of CARDAMYST for commercial sale in the United States. In addition, we expect to continue to incur significant research and development and general administrative expenses related to our operations as we advance etripamil nasal spray for the treatment AFib-RVR and continue to invest in the resources needed to support a developing commercialized public company.
Since inception, we have incurred significant operating losses. For the years ended December 31, 2025 and 2024, we recorded net losses of $63.1 million and $41.5 million, respectively. As of December 31, 2025, we had an accumulated deficit of $430.6 million. We expect to continue to incur significant losses for the foreseeable future. We anticipate that a substantial portion of our capital resources and efforts in the foreseeable future will be focused on commercialization of CARDAMYST and the development of an additional etripamil indication. We had $73.0 million of cash and cash equivalents and $32.9 million of short-term investments at December 31, 2025.
We expect to continue to incur significant expenses and increasing operating losses for the foreseeable future. Our net losses may fluctuate significantly from period to period, depending on the timing of our planned clinical trials and expenditures on other research and development activities. We expect our expenses will increase over time as we:
| • | continue our ongoing and planned development of etripamil, including future Phase 3 clinical trials for the treatment of AFib-RVR and potential Phase 4 clinical trials for treatment of PSVT; |
| • | seek, either directly or through collaboration with our partners, marketing approval for etripamil nasal spray for the treatment of PSVT from regulatory agencies responsible for regions outside the United States; |
| • | seek marketing approvals for etripamil for the treatment of AFib-RVR and other cardiovascular indications; |
| • | increase our sales, marketing, manufacturing, and distribution capability, either directly or indirectly through third parties; |
83
| • | build a portfolio of product candidates through development, or the acquisition of in-license of drugs, product candidates or technologies; |
| • | initiate preclinical studies and clinical trials for etripamil for any additional indications we may pursue, including the clinical trials for the treatment of atrial fibrillation and rapid ventricular rate as well as other areas of unmet medical need, and for any additional product candidates that we may pursue in the future; |
| • | maintain, protect, and expand our intellectual property portfolio; |
| • | hire additional clinical, regulatory, and scientific personnel; |
| • | add operational, financial, and management information systems and personnel, or other general & administrative personnel, including personnel to support our product development and potential expansion of our future commercialization efforts; and |
| • | incur additional legal, accounting, insurance and other expenses associated with operating as a public company. |
Recent Developments
As previously disclosed, on March 27, 2023, we entered into a Purchase and Sale Agreement, or the “Royalty Purchase Agreement,” with RTW Royalty I DAC, an affiliate of RTW Investments, LP, or “RTW,” pursuant to which RTW agreed to purchase, following U.S. Food and Drug Administration approval of etripamil for the treatment of PSVT (subject to certain conditions), at a purchase price of $75.0 million, the right to receive tiered quarterly royalty payments, or the “royalty interest,” on net product sales of CARDAMYST (etripamil) in the United States.
On January 12, 2026, we closed the sale of the royalty interest under the Royalty Purchase Agreement and received cash of $75.0 million from RTW.
The Macroeconomic Climate
Inflation rates may materially adversely affect our business and corresponding financial position and cash flows. Inflationary factors, changes to interest rates, and overhead costs may adversely affect our operating results. Interest and inflation rates also present a recent challenge impacting the U.S. economy and could make it more difficult for us to obtain traditional financing on acceptable terms, if at all, in the future. Additionally, geopolitical events, including war and terrorism, banking instabilities, ongoing changes to U.S. and international tariffs and other trade restrictions and trade barriers, renegotiation of international trade agreements, or further escalations of trade tensions, and other U.S. geopolitical issues affecting other territories and employee availability and wage increases, and economic markets all of which may result in additional stress on our working capital resources. The ongoing trade tensions between the U.S. and other jurisdictions have resulted in multiple rounds of tariffs and anticipated tariffs affecting pharmaceuticals and pharmaceutical ingredients, including finished drug products, manufacturing equipment, and related supplies. In any event, the dynamic and unpredictable tariff and trade landscape may create substantial uncertainty and planning challenges for our operations.
84
Components of Results of Operations
License revenue
We generated $1.5 million of license revenue for the year ended December 31, 2025, compared to no revenue generated for the year ended December 31, 2024. The current year license revenue was the result of having reached a milestone pursuant to our License and Collaboration Agreement, dated May 15, 2021, with Corxel Pharmaceuticals, or “Corxel,” formerly known as Ji Xing Pharmaceuticals Limited, such party being referred to as “Ji Xing” and, such agreement, as the “Ji Xing License Agreement,” due upon the successful NDA approval by the FDA in the U.S. of etripamil for the treatment of PSVT. For additional information about our Revenue, see Note 2, “Summary of Significant Accounting Policies”, and Note 3, “Revenue” in the accompanying notes to our consolidated financial statements.
Research and Development Expenses
Research and development expenses consist primarily of salaries and fees paid to external service providers, as well as personnel costs, including share-based compensation expense, and other related compensation expenses. We expense research and development costs in the periods in which they are incurred. Costs for certain development activities are recognized based on an evaluation of the progress to completion of specific tasks using information and data provided to us by our vendors, collaborators and third-party service providers.
To date, the majority of our research and development expenses have been related to the preclinical and clinical development of etripamil inclusive of manufacturing costs pre-FDA approval. As we advance etripamil or other product candidates for other indications, we expect to allocate our direct external research and development costs across each of the indications or product candidates. Further, we expect our research and development costs to increase for the development of etripamil in AFib-RVR, and we expect our research and development expenses related to the development of etripamil for PSVT to decrease as a percentage of our total research and development expenses.
The process of conducting the necessary clinical research to obtain regulatory approval is costly and time-consuming and is subject to uncertainties and delays. As a result of the uncertainties discussed above, we are unable to determine the duration and completion costs of our research and development projects or when and to what extent we will generate revenue from the commercialization and sale of our product candidates, if at all.
We recognize the benefit of Canadian research and development tax credits as a reduction of research and development costs for fully refundable investment tax credits.
General and Administrative Expenses
General and administrative expenses include personnel and related compensation costs, expenses for outside professional services, lease expenses, insurance expenses and other general administrative expenses. Personnel costs consist of salaries, bonuses, benefits, related payroll taxes, and share-based compensation. Outside professional services consist of legal, accounting, and audit services and other consulting fees.
We expect to continue to incur expenses as a public company, including expenses related to compliance with the rules and regulations of the Securities and Exchange Commission, or “SEC,” and those of any national securities exchange on which our securities are traded, additional insurance expenses, investor relations activities, and other administrative and professional services.
Commercial Expenses
Commercial expenses consist primarily of personnel and related compensation costs, market and health economic research, and market development activities for PSVT and, to a lesser extent, AFib-RVR. The focus of these expenses is three-fold: first, we want to leverage rigorous primary and secondary research to fully understand our target disease states from the perspective of the patient, healthcare provider, and payer; second, we want to understand and document the burden of
85
disease posed by PSVT and AFib-RVR from an epidemiology, healthcare resource use, and cost perspective; and third, we want to engage our target patient, physician, and payer stakeholders with evidence-based and compliant educational materials that serve to increase the awareness and understanding of the impact of PSVT and AFib-RVR on patients and the overall healthcare system.
We anticipate our commercial expenses will increase as we invest in the infrastructure, personnel, and operational expenses required to commercialize CARDAMYST in the United States.
Interest Income
Interest income primarily consists of interest income from our cash equivalents and short-term investments.
Interest Expense
Interest expense primarily consists of contractual debt interest expense and the amortization of debt costs.
Results of Operations
Comparison of the Years Ended December 31, 2025 and 2024
Year ended December 31, | |||||||||||
(in thousands) | | 2025 | | 2024 | $ Change | | % Change | ||||
Revenue | $ | 1,546 | $ | — | $ | 1,546 |
| 100.0% | |||
Operating expenses | |||||||||||
Research and development, net of tax credits | $ | 18,108 | $ | 14,357 | $ | 3,751 |
| 26.1% | |||
General and administrative |
| 17,274 |
| 16,742 |
| 532 |
| 3.2% | |||
Commercial |
| 28,304 |
| 11,003 | 17,301 |
| 157.2% | ||||
Total operating expenses |
| 63,686 |
| 42,102 |
| 21,584 |
| 51.3% | |||
Loss from operations |
| (62,140) |
| (42,102) |
| (20,038) |
| 47.6% | |||
Interest income |
| 2,920 |
| 4,164 |
| (1,244) |
| (29.9)% | |||
Interest expense | (3,838) | (3,581) | (257) | 7.2% | |||||||
Net loss | $ | (63,058) | $ | (41,519) | $ | (21,539) |
| 51.9% | |||
License revenue
We recorded $1.5 million of license revenue for the year ended December 31, 2025, compared to no revenue recorded for the year ended December 31, 2024. The license revenue for the year ended December 31, 2025 was the result of having reached a milestone pursuant to our License and Collaboration Agreement, dated May 15, 2021, with Corxel Pharmaceuticals, or “Corxel,” formerly known as Ji Xing Pharmaceuticals Limited, such party being referred to as “Ji Xing” and, such agreement, as the “Ji Xing License Agreement,” due upon the successful NDA approval by the FDA for CARDAMYST for the treatment of PSVT in the United States.
86
Research and Development Expenses
The following table shows our research and development expenses by type of activity for the periods indicated.
Year ended December 31, | |||||||||||
(in thousands) | | 2025 | | 2024 | $ Change | | % Change | ||||
Clinical | $ | 7,356 | $ | 6,596 | $ | 760 |
| 11.5% | |||
Drug manufacturing and formulation |
| 5,772 |
| 4,145 |
| 1,627 |
| 39.3% | |||
Regulatory and other costs |
| 5,325 |
| 3,875 |
| 1,450 |
| 37.4% | |||
Less: R&D tax credits |
| (345) |
| (259) |
| (86) |
| 33.2% | |||
Total R&D expenses | $ | 18,108 | $ | 14,357 | $ | 3,751 |
| 26.1% | |||
Research and development expenses increased by $3.8 million, or 26.1%, for the year ended December 31, 2025, compared to the year ended December 31, 2024. The increase was primarily due to higher consulting and outside service costs. These higher costs were partially offset by lower personnel-related costs.
General and Administrative
General and administrative expenses increased $0.5 million, or 3.2%, for the year ended December 31, 2025, compared to the year ended December 31, 2024. This increase was driven primarily by an increase in outside service costs and higher personnel costs.
Commercial
Commercial expenses increased by $17.3 million, or 157.2%, for the year ended December 31, 2025, compared to the year ended December 31, 2024. This increase is primarily a result of additional personnel costs, professional costs, and other operational expenses related to preparation for the launch of CARDAMYST.
Interest Income
Interest income was $2.9 million and $4.2 million for the year ended December 31, 2025 and 2024, respectively. The decrease in interest income in 2025 was driven decreasing interest rates and a smaller average investment balance in 2025 when compared to 2024.
Interest Expense
Interest expense remained materially consistent for the year ended December 31, 2025 compared to the year ended December 31, 2024.
Liquidity and Capital Resources
Sources of Liquidity
We have incurred operating losses and experienced negative operating cash flows since our inception, and we anticipate continuing to incur losses for the foreseeable future. As of December 31, 2025, we had cash, cash equivalents, and short-term investments of $106.0 million and an accumulated deficit of $430.6 million.
On July 11, 2025, we entered into an underwriting agreement, or the “Underwriting Agreement,” related to an underwritten public offering, or the “2025 Offering,” of (i) 31,500,000 of our common shares, accompanying Series A common warrants, or the “Series A Common Warrants,” to purchase an aggregate of 31,500,000 common shares and accompanying Series B common warrants, or the “Series B Common Warrants,” to purchase an aggregate of 31,500,000 common shares, at a combined public offering price of $1.50 per share and accompanying Series A Common Warrant and Series B Common Warrant and (ii) in lieu of common shares to certain investors that so choose, pre-funded warrants or the “2025
87
Pre-Funded Warrants” and, together with the Series A Common Warrants and the Series B Common Warrants, the “Warrants,” to purchase 3,502,335 common shares, accompanying Series A Common Warrants to purchase an aggregate of 3,502,335 common shares and accompanying Series B Common Warrants to purchase an aggregate of 3,502,335 common shares, at a combined public offering price of $1.499 per Pre-Funded Warrant and accompanying Series A Common Warrant and Series B Common Warrant, which represented the combined public offering price for the Common Shares and accompanying Common Warrants less the $0.001 per share exercise price for each such Pre-Funded Warrant.
The net proceeds to the Company from the 2025 Offering were $48.6 million after deducting underwriting commissions and other offering expenses payable by us, in the amount of $3.9 million.
On February 28, 2024, we entered into an underwriting agreement related to an underwritten public offering, or the “February 2024 Offering,” of 16,666,667 of our common shares, without par value, at a public offering price of $1.50 per share and, in lieu of common shares to certain investors, pre-funded warrants to purchase 3,333,333 common shares, or the “2024 Pre-Funded Warrants,” at a public offering price of $1.499 per 2024 Pre-Funded Warrants. Each of the 2024 Pre-Funded Warrants has an exercise price of $0.001 per share. The 2024 Pre-Funded Warrants were exercisable immediately upon issuance, subject to certain beneficial ownership limitations. Under the terms of the agreement, we granted the underwriters party thereto an option to purchase up to an additional 3,000,000 common shares at the same price per share as the other common shares sold in this offering, which was exercised by such underwriters in full on February 29, 2024.
The net proceeds to us from the February 2024 Offering, including the proceeds from the exercise by the underwriters of their option to purchase the additional 3,000,000 common shares in full, was $31.9 million after deducting underwriting commissions and offering expenses payable by us.
On July 29, 2020, we entered into an Open Market Sale AgreementSM, or the “Original Sale Agreement,” with respect to an at-the-market offering program, or the “ATM Program,” under which we could issue and sell our common shares having an aggregate offering price of up to $50.0 million through Jefferies LLC, or “Jefferies,” as sales agent or principal. On March 18, 2025, we entered into an Amended and Restated Open Market Sale AgreementSM, or the “Amended Agreement,” with Jefferies. The Amended Agreement amends and restates, in its entirety, the Original Sale Agreement. Under the Amended Agreement, we may issue and sell its common shares, no par value per share, for an aggregate offering price of up to $77.8 million (which includes the $2.8 million of sales previously made pursuant to the Original Sale Agreement through the date the Amended Agreement was entered into), or the “ATM Shares.” The ATM Shares will be sold pursuant to our shelf registration statement on Form S-3 (File No. 333-283162). We previously issued 361,236 common shares under the Original Sale Agreement, resulting in net proceeds of approximately $2.6 million (net of issuance costs of approximately $0.1 million). We issued 5,690,920 shares under the amended agreement resulting in net proceeds of $14.5 million, after deducting sales agent commissions payable by us of $0.4 million, during the year ended December 31, 2025.
We expect that our operating plan, existing cash and cash equivalents and short-term investments to be sufficient to fund our operations for at least the next 12 months from the date of issuance of this Annual Report on Form 10-K for the year ending December 31, 2025 and that there are no known events or conditions that may cast substantial doubt on our ability to continue as a going concern for at least the next 12 months from the date of this filing.
Royalty Purchase Agreement
Pursuant to the Royalty Purchase Agreement, RTW agreed to purchase, following FDA approval of etripamil (subject to certain conditions), in exchange for a purchase price of $75.0 million, the right to receive a tiered quarterly royalty payments, or “royalty interest,” on the annual net product sales of etripamil in the United States.
On January 12, 2026, we closed the sale of the royalty interest under the Royalty Purchase Agreement and received the $75.0 million purchase price from certain funds managed by RTW in exchange for the future royalty interest.
88
Funding Requirements
We anticipate that we will use our cash and cash equivalents primarily to fund commercialization and research and development expenditures. We expect our expenses to increase as we continue the development of etripamil and invest in the infrastructure, personnel, and operational expenses required to commercialize CARDAMYST in the United States. We expect to incur increasing operating losses for the foreseeable future as we continue the clinical development of subsequent etripamil indications or any future product candidates. At this time, due to the inherently unpredictable nature of clinical development, we cannot reasonably estimate the costs we will incur and the timelines that will be required to complete development, obtain marketing approval, and commercialize subsequent etripamil indications or any future product candidates, if at all. For the same reasons, we are also unable to predict when, if ever, we may achieve profitability generated from product sales. Clinical and preclinical development timelines, the probability of success, and development costs can differ materially from expectations.
In addition, we have exclusive development and commercialization rights for etripamil for all indications that we may pursue, and as such, have the potential to license development and or commercialization rights for etripamil to a potential partner in regions outside of Greater China. We have established commercialization and marketing capabilities using a direct sales force to commercialize CARDAMYST in the United States. Outside of the United States, we are considering commercialization strategies that may include collaborations with other companies.
For other new product candidates, our efforts are focused on licensing development and/or commercialization rights from potential partners. In the case of either in-licensing or out-licensing, we cannot forecast when such arrangements will be secured, if at all, and to what degree such arrangements would affect our development and commercialization plans and capital requirements.
The timing and amount of our operating expenditures will depend largely on:
| • | the timing, progress and results of our ongoing and planned clinical trials and other development activities of etripamil in AFib-RVR and in other cardiovascular indications; |
| • | the scope, progress, results, and costs of preclinical development, laboratory testing, and clinical trials of etripamil for additional indications or any future product candidates that we may pursue; |
| • | our ability to establish additional collaborations on favorable terms, if at all; |
| • | the ability of vendors and third-party service providers to accurately forecast expenses and deliver on expectations; |
| • | the costs, timing, and outcome of regulatory review of subsequent etripamil indications and any future product candidates; |
| • | the costs and timing of CARDAMYST commercialization activities, including product manufacturing, marketing, sales, and distribution for CARDAMYST and any future product candidates for which we receive marketing approval; |
| • | the costs and timing of preparing, filing and prosecuting patent applications, maintaining and enforcing our intellectual property rights and defending any intellectual property-related claims; and |
| • | the extent to which we acquire or in-license other product candidates and technologies. |
Until such time, if ever, as we can generate substantial revenue from product sales, we expect to fund our operations and capital funding needs through equity and/or debt financing. We may also consider entering into collaboration arrangements or selectively partnering for clinical development and commercialization. The sale of additional equity would result in additional dilution to our shareholders. The incurrence of debt financing would result in debt service obligations and the
89
instruments governing such debt could provide for operating and financing covenants that restrict our operations or our ability to incur additional indebtedness or pay dividends, among other items. If we are not able to secure adequate additional funding, we may be forced to make reductions in spending, extend payment terms with suppliers, liquidate assets where possible, and/or suspend or curtail planned programs. Any of these actions could materially and adversely affect our business, financial condition and results of operations.
Cash Flows
The following table summarizes our cash flows for the periods indicated:
Year ended December 31, | |||||||||||
(in thousands) | 2025 | | 2024 | $ Change | % Change | ||||||
Net cash (used in) provided by: | |||||||||||
Operating activities | $ | (49,041) | $ | (28,848) | $ | (20,193) |
| 70.0% | |||
Investing activities | 11,246 | 8,283 | 2,963 |
| 35.8% | ||||||
Financing activities | 85,527 | 32,119 | 53,408 |
| 166.3% | ||||||
Net increase in cash and cash equivalents during the period | $ | 47,732 |
| $ | 11,554 | $ | 36,178 |
| |||
Operating Activities
Net cash used in operating activities during the year ended December 31, 2025, was $49.0 million, which consisted primarily of a net loss of $63.1 million. The net loss was partially offset by a net cash increase of $2.7 million related to the change in assets and liabilities, non-cash charges of $7.5 million related to share-based compensation and non-cash interest and debt charges of $3.8 million related to the 2029 Convertible Notes.
Net cash used in operating activities during the year ended December 31, 2024, was $28.8 million, which consisted primarily of a net loss of $41.5 million. The net loss was partially offset by a net cash increase of $3.7 million related to the change in assets and liabilities, non-cash charges of $5.8 million related to share-based compensation and non-cash interest charges of $3.2 million related to the 2029 Convertible Notes.
Investing Activities
During the year ended December 31, 2025, we redeemed $94.5 million of short-term investments, and we acquired $82.9 million of short-term investments. In addition, we acquired $0.3 million in property and equipment.
During the year ended December 31, 2024, we redeemed $121.9 million of short-term investments, and we acquired $113.6 million of short-term investments. In addition, we acquired $0.1 million in property and equipment.
Financing Activities
During the year ended December 31, 2025, our financing activities provided cash proceeds of $85.5 million. These proceeds were primarily a result of $48.6 million received, net of issuance costs, from the issuance of common shares, pre-funded warrants, and common warrants under the 2025 Offering, proceeds of $21.7 million, net of issuance costs, for the exercise of common warrants, and proceeds of $14.5 million, net of issuance costs, for shares sold under the ATM Program. We also had proceeds of $0.4 million and $0.3 million for the exercise of employee options and shares issued as part of the employee stock purchase plan, respectively.
During the year ended December 31, 2024, our financing activities provided cash proceeds of $32.1 million. These proceeds were primarily a result of the $31.9 million received from the issuance of common shares and pre-funded warrants, net of $2.6 million in issuance costs paid under the February 2024 Offering.
Contractual Obligations
We enter into contracts in the normal course of business with clinical research organizations, or “CROs,” contract
90
manufacturing organizations, or “CMOs,” and other third parties for clinical trials, preclinical research studies, and testing and manufacturing services. These contracts are generally cancelable at our option with various notice requirements as defined in the contract. Payments due upon cancellation consist of payments for services provided or expenses incurred, including noncancelable obligations of our service providers, up to and through the date of cancellation. These payments are not included as the amount and timing of these payments are not known.
Critical Accounting Estimates
Our management’s discussion and analysis of our financial condition and results of operations is based on our consolidated financial statements as at December 31, 2025, which have been prepared in accordance with United States generally accepted accounting principles, or “U.S. GAAP,” and on a basis consistent with those accounting principles followed by us. The preparation of these consolidated financial statements requires our management to make judgments and estimates that affect the reported amounts of assets and liabilities and the disclosure of contingent assets and liabilities at the date of the financial statements, as well as the reported revenue generated and expenses incurred during the reporting periods. Our estimates are based on our historical experience and on various other factors that we believe are reasonable under the circumstances, the results of which form the basis for making judgments about the carrying value of assets and liabilities that are not readily apparent from other sources. Significant estimates and judgments include, but are not limited to:
| • | Estimates of the percentage of work completed of the total work over the life of the individual trial in accordance with agreements established with CROs, CMOs, and clinical trial sites which in turn impact the research & development expenses. |
| • | Estimate of the grant date fair value share options granted to employees, consultants and directors, and the resulting share-based compensation expense, using the Black Scholes option pricing model. |
Accordingly, actual results may differ from these judgments and estimates under different assumptions or conditions and any such differences may be material. We believe that the accounting policies discussed below are critical to understanding our historical and future performance, as these policies relate to the more significant areas involving management’s judgments and estimates.
a) Research & Development Expenses — Accruals
Research and development costs are charged against income in the period of expenditure. Our research and development costs consist primarily of salaries and fees paid to CROs and to CMO for clinical trial expenses and manufacturing costs prior to FDA approval.
Clinical trial expenses include direct costs associated with CROs, direct CMO costs for the formulation and packaging of clinical trial material, as well as investigator and patient-related costs at sites at which our trials are being conducted. Direct costs associated with our CROs and CMOs are generally payable on a time-and-materials basis, or when milestones are achieved. The invoicing from clinical trial sites can lag several months. We record expenses for our clinical trial activities performed by third parties based upon estimates of the percentage of work completed of the total work over the life of the individual trial in accordance with agreements established with CROs and clinical trial sites. We determine the estimates through discussions with internal clinical personnel, CROs, and CMOs as to the progress or stage of completion of trials or services and the agreed-upon fee to be paid for such services based on facts and circumstances known to us as of each consolidated balance sheet date. The actual costs and timing of clinical trials are highly uncertain, subject to risks and may change depending upon a number of factors, including our clinical development plan. If the actual timing of the performance of services and of the level of effort varies from the estimate, we will adjust the accrual accordingly. Adjustments to prior period estimates have not been material. We recognize the benefit of Canadian research and development tax credits as a reduction of research and development costs for fully refundable investment tax credits and as a reduction of income taxes for investment tax credits that can only be claimed against income taxes payable when there is reasonable assurance that the claim will be recovered.
91
b) Share-Based Compensation
We recognize compensation costs related to share options granted to employees, consultants, and directors based on the estimated fair value of the awards on the date of grant. We estimate the grant date fair value, and the resulting share-based compensation expense, using the Black Scholes option pricing model. This Black Scholes option pricing model uses various inputs to measure fair value, including estimated fair value of our underlying common shares at the grant date, expected term, estimated volatility, risk-free interest rate, and expected dividend yields of our common shares. The estimated volatility creates a critical estimate because we have not been a public company long enough to demonstrate our own historical volatility. The grant date fair value of the share-based awards is recognized on a straight-line basis over the requisite service periods, which are generally the vesting period of the respective awards. Forfeitures are accounted for as they occur.
The following table summarizes, by grant date, the number of underlying common shares and the associated per-share exercise price, which was the fair value per share as determined by our board of directors on the applicable grant date, for share options granted during the years ended December 31, 2025 and 2024:
Estimated | |||||||||||
Number of | Fair Value | Estimated | |||||||||
Common Shares | Exercise | per Common | Per-Share | ||||||||
Subject to | Price Per | Share at | Fair Value | ||||||||
| Options Granted | | Common Share | | Grant Date | | of Options | ||||
March 19, 2024 | 45,000 | $ | 1.45 | $ | 1.45 | $ | 1.12 | ||||
May 6, 2024 | 924,000 | $ | 1.74 | $ | 1.74 | $ | 1.38 | ||||
July 1, 2024 | 8,000 | $ | 1.34 | $ | 1.34 | $ | 1.03 | ||||
July 15, 2024 | 160,000 | $ | 1.59 | $ | 1.59 | $ | 1.18 | ||||
July 17, 2024 | 22,000 | $ | 1.55 | $ | 1.55 | $ | 1.19 | ||||
August 28, 2024 | 320,000 | $ | 1.45 | $ | 1.45 | $ | 1.07 | ||||
September 3, 2024 | 80,000 | $ | 1.39 | $ | 1.39 | $ | 1.02 | ||||
October 1, 2024 | 8,000 | $ | 1.50 | $ | 1.50 | $ | 1.19 | ||||
October 4, 2024 | 10,000 | $ | 1.50 | $ | 1.50 | $ | 1.14 | ||||
November 1, 2024 | 75,000 | $ | 1.44 | $ | 1.44 | $ | 1.15 | ||||
January 2, 2025 | 113,000 | $ | 2.17 | $ | 2.17 | $ | 1.76 | ||||
January 26, 2025 | 1,151,400 | $ | 2.02 | $ | 2.02 | $ | 1.64 | ||||
January 31, 2025 | 40,000 | $ | 1.97 | $ | 1.97 | $ | 1.65 | ||||
March 3, 2025 | 94,000 | $ | 1.65 | $ | 1.65 | $ | 1.33 | ||||
March 17, 2025 | 37,000 | $ | 2.41 | $ | 2.41 | $ | 1.95 | ||||
June 10, 2025 | 320,000 | $ | 1.75 | $ | 1.75 | $ | 1.54 | ||||
August 29, 2025 | 3,000 | $ | 1.84 | $ | 1.84 | $ | 1.64 | ||||
October 1, 2025 | 65,000 | $ | 2.05 | $ | 2.05 | $ | 1.83 | ||||
October 20, 2025 | 4,000 | $ | 1.99 | $ | 1.99 | $ | 1.78 | ||||
November 24, 2025 | 15,000 | $ | 2.37 | $ | 2.37 | $ | 2.11 | ||||
December 1, 2025 | 15,000 | $ | 2.69 | $ | 2.69 | $ | 2.40 | ||||
December 15, 2025 | 8,500 | $ | 2.35 | $ | 2.35 | $ | 2.10 | ||||
Recent Accounting Pronouncements
Refer to Note 2, “Summary of Significant Accounting Policies,” in the accompanying notes to our consolidated financial statements for a discussion of recent accounting pronouncements.
92
ITEM 7A. QUANTITATIVE AND QUALITATIVE DISCLOSURES ABOUT MARKET RISK.
Quantitative and Qualitative Disclosures about Market Risk
We are exposed to market risks in the ordinary course of our business. These risks primarily relate to interest rate risks. For the year ended December 31, 2025, we had cash and cash equivalents of $73.0 million and short-term investments of $32.9 million. For the year ended December 31, 2024, we had cash and cash equivalents of $25.3 million and short-term investments of $44.4 million. These cash and cash equivalents and short-term investments consist primarily of bank deposits, guaranteed investment certificates, and U.S. treasury bills. The primary objective of our investment activities is to preserve principal and liquidity while maximizing income without significantly increasing risk. We do not enter into investments for trading or speculative purposes. Due to the short-term nature of our investment portfolio, we do not believe an immediate 10% increase or decrease in interest rates would have a material effect on the fair market value of our portfolio, and accordingly we do not expect our operating results or cash flows to be materially affected by a sudden change in market interest rates.
We undertake certain transactions in Canadian dollars and as such are subject to risk due to fluctuations in exchange rates. Canadian dollar denominated payables are paid at the converted rate as due. We do not use derivative instruments to hedge exposure to foreign exchange rate risk due to the low volume of transactions denominated in foreign currencies. At December 31, 2025 and 2024, our net monetary exposure denominated in Canadian dollars was $2.8 million and $1.7 million, respectively.
Our operating results and financial position are reported in U.S. dollars in our financial statements. The fluctuation of the Canadian dollar in relation to the U.S. dollar might, consequently, have an impact upon our loss and may also affect the value of our assets and the amount of shareholders’ equity.
93
ITEM 8. FINANCIAL STATEMENTS
INDEX TO CONSOLIDATED FINANCIAL STATEMENTS
Report of Independent Registered Public Accounting Firm (PCAOB ID: | 95 |
96 | |
97 | |
98 | |
99 | |
100 |
94
Report of Independent Registered Public Accounting Firm
To the Board of Directors and Shareholders of Milestone Pharmaceuticals Inc.
Opinion on the Financial Statements
We have audited the accompanying consolidated balance sheets of Milestone Pharmaceuticals Inc. and its subsidiary (the Company) as of December 31, 2025 and 2024, and the related consolidated statements of loss, of shareholders’ equity and of cash flows for the years then ended, including the related notes (collectively referred to as the consolidated financial statements). In our opinion, the consolidated financial statements present fairly, in all material respects, the financial position of the Company as of December 31, 2025 and 2024, and the results of its operations and its cash flows for the years then ended in conformity with accounting principles generally accepted in the United States of America.
Basis for Opinion
These consolidated financial statements are the responsibility of the Company’s management. Our responsibility is to express an opinion on the Company’s consolidated financial statements based on our audits. We are a public accounting firm registered with the Public Company Accounting Oversight Board (United States) (PCAOB) and are required to be independent with respect to the Company in accordance with the U.S. federal securities laws and the applicable rules and regulations of the Securities and Exchange Commission and the PCAOB.
We conducted our audits of these consolidated financial statements in accordance with the standards of the PCAOB. Those standards require that we plan and perform the audits to obtain reasonable assurance about whether the consolidated financial statements are free of material misstatement, whether due to error or fraud. The Company is not required to have, nor were we engaged to perform, an audit of its internal control over financial reporting. As part of our audits, we are required to obtain an understanding of internal control over financial reporting but not for the purpose of expressing an opinion on the effectiveness of the Company’s internal control over financial reporting. Accordingly, we express no such opinion.
Our audits included performing procedures to assess the risks of material misstatement of the consolidated financial statements, whether due to error or fraud, and performing procedures that respond to those risks. Such procedures included examining, on a test basis, evidence regarding the amounts and disclosures in the consolidated financial statements. Our audits also included evaluating the accounting principles used and significant estimates made by management, as well as evaluating the overall presentation of the consolidated financial statements. We believe that our audits provide a reasonable basis for our opinion.
Critical Audit Matters
Critical audit matters are matters arising from the current period audit of the consolidated financial statements that were communicated or required to be communicated to the audit committee and that (i) relate to accounts or disclosures that are material to the consolidated financial statements and (ii) involved our especially challenging, subjective, or complex judgments. We determined there are no critical audit matters.
/s/
March 20, 2026
We have served as the Company's auditor since 2016
95
Milestone Pharmaceuticals Inc.
Consolidated Balance Sheets
(in thousands of US dollars, except share data)
| December 31, 2025 | | December 31, 2024 | |||
Assets | |
| | |||
Current assets |
| | ||||
Cash and cash equivalents | $ | |
| $ | | |
Short-term investments | | | ||||
License receivable | | — | ||||
Research and development tax credits receivable | |
| | |||
Prepaid expenses | |
| | |||
Inventory, net | | — | ||||
Other receivables | |
| | |||
Total current assets | |
| | |||
Operating lease right-of-use assets | | | ||||
Property and equipment, net | |
| | |||
Total assets | $ | |
| $ | | |
Liabilities, and Shareholders' Equity | |
| | |||
Current liabilities | |
| | |||
Accounts payable and accrued liabilities | $ | |
| $ | | |
Operating lease liabilities | |
| | |||
Other current liabilities | | — | ||||
Total current liabilities | |
| | |||
Operating lease liabilities, net of current portion | |
| | |||
Senior secured convertible notes | | | ||||
Other long-term liabilities | | — | ||||
Total liabilities | |
| | |||
Shareholders’ Equity |
| | ||||
Common shares, | |
| | |||
Pre-funded warrants - | | | ||||
Additional paid-in capital | |
| | |||
Accumulated deficit | ( |
| ( | |||
Total shareholders’ equity | |
| | |||
Total liabilities and shareholders’ equity | $ | |
| $ | | |
The accompanying notes are an integral part of these consolidated financial statements.
96
Milestone Pharmaceuticals Inc.
Consolidated Statements of Loss
(in thousands of US dollars, except share and per share data)
Years Ended | |||||||
December 31, | |||||||
| 2025 | | 2024 | ||||
Revenue | $ | | $ | — | |||
Operating expenses |
| ||||||
Research and development, net of tax credits | $ | |
| $ | | ||
General and administrative | |
| | ||||
Commercial | |
| | ||||
Loss from operations | ( |
| ( | ||||
Interest income | |
| | ||||
Interest expense | ( | ( | |||||
Net loss and comprehensive loss | $ | ( |
| $ | ( | ||
Weighted average number of shares and pre-funded warrants outstanding, basic and diluted | | | |||||
Net loss per share, basic and diluted | $ | ( |
| $ | ( | ||
The accompanying notes are an integral part of these consolidated financial statements.
97
Milestone Pharmaceuticals Inc.
Consolidated Statements of Shareholders’ Equity
(in thousands of US dollars, except share data)
Common Shares | Pre-funded warrants | ||||||||||||||||||
| Number | | Amount | | Number | | Amount | | Additional | | Accumulated | | Total | ||||||
Balance as of December 31, 2023 | | $ | | | $ | | $ | | $ | ( | $ | | |||||||
Transactions during 2024 | |||||||||||||||||||
Net loss | — | — | — | — | — | ( | ( | ||||||||||||
Exercise of stock options | | | — | — | ( | — | | ||||||||||||
Pre-funded warrants, net of issuance costs | — | — | | | — | — | | ||||||||||||
Share-based compensation | — | — | — | — | | — | | ||||||||||||
Issuance of common shares, net of issuance costs | | | — | — | — | — | | ||||||||||||
Employee stock purchase plan purchases | | | — | — | — | — | | ||||||||||||
Balance as of December 31, 2024 | | $ | | | $ | | $ | | $ | ( | $ | | |||||||
Balance as of December 31, 2024 | | $ | | | $ | | $ | | $ | ( | $ | | |||||||
Transactions during 2025 | |||||||||||||||||||
Net loss | — | — | — | — | — | ( | ( | ||||||||||||
Exercise of stock options | | | — | — | ( | — | | ||||||||||||
Pre-funded warrants, net of issuance costs | — | — | | | — | — | | ||||||||||||
Share-based compensation | — | — | — | — | | — | | ||||||||||||
Issuance of common shares, vesting of performance stock units | | | — | — | ( | — | — | ||||||||||||
Issuance of common shares, net of issuance costs | | | — | — | — | — | | ||||||||||||
Issuance of Series A common stock warrants, net of issuance costs | — | — | — | — | | — | | ||||||||||||
Issuance of Series B common stock warrants, net of issuance costs | — | — | — | — | | — | | ||||||||||||
Exercise of Series A common stock warrants, net of issuance costs | | | — | — | ( | — | | ||||||||||||
Exercise of Series B common stock warrants, net of issuance costs | | | — | — | ( | — | | ||||||||||||
Employee stock purchase plan purchases | | | — | — | — | — | | ||||||||||||
Balance as of December 31, 2025 | | $ | | | $ | | $ | | $ | ( | $ | | |||||||
The accompanying notes are an integral part of these consolidated financial statements.
98
Milestone Pharmaceuticals Inc.
Consolidated Statements of Cash Flows
(in thousands of US dollars)
Year ended December 31, | ||||||
2025 | | 2024 | ||||
Cash flows used in operating activities | ||||||
Net loss | $ | ( | $ | ( | ||
Adjustments to reconcile net loss to net cash used in operating activities: | ||||||
Depreciation of property and equipment | | | ||||
Amortization of debt costs | | | ||||
Accretion of investment discount | ( | ( | ||||
Non-cash interest expense related to Senior Secured Convertible Note | | | ||||
Share-based compensation expense | | | ||||
Loss on disposals of property and equipment | — | | ||||
Changes in operating assets and liabilities: | ||||||
License receivable | ( | — | ||||
Other receivables | ( | | ||||
Research and development tax credits receivable | | ( | ||||
Prepaid expenses | | | ||||
Inventory | ( | — | ||||
Operating lease assets and liabilities | ( | ( | ||||
Accounts payable and accrued liabilities | | | ||||
Net cash used in operating activities | ( | ( | ||||
Cash provided by investing activities | ||||||
Acquisition of property and equipment | ( | ( | ||||
Acquisition of short-term investments | ( | ( | ||||
Redemption of short-term investments | | | ||||
Net cash provided by investing activities | | | ||||
Cash provided by financing activities | ||||||
Proceeds from exercise of options | | | ||||
Proceeds from issuance of common shares, net of issuance costs | | | ||||
Proceeds from issuance of pre-funded warrants, net of issuance costs | | | ||||
Proceeds from issuance of series A common stock warrants, net of issuance costs | | — | ||||
Proceeds from issuance of series B common stock warrants, net of issuance costs | | — | ||||
Proceeds from exercise of series A warrants, net of issuance costs | | — | ||||
Proceeds from exercise of series B warrants, net of issuance costs | | — | ||||
Proceeds from employee stock purchase plan | | | ||||
Cash provided by financing activities | | | ||||
Net increase in cash and cash equivalents | | | ||||
Cash and cash equivalents – Beginning of period | | | ||||
Cash and cash equivalents – End of period | $ | | $ | | ||
The accompanying notes are an integral part of these consolidated financial statements.
99
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
1. Organization and Nature of Operations
Milestone Pharmaceuticals Inc., or “Milestone,” or the “Company,” is a biopharmaceutical company incorporated under the Business Corporations Act (Québec). Milestone’s headquarters is currently located in Montréal (Québec), Canada. The Company’s common shares began trading on The Nasdaq Global Select Market on May 9, 2019, and trade under the symbol “MIST”. Milestone is focused on the development and commercialization of cardiovascular medicines. Milestone’s lead product candidate, etripamil, is a novel, potent rapid-onset calcium channel blocker that the Company designed and is developing as a rapid-onset nasal spray to be administered by patients. On December 12, 2025, CARDAMYST™ (etripamil) was approved by the U.S. Food and Drug Administration, or “FDA,” to treat paroxysmal supraventricular tachycardia, or “PSVT.” The Company is also developing etripamil for the treatment of atrial fibrillation, or “AFib,” and other cardiovascular indications.
2. Summary of Significant Accounting Policies
a) Basis of consolidation
The consolidated financial statements include the accounts of the Company and Milestone Pharmaceuticals USA, Inc. All intercompany transactions and balances have been eliminated.
b) Basis of Presentation and Use of Accounting Estimates
These consolidated financial statements of the Company have been presented in United States dollars (USD) and have been prepared in accordance with accounting principles generally accepted in the United States of America, or “U.S. GAAP,” including the applicable rules and regulations of the Securities and Exchange Commission (SEC) regarding financial reporting.
The preparation of consolidated financial statements in conformity with U.S. GAAP requires the Company to make estimates and judgments that affect certain reported amounts of assets and liabilities and disclosure of contingent assets and liabilities at the date of the consolidated financial statements and the reported amounts of revenue and expenses during the year. The Company bases its estimates and assumptions on current facts, historical experience and various other factors that it believes are reasonable under the circumstances, to determine the carrying values of assets and liabilities that are not readily apparent from other sources. Significant estimates and judgments include, but are not limited to,
| • | Estimates of the percentage of work completed of the total work over the life of the individual trial in accordance with agreements established with clinical research organizations, or “CROs,” contract manufacturing organizations, or “CMOs,” and clinical trial sites which in turn impact the research & development expenses. |
| • | Estimate of the grant date fair value of share options granted to employees, consultants and directors, and the resulting share-based compensation expense, using the Black Scholes option pricing model. |
Estimates and assumptions about future events and their effects cannot be determined with certainty and therefore require the exercise of judgment. As of the date of issuance of these consolidated financial statements, the Company is not aware of any specific event or circumstance that would require the Company to update its estimates, assumptions and judgments. These estimates may change as new events occur and additional information is obtained and are recognized in the consolidated financial statements as soon as they become known. Actual results could differ from those estimates, and any such differences may be material to the Company’s consolidated financial statements.
c) Segment Information
The Company manages its operations as a single operating segment for the purposes of assessing performance and making operating decisions. See Note 20, “Segment Reporting.”
100
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
d) Revenue Recognition
Collaborative Arrangements
The Company considers the nature and contractual terms of arrangements and assesses whether an arrangement involves a joint operating activity pursuant to which the Company is an active participant and is exposed to significant risks and rewards dependent on the commercial success of the activity. If the Company is an active participant and is exposed to significant risks and rewards dependent on the commercial success of the activity, the Company accounts for such an arrangement as a collaborative arrangement under Accounting Standards Codification, or “ASC” 808, Collaborative Arrangements, or “ASC 808,” which requires that certain transactions between the Company and collaborators be recorded in its consolidated statements of loss on either a gross basis or net basis, depending on the characteristics of the collaborative relationship, and requires enhanced disclosure of collaborative relationships. The Company evaluates its collaboration agreements for proper classification in its consolidated statements of loss based on the nature of the underlying activity. If payments to and from collaborative partners are not within the scope of other authoritative accounting literature, the consolidated statements of loss classification for the payments is based on a reasonable, rational analogy to authoritative accounting literature that is applied in a consistent manner. If the Company concludes that it has a customer relationship with one of its collaborators, the Company follows the guidance in ASC Topic 606, Revenue From Contracts With Customers, or “ASC 606.”
Revenue from Contracts with Customers
In accordance with ASC 606, revenue is recognized when a customer obtains control of promised goods or services. The amount of revenue recognized reflects the consideration to which the Company expects to be entitled in exchange for these goods and services. To achieve this core principle, the Company applies the following five steps: 1) identify the customer contract; 2) identify the contract’s performance obligations; 3) determine the transaction price; 4) allocate the transaction price to the performance obligations; and 5) recognize revenue when or as a performance obligation is satisfied. The Company evaluates all promised goods and services within a customer contract and determines which of such goods and services are separate performance obligations. This evaluation includes an assessment of whether the good or service is capable of being distinct and whether the good or service is separable from other promises in the contract.
In assessing whether promised goods or services in licensing arrangements are distinct, the Company considers factors such as the stage of development of the underlying intellectual property and the capabilities of the customer to develop the intellectual property on their own or whether the required expertise is readily available. Licensing arrangements are analyzed to determine whether the promised goods or services, which often include licenses, research and development services and governance committee services, are distinct or whether they must be accounted for as part of a combined performance obligation. If the license is considered not to be distinct, the license would then be combined with other promised goods or services as a combined performance obligation. If the Company is involved in a governance committee, it assesses whether its involvement constitutes a separate performance obligation. When governance committee services are determined to be separate performance obligations, the Company determines the fair value to be allocated to this promised service. Certain contracts contain optional and additional items, which are considered marketing offers and are accounted for as separate contracts with the customer if such option is elected by the customer, unless the option provides a material right which would not be provided without entering into the contract. An option that is considered a material right is accounted for as a separate performance obligation. The transaction price is determined based on the consideration to which the Company will be entitled in exchange for transferring goods and services to the customer. A contract may contain variable consideration, including potential payments for both milestone and research and development services. For certain potential milestone payments, the Company estimates the amount of variable consideration by using the most likely amount method. In making this assessment, the Company evaluates factors such as the clinical, regulatory, commercial and other risks that must be overcome to achieve the milestone. Each reporting period the Company re-evaluates the probability of achievement of such variable consideration and any related constraints. Milestone will include variable consideration, without constraint, in the transaction price to the extent it is probable that a significant reversal in the amount of cumulative revenue recognized will not occur when the uncertainty associated with the variable consideration is subsequently resolved.
101
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
If the contract contains a single performance obligation, the entire transaction price is allocated to the single performance obligation. Contracts that contain multiple performance obligations require an allocation of the transaction price among the performance obligations on a relative standalone selling price basis unless a portion of the transaction price is variable and meets the criteria to be allocated entirely to a performance obligation or to a distinct good or service that forms part of a single performance obligation.
The Company allocates the transaction price based on the estimated standalone selling price of the underlying performance obligations or in the case of certain variable consideration to one or more performance obligations. The Company must develop assumptions that require judgment to determine the stand-alone selling price for each performance obligation identified in the contract. The Company utilizes key assumptions to determine the stand-alone selling price, which may include other comparable transactions, pricing considered in negotiating the transaction and the estimated costs to complete the respective performance obligation. Certain variable consideration is allocated specifically to one or more performance obligations in a contract when the terms of the variable consideration relate to the satisfaction of the performance obligation and the resulting amounts allocated to each performance obligation are consistent with the amount the Company would expect to receive for each performance obligation.
When a performance obligation is satisfied, revenue is recognized for the amount of the transaction price, excluding estimates of variable consideration that are constrained, that is allocated to that performance obligation on a relative standalone selling price basis. Significant management judgment is required in determining the level of effort required under an arrangement and the period over which the Company is expected to complete its performance obligations under an arrangement.
For performance obligations consisting of licenses and other promises, the Company utilizes judgment to assess the nature of the combined performance obligation to determine whether the combined performance obligation is satisfied over time or at a point in time and, if over time, the appropriate method of measuring progress for purposes of recognizing revenue from non- refundable, up-front fees. The Company evaluates the measure of progress each reporting period and, if necessary, adjusts the measure of performance and related revenue recognition. If the license to the Company’s intellectual property is determined to be distinct from the other performance obligations identified in the arrangement, the Company will recognize revenue from non-refundable, up-front fees allocated to the license at the point in time when the license is transferred to the customer and the customer is able to use and benefit from the license.
e) Cash and Cash Equivalents
Cash and cash equivalents consist of cash and highly liquid investments that are readily convertible into cash with original maturities of 90 days or less at acquisition date.
f) Short-Term Investments
Short-term investments are classified as held-to-maturity, are initially recognized at fair value and are subsequently accounted for at amortized cost. They are comprised of guaranteed investment certificates with a maturity greater than 90 days but less than one year and, as such, are classified as current assets.
g) Concentration Risks
Concentration of Credit Risk
Financial instruments that potentially subject the Company to concentration of credit risk consist primarily of cash and cash equivalents and investment securities classified as held-to-maturity. The Company maintains deposits at major financial institutions and such amounts may exceed those amounts insured by the Federal Deposit Insurance Corporation, or the “FDIC.” The Company has not experienced any losses on its deposits since inception, and the Company believes
102
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
that minimal credit risk exists with respect to these financial institutions. Additionally, the Company has adopted an investment policy that includes guidelines relative to credit quality, diversification of maturities and liquidity.
Concentration of Supplier Risk
The Company relies on a single source manufacturer for starting materials and active pharmaceutical ingredient, or “API,” packaging components, and packaged pharmaceutical product ready for commercial sale.
h) Currency Risk
The Company is exposed to currency risk due to financial instruments denominated in foreign currencies. The Company is exposed to the Canadian dollar currency risk and does not enter into arrangements to hedge its currency risk exposure.
i) Inventory
Inventory is stated at the lower of cost or net realizable value, with cost determined on a first‑in, first‑out, or “FIFO,” basis. Inventory costs include raw materials, work‑in‑process, finished goods, and applicable manufacturing overhead. Raw materials include starting materials used to manufacture the API. Work‑in‑process includes API, intermediate compounding steps, and bulk drug product. Finished goods include packaging components and the packaged pharmaceutical product ready for commercial sale.
Prior to the FDA’s approval of CARDAMYST on December 12, 2025, costs associated with manufacturing pre‑approval commercial batches and clinical materials were expensed as research and development expenses in accordance with our accounting policy and SAB Topic 5A, as the product had not yet received regulatory approval and therefore did not meet the criteria for capitalization. Following FDA approval, we began capitalizing inventory related to CARDAMYST manufactured after the approval date. Inventory produced prior to approval and previously expensed as research and development remains recorded at a zero‑cost basis and is excluded from the inventory balance.
Because our inventory is subject to expiration, the Company evaluates its carrying value each reporting period and records allowances for any estimated excess, obsolete, short‑dated, or otherwise unmarketable inventory. In accordance with regulatory requirements and current Good Manufacturing Practices, or “cGMP,” our inventory is also subject to strict quality control and monitoring throughout the manufacturing process. If certain batches or units do not meet quality specifications, we would write down the related inventory to its estimated net realizable value and record the expense as a cost of production in the Consolidated Statement of Loss. Inventory valuation adjustments require judgment and consideration of various factors, including forecasted demand, remaining shelf life, and quality assessments, among others.
j) Property and Equipment
Property and equipment is stated at historical cost less accumulated depreciation. Expenditures for maintenance and repairs are recorded to expense as incurred. The Company reviews its property and equipment whenever events or changes in circumstances indicate that the carrying value of certain assets might not be recoverable and recognizes an impairment loss when it is probable that an asset’s realizable value is less than the carrying value. To date,
103
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
have been recorded. Depreciation is calculated using the straight-line method over the following estimated useful lives of the assets:
Computer hardware and software | | |
Office equipment |
| |
Furniture and fixtures |
| |
Leasehold improvements |
| over the lease term |
k) Leases
At the inception of an arrangement, the Company determines whether the arrangement is or contains a lease based on the unique facts and circumstances present in the arrangement. Leases with a term greater than one year are recognized on the balance sheet as right-of-use assets and short-term and long-term lease liabilities, as applicable. The Company does not have financing leases.
Operating lease liabilities and their corresponding right-of-use assets are initially recorded based on the present value of lease payments over the expected remaining lease term. Right-out-use assets are subsequently accounted for as long-lived assets, including evaluating for indicators of impairment. Certain adjustments to the right-of-use asset may be required for items such as incentives received. The interest rate implicit in lease contracts is typically not readily determinable. As a result, the Company utilizes its incremental borrowing rate to discount lease payments, which reflects the fixed rate at which the Company could borrow on a collateralized basis the amount of the lease payments in the same currency, for a similar term, in a similar economic environment. Prospectively, the Company will adjust the right-of-use assets for straight-line rent expense, or any incentives received, and remeasure the lease liability at the net present value using the same incremental borrowing rate that was in effect as of the lease commencement or transition date.
The Company has elected not to recognize leases with an original term of one year or less on the balance sheet. The Company typically only includes an initial lease term in its assessment of a lease arrangement. Options to renew a lease are not included in the Company’s assessment unless there is reasonable certainty that the Company will renew.
l) Pre-funded Warrants
Pre-funded warrants allow the holder to pay little or no consideration to receive the shares upon exercise of the warrant. The pre-funded warrants do not meet the definition of a derivative under ASC 815, Derivatives and Hedging, because their fair value at issuance is equal to the fair value of the shares underlying the warrant. As such, they have the characteristics of a prepaid forward sale of equity. As a result, the pre-funded warrants are accounted for as equity instruments.
m) Share and Debt Issuance Costs
Share issuance costs include legal fees, accounting fees, sales agent commissions, broker and underwriting fees, and other direct costs incurred in connection with the issuance of equity instruments. Share issuance costs are recorded as a reduction of the financing equity proceeds.
Debt issuance costs include legal fees, accounting fees, and other direct costs incurred in connection with the execution of our debt financing. Debt discounts represent costs paid to the lenders. Debt issuance costs and debt discounts are deducted from the carrying amount of the debt liability and are amortized to interest expense over the term of the related debt using the effective interest method.
104
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
n) Research and Development and Investment Tax Credits
Research and development costs are charged to expense as costs are incurred in performing research and development activities. The Company’s research and development costs consist primarily of salaries and fees paid to contract research organizations, or “CROs” and non-capitalizable costs paid to contract manufacturing organizations, or “CMOs.”
Clinical trial expenses include direct costs associated with CROs, direct CMO costs for the formulation and packaging of clinical trial material, as well as investigator and patient related costs at sites at which the Company’s trials are being conducted. Direct costs associated with the Company’s CROs and CMOs are generally payable on a time and materials basis, or when milestones are achieved. The invoicing from clinical trial sites can lag several months. The Company records expenses for its clinical trial activities performed by third parties based upon estimates of the percentage of work completed of the total work over the life of the individual study in accordance with agreements established with CROs and clinical trial sites. The Company determines the estimates through discussions with internal clinical personnel, CROs and CMOs as to the progress or stage of completion of trials or services and the agreed upon fee to be paid for such services based on facts and circumstances known to the Company as of each consolidated balance sheet date. The actual costs and timing of clinical trials are highly uncertain, subject to risks and may change depending upon a number of factors, including the Company’s clinical development plan. If the actual timing of the performance of services of the level of effort varies from the estimate, the Company will adjust the accrual accordingly.
The Company recognizes the benefit of Canadian research and development tax credits as a reduction of research and development costs for fully refundable investment tax credits and as a reduction of income taxes for investment tax credits that can only be claimed against income taxes payable when there is reasonable assurance that the claim will be recovered.
o) Income Taxes
The provision for income taxes is computed using the liability method. Under this method, deferred tax assets and liabilities are determined based on differences between financial reporting and tax bases of assets and liabilities. Deferred tax assets and liabilities are measured using enacted tax rates and laws that will be in effect when the differences are expected to reverse. A valuation allowance is recorded to reduce the carrying amount of deferred income tax assets until when it is more likely than not that these assets will be realized. Tax benefits related to tax positions not deemed to meet the “more likely than not” threshold are not permitted to be recognized in the consolidated financial statements.
p) Foreign Currency Translation and Transactions
The functional currency of the Company is the U.S. dollar. Accordingly, transactions denominated in currencies other than the functional currency are measured and recorded in the functional currency at the exchange rate in effect on the date of the transactions. At each consolidated balance sheet date, monetary assets and liabilities denominated in currencies other than the functional currency are remeasured using the exchange rate in effect at that date. Non-monetary assets and liabilities and revenue and expense items denominated in foreign currencies are translated into the functional currency using the exchange rate prevailing at the dates of the respective transactions. Any gains or losses arising on remeasurement are included in the consolidated statement of loss.
q) Share-Based Compensation
The Company has a share-based compensation plan which is described in detail in Note 9, “Shareholders’ Equity,” and records all share-based payments, including grants of employee share options, performance stock options, or “PSOs,” restricted stock units, or “RSUs,” and performance stock units, or “PSUs, based on the fair value of the awards as of the grant date. The fair value of share options granted to employees and non-employees is estimated at the date of grant using the Black-Scholes option pricing model. The fair value of RSUs and PSUs with performance-based vesting conditions is calculated using the closing price of the Company’s common stock on the date of grant.
105
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
The Company recognizes share-based compensation expense over the requisite service period of the award, which equals the vesting period, using the straight-line method, with expense for PSUs and PSOs with performance-based vesting conditions, adjusted based on the likelihood of future achievement of the performance metrics.
Forfeitures, if any, are recorded as they occur. Any consideration paid by employees on exercising share options and the corresponding portion previously credited to contributed surplus are credited to share capital. The Black-Scholes option pricing model used by the Company to calculate option values was developed to estimate fair value.
The Company approved an employee share purchase plan in April 2019, which became effective on May 8, 2019, and is described in Note 10, “Share-Based Compensation.” The plan provides a means by which eligible employees of the Company may be given an opportunity to purchase common shares. The plan permits the Company to grant a series of purchase rights to eligible employees under an employee stock purchase plan.
r) Recent Accounting Pronouncements
New Accounting Pronouncements - Issued and Adopted
In December 2023, the FASB issued ASU 2023-09, Income Taxes (Topic 740): Improvements to Income Tax Disclosures, or “ASU 2023-09.” The amendments in this update require that public business entities on an annual basis (1) disclose specific categories in the rate reconciliation and (2) provide additional information for reconciling items that meet a quantitative threshold (if the effect of those reconciling items is equal to or greater than 5 percent of the amount computed by multiplying pretax income (or loss) by the applicable statutory income tax rate). The amendments also require entities on an annual basis to disclose disaggregated amounts of income taxes paid. ASU 2023-09 is effective for annual periods beginning after December 15, 2024. The Company adopted ASU 2023-09 for the year ended December 31, 2025, as documented in Note 13, “Income Taxes.”
New Accounting Pronouncements - Issued but Not Yet Adopted
In November 2024, the FASB issued ASU 2024-03 “Income Statement: Reporting Comprehensive Income— Expense Disaggregation Disclosures,” which requires more detailed information about specified categories of expenses (purchases of inventory, employee compensation, depreciation, amortization, and depletion) included in certain expense captions presented on the face of the income statement, as well as disclosures about selling expenses. This ASU is effective for fiscal years beginning after December 15, 2026, and for interim periods within fiscal years beginning after December 15, 2027. Early adoption is permitted. The amendments may be applied either (1) prospectively to financial statements issued for reporting periods after the effective date of this ASU or (2) retrospectively to all prior periods presented in the financial statements. The Company is currently evaluating this guidance to determine the impact it may have on its financial statement disclosures.
s) Significant Risks and Uncertainties
The Company is subject to challenges and risks specific to its business and its ability to execute on its strategy, as well as risks and uncertainties common to companies in the pharmaceutical industry, including, without limitation, risks and uncertainties associated with: commercializing CARDAMYST; procuring inventory given our reliance on a concentrated supplier base, delays or problems in the supply of its study drug or failure to comply with manufacturing regulations; identifying, acquiring or in-licensing product candidates; pharmaceutical product development and the inherent uncertainty of clinical success; and the challenges of protecting and enhancing its intellectual property rights; and complying with applicable regulatory requirements.
Further, the Company may be impacted by general economic, political, and market conditions, including deteriorating market conditions due to investor concerns regarding inflation, armed conflicts, and overall fluctuations in the financial markets in the U.S. and abroad.
106
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
t) Sources of Liquidity and Funding Requirements
The Company has incurred operating losses and experienced negative operating cash flows since its inception and expect to incur additional costs in connection with commercialization in 2026. The Company intends to finance its commercialization and working capital needs from existing cash, potential proceeds from the commercialization of CARDAMYST, royalty revenue, the exercise of outstanding common stock options and warrants to purchase common stock, and existing and future licensing and commercial partnership agreements. As of December 31, 2025, the Company had cash and cash equivalents and short-term investments of $
The Company has historically financed its operations primarily through the sale of equity securities, convertible notes, short-term investments, and from cash received pursuant to its license agreement. To date, the Company has not generated any revenue from product sales. Management expects operating losses and negative cash flows from operations to continue for the foreseeable future. There can be no assurance that, in the event the Company requires additional financing, such financing will be available at terms acceptable to the Company, if at all. Failure to generate sufficient cash flows from operations, raise additional capital and reduce discretionary spending should additional capital not become available could have a material adverse effect on the Company’s ability to achieve its business objectives.
3. Revenue
The Company recorded license revenue of $
Strategic Partnerships
Corxel Pharmaceuticals
On May 15, 2021, the Company entered into the License Agreement with Corxel, which is an entity affiliated with RTW Investments, LP, or “RTW,” a beneficial owner of approximately
Management evaluated all of the promised goods or services within the contract and determined that such goods and services were separate performance obligations. The Company determined that the license granted was a separate performance obligation as Corxel can benefit from the license granted on its own after the transfer of the license, as it does not require any significant development, regulatory or commercialization activities from Milestone. Corxel is responsible for all development, regulatory and commercialization activities in the Territory, including the performance of clinical trials necessary for regulatory approval, and is responsible for all such related costs. Supply of the product can be provided by another entity, as the Company currently uses a CMO for the production of etripamil without subsequent significant
107
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
modification or customization by the Company, therefore the Company determined the obligation to supply product is a separate and distinct obligation. The Company concluded that the obligation for participation on the various governance committees was distinct as the services could be performed by an outside party, however it was determined to be immaterial after estimating the stand-alone cost compared to the License Agreement as a whole. As a result, the Company concluded there were
4. Short-term Investments
As of December 31, 2025, short-term investments of $
As of December 31, 2025, $
5. Inventory
Following FDA approval in December 2025, the Company began capitalizing inventory costs related to CARDAMYST manufactured after the approval date. The components of inventory are as follows:
| December 31, 2025 | | December 31, 2024 | |||
Raw materials |
| $ | — | $ | — | |
Work-in-process |
|
| |
| — | |
Finished goods | | — | ||||
Total inventory | | — | ||||
Inventory manufactured prior to FDA approval was expensed through research and development costs, and therefore, this inventory has a zero-cost basis and is not included in the inventory balance.
6. Leases
On May 20, 2022, the Company entered into a lease arrangement for a term for new office space located in Charlotte, NC. The Company recognized the operating lease right-of-use asset and operating lease liabilities at the lease commencement date on August 1, 2022. The interest rate implicit in lease contracts is not readily determinable and the Company does not have a public credit rating and carries no debt. As such, several factors were considered in the determination of the Company’s incremental borrowing rate used in determining the present value of lease payments. The Company’s examined credit ratings for similar companies, assumed equivalency between the Canadian and U.S. markets for collateralized debt and used rates near the period. This resulted in an incremental borrowing rate of
On August 12, 2025, the Company entered into an arrangement for the lease renewal for its headquarters located in Ville Saint-Laurent, Quebec. The lease term is from December 1, 2025, expiring on November 30, 2027. The Company recorded the operating lease right-of-use asset and operating lease liabilities at the effective lease arrangement date of August 12, 2025. The Company’s examined credit ratings for similar companies, assumed equivalency between the Canadian and U.S. markets for collateralized debt and used rates for the remaining lease term of
108
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
in an incremental borrowing rate of
The Company's operating office leases right-of-use assets as at December 31 were as follows:
2025 | | 2024 | ||||
Opening balance | | $ | | $ | | |
Right-of-use adjustment, renewal August 2025 | | — | ||||
Amortization of right-of-use asset | ( | ( | ||||
Closing balance | $ | | $ | | ||
Operating lease expenses of $
The following table summarizes the future minimum lease payments of right-of-use assets operating leases as of December 31, 2025:
January 1, 2026 to December 31, 2026 | $ | | |
January 1, 2027 to November 30, 2027 | | ||
Total | | ||
Less interest | ( | ||
Total net of interest | $ | |
7. Property and equipment
Property and equipment consist of the following at December 31:
| 2025 | | 2024 | |||
Computer hardware and software | $ | | $ | | ||
Office equipment |
| |
| | ||
Leasehold improvements |
| |
| | ||
Total | $ | | $ | | ||
Less accumulated depreciation |
| ( |
| ( | ||
Property and equipment, net | $ | | $ | | ||
109
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
8. Accounts payable and accrued liabilities
Accounts payable and accrued liabilities comprised the following as of December 31:
| 2025 | 2024 | ||||
| |
| | |||
Trade accounts payable |
| $ | | $ | | |
Accrued compensation and benefits payable |
| | | |||
Accrued research and development liabilities |
| | | |||
Accrued commercial liabilities | | | ||||
Accrued underwriting fees | | — | ||||
Accrued legal liabilities | | | ||||
Other accrued liabilities |
| | | |||
Total |
| $ | | $ | | |
9. Shareholders’ Equity
Authorized Share Capital
The Company has authorized and issued common shares, voting and participating, without par value, of which unlimited shares were authorized, and
As of December 31, 2025, there were
On July 11, 2025, the Company entered into an underwriting agreement, or the “Underwriting Agreement,” related to an underwritten public offering, or the “2025 Offering,” of (i)
The net proceeds to the Company from the 2025 Offering were $
On February 28, 2024, the Company entered into an underwriting agreement, or the “Underwriting Agreement,” related to an underwritten public offering, or the “February 2024 Offering,” of
110
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
On July 29, 2020, the Company entered into an Open Market Sale AgreementSM, or the “Original Sale Agreement,” with respect to an at-the-market offering program, or the “ATM Program,” under which the Company could issue and sell its common shares having an aggregate offering price of up to $
The Company determines the accounting classification of warrants that are issued, as either liability or equity, by first assessing whether the warrants meet liability classification in accordance with ASC 480-10, Accounting for Certain Financial Instruments with Characteristics of both Liabilities and Equity, and then in accordance with ASC 815-40, Accounting for Derivative Financial Instruments Indexed to, and Potentially Settled in, a Company’s Own Stock. Under ASC 480, warrants are considered liability classified if the warrants are mandatorily redeemable, obligate the issuer to settle the warrants or the underlying shares by paying cash or other assets, or must or may require settlement by issuing a variable number of shares.
If warrants do not meet liability classification under ASC 480-10, the Company assesses the requirements under ASC 815-40, which states that contracts that require or may require the issuer to settle the contract for cash are liabilities recorded at fair value, irrespective of the likelihood of the transaction occurring that triggers the net cash settlement feature. If the warrants do not require liability classification under ASC 815-40, in order to conclude equity classification, the Company assesses whether the warrants are indexed to its common stock and whether the warrants are classified as equity under ASC 815-40 or other applicable U.S. GAAP. After all relevant assessments are made, the Company concludes whether the warrants are classified as liability or equity. The Company's outstanding common stock warrants are classified as equity and recorded in additional paid-in capital, or “APIC,” based on an allocation of the proceeds from the 2025 Offering, which was based on the relative fair value of the Series A and B Common Warrants at the issuance date and is not subject to change after the issuance date. The fair value used for the relative fair value allocation was calculated using the Black-Scholes Model for the Series A Warrants and the Monte-Carlo simulation method for the Series B Warrants.
During the year ended December 31, 2025,
10. Share-Based Compensation
Stock Options
Under the Company’s 2019 Equity Incentive Plan, or the “2019 Plan,” and the Company’s Stock Option Plan, or the “2011 Plan,” unless otherwise decided by the Board of Directors, options vest and are exercisable as follows:
On January 1, 2025, the number of the Company’s common shares reserved for issuance under the 2019 Plan automatically increased by
111
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
were cancelled and were made available for future grants. As of December 31, 2025, there were
On November 10, 2021, the Company established a 2021 Inducement Plan, or the “Inducement Plan.” The Inducement Plan is intended to help the Company provide an inducement for certain individuals to enter employment with the Company, incentives for such persons to exert maximum efforts for the success of the Company and a means by which employees may benefit from increases in value of the common shares. As of December 31, 2025, there were
112
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
The total outstanding and exercisable options from the 2011 Plan, 2019 Plan, and Inducement Plan as of December 31 were as follows:
2025 | ||||||||||||||
Weighted | ||||||||||||||
Number | average | |||||||||||||
of shares | exercise | |||||||||||||
| 2019 Plan | Inducement Plan | 2011 Plan | | Total | | price | |||||||
Outstanding at beginning of year - 2011 Plan |
| — | — | | | | | | $ | | ||||
Outstanding at beginning of year - 2019 Plan | | — | — | | | |||||||||
Outstanding at beginning of year - Inducement Plan | — | | — | | | |||||||||
Granted - 2019 Plan | | — | — | | | |||||||||
Granted - Inducement Plan | — | | — | | | |||||||||
Exercised - 2011 Plan | — | — | ( | ( | | |||||||||
Forfeited - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - 2019 Plan | ( | — | — | ( | | |||||||||
Expired - 2019 Plan | ( | — | — | ( | | |||||||||
Expired - 2011 Plan | — | — | ( | ( | | |||||||||
Outstanding at end of year |
| | | | | $ | | |||||||
Outstanding at end of year - Weighted average exercise price | $ | | $ | | $ | | ||||||||
Exercisable at end of year | | | | | $ | | ||||||||
Exercisable at end of year - Weighted average exercise price |
| $ | | $ | | $ | | |||||||
2024 | ||||||||||||||
Weighted | ||||||||||||||
Number | average | |||||||||||||
of shares | exercise | |||||||||||||
2019 Plan | Inducement Plan | 2011 Plan | | Total | | price | ||||||||
Outstanding at beginning of year - 2011 Plan | | — | — | | | $ | | |||||||
Outstanding at beginning of year - 2019 Plan |
| | — | — | | | ||||||||
Outstanding at beginning of year - Inducement Plan | — | | — | | | |||||||||
Granted - 2019 Plan | | — | — | | | |||||||||
Exercised - 2011 Plan | — | — | ( | ( | | |||||||||
Forfeited - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - 2019 Plan | ( | — | — | ( | | |||||||||
Expired - 2019 Plan | ( | — | — | ( | | |||||||||
Expired - 2011 Plan | — | — | ( | ( | | |||||||||
Expired - Inducement Plan | — | ( | — | ( | | |||||||||
Outstanding at end of period | | | | | $ | | ||||||||
Outstanding at end of year - Weighted average exercise price | $ | | $ | | $ | | ||||||||
Exercisable at end of year | | | | | $ | | ||||||||
Exercisable at end of year - Weighted average exercise price | $ | | $ | | $ | | ||||||||
The weighted average remaining contractual life was
There was $
Options granted are valued using the Black-Scholes option pricing model. This model also requires assumptions, including expected option life, volatility, risk-free interest rate and dividend yield, which greatly affect the calculated values.
113
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
Amortization of the fair value of the options over vesting years has been expensed and credited to additional paid-in capital in shareholders’ equity. The non-vested options as of December 31 were as follows:
2025 | ||||||||||||||
Number | Weighted | |||||||||||||
of options | average | |||||||||||||
2019 Plan | | Inducement Plan | | 2011 Plan | | Total | | fair value | ||||||
Non-vested share options at beginning of year - 2019 Plan | | — | — | |
| $ | | |||||||
Non-vested share options at beginning of year - Inducement Plan | — | | — | | | |||||||||
Granted - 2019 Plan |
| | — | — | | | ||||||||
Granted - Inducement Plan | — | | — | | | |||||||||
Vested, outstanding - 2019 Plan | ( | — | — | ( | | |||||||||
Vested, outstanding - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - 2019 Plan | ( | — | — | ( | | |||||||||
Non-vested share options at end of year |
| | | — | |
| $ | | ||||||
Non-vested share options at end of period - Weighted average fair value | $ | | $ | | $ | — | ||||||||
2024 | ||||||||||||||
Number | Weighted | |||||||||||||
of options | average | |||||||||||||
| 2019 Plan | | Inducement Plan | | 2011 Plan | | Total | | fair value | |||||
Non-vested share options at beginning of year - 2019 Plan | | — | — | |
| $ | | |||||||
Non-vested share options at beginning of year - Inducement Plan | — | | — | | | |||||||||
Granted - 2019 Plan |
| | — | — | | | ||||||||
Vested, outstanding - 2019 Plan | ( | — | — | ( | | |||||||||
Vested, outstanding - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - Inducement Plan | — | ( | — | ( | | |||||||||
Forfeited - 2019 Plan | ( | — | — | ( | | |||||||||
Non-vested share options at end of year |
| | | — | |
| $ | | ||||||
Non-vested share options at end of year - Weighted average fair value | $ | | $ | | $ | — | ||||||||
The following table summarizes information with respect to share options outstanding as of December 31, 2025:
Options outstanding | Options exercisable | |||||||||||||
| | Weighted | | | | Weighted | | |||||||
average | Weighted | average | Weighted | |||||||||||
remaining | average | remaining | average | |||||||||||
Number | contractual | exercise | Number | contractual | exercise | |||||||||
Exercise price | of options | life (years) | price | of options | life (years) | price | ||||||||
$ |
| | | | | |||||||||
$ |
| | | - | - | - | ||||||||
$ |
| | | | | |||||||||
$ |
| | | | | |||||||||
$ |
| | | | | |||||||||
$ | | | | | ||||||||||
$ | | | | | ||||||||||
$ | | | | | ||||||||||
Total |
| |
| $ | |
| |
| $ | | ||||
114
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
The fair value of options granted for the 2011 Plan, 2019 Plan, and Inducement Plan were estimated using Black-Scholes option pricing model, resulting in the following weighted average assumptions for the options granted:
Year ended December 31, | |||||||
2025 | | 2024 |
| ||||
Exercise price | $ | |
| $ | | ||
Share price | $ | |
| $ | | ||
Volatility |
| | % | | % | ||
Risk-free interest rate |
| | % | | % | ||
Expected life |
|
| |||||
Dividend |
| % | % | ||||
Expected volatility is determined using the Company’s historical volatility. Prior to establishing sufficient historical volatility, the Company used comparable companies for which the information is publicly available. The risk-free interest rate is determined based on the U.S. sovereign rates benchmark in effect at the time of grant with a remaining term equal to the expected life of the option. Expected option life is determined based on the simplified method as the Company does not have sufficient historical exercise data to provide a reasonable basis upon which to estimate expected term. The simplified method is an average of the contractual term of the options and its ordinary vesting period. Dividend yield is based on the share option’s exercise price and expected annual dividend rate at the time of grant. The total grant date fair value for options granted during the year ended December 31, 2025 and 2024 was $
Performance Stock Options
On May 6, 2024, the Company, pursuant to the 2019 Plan, awarded
The original grant-date fair value of each option was estimated on the date of grant using the same option valuation model used for the options outlined above. The original grant-date fair value of $
The vesting of the performance-based stock options is conditional upon the FDA approval of etripamil for the treatment of PSVT. Subject to the option holders continuous service as of each such date,
Employee Stock Purchase Plan
On July 15, 2022, the Company offered an employee stock purchase plan, or “ESPP,” in which participation is available to our employees in the United States and Canada who meet certain service eligibility requirements. Eligible employees may authorize an amount up to
On January 1, 2025, the number of common shares reserved for issuance under the ESPP automatically increased by
115
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
ESPP related to the purchase discount and the “look-back” option was determined using a Black-Scholes option pricing model.
Performance Share Units
On May 6, 2024, the Company, pursuant to the 2019 Plan, awarded
The number of PSUs granted represents the total number of common shares that may be earned. However, the actual number of shares earned will be based on the satisfaction of the performance criteria. Upon satisfaction of the performance criteria,
Restricted Stock Units
Pursuant to the 2019 Plan, the Company issues Restricted Stock Units, or “RSUs,” to employees which vest based on a service criteria. When vested, the RSUs represent the right to be issued a number of shares of the Company’s common stock equal to the number of RSUs granted. The grant date fair value for RSUs is based on the market price of the Company’s common stock on the date of the grant. The fair value is then amortized to compensation expense over the requisite service period or vesting term. The Company issued
The total unrecognized compensation cost related to the non-vested RSUs as of December 31, 2025, was $
Share-Based Compensation Expense
The Company recognized share-based compensation expense for all plans as follows for the years ended December 31:
| 2025 | | 2024 | |||
Administration | $ | |
| $ | | |
Research and development | |
| | |||
Commercial activities | |
| | |||
Total | $ | |
| $ | | |
11. Debt
On March 27, 2023, the Company entered into a note purchase agreement, or the “Note Purchase Agreement,” with RTW Investments LP and certain of its affiliates, or collectively, “RTW.”
On March 29, 2023, the Company closed the transactions contemplated by the Note Purchase Agreement, and issued and sold $
The 2029 Convertible Notes are senior secured obligations and are guaranteed on a senior secured basis by the Company’s wholly owned subsidiary, Milestone Pharmaceuticals USA, Inc. Interest at the annual rate of
116
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
cash or, at our option, payable in kind for the first
Each $
On or after March 27, 2027, the 2029 Convertible Notes are redeemable by the Company, subject to certain conditions, if the closing sale price of the common shares exceeds
In accounting for the issuance of the 2029 Convertible Notes, the Company determined there were no embedded features, which require bifurcation between debt and equity components. As a result, the 2029 Convertible Notes are accounted for as a liability. As of December 31, 2025, the estimated fair value of the 2029 Convertible Notes was approximately $
The net carrying amount of the 2029 Convertible Note were as follows:
| December 31, 2025 | December 31, 2024 | ||||
Original principal |
| $ | | $ | | |
Paid in kind (PIK) interest | | | ||||
Unamortized debt discount | ( | ( | ||||
Unamortized debt issuance costs |
| ( | ( | |||
Total |
| $ | | $ | | |
The following table presents the total amount of interest cost recognized relating to the 2029 Convertible Notes:
Year ended December 31, | ||||||||
2025 | | 2024 | ||||||
Contractual interest expense | $ | |
| $ | | |||
Amortization of debt discount | | | ||||||
Amortization of debt issuance costs | |
| | |||||
Total interest expense | $ | |
| $ | | |||
117
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
12. Net loss per share
Basic and diluted net loss per common share is determined by dividing net loss applicable to common shareholders by the weighted average number of common shares and pre-funded warrants outstanding during the period. In addition to the conversion feature on the 2029 Convertible Notes described above, which the Company reviewed and concluded that if-converted would be anti-dilutive due to the facts surrounding the feature, the following potentially dilutive securities have also been excluded from the computation of diluted weighted average shares outstanding as of December 31, as they would be anti-dilutive:
| 2025 | | 2024 | |
Stock options and RSUs |
| |
| |
Amounts in the table above reflect the common share equivalents of the noted instruments.
13. Income taxes
Upon adoption of ASU No. 2023-09, for the year ended December 31, 2025, the provision for income taxes differs from the expense that would be obtained by applying the Canadian federal statutory income tax rate as a result of the following:
2025 | ||||||
Amount | Percent | |||||
Loss before income taxes | $ | ( | ||||
Canadian federal statutory tax rate(1) |
| ( |
| | % | |
Provincial and local taxes(2) |
| ( |
| | % | |
Foreign tax effects | ||||||
United States | ||||||
Statutory tax rate difference | ( | | % | |||
Non-taxable item: share based compensation | | ( | % | |||
Other | | ( | % | |||
Changes in Valuation Allowances | | ( | % | |||
Changes in unrecognized tax benefits | | ( | % | |||
Non-taxable or non-deductible items | ||||||
Share-based compensation | | ( | % | |||
Other | ( | | % | |||
Effective Tax Rate | $ | — | - | % | ||
(1) We apply the federal tax rate of
(2) We apply the provincial tax rate in Quebec of
For the year ended December 31, 2025, loss before income taxes was $
118
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
Prior to the adoption of ASU No. 2023-09, for the year ended December 31, 2024, the provision for income taxes differs from the expense that would be obtained by apply the Canadian statutory income tax rate as result of the following:
| 2024 | |||||
Amount | Percent | |||||
Loss before income taxes | $ | ( | ||||
Canadian statutory rate(1) |
| | % | |||
Computed income tax recovery |
| ( |
|
| ||
Effect on income tax rate resulting from |
|
|
| |||
Non‑deductible share‑based compensation | | ( | % | |||
Other permanent differences | | ( | % | |||
Tax benefits of current period losses and other tax assets, subject to full valuation allowance | | ( | % | |||
Valuation allowance for prior year adjustment | | ( | % | |||
Foreign income tax rate difference | | ( | % | |||
Other | | ( | % | |||
Income tax expense (recovery) reported in the consolidated statements of loss | $ | — | - | % | ||
(1) Our Canadian corporate tax rate is comprised of a basic Part I federal tax rate of
The Company has incurred Canadian federal and provincial net operating losses, or “NOLs,” from inception. As of December 31, 2025, the Company has NOL carry-forwards of approximately $
The Company has incurred NOLs for U.S. tax purposes. As of December 31, 2025, the Company has carry-forwards of approximately $
Deferred income taxes reflect the net tax effects of temporary differences between the carrying amounts of assets and liabilities for financial reporting purposes and the amounts used for income tax purposes. The net deferred tax assets have not been recognized in these financial statements because the criteria for recognition of these assets were not met.
119
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
The Company’s net deferred tax assets consist of the following for the years ended December 31:
| 2025 | 2024 | ||
Net operating loss carry‑forwards | $ | | $ | |
Tax basis of property and equipment in excess of carrying values |
| | | |
Tax basis of right of use assets | ( | ( | ||
Tax basis of lease liability | | | ||
Tax basis of reserves | | | ||
Federal SR&ED investment tax credits |
| | | |
Taxation of federal SR&ED investment tax credits |
| ( | ( | |
Research and development expenditures |
| | | |
Financing costs |
| | | |
Stock based compensation | | | ||
Others |
| | | |
Total Net deferred tax assets |
| | | |
Less Valuation allowance | ( | ( | ||
Net deferred tax assets | $ | — | $ | — |
The Company files income tax returns in Canada and in the United States. The Company is subject to Canada Revenue Agency and Revenu Québec examination for fiscal years 2020 to 2025 due to unexpired statute of limitation periods and is subject to US Federal and state income tax examination for fiscal years 2022 to 2025.
On July 4, 2025, the President of the United States signed H.R. 1, the “One Big Beautiful Bill Act,” into law. The legislation includes several changes to federal tax law that generally allow for more favorable deductibility of certain business expenses beginning in 2025, including the restoration of immediate expensing of domestic R&D expenditures, reinstatement of 100% bonus depreciation, and more favorable rules for determining the limitation on business interest expense. These changes were reflected in the income tax provision for the period ended December 31, 2025, as enactment occurred after the balance sheet date. The Company has elected to continue capitalizing domestic R&D expenditures and will continue to amortize prior year R&D expenditures that were capitalized.
14. Government assistance
The Company incurs research and development expenditures that are eligible for investment tax credits. The investment tax credits recorded are based on management’s estimates of amounts expected to be recovered and are subject to audit by the taxation authorities. These amounts have been recorded as a reduction of research and development expenditures in the years ended December 31, 2025 and 2024 for an amount of $
15. Commitments
In the normal course of business, the Company enters into contracts with clinical research organizations, drug manufacturers and other vendors for preclinical and clinical research studies, research and development supplies and other services and products for operating purposes. These contracts generally provide for termination with reasonable notice or upon certain circumstances, and therefore are cancellable contracts. Therefore, as of December 31, 2025, there are
16. Currency risk
The Company is exposed to the financial risk related to the fluctuation of foreign exchange rates and the degree of volatility of those rates. The foreign currency risk is limited to the portion of the Company’s business transactions denominated in
120
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
currency other than US dollars. The following table provides an indication of the Company’s exposure to the Canadian dollar, which is expressed in US dollars as of December 31:
| 2025 | | 2024 | |||
Cash and cash equivalents | $ | | $ | | ||
Short-term investments | | | ||||
Other receivables | | | ||||
Operating lease assets | | | ||||
Accounts payable and accrued liabilities | ( | ( | ||||
Operating lease liabilities | ( | ( | ||||
Net financial position exposure | $ | | $ | | ||
The Company does not enter into arrangements to hedge its currency risk exposure.
17. Fair value of financial instruments
Pursuant to the accounting guidance for fair value measurement and its subsequent updates, fair value is defined as the price that would be received to sell an asset or paid to transfer a liability (i.e. the exit price) in an orderly transaction between market participants at the measurement date. The accounting guidance establishes a hierarchy for inputs used in measuring fair value that minimizes the use of unobservable inputs by requiring the use of observable market data when available. Observable inputs are inputs that market participants would use in pricing the asset or liability based on active market data. Unobservable inputs are inputs that reflect the assumptions market participants would use in pricing the asset or liability based on the best information available in the circumstances.
The fair value hierarchy is broken down into the three input levels summarized below:
Level 1 | — | Valuations are based on quoted prices in active markets for identical assets or liabilities and readily accessible by the Company at the reporting date. |
Level 2 | — | Valuations based on inputs other than the quoted prices in active markets that are observable either directly or indirectly in active markets. |
Level 3 | — | Valuations based on unobservable inputs in which there is little or no market data, which requires the Company to develop its own assumptions. |
For the years ending December 31, 2025 and December 31, 2024, there were
18. Royalty Purchase Agreement
On March 27, 2023, the Company entered into a purchase and sale agreement, or the “Royalty Purchase Agreement,” with RTW and certain of its affiliates.
Pursuant to the Royalty Purchase Agreement, RTW agreed to purchase, following the U.S. Food and Drug Administration approval of etripamil (subject to certain conditions), at a purchase price of $
121
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
19. Other receivables
Other receivables comprised the following as of December 31:
| December 31, 2025 | | December 31, 2024 | |||
Interest receivable |
| $ | | $ | | |
Sales tax receivable |
|
| |
| | |
Clinical receivable | — | | ||||
Employee withholding taxes receivable | | — | ||||
Other current receivable | | | ||||
Total |
| $ | | $ | | |
20. Segment Reporting
The Company manages its operations as a operating segment for the purpose of assessing performance and making operating decisions while focusing on the development and commercialization of innovative cardiovascular medicines. These operations are focused on a single product, which are reported on a consolidated basis. The accounting policies of the single operating segment are the same as those described in the summary of significant accounting policies. The chief operating decision maker, or “CODM,” assesses performance of the Company’s single operating segment based on consolidated net loss. Net loss is used by the CODM to evaluate budget to actual analytics, which are used to monitor the single segment spend and confirm the Company is meeting established budgetary goals. The CODM is the principal officer group, which includes the Company’s chief executive officer and chief financial officer.
The following table presents information about the Company’s significant expenses, as provided to the Company’s CODM, and includes a reconciliation to consolidated net loss:
Year ended December 31, | ||||||||
(in thousands) | | 2025 | | 2024 | ||||
Revenue | $ | | $ | — | ||||
Less: | ||||||||
Research and development, net of tax credits, excluding share-based compensation | | | ||||||
General and administrative, excluding share-based compensation |
| |
| | ||||
Commercial, excluding share-based compensation |
| |
| | ||||
Share-based compensation expense | | | ||||||
Interest income |
| ( | ( | |||||
Interest expense | | | ||||||
Net loss | $ | ( | $ | ( | ||||
The measure of segment assets is reported on the balance sheet as total consolidated assets.
122
Milestone Pharmaceuticals Inc.
Notes to Consolidated Financial Statements
(in thousands of US dollars, except share and per share data)
21. Subsequent Events
Closing of sale of royalty interest
As previously disclosed, on March 27, 2023, the Company entered into the Royalty Purchase Agreement with RTW, pursuant to which RTW agreed to purchase, following FDA approval of etripamil (subject to certain conditions) for the treatment of PSVT, at a purchase price of $
On January 12, 2026, the Company closed the sale of Royalty interest under the Royalty Purchase Agreement and received cash of $
Sale of common shares under 2025 Offering and Open Market Sale AgreementSM
Subsequent to the year ended December 31, 2025, through to the date of issuance of the financial statements, the Company raised aggregate gross proceeds of $
Specifically,
123
ITEM 9. CHANGES IN AND DISAGREEMENTS WITH ACCOUNTANTS ON ACCOUNTING AND FINANCIAL DISCLOSURE.
None.
ITEM 9A. CONTROLS AND PROCEDURES.
Evaluation of Disclosure Controls and Procedures
We maintain disclosure controls and procedures, as defined in Rules 13a-15(e) and 15d-15(e) under the Securities Exchange Act of 1934 (the Exchange Act) that are designed to ensure that information required to be disclosed in our periodic and current reports that we file with the SEC is recorded, processed, summarized and reported within the time periods specified in the SEC’s rules and forms, and that such information is accumulated and communicated to our management, including our principal executive officer and principal financial officer, as appropriate, to allow timely decisions regarding required disclosure. In designing and evaluating the disclosure controls and procedures, management recognizes that any controls and procedures, no matter how well designed and operated, can provide only reasonable and not absolute assurance of achieving the desired control objectives. In reaching a reasonable level of assurance, management necessarily is required to apply its judgment in evaluating the cost-benefit relationship of possible controls and procedures. In addition, the design of any system of controls also is based in part upon certain assumptions about the likelihood of future events, and there can be no assurance that any design will succeed in achieving its stated goals under all potential future conditions; over time, control may become inadequate because of changes in conditions, or the degree of compliance with policies or procedures may deteriorate. Because of the inherent limitations in a cost-effective control system, misstatements due to error or fraud may occur and not be detected.
We carried out an evaluation, under the supervision and with the participation of our management, including our principal executive officer and principal financial officer, of the effectiveness of the design and operation of our disclosure controls and procedures, as defined in Rules 13a-15(e) and 15d-15(e) under the Exchange Act. Based on this evaluation, our principal executive officer and principal financial officer concluded that our disclosure controls and procedures were effective at the reasonable assurance level as of December 31, 2025.
Management’s Report on Internal Control Over Financial Reporting
Our management is responsible for establishing and maintaining adequate internal control over financial reporting as such term is defined in Rules 13a-15(f) and 15-d-15(f) of the Exchange Act. Internal control over financial reporting is a process designed under the supervision and with the participation of our management, including our principal executive officer and principal financial officer, to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with accounting principles generally accepted in the United States of America.
As of December 31, 2025, management assessed and management concluded the effectiveness of internal control over financial reporting using the criteria set forth by the Committee of Sponsoring Organizations of the Treadway Commission in Internal Control – 2013 Integrated Framework (2013 Framework). Based on this assessment, management concluded that our internal control over financial reporting was effective as of December 31, 2025.
This Annual Report on Form 10-K does not include an attestation report of our independent registered public accounting firm due to our status as a non-accelerated filer.
Inherent Limitations of Internal Controls
Our management, including our principal executive officer and principal financial officer, does not expect that our disclosure controls and procedures or our internal controls will prevent all errors and all fraud. A control system, no matter how well conceived and operated, can provide only reasonable, not absolute, assurance that the objectives of the control system are met. Because of the inherent limitations in all control systems, no evaluation of controls can provide absolute assurance that all control issues and instances of fraud, if any, within the Company have been detected. These inherent
124
limitations include the realities that judgments in decision-making can be faulty, and that breakdowns can occur because of a simple error or mistake. Additionally, controls can be circumvented by the individual acts of some persons, by collusion of two or more people, or by management override of the control. The design of any system of controls also is based in part upon certain assumptions about the likelihood of future events, and there can be no assurance that any design will succeed in achieving its stated goals under all potential future conditions. Over time, controls may become inadequate because of changes in conditions, or the degree of compliance with the policies or procedures may deteriorate. Because of the inherent limitations in a cost-effective control system, misstatements due to error or fraud may occur and not be detected.
Changes in Internal Control Over Financial Reporting
During the quarter ended December 31, 2025, there have been no changes in our internal control over financial reporting, as such term is defined in Rules 13a-15(f) and 15d-15(f) promulgated under the Exchange Act, that have materially affected, or are reasonably likely to materially affect, our internal control over financial reporting.
ITEM 9B. OTHER INFORMATION.
Rule 10b5-1 Trading Arrangements
ITEM 9C. DISCLOSURE REGARDING FOREIGN JURISDICTIONS THAT PREVENT INSPECTIONS.
Not applicable.
125
PART III
Certain information required by Part III is omitted from this report because we will file with the SEC a definitive proxy statement pursuant to Regulation 14A, or the “2026 Proxy Statement,” no later than 120 days after the end of our fiscal year, and certain information included therein is incorporated herein by reference.
ITEM 10. DIRECTORS, EXECUTIVE OFFICERS AND CORPORATE GOVERNANCE
The information required by this item is incorporated by reference to the information set forth in the sections titled “Proposal No. 1: Election of Directors,” “Proposal No. 1: Election of Directors─Information Regarding the Board of Directors and Corporate Governance,” “Executive Officers,” and “Delinquent Section 16(a) Reports,” if applicable, in our 2026 Proxy Statement. Such information to be included in our 2026 Proxy Statement is incorporated herein by reference.
The information required by Item 408(b) of Regulation S-K will be set forth in the section titled “Insider Trading Arrangements and Policies” in our 2026 Proxy Statement and is incorporated by reference herein.
Information regarding our Code of Business Conduct and Ethics, or the Code of Conduct, required by this item will be contained in our 2026 Proxy Statement under the caption “Proposal No. 1: Election of Directors─Information Regarding the Board of Directors and Corporate Governance─Code of Business Conduct and Ethics,” and is hereby incorporated by reference. We intend to promptly disclose on our website or in a Current Report on Form 8-K in the future (i) the date and nature of any amendment (other than technical, administrative or other non-substantive amendments) to the Code of Conduct that applies to our principal executive officer, principal financial officer, principal accounting officer or controller or persons performing similar functions and relates to any element of the code of ethics definition enumerated in Item 406(b) of Regulation S-K and (ii) the nature of any waiver, including an implicit waiver, from a provision of the Code of Conduct that is granted to one of these specified individuals that relates to one or more of the elements of the code of ethics definition enumerated in Item 406(b) of Regulation S-K, the name of such person who is granted the waiver and the date of the waiver. The full text of our Code of Conduct is available at the investors section of our website at www.milestonepharma.com. The reference to our website address does not constitute incorporation by reference of the information contained at or available through our website, and you should not consider it to be a part of this Annual Report.
ITEM 11. EXECUTIVE COMPENSATION.
The information required by this item is incorporated by reference to the information set forth in the section titled “Executive Compensation” in our 2026 Proxy Statement and is incorporated by reference herein, provided that the information required by Item 402(x) of Regulation S-K shall be set forth in the section titled "Policies and practices related to the grant of certain equity awards close in time to the release of material nonpublic information" in our 2026 Proxy Statement and is incorporated by reference herein.
ITEM 12. SECURITY OWNERSHIP OF CERTAIN BENEFICIAL OWNERS AND MANAGEMENT AND RELATED STOCKHOLDER MATTERS.
The information required by this item is incorporated by reference to the information set forth in the section titled “Security Ownership of Certain Beneficial Owners and Management” in our 2026 Proxy Statement.
ITEM 13. CERTAIN RELATIONSHIPS AND RELATED TRANSACTIONS, AND DIRECTOR INDEPENDENCE.
The information required by this item is incorporated by reference to the information set forth in the sections titled “Certain Relationships and Related Transactions” and “Proposal No. 1: Election of Directors─Information Regarding the Board of Directors and Corporate Governance─Director Independence” in our 2026 Proxy Statement.
126
ITEM 14. PRINCIPAL ACCOUNTANT FEES AND SERVICES.
The information required by this item is incorporated by reference to the information set forth in the section titled “Proposal No. 2: Appointment of Auditor─Auditor Fees” in our 2026 Proxy Statement.
127
PART IV
ITEM 15. EXHIBITS, FINANCIAL STATEMENT SCHEDULES.
(a)(1) Financial Statements
See Index to Consolidated Financial Statements on page 103 of this Annual Report on Form 10-K, which is incorporated into this item by reference.
(a)(2) Financial Statement Schedules
All financial statement schedules are omitted because they are not applicable or the required information is shown in the financial statements or notes thereto.
(b) Exhibits
The following list of exhibits includes exhibits submitted with this Annual Report on Form 10-K as filed with the SEC and others incorporated by reference to other filings.
EXHIBIT | DESCRIPTION |
3.1 | |
3.2 | |
4.1 | |
4.2 | |
4.3 | |
4.4 | |
4.5 | |
4.6 | |
4.7 | |
4.8 | |
4.9 | |
4.10 | |
10.1+ | |
10.2+ | |
10.3+ | |
10.4+ |
128
129
10.20* | |
10.21*♦ | |
10.22 | |
10.23 | |
10.24*♦ | |
10.25*♦ | |
10.26 | |
10.27 | |
23.1 | Consent of PricewaterhouseCoopers LLP, Independent Registered Public Accounting Firm. |
24.1 | Power of Attorney (included on the signature page to this registration statement). |
31.1 | |
31.2 | |
32.1˄ | |
97.1 | |
101.INS | Inline XBRL Instance Document |
101.SCH | Inline XBRL Taxonomy Extension Schema Document |
101.CAL | Inline XBRL Taxonomy Extension Calculation Linkbase Document |
101.DEF | Inline XBRL Taxonomy Extension Definition Linkbase Document |
101.LAB | Inline XBRL Taxonomy Extension Label Linkbase Document |
101.PRE | Inline XBRL Taxonomy Extension Presentation Linkbase Document |
104 | Cover Page Interactive Data File (formatted as inline XBRL with applicable taxonomy extension information contained in Exhibit 101) |
* | Certain portions of this exhibit have been omitted pursuant to Item 601(b)(10) of Regulation S-K. The Registrant hereby undertakes to furnish to the SEC, upon request, copies of any such instruments. |
♦ | In accordance with Item 601(b)(10)(iv) of Regulation S-K, certain information (indicated by “[***]”) has been excluded from this exhibit. |
+ | Indicates a management contract or compensatory plan |
130
˄ These certifications are being furnished solely to accompany this Annual Report pursuant to 18 U.S.C. Section 1350, and are not being filed for purposes of Section 18 of the Securities Exchange Act of 1934, as amended, and are not to be incorporated by reference into any filing of the Registrant, whether made before or after the date hereof, regardless of any general incorporation language in such filing.
ITEM 16. FORM 10-K SUMMARY
Not applicable
131
SIGNATURES
Pursuant to the requirements of Section 13 or 15(d) of the Securities Exchange Act of 1934, the Company has duly caused this report to be signed on its behalf by the undersigned, thereunto duly authorized.
Milestone Pharmaceuticals Inc. | |
Dated: March 20, 2026 | /s/ Joseph Oliveto |
Joseph Oliveto | |
Chief Executive Officer | |
POWER OF ATTORNEY
KNOW ALL BY THESE PRESENTS, that each person whose signature appears below constitutes and appoints Joseph Oliveto and Amit Hasija, and each of them, as his or her true and lawful attorneys-in-fact and agents, each with the full power of substitution, for him or her and in his or her name, place or stead, in any and all capacities, to sign any and all amendments to this report, with exhibits thereto and other documents in connection therewith, with the Securities and Exchange Commission, granting unto said attorneys-in-fact and agents, and each of them, full power and authority to do and perform each and every act and thing requisite and necessary to be done in and about the premises, as fully to all intents and purposes as he or she might or could do in person, hereby ratifying and confirming all that said attorneys-in-fact and agents, or their, his substitute or substitutes, may lawfully do or cause to be done by virtue hereof.
Pursuant to the requirements of the Securities Exchange Act of 1934, this report has been signed below by the following persons on behalf of the Company and in the capacities indicated on the 20th of March 2026.
/s/ Joseph Oliveto | | Chief Executive Officer and Director | |
Joseph Oliveto | (principal executive officer) | ||
/s/ Amit Hasija | Chief Financial Officer | ||
Amit Hasija | (principal financial officer and principal accounting officer) | ||
/s/ Robert J. Wills | Chairman of the Board | ||
Robert Wills | |||
/s/ Seth Fischer | Director | ||
Seth Fischer | |||
/s/ Lisa Giles | Director | ||
Lisa M. Giles | |||
/s/ Michael Tomsicek | Director | ||
Michael Tomsicek | |||
/s/ Andrew Saik | Director | ||
Andrew Saik | |||
/s/ Stuart Duty | Director | ||
Stuart Duty | |||
/s/ Joseph Papa | Director | ||
Joseph Papa | |||
132
Exhibit 4.5
DESCRIPTION OF THE REGISTRANT’S SECURITIES
REGISTERED PURSUANT TO SECTION 12 OF THE
SECURITIES EXCHANGE ACT OF 1934
The following description sets forth certain material terms and provisions of the securities of Milestone Pharmaceuticals Inc. (the “Company”) that are registered under Section 12 of the Securities Exchange Act of 1934, as amended. The following description of our securities is intended as a summary only and is qualified in its entirety by reference to our articles of incorporation and amendments thereto and our bylaws, each of which are filed as exhibits to the Annual Report on Form 10-K of which this description is a part, and to the applicable provisions of the Business Corporations Act (Québec) (BCA).
General
Our authorized share capital consists of an unlimited number of common shares, no par value per share, and an unlimited number of preferred shares, no par value per share, which are issuable in one or more series.
Common Shares
Voting Rights
Under our articles of incorporation, the holders of common shares are entitled to one vote for each share held at any meeting of our shareholders.
Dividends
Subject to the prior rights of holders of our preferred shares, if applicable, the holders of common shares are entitled to receive dividends as and when declared by our board of directors. We have never declared or paid cash dividends on our share capital, and we do not currently intend to pay any cash dividends on our share capital in the foreseeable future. We currently intend to retain all available funds and any future earnings, if any, to fund the development and expansion of our business. Any future determination related to dividend policy will be made at the discretion of our board of directors, subject to applicable laws, and will depend upon, among other factors, our results of operations, financial condition, contractual restrictions and capital requirements. In addition, our ability to pay cash dividends on our share capital in the future may be limited by the terms of any future debt or preferred securities we issue or any credit facilities we enter into.
Liquidation
Subject to the prior payment to holders of our preferred shares, if any, in the event of our liquidation, dissolution or winding-up or other distribution of our assets among our shareholders, the holders of common shares are entitled to share pro rata in the distribution of the balance of our assets.
Rights and Preferences
The holders of common shares have no preemptive, conversion rights or other subscription rights. There are no redemption or sinking fund provisions applicable to our common shares. There is no provision in our articles of incorporation requiring the holders of common shares to contribute additional capital or permitting or restricting the issuance of additional securities or any other material restrictions. The rights, preferences and privileges of the holders of common shares may be subject to, and adversely affected by, the rights of the holders of any series of preferred shares that we may designate in the future.
Preferred Shares
We do not have any preferred shares outstanding. Under our articles of incorporation, we are authorized to issue, without shareholder approval, an unlimited number of preferred shares, issuable in one or more
series, and, subject to the provisions of the BCA, having such designations, rights, privileges, restrictions and conditions, including dividend and voting rights, as our board of directors may determine, and such rights and privileges, including dividend and voting rights, may be superior to those of the common shares. The issuance of preferred shares, while providing flexibility in connection with possible acquisitions and other corporate purposes, could, among other things, have the effect of delaying, deferring or preventing a change in control of our company and might adversely affect the market price of our common shares and the voting and other rights of the holders of common shares. We have no current plans to issue any preferred shares.
Warrants to Purchase Common Shares
Common Stock Warrants
On July 11, 2025, we issued (i) Series A common warrants (the “Series A Common Warrants”) to purchase an aggregate of 31,500,000 common shares and (ii) Series B common warrants (the “Series B Common Warrants” and, together with the Series A Common Warrants, the “Common Warrants”) to purchase an aggregate of 31,500,000 common shares.
Form
The form of Series A Common Warrants was filed as Exhibit 4.2 to our Current Report on Form 8-K filed with the SEC on July 14, 2025. The form of Series B Common Warrants was filed as Exhibit 4.3 to our Current Report on Form 8-K filed with the SEC on July 14, 2025.
Fractional Shares
No fractional shares will be issued in connection with any the exercise the Common Warrants. In lieu of any fractional shares that would otherwise be issuable, the number of common shares to be issued shall be rounded down to the next whole number and we shall pay the holder of the Common Warrants in cash the fair market value based on the closing sale price for any such fractional shares.
Exercise Limitations
Under the terms of the Common Warrants, we may not give effect to the exercise of any such Common Warrant, and a holder will not be entitled to exercise any portion of any such Common Warrants, if, upon giving effect to such exercise, the aggregate number of common shares beneficially owned by the holder (together with its affiliates, any other persons acting as a group together with the holder or any of the holder’s affiliates, and any other persons whose beneficial ownership of common shares would or could be aggregated with the holder’s for purposes of Section 13(d) or Section 16 of the Securities Exchange Act of 1934, as amended, or the Exchange Act) would exceed 4.99% or 9.99%, as elected by the purchaser, of the number of common shares outstanding immediately after giving effect to such exercise. By giving written notice to us, a holder may increase or decrease the maximum percentage to any other percentage specified not in excess of 19.99%; provided that any such increase will not be effective until the sixty-first day after such notice is delivered to us.
Transferability
Subject to applicable laws, the Common Warrants may be offered for sale, sold, transferred or assigned without our consent.
Exchange Listing
There is no established trading market for the Common Warrants, and we do not expect a market to develop. We do not intend to apply for the listing of Common Warrants on the Nasdaq Stock Market, any other national securities exchange or any other nationally recognized trading system.
No Rights as a Shareholder
Except by virtue of such holder’s ownership of shares of our common shares, the holder of a Common Warrant does not have the rights or privileges of a holder of our common shares, including any voting rights, until such holder exercises the Common Warrant.
Registration of Shares Underlying Common Warrants
The common shares underlying the Common Warrants were registered on a Prospectus Supplement, which was filed on July 14, 2025, to the Prospectus included in the Form S-3 registration statement, which was declared effective on November 22, 2024.
Exercise Price and Term
Each Series A Common Warrant entitles the holder thereof to purchase one common share at an exercise price equal to $1.50 per share (or $1.499 per Pre-Funded Warrant), will be immediately exercisable and will expire on the date that is one year from the date of issuance. Each Series B Common Warrant entitles the holder thereof to purchase one common share at an exercise price equal to $1.875 per share (or $1.874 per Pre-Funded Warrant), will be immediately exercisable and will expire five years from the date of issuance. In addition, for so long as the volume weighted average price of the our common shares on each of the preceding 10 consecutive trading days is at least $3.50, we may force the exercise of the Series B Common Warrants, in whole or in part, in cash, by delivering a notice of mandatory exercise to the holders. In the event that we force the exercise of the Series B Common Warrants, to the extent that the holder would own more than the Beneficial Ownership Limitation (as defined in the Common Warrants), the holder would receive a Pre-Funded Warrant for those number of common shares underlying such Series B Common Warrant which would cause the holder to otherwise exceed the Beneficial Ownership Limitation.
Fundamental Transactions
Upon the consummation of a fundamental transaction (as described in the Common Warrants, and, among other things, including the sale, transfer or other disposition of all or substantially all of our properties or assets, our consolidation or merger with or into another person in which the Company is not the surviving person, the acquisition of more than 50% of our outstanding common shares, any person or group becoming the beneficial owner of 50% of the voting power of our outstanding common shares or any reorganization, recapitalization or reclassification of our common shares), the holders of the Common Warrants will be entitled to receive, upon exercise of the Common Warrant, the same amount and kind of securities, cash or property as the holder would have been entitled to receive upon the occurrence of such fundamental transaction if the holder had been, immediately prior to such fundamental transaction, the holder of the number of common shares underlying the Common Warrant then issuable upon exercise in full of the Common Warrant without regard to any limitations on exercise contained herein. Notwithstanding the foregoing, in the event of a fundamental transaction, and subject to certain exceptions, the holders of the Common Warrants have the right to require us or a successor entity to redeem the Common Warrants for cash in the amount of the Black-Scholes Value (as defined in the Common Warrants) of the unexercised portion of the Common Warrants. However, in the event of a fundamental transaction which is not in the Company’s control, including a fundamental transaction not approved by the Company’s board of directors, the holders of the Common Warrants will only be entitled to receive from the Company or its successor entity the same type or form of consideration (and in the same proportion) at the Black Scholes Value of the unexercised portion of the Common Warrants that is being offered and paid to the holders of the Company’s common shares in connection with the fundamental transaction, whether that consideration be in the form of cash, shares or any combination thereof, or whether the holders of the Company’s common shares are given the choice to receive from among alternative forms of consideration in connection with the fundamental transaction.
Pre-Funded Warrants
On March 4, 2024, we issued pre-funded warrants (the “2024 Pre-Funded Warrants”) to purchase 3,333,333 common shares and on July 11, 2025, we issued pre-funded warrants (the “2025 Pre-Funded Warrants”) to purchase 3,502,335 common stock.
Form
The Pre-Funded Warrants were issued as individual warrant agreements to certain investors. The form of 2024 Pre-Funded Warrant was filed as Exhibit 4.1 to our Current Report on Form 8-K filed with the SEC on March 4, 2024. The form of 2025 Pre-Funded Warrant was filed as Exhibit 4.1 to our Current Report on Form 8-K filed with the SEC on July 14, 2025.
Fractional Shares
No fractional shares will be issued in connection with any the exercise the Pre-Funded Warrants. In lieu of any fractional shares that would otherwise be issuable, the number of common shares to be issued shall be rounded down to the next whole number and we shall pay the holder of the Pre-Funded Warrant in cash the fair market value based on the closing sale price for any such fractional shares.
Exercise Limitations
Under the terms of the Pre-Funded Warrants, we may not give effect to the exercise of any such Pre-Funded Warrant, and a holder will not be entitled to exercise any portion of any such Pre-Funded Warrant, if, upon giving effect to such exercise, the aggregate number of common shares beneficially owned by the holder (together with its affiliates, any other persons acting as a group together with the holder or any of the holder’s affiliates, and any other persons whose beneficial ownership of common shares would or could be aggregated with the holder’s for purposes of Section 13(d) or Section 16 of the Securities Exchange Act of 1934, as amended, or the Exchange Act, would exceed 4.99% or 9.99%, as elected by the purchaser, of the number of common shares outstanding immediately after giving effect to such exercise. By giving written notice to us, a holder may increase or decrease the maximum percentage to any other percentage specified not in excess of 19.99%; provided that any such increase will not be effective until the sixty-first day after such notice is delivered to us.
Transferability
Subject to applicable laws, the Pre-Funded Warrants may be offered for sale, sold, transferred or assigned without our consent.
Exchange Listing
There is no established trading market for the Pre-Funded Warrants, and we do not expect a market to develop. We do not intend to apply for the listing of Pre-Funded Warrants on the Nasdaq Stock Market, any other national securities exchange or any other nationally recognized trading system.
No Rights as a Shareholder
Except by virtue of such holder’s ownership of shares of our common shares, the holder of a Pre-Funded Warrant does not have the rights or privileges of a holder of our common shares, including any voting rights, until such holder exercises the Pre-Funded Warrant.
Registration of Shares Underlying Pre-Funded Warrants
The common shares underlying the 2024 Pre-Funded Warrants were registered on a Prospectus Supplement, which was filed on February 28, 2024, to the Prospectus included in the Form S-3 registration statement, which was declared effective on February 2, 2022. The common shares underlying the 2025 Pre-Funded Warrants were registered on a Prospectus Supplement, which was filed on July 14, 2025, to the Prospectus included in the Form S-3 registration statement, which was declared effective on November 22, 2024.
Exercise Price
Each Pre-Funded Warrant has an exercise price of $0.001 per share. In lieu of making the cash payment otherwise contemplated to be made to the Company upon exercise of a Pre-Funded Warrant in payment
of the aggregate exercise price, the holder may elect instead to receive upon such exercise (either in whole or in part) the net number of common shares determined according to a formula set forth in the Pre-Funded Warrants.
Term
Each Pre-Funded Warrant is exercisable immediately and is exercisable until the Pre-Funded Warrant is exercised in full.
Fundamental Transactions
Upon the consummation of a fundamental transaction (as described in the Pre-Funded Warrants, and, among other things, including the sale, transfer or other disposition of all or substantially all of our properties or assets, our consolidation or merger with or into another person in which the Company is not the surviving person, the acquisition of more than 50% of our outstanding common shares, any person or group becoming the beneficial owner of 50% of the voting power of our outstanding common shares or any reorganization, recapitalization or reclassification of our common shares), the holders of the Pre-Funded Warrants will be entitled to receive, upon exercise of the Pre-Funded Warrant, the same amount and kind of securities, cash or property as the holder would have been entitled to receive upon the occurrence of such fundamental transaction if the holder had been, immediately prior to such fundamental transaction, the holder of the number of common shares underlying the Pre-Funded Warrant then issuable upon exercise in full of the Pre-Funded Warrant without regard to any limitations on exercise contained herein.
Transfer Agent and Registrar
Our transfer agent and registrar for our common shares is Computershare Investor Services Inc., with an address of 650 de Maisonneuve Blvd W., 7th Floor, Montréal, Quebec H3A 3T2.
Nasdaq Global Market Listing
Our common shares are listed on The Nasdaq Global Market under the trading symbol “MIST.”
Advance Notice Procedures and Shareholder Proposals
Under the BCA, shareholders may make proposals for matters to be considered at the annual general meeting of shareholders. Such proposals must be sent to us in advance of any proposed meeting by delivering a timely written notice in proper form to our registered office in accordance with the requirements of the BCA. The notice must include information on the business the shareholder intends to bring before the meeting.
In addition, our bylaws require that shareholders provide us with advance notice of their intention to nominate any persons, other than those nominated by management, for election to our board of directors at a meeting of shareholders.
These provisions could have the effect of delaying the nomination of certain persons for director that are favored by the holders of a majority of our outstanding voting securities.
Exhibit 10.22
AMENDMENT NO. 1 TO
PURCHASE AND SALE AGREEMENT
This Amendment No. 1 to the Purchase and Sale Agreement ("Amendment"), dated August 12, 2024 (the "Amendment Date"), is between RTW Royalty I DAC, a designated activity company incorporated under the laws of Ireland with registered number 734869 (the "Buyer"), and Milestone Pharmaceuticals Inc., a Quebec corporation (the "Company"), and amends that certain Purchase and Sale Agreement by and between the Buyer and the Company, dated as of March 27, 2023 (the "Agreement").
WHEREAS, in accordance with Section 3.1 of the Agreement, the parties hereto mutually wish to amend the Agreement to revise the Closing Date;
WHEREAS, Section 3.1 of the Agreement provides that the Closing can take place during such period as mutually agreed upon by the parties to the Agreement; and
WHEREAS, Section 10.4 of the Agreement provides that the Agreement may be amended, modified or supplemented only in a writing signed by each of the parties to the Agreement.
NOW THEREFORE, in consideration of the covenants and agreements set forth herein and for good and valuable consideration, the receipt and adequacy of which are hereby acknowledged, the Company and the Buyer hereby agree as follows:
1. | Defined Terms. Capitalized terms that are used in this Amendment have the meaning set forth in the Agreement, unless otherwise defined in this Amendment. |
2. | Closing Date. Effective as of the Amendment Date, Section 3.1 of the Agreement is amended and restated to read in its entirety as follows: |
Closing. The Closing shall take place remotely via the exchange of documents and signatures on the date that is thirty (30) calendar days, or such period as mutually agreed upon by the parties hereto, after the date on which the conditions set forth in ARTICLE 5 are satisfied or waived.
3. | Entire Agreement. The provisions of this Amendment supersede all provisions of the Agreement that are inconsistent with the provisions of this Amendment. The Agreement, as modified by this Amendment, remains in full force and effect and, with the Exhibits annexed thereto and the Disclosure Schedule constitute the entire understanding between the parties hereto with respect to the subject matter hereof and thereof and supersede all other understandings and negotiations with respect hereto and thereto. Unless the context otherwise requires, the term "Agreement" as used in the Agreement shall be deemed to refer to the Agreement as amended hereby. |
4. | Governing Law. This Amendment shall be exclusively governed by, and construed in accordance with, the laws of the State of New York without giving effect to any choice or |
1
conflict of law provision or rule that would cause the application of the laws of any other jurisdiction.
5. | Waiver. This Amendment shall not be deemed a waiver by any party of any of its rights or remedies under the Agreement, except to the extent expressly set forth in this Amendment. |
6. | Counterparts. This Amendment may be executed in any number of counterparts and by the parties hereto in separate counterparts, each of which when so executed shall be deemed to be an original and all of which taken together shall constitute one and the same agreement. Copies of executed counterparts transmitted by telecopy, facsimile or other similar means of electronic transmission, including "PDF," shall be considered original executed counterparts, provided receipt of such counterparts is confirmed. |
[Signature Page Follows]
2
IN WITNESS WHEREOF, the parties hereto have caused this Amendment to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| COMPANY | |
| | |
| MILESTONE PHARMACEUTICALS INC. | |
| | |
| By: | /s/ Amit Hasija |
| | Name: Amit Hasija |
| | Title: Chief Financial Officer |
| | |
| BUYER | |
| | |
| RTW ROYALTY I DAC | |
| | |
| By: | /s/ Roderick Wong |
| | Name: Roderick Wong, M.D. |
| | Title: Authorized Attorney |
[Signature Page to Amendment No. 1 to Purchase and Sale Agreement]
Exhibit 10.23
AMENDMENT NO. 2 TO
PURCHASE AND SALE AGREEMENT
This Amendment No. 2 to the Purchase and Sale Agreement (“Amendment”), dated July 10, 2025 (the “Amendment Date”), is between RTW Royalty I DAC, a designated activity company incorporated under the laws of Ireland with registered number 734869 (the “Buyer”), and Milestone Pharmaceuticals Inc., a Quebec corporation (the “Company”), and amends that certain Purchase and Sale Agreement by and between the Buyer and the Company, dated as of March 27, 2023, as amended by Amendment No. 1, dated August 12, 2024 (as amended, the “Agreement”).
WHEREAS, Section 10.4 of the Agreement provides that the Agreement may be amended, modified or supplemented only in writing signed by each of the parties to the Agreement.
NOW THEREFORE, in consideration of the covenants and agreements set forth herein and for good and valuable consideration, the receipt and adequacy of which are hereby acknowledged, the Company and the Buyer hereby agree as follows:
1. | Defined Terms. Capitalized terms that are used in this Amendment have the meaning set forth in the Agreement, unless otherwise defined in this Amendment. |
1.Amendment to Defined Terms. Effective as of the Amendment Date, Section 1.1 of the Agreement is amended by removing the definition of “Marketing Approval Deadline” and replacing such definition in its entirety as follows:
“Marketing Approval Deadline” means 11:59 pm P.T. December 31, 2025.
2. | Entire Agreement. The provisions of this Amendment supersede all provisions of the Agreement that are inconsistent with the provisions of this Amendment. The Agreement, as modified by this Amendment, remains in full force and effect and, with the Exhibits annexed thereto and the Disclosure Schedule constitute the entire understanding between the parties hereto with respect to the subject matter hereof and thereof and supersede all other understandings and negotiations with respect hereto and thereto. Unless the context otherwise requires, the term “Agreement” as used in the Agreement shall be deemed to refer to the Agreement as amended hereby. |
3. | Governing Law. This Amendment shall be exclusively governed by, and construed in accordance with, the laws of the State of New York without giving effect to any choice or conflict of law provision or rule that would cause the application of the laws of any other jurisdiction. |
4. | Waiver. This Amendment shall not be deemed a waiver by any party of any of its rights or remedies under the Agreement, except to the extent expressly set forth in this Amendment. |
5. | Counterparts. This Amendment may be executed in any number of counterparts and by the parties hereto in separate counterparts, each of which when so executed shall be deemed to |
1
be an original and all of which taken together shall constitute one and the same agreement. Copies of executed counterparts transmitted by telecopy, facsimile or other similar means of electronic transmission, including “PDF,” shall be considered original executed counterparts, provided receipt of such counterparts is confirmed.
[Signature Page Follows]
2
IN WITNESS WHEREOF, the parties hereto have caused this Amendment to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| COMPANY | |
| | |
| MILESTONE PHARMACEUTICALS INC. | |
| | |
| By: | /s/ Amit Hasija |
| | Name: Amit Hasija |
| | Title: Chief Financial Officer |
| | |
| BUYER | |
| | |
| RTW ROYALTY I DAC | |
| | |
| By: | /s/ Roderick Wong |
| | Name: Roderick Wong, M.D. |
| | Title: Authorized Attorney |
[Signature Page to Amendment No. 2 to Purchase and Sale Agreement]
Exhibit 10.24
CERTAIN CONFIDENTIAL INFORMATION CONTAINED IN THIS DOCUMENT, MARKED BY [***], HAS BEEN OMITTED BECAUSE IT IS BOTH (I) NOT MATERIAL AND (II) IS THE TYPE THAT MILESTONE PHARMACEUTICALS INC. TREATS AS PRIVATE OR CONFIDENTIAL
AMENDMENT NO. 3 TO AND ASSIGNMENT OF
PURCHASE AND SALE AGREEMENT
This Amendment No. 3 to and Assignment of the Purchase and Sale Agreement (this “Amendment”), dated December 12, 2025 (the “Amendment Date”), is among RTW Royalty I DAC, a designated activity company incorporated under the laws of Ireland with registered number 734869 (the “Buyer”), Milestone Pharmaceuticals Inc., a Quebec corporation (“Holdings”) and Milestone Pharmaceuticals USA, Inc., a Delaware corporation (the “US Subsidiary”), and amends that certain Purchase and Sale Agreement by and between the Buyer and Holdings, dated as of March 27, 2023, as amended by Amendment No. 1, dated August 12, 2024 and Amendment No. 2, dated July 10, 2025 (as amended, the “Existing Agreement”; and as further amended by this Amendment, the “Agreement”).
WHEREAS, Section 10.3 of the Existing Agreement provides that Holdings as the “Seller” under the Existing Agreement must obtain the prior written consent of Buyer to assign the obligations to make Royalty Payments and the CoC Payment to the US Subsidiary prior to the Closing Date; and
WHEREAS, Section 10.4 of the Existing Agreement provides that the Existing Agreement may be amended, modified or supplemented only in writing signed by each of the parties to the Existing Agreement.
NOW THEREFORE, in consideration of the covenants and agreements set forth herein and for good and valuable consideration, the receipt and adequacy of which are hereby acknowledged, the Buyer, Holdings and the US Subsidiary hereby agree as follows:
1. | Defined Terms. Capitalized terms that are used in this Amendment have the meaning set forth in the Agreement, unless otherwise defined in this Amendment. |
2. | Condition to Amendment. At or prior to the Amendment Date, the Seller shall have paid the Transaction Expenses incurred prior to or on the Amendment Date to the extent invoiced two (2) Business Days prior to the Amendment Date. |
3. | Amendments to Existing Agreement. Effective as of the Amendment Date and concurrent with the assignment set forth in Section 4 below, the Existing Agreement is hereby amended to delete all stricken text (indicated textually in the same manner as the following example: stricken text) and to add all double-underlined text (indicated textually in the same manner as the following example: double-underlined text) as set forth in the conformed Agreement attached hereto as Exhibit A, |
4. | Assignments of the Agreement. Effective as of the Amendment Date and concurrent with the amendment set forth in Section 3 above, Holdings hereby irrevocably assigns to the US Subsidiary, and the US Subsidiary hereby irrevocably assumes from Holdings, all of Holdings’ rights and obligations in its capacity as the “Seller” under the Agreement and any other documents or instruments delivered pursuant thereto. Such assignment is without recourse to the Buyer or Holdings, as the assignor, and, except as expressly provided in the Agreement, without representation or warranty by Holdings. |
5. | Entire Agreement. The provisions of this Amendment supersede all provisions of the Existing Agreement that are inconsistent with the provisions of this Amendment. The Existing Agreement, as modified by this Amendment, remains in full force and effect and, with the Exhibits annexed thereto and the Disclosure Schedule constitute the entire understanding between the parties hereto with respect to the subject matter hereof and thereof and supersede all other understandings and negotiations with respect hereto and thereto. Unless the context otherwise requires, the term “Agreement” as used in the Agreement shall be deemed to refer to the Existing Agreement as amended hereby. |
6. | Governing Law. This Amendment shall be exclusively governed by, and construed in accordance with, the laws of the State of New York without giving effect to any choice or conflict of law provision or rule that would cause the application of the laws of any other jurisdiction. |
7. | Waiver. This Amendment shall not be deemed a waiver by any party of any of its rights or remedies under the Agreement, except to the extent expressly set forth in this Amendment. |
8. | Counterparts. This Amendment may be executed in any number of counterparts and by the parties hereto in separate counterparts, each of which when so executed shall be deemed to be an original and all of which taken together shall constitute one and the same agreement. Copies of executed counterparts transmitted by telecopy, facsimile or other similar means of electronic transmission, including “PDF,” shall be considered original executed counterparts, provided receipt of such counterparts is confirmed. |
[Signature Page Follows]
2
IN WITNESS WHEREOF, the parties hereto have caused this Amendment to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| HOLDINGS: | ||
| | ||
| Milestone Pharmaceuticals Inc. | ||
| | ||
| By: | /s/Amit Hasija | |
| Name: | Amit Hasija | |
| Title: | Chief Financial Officer | |
| | ||
| US SUBSIDIARY: | ||
| | ||
| Milestone Pharmaceuticals USA, Inc. | ||
| | ||
| By: | /s/Amit Hasija | |
| Name: | Amit Hasija | |
| Title: | Chief Financial Officer | |
[Signature Page to Amendment No. 3 to Purchase and Sale Agreement]
IN WITNESS WHEREOF, the parties hereto have caused this Amendment to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| BUYER: | |
| | |
| RTW Royalty I DAC | |
| | |
| By: | /s/Roderick Wong |
| Name: Roderick Wong, M.D. | |
| Title: Authorized Attorney | |
[Signature Page to Amendment No. 3 to Purchase and Sale Agreement]
EXHIBIT A
Purchase and Sale Agreement as amended through Amendment No. 3
[Attached]
Execution Version
As amended by Amendment No.1, Amendment No.2 and Amendment No. 3
Purchase and Sale Agreement
By and Between
Milestone Pharmaceuticals USA, Inc. as assignee of Milestone Pharmaceuticals Inc.
AND
RTW Royalty I DAC
Dated as of March 27, 2023
TABLE OF CONTENTS
| Page | ||
ARTICLE 1. | DEFINITIONS | 1 | |
Section 1.1 | Definitions | 1 | |
Section 1.2 | Interpretation (Quebec). | 16 | |
ARTICLE 2. | PURCHASE, SALE AND ASSIGNMENT OF THE REVENUE PARTICIPATION RIGHT | 17 | |
Section 2.1 | Purchase, Sale and Assignment. | 17 | |
Section 2.2 | Purchase Price | 17 | |
Section 2.3 | No Assumed Obligations, Etc | 18 | |
ARTICLE 3. | CLOSING | 18 | |
Section 3.1 | Closing | 18 | |
Section 3.2 | Payment of Purchase Price | 18 | |
Section 3.3 | Bill of Sale | 18 | |
ARTICLE 4. | REPRESENTATIONS AND WARRANTIES | 18 | |
Section 4.1 | Seller’s Representations and Warranties | 18 | |
Section 4.2 | Buyer’s Representations and Warranties | 22 | |
Section 4.3 | No Implied Representations and Warranties | 22 | |
ARTICLE 5. | CONDITIONS TO CLOSING | 23 | |
Section 5.1 | Conditions to the Buyer’s Obligations | 23 | |
Section 5.2 | Conditions to the Seller’s Obligations | 24 | |
ARTICLE 6. | COVENANTS | 24 | |
Section 6.1 | Reports; Meetings | 25 | |
Section 6.2 | Royalty Payments; Royalty Reports; Buy-Back Option. | 25 | |
Section 6.3 | Disclosures | 26 | |
Section 6.4 | Inspections and Audits of the Seller | 27 | |
Section 6.5 | Intellectual Property Matters. | 27 | |
Section 6.6 | In-Licenses. | 28 | |
Section 6.7 | Permitted Licenses; Sales | 29 | |
Section 6.8 | Restricted Indebtedness | 29 | |
Section 6.9 | Diligence. | 30 | |
Section 6.10 | Efforts to Consummate Transactions | 30 | |
Section 6.11 | Continuing Efforts; Further Assurances. | 30 | |
Section 6.12 | Non-Impairment; Back-Up Security Interest | 30 | |
Section 6.13 | Certain Tax Matters. | 31 | |
Section 6.14 | Use of Proceeds | 31 | |
Section 6.15 | Intercompany License | 31 | |
Section 6.16 | Ownership of Seller | 32 | |
ARTICLE 7. | INDEMNIFICATION | 32 | |
i
Section 7.1 | General Indemnity | 32 | |
Section 7.2 | Notice of Claims | 32 | |
Section 7.3 | Claim Procedures | 32 | |
Section 7.4 | Limitations on Liability. | 33 | |
Section 7.5 | Exclusive Remedy | 33 | |
Section 7.6 | Tax Treatment of Indemnification Payments | 33 | |
Section 7.7 | Survival | 33 | |
ARTICLE 8. | CONFIDENTIALITY | 33 | |
Section 8.1 | Confidentiality | 33 | |
Section 8.2 | Authorized Disclosure. | 34 | |
ARTICLE 9. | TERMINATION | 35 | |
Section 9.1 | Mutual Termination | 35 | |
Section 9.2 | Termination upon Failure to Achieve U.S. Marketing Approval | 35 | |
Section 9.3 | Buyer Termination for Unexpected Contraindications | 35 | |
Section 9.4 | Automatic Termination | 35 | |
Section 9.5 | Survival | 35 | |
ARTICLE 10. | MISCELLANEOUS | 35 | |
Section 10.1 | Headings | 35 | |
Section 10.2 | Notices | 35 | |
Section 10.3 | Assignment | 36 | |
Section 10.4 | Amendment and Waiver. | 36 | |
Section 10.5 | Entire Agreement | 37 | |
Section 10.6 | No Third-Party Beneficiaries. | 37 | |
Section 10.7 | Governing Law | 37 | |
Section 10.8 | Jurisdiction; Venue. | 37 | |
Section 10.9 | Severability | 37 | |
Section 10.10 | Specific Performance | 38 | |
Section 10.11 | Counterparts | 38 | |
Section 10.12 | Relationship of the Parties | 38 | |
Section 10.13 | Expenses | 38 | |
Section 10.14 | Limited Recourse and Non-Petition. | 38 | |
Section 10.15 | Subordination of Obligations to Holdings. | 39 | |
ARTICLE 11. | GUARANTEE | 40 | |
Section 11.1 | Guarantee | 40 | |
Section 11.2 | Guarantee of Payment | 40 | |
Section 11.3 | No Limitations, Etc. | 40 | |
ii
Section 11.4 | Reinstatement | 40 | |
Section 11.5 | Agreement To Pay; Subrogation | 40 | |
Section 11.6 | Information | 40 | |
Section 11.7 | Subordination. | 41 | |
Section 11.8 | Termination | 41 | |
Index of Exhibits
Exhibit A:Description of Etripamil
Exhibit B:Bill of Sale
Exhibit C:Form of Royalty Report
Exhibit D:Assignment Agreement
Index of Schedules
Schedule 4.1(k)(i)Patents
iii
PURCHASE AND SALE AGREEMENT
This PURCHASE AND SALE AGREEMENT, dated as of March 27, 2023 (as amended, supplemented or modified from time to time, this “Agreement”), is made and entered into by and between RTW Royalty I DAC, a designated activity company incorporated under the laws of Ireland with registered number 734869 and registered address at 2nd Floor, Palmerston House, Denzille Lane, Dublin 2, D02 WD37, Ireland (the “Buyer”), and Milestone Pharmaceuticals USA, Inc., a Delaware corporation (the “Seller”), as assignee to Milestone Pharmaceuticals Inc., a Quebec corporation (“Holdings”) and Holdings.
RECITALS:
WHEREAS, the Seller is in the business of, among other things, developing and commercializing the Product; and WHEREAS, the Buyer desires to purchase the Revenue Participation Right from the Seller in exchange for payment of the Purchase Price, and the Seller desires to sell the Revenue Participation Right to the Buyer in exchange for the Buyer’s payment of the Purchase Price, in each case on the terms and conditions set forth in this Agreement.
NOW THEREFORE, in consideration of the representations, warranties, covenants and agreements set forth herein and for good and valuable consideration, the receipt and adequacy of which are hereby acknowledged, the Seller and the Buyer hereby agree as follows:
Article 1.
DEFINITIONS
Section 1.1Definitions. The following terms, as used herein, shall have the following meanings:
“Acceptable Intercreditor Agreement” means an intercreditor agreement on customary terms among the Buyer, the Seller, Holdings and the Senior Debt Provider, to be mutually agreed by the parties thereto.
“Affiliate” means, (a) with respect to any particular Person, any other Person directly or indirectly controlling, controlled by or under common control with such particular Person or (b) with respect to the Buyer, any Person now or hereafter existing that is managed or otherwise controlled by RTW Investments, LP. For purposes of the foregoing sentence, the term “control” means direct or indirect ownership of (i) fifty percent (50%) or more, including ownership by trusts with substantially the same beneficial interests, of the voting and equity rights of such Person, firm, trust, corporation, partnership or other entity or combination thereof, or (ii) the power to direct the management of such person, firm, trust, corporation, partnership or other entity or combination thereof, by contract or otherwise.
“Agreement” is defined in the preamble.
“Amendment Date” means December 12, 2025.
“Approved Indications” means the treatment of paroxysmal supraventricular tachycardia (“PSVT”) in adults with a confirmed diagnosis of PSVT, which may include expected contraindications including but not limited to: hypersensitivity to Etripamil; heart failure - NYHA Class II to IV; known Wolff-Parkinson-White (WPW), Lown-Ganong-Levine (LGL) syndromes, sick sinus syndrome (except in patients with a functioning artificial ventricular pacemaker); second degree atrioventricular (AV) Mobitz 2 block and higher; or first degree AV block plus bundle branch block (BBB)/intraventricular conduction delay (IVCD), but shall exclude any and all Unexpected Contraindications.
“Back-Up Security Interest” is defined in Section 2.1(b).
“Bankruptcy Event” means the occurrence of any of the following in respect of a Person: (a) such Person shall generally not, shall be unable to, or an admission in writing by such Person of its inability to, pay its debts as
1
they come due or a general assignment by such Person for the benefit of creditors; (b) the filing of any petition or answer by such Person seeking to adjudicate itself as bankrupt or insolvent, or seeking for itself any liquidation, winding-up, reorganization, arrangement, adjustment, protection, relief or composition of such Person or its debts under any applicable law relating to bankruptcy, insolvency, examinership, receivership, winding-up, liquidation, reorganization, examination, relief of debtors or other similar applicable law now or hereafter in effect, or seeking, consenting to or acquiescing in the entry of an order for relief in any case under any such applicable law, or the appointment of or taking possession by a receiver, trustee, custodian, liquidator, examiner, assignee, sequestrator or other similar official for such Person or for any substantial part of its property; (c) corporate or other entity action taken by such Person to authorize any of the actions set forth in clause (a) or clause (b) above; or (d) without the consent or acquiescence of such Person, the commencement of an action seeking entry of an order for relief or approval of a petition for relief or reorganization or any other petition seeking any reorganization, arrangement, composition, readjustment, liquidation, dissolution or other similar relief under any present or future bankruptcy, insolvency or similar applicable law, or the filing of any such petition against such Person, or, without the consent or acquiescence of such Person, the commencement of an action seeking entry of an order appointing a trustee, custodian, receiver or liquidator of such Person or of all or any substantial part of the property of such Person, in each case where such petition or order shall remain unstayed or shall not have been stayed or dismissed within ninety (90) calendar days from entry thereof.
“Bankruptcy Laws” means, collectively, bankruptcy, insolvency, reorganization, moratorium, fraudulent conveyance, fraudulent transfer or other similar laws affecting the enforcement of creditors’ rights generally.
“Business Day” means any day other than (a) a Saturday or Sunday or (b) a day on which banking institutions located in New York or Ireland are permitted or required by applicable law or regulation to remain closed.
“Buy-Back Option” is defined in Section 6.2(d).
“Buy-Back Requirement” is defined in Section 6.2(d).
“Buyer” is defined in the preamble.
“Buyer Indemnified Parties” is defined in Section 7.1(a).
“Buyer Representative” means (a) if there is more than one Buyer, one Buyer acting on behalf of all Buyers, as designated in writing from time to time by the Buyers to the Seller, and (b) if there is only one Buyer, the Buyer.
“Calendar Quarter” means, for the first calendar quarter, the period beginning on the first day of the calendar quarter in which the Closing Date falls and ending on the last day of such calendar quarter, and thereafter, each successive period of three (3) consecutive calendar months ending on March 31, June 30, September 30 or December 31.
“Cash Management Obligations” means obligations owed by the Seller to any provider of Treasury Management Arrangement pursuant to such arrangement.
“Change of Control” means the occurrence of any one or more of the following: (a) the acquisition, whether directly, indirectly, beneficially or of record, whether by merger, consolidation, sale or other transfer of securities in a single transaction or series of related transactions, by any Person of any voting securities of Holdings, or if the percentage ownership of any Person in the voting securities of Holdings is increased through stock redemption, cancellation, or other recapitalization, and immediately after such acquisition or increase such Person is, directly or indirectly, the beneficial owner of voting securities representing fifty percent (50%) or more of the total voting power of all of the then outstanding voting securities of Holdings; (b) a merger, consolidation, recapitalization, or reorganization of Holdings is consummated that would result in shareholders or equity holders of Holdings immediately prior to such transaction that did not own more than fifty percent (50%) of the outstanding voting securities of Holdings immediately prior to such transaction, owning more than fifty percent (50%) of the outstanding voting securities of the surviving entity (or its parent entity) immediately following such transaction; (c) the sale, lease, transfer, license or other disposition, in a single transaction or series of related transactions, by the Seller or any
2
Subsidiary of the Seller of all or substantially all the assets of the Seller and its Subsidiaries taken as a whole, or the sale or disposition (whether by merger, consolidation or otherwise) of one or more Subsidiaries of the Seller if substantially all of the assets of the Seller and its Subsidiaries taken as a whole are held by such Subsidiary or Subsidiaries, except where such sale, lease, transfer, license or other disposition is to a wholly owned Subsidiary of Holdings; and (d) Holdings ceases to own directly or indirectly 100% of outstanding voting securities of the Seller or any successor or assignee of the Seller.
“Clinical Trial” means a clinical trial or study intended to support or maintain Marketing Approval or Commercialization of a Product, including post-approval clinical trials or studies.
“Clinical Updates” means, [***].
“Closing” means the closing of the sale, transfer, assignment and conveyance of the Revenue Participation Right hereunder.
“Closing Date” is defined in Section 3.1.
“CMC” means chemistry, manufacturing and controls with respect to a Product.
“CoC Agreement” is defined in Section 6.2(d).
“CoC Date” is defined in Section 6.2(d).
“CoC Payment” is defined in Section 6.2(d).
“Combination Product” means:
(a)a single pharmaceutical formulation (whether co-formulated or administered together via the same administration route) containing as its active ingredients both a Product and one or more other therapeutically or prophylactically active pharmaceutical or biologic ingredients (each an “Other Component”), or
(b)a combination therapy comprised of a Product and one or more Other Component(s), whether priced and sold in a single package containing such multiple products, packaged separately but sold together for a single price, or sold under separate price points but labeled for use together, in each case, including all dosage forms, formulations, presentations, and package configurations. Drug delivery vehicles, adjuvants and excipients will not be deemed to be “active ingredients,” except in the case where such delivery vehicle, adjuvant or excipient is recognized by the FDA as an active ingredient in accordance with 21 C.F.R. 210.3(b)(7). All references to Products in this Agreement shall be deemed to include Combination Products.
“Commercial Updates” means, [***].
“Commercialization” means any and all activities directed to the distribution, marketing, detailing, promotion, selling and securing of reimbursement of a Product (including the using, importing, selling and offering for sale of such Product), and shall include post-Marketing Approval studies to the extent required by a Regulatory Authority, post-launch marketing, promoting, detailing, distributing, selling such Product, importing, exporting or transporting such Product for sale, and regulatory compliance with respect to the foregoing. When used as a verb, “Commercialize” shall mean to engage in Commercialization. Except with respect to post-Marketing Approval studies required by a Regulatory Authority, “Commercialization” shall not include any activities directed to the research or development (including pre-clinical and clinical development) or manufacture of a Product.
“Commercially Reasonable Efforts” means the level of efforts and resources (measured as of the time that such efforts and resources are required to be used under this Agreement) that are commonly used by a biotechnology company of similar size and resources to Holdings (provided that, for purposes of this definition, such size and resources shall not be less than the size and resources of Holdings as of the Closing Date), to develop, manufacture or
3
commercialize, as the case may be, a comparable product for a comparable clinical indication (with respect to market size and commercial opportunity) at a similar stage in its development or product life and of a similar market and potential to a Product, but without regard to the Seller’s financial obligations under this Agreement.
“Confidential Information” is defined in Section 8.1.
“Deposit Account” means a “deposit account” (as defined in Article 9 of the UCC), investment account or other account in which funds are held or invested to or for the credit or account of the Seller.
“Disclosing Party” is defined in Section 8.1.
“Disclosure Schedule” means the Disclosure Schedule, dated as of the date hereof, delivered to the Buyer by the Seller concurrently with the execution of this Agreement.
“Distributor” means a Third Party that (a) purchases or has the option to purchase any Product in finished form from or at the direction of the Seller or any of its Affiliates or Licensees, and (b) has the right, option or obligation to distribute, market and sell such Product (with or without packaging rights) in one or more regions, but does not otherwise obtain a license or other rights to Intellectual Property Rights with respect to such Product. The term “packaging rights” in this definition will mean the right for the Distributor to package or have packaged Product supplied in unpackaged bulk form into individual ready-for-sale packs.
“EMA” means the European Medicines Agency, or any successor agency thereto. “ERISA” means the Employee Retirement Income Security Act of 1974.
“Etripamil” means the calcium channel blocker referred to by Seller as of the date hereof and as of the Amendment Date as etripamil and described on Exhibit A hereto.
“Excluded Collateral” means, each of (a) any “intent to use” trademark applications for which a statement of use has not been filed (but only until such statement is filed), (b) equipment, (c) investment property and securities and bank accounts, and (d) any permit, license or agreement entered into by the Seller or Holdings (except the Intercompany License) (i) to the extent that any such permit, license or agreement or any law applicable thereto prohibits the creation of a Lien thereon, but only to the extent, and for as long as, such prohibition is not terminated or rendered unenforceable or otherwise deemed ineffective by the UCC or any other applicable law, or (ii) to the extent that the creation of a Lien in favor of the Buyer would result in a breach or termination pursuant to the terms of or a default under any such permit, license, or agreement (other than to the extent that any such term would be rendered ineffective pursuant to the Sections 9-406, 9-407, 9-408, or 9-409 of the UCC or any other applicable law (including the Bankruptcy Laws) or principles of equity).
“Existing Patent Rights” is defined in Section 4.1(k)(i).
“FD&C Act” means the United States Federal Food, Drug, and Cosmetic Act.
“FDA” means the U.S. Food and Drug Administration, or any successor agency thereto. “First Commercial Sale” means, with respect to a Product, the first sale for use or consumption by an end user of such Product in the Territory after Marketing Approval of such Product has been granted in the Territory.
“GAAP” means generally accepted accounting principles in the United States in effect from time to time.
“Governmental Entity” means any: (a) nation, principality, republic, state, commonwealth, province, territory, county, municipality, district or other jurisdiction of any nature; (b) federal, state, local, municipal, foreign or other government; (c) governmental or quasi-governmental authority of any nature (including any governmental division, subdivision, department, agency, bureau, branch, office, commission, council, board, instrumentality, officer, official, representative, organization, unit, body or other entity and any court, arbitrator or other tribunal); (d) multi-national organization or body; or (e) individual, body or other entity exercising, or entitled to exercise, any executive, legislative, judicial, administrative, regulatory, police, military or taxing authority or power of any nature.
4
“Gross Sales” is defined in the definition of “Net Sales”. “Holdings” is defined in the preamble.
“Improvements” means any improvement, invention or discovery necessary for the development, manufacture, use, or Commercialization of a Product or primarily directed to a Product (other than with respect to a new composition of matter), including the formulation, or the method of manufacture of a Product.
“In-License” means (i) any license, settlement agreement or other agreement or arrangement between the Seller or any of its Affiliates and any Third Party pursuant to which the Seller or any of its Affiliates obtains a license or a covenant not to sue or similar grant of rights to any Patents or other intellectual property rights of such Third Party that is necessary for the research, development, manufacture, use or Commercialization of a Product in the Territory, other than non-exclusive license or other rights ancillary to the provision of services by any Third Party, and (ii) the Intercompany License.
“Indebtedness” of any Person means any indebtedness for borrowed money, any obligation evidenced by a note, bond, debenture, or similar instrument, or any guarantee of any of the foregoing.
“Indemnified Party” is defined in Section 7.2.
“Indemnified Taxes” means all Canadian and/or U.S. federal withholding Taxes imposed on or with respect to any payment made by (or on account of any obligation of) the Seller and/or Holdings under this Agreement.
“Indemnifying Party” is defined in Section 7.2.
“Initial Tier Royalty” is defined in the definition of “Royalty Rate.”
“Intellectual Property Product Rights” means any and all of the following as they exist in the Territory at any time: (a) the Intellectual Property Rights; (b) rights in registered and unregistered trademarks, service marks, trade names, trade dress, logos, packaging design, slogans and Internet domain names, and registrations and applications for registration of any of the foregoing, in each case, that are (i) owned or in-licensed by the Seller or any of its Affiliates and (ii) used by Seller or any of its Affiliates in connection with the development, manufacture, use, or Commercialization of any Product; and (c) any and all other intellectual property rights and/or proprietary rights, whether or not patentable, relating to any of the foregoing, in each case, (i) owned or in-licensed by the Seller or any of its Affiliates (other than software licenses or non-exclusive licenses granted by service providers in connection with the development or manufacture of the Product), and (ii) that are necessary for the development, manufacture, use, or Commercialization of a Product or primarily directed to a Product.
“Intellectual Property Rights” means any and all of the following as they exist in the Territory at any time: (a) the Patent Rights and (b) the Know-How Rights.
“Intellectual Property Updates” means an [***].
“Intercompany License” means the [***].
“Judgment” means any judgment, order, writ, injunction, citation, award or decree of any nature.
“Know-How” means any and all proprietary or confidential information, know-how and trade secrets, including processes, formulae, models and techniques (but excluding rights in research in progress, algorithms, data, databases, data collections, chemical and biological materials and the results of experimentation and testing).
“Know-How Rights” means any and all Know-How owned or in-licensed by the Seller or any of its Affiliates under which the Seller or any of its Affiliates is or may become empowered to grant licenses and are necessary for the development, manufacture, use, or Commercialization of a Product or primarily directed to a Product (other than software licenses or non-exclusive licenses granted by service providers in connection with the development or manufacture of the Product).
5
“Knowledge of the Seller and Holdings” means the actual knowledge of the individuals listed on Schedule 1.1 of the Disclosure Schedule, after reasonable due inquiry.
“Licensee” means, with respect to any Product, a Third Party to whom the Seller or any Affiliate of the Seller has granted a license or sublicense to Commercialize such Product. For clarity, a Distributor shall not be deemed to be a “Licensee.”
“Licensee Reports” is defined in Section 6.7(c).
“Lien” means any mortgage, lien, pledge, participation interest, charge, adverse claim, security interest, encumbrance or restriction of any kind, including any restriction on use, transfer or exercise of any other attribute of ownership of any kind.
“Loss” means any and all Judgments, damages, losses, claims, costs, liabilities and expenses, including reasonable and documented fees and out-of-pocket expenses of counsel.
“Loss of Market Exclusivity” shall mean the later to occur of: (a) the expiration of the last-to-expire Valid Claim of a Patent Right covering such Product in the Territory; and (b) the expiry of all Regulatory Exclusivity Periods for such Product in the Territory.
“Major Stock Exchange” means the NYSE, NASDAQ, Tokyo Stock Exchange, Euronext, or the stock exchanges of Toronto, Frankfurt, or London.
“Marketing Approval” means an NDA approved by the FDA (including accelerated approval). For clarity, “Marketing Approval” shall not include pricing and reimbursement approvals.
“Marketing Approval Deadline” means 11:59 pm P.T. December 31, 2025.
“Material Adverse Effect” means (a) a material adverse effect on (i) a Product, (ii) any of the Intellectual Property Rights, including Holdings’ or the Seller’s rights in or to any Intellectual Property Rights, (iii) any Marketing Approval of a Product or the timing thereof, (iv) the legality, validity or enforceability of any provision of this Agreement, (v) the ability of the Seller or Holdings to perform any of its obligations under this Agreement, (vi) the rights or remedies of the Buyer under this Agreement, or (vii) the business of Holdings or its Affiliates; or (b) an adverse effect in any material respect on (i) the timing, duration or amount of the Royalty Payments, or (ii) the Revenue Participation Right, the Product Collateral, or the Back-Up Security Interest.
“NDA” means a New Drug Application submitted to the FDA in the United States in accordance with the FD&C Act with respect to a pharmaceutical product.
“Net Sales” means, with respect to each Product, the gross amount invoiced, billed or otherwise recorded for sales of such Product in the Territory by or on behalf of the Seller, its Affiliates, or any Licensee of the Seller or any of the Seller’s Affiliates (each of the foregoing Persons, for purposes of this definition, shall be considered a “Related Party”) to a Third Party (including Distributors) in an arms-length transaction (“Gross Sales”), less the following amounts incurred in connection with such Product sales in the Territory, to the extent actually incurred or accrued in accordance with GAAP consistently applied, and not reimbursed by such Third Party, provided that any given amount may be taken as a permitted deduction only once:
(a)reasonable and customary rebates, chargebacks, quantity, trade and similar discounts, credits and allowances and other price reductions reasonably granted, allowed, incurred or paid in so far as they are applied to sales of a Product;
(b)discounts (including cash, quantity, trade, governmental, and similar discounts), coupons, retroactive price reductions, charge back payments and rebates granted to wholesalers, pharmacies and other retailers, buying groups (including group purchasing organizations), and other purchasers, including trade customers, Third Party payers, pharmacy benefit management companies (or equivalents thereof), health care
6
insurance carriers, health maintenance organizations, managed care organizations or to federal, state and local governments, or to their respective agencies, purchasers or reimbursers (including payments made under the new “Medicare Part D Coverage Gap Discount Program” and the “Annual Fee for Branded Pharmaceutical Manufacturers” specific to a Product), in each case, as applied to sales of a Product and actually given to customers;
(c)reasonable and customary credits, adjustments, and allowances, including those granted on account of price adjustments, billing errors, and damage, Product otherwise not in saleable condition, and rejection, return or recall of a Product;
(d)reasonable and customary freight, postage, shipping, and insurance costs incurred with respect to the shipment of a Product to customers, in each case if charged and invoiced to the customer;
(e)customs duties, surcharges and other similar governmental charges incurred in connection with the exportation or importation of a Product to the extent included in the gross amount invoiced;
(f)sales, use, value-added, excise, turnover, inventory and other similar Taxes (excluding income or franchise Taxes of any kind), and that portion of annual fees due under Section 9008 of the United States Patient Protection and Affordable Care Act of 2010 (Pub. L. No. 111-48) and any other fee imposed by any equivalent applicable law, in each of the foregoing cases, that Seller allocates to sales of a Product in accordance with Seller’s standard policies and procedures consistently applied across its products, as adjusted for offsets, rebates and refunds, imposed in connection with the sales of a Product to any Third Party, to the extent such Taxes are not paid by the Third Party;
(g)actual copayment waiver amounts and uncollected or uncollectible debt amounts with respect to sales of a Product, provided that, if the debt is thereafter paid, the corresponding amount shall be added to the Net Sales of the period during which it is paid;
(h)reasonable, customary and documented out-of-pocket amounts directly relating to co-pay programs, bridging programs or other similar patient assistance programs which may be implemented from time to time by the Seller; and
(i)other similar or customary deductions taken in the ordinary course of business as permitted in calculating net sales or net revenue (as applicable) under GAAP consistently applied. For the avoidance of doubt, such deductions shall not include any amount that is treated as an operating expense for the U.S. federal income tax purposes.
“Net Sales” will include monetary damages, but not punitive damages, recovered by Seller, its Affiliates or Holdings from a Third Party to the extent permitted, and treated as “Net Sales,” under Section 6.5(e)(iii). “Net Sales” shall be consistent with the meaning ascribed to “net sales” for GAAP, provided that, notwithstanding anything to the contrary in this definition, in the case of any sales by any Licensee (or any of its Affiliates or sublicensees), “Net Sales” (including Combination Product allocations in connection with such Net Sales) will be defined in the same manner as such term or comparable term in the applicable license agreement with such Licensee, provided that any such change to the definition of “Net Sales” does not cause a Material Adverse Effect. For clarity, “Net Sales” will not include (i) sales or dispositions for charitable, promotional, pre-clinical, clinical, regulatory, compassionate use, named patient use or indigent or other similar programs, reasonable quantities of Products used as samples, and Products used in the development of Products, (ii) sales or dispositions between any of the Related Parties (unless a Related Party is the final end-user of such Product), but will include subsequent sales or dispositions of Products to a non-Related Party, or (iii) any amounts or other consideration received by a Related Party from a Licensee, Distributor, or a non-Related Party in consideration of the grant of a (sub)license or co-promotion or distribution right to such non-Related Party, including any upfront or milestone payments (whether or not such milestones are based on net sales of a Product).
In calculating Net Sales, no deduction from Gross Sales may be made with respect to any monies paid or owed by Seller to Holdings pursuant to the Intercompany License, nor any fees or charges paid to Seller or Holdings
7
in connection therewith, including any Taxes or tariffs, except for Taxes and tariffs otherwise deductible under the definition of Net Sales. For clarity, no deductions from Gross Sales are permitted with respect to any charges, costs, fees, payments, or other adverse impacts in any way associated with the Intercompany License.
With respect to sales of a Product invoiced in U.S. dollars, Net Sales shall be determined in U.S. dollars. If applicable, with respect to sales of a Product invoiced in a currency other than U.S. dollars, Net Sales shall be determined by converting the currencies at which the sales are made into U.S. dollars, at rates of exchange determined in a manner consistent with Holdings’, the Seller’s or a Licensee’s, as applicable, method for calculating rates of exchange in the preparation of the Seller’s or such Licensee’s annual financial statements in accordance with GAAP consistently applied.
Net Sales for any Combination Product in the Territory shall be calculated by multiplying actual Net Sales of such Combination Product by the fraction A/(A+B) where “A” is the weighted average invoice price of a Product contained in such Combination Product when sold separately in the Territory during the applicable accounting period in which the sales of the Combination Product were made, and “B” is the combined weighted average invoice prices of all of the Other Components contained in such Combination Product sold separately in the Territory during such same accounting period. If the weighted average invoice price of a Product in the Territory can be determined but combined weighted average invoice prices of all of the Other Components contained in such Combination Product cannot be determined, Net Sales for such Combination Product in the Territory shall be calculated by multiplying actual Net Sales of such Combination Product by the fraction A/C, where “A” is the weighted average invoice price of a Product contained in such Combination Product when sold separately in the Territory during the applicable accounting period in which the sales of the Combination Product were made, and “C” is the weighted average invoice price of such Combination Product sold in the Territory during the applicable accounting period. If a Product contained in such Combination Product is not sold separately in finished form in the Territory but the Other Components included in such Combination Product are sold separately in finished form in the Territory, Net Sales for such Combination Product in the Territory shall be calculated by multiplying actual Net Sales of such Combination Product by the fraction determined by the following formula: one (1) minus (B/C), where “B” is the combined weighted average invoice prices of all of the Other Components contained in such Combination Product when sold separately in the Territory during the applicable accounting period in which the sales of the Combination Product were made and “C” is the weighted average invoice price of such Combination Product sold in the Territory during the applicable accounting period. If neither the Product nor the Other Components contained in such Combination Product is sold separately in finished form in the Territory, the Seller shall determine Net Sales for such Product in good faith based on the relative contribution of such Product and each such Other Component in such Combination Product, and shall take into account in good faith any applicable allocations and calculations that may have been made for the same period in the Territory.
“Note Closing” shall have the meaning ascribed to the term “Closing” set forth in the Note Purchase Agreement.
“Note Closing Date” means the date on which the Note Closing occurs.
“Note Purchase Agreement” means that certain Note Purchase Agreement dated as of the date hereof among Holdings and the Buyer.
“Orange Book” means the FDA publication “Approved Drug Products with Therapeutic Equivalence Evaluations,” as may be amended from time to time.
“Orange Book Patents” means the Patents listed in the Orange Book by Seller, its Affiliates or Licensees in connection with Etripamil.
“Other Component” is defined in the definition of “Combination Products”. “Other License” is defined in Section 6.7(a).
8
“Out-License” means each license or other agreement between the Seller or any of its Affiliates and any Third Party (other than Distributors) pursuant to which the Seller or any of its Affiliates grants a license, sublicense or other rights to practice any Intellectual Property Right to Commercialize a Product in the Territory.
“Patents” means any and all patents and patent applications existing as of the date of this Agreement and all patent applications filed hereafter, including any continuation, continuation-in-part, division, provisional or any substitute applications, any patent issued with respect to any of the foregoing patent applications, any certificate, reissue, reexamination, renewal or patent term extension or adjustment (including any supplementary protection certificate) of any such patent or other governmental actions which extend any of the subject matter of a patent, and any substitution patent, confirmation patent or registration patent or patent of addition based on any such patent, and all foreign counterparts of any of the foregoing.
“Patent Rights” means any and all Patents owned or in-licensed by the Seller or any of its Affiliates under which the Seller or any of its Affiliates is or may become empowered to grant licenses that covers the composition of matter, method of using or making, delivery systems for, or formulation of the Product, in each case that is necessary or used in the development, manufacture, use, or Commercialization of a Product in the Territory, as well as existing or future Patents covering any Improvements of the foregoing in the Territory.
“Permitted Indebtedness” is defined in the definition of “Restricted Indebtedness”.
“Permitted License” is defined in Section 6.7(a).
“Permitted Liens” means the following:
(a)Liens for Taxes, assessments or governmental charges or levies not yet due or which are being contested in good faith and by appropriate proceedings diligently conducted, if adequate reserves with respect thereto are maintained on the books of the applicable Person in accordance with GAAP;
(b)statutory Liens of landlords and Liens of carriers, warehousemen, mechanics, materialmen and suppliers and other Liens imposed by law or pursuant to customary reservations or retentions of title arising in the ordinary course of business, provided that such Liens secure only amounts not yet due and payable or, if due and payable, are unfiled and no other action has been taken to enforce the same or are being contested in good faith by appropriate proceedings for which adequate reserves determined in accordance with GAAP have been established;
(c)Liens of any judgment rendered or claim filed against the Seller or Holdings, which the Seller, Holdings or others on its behalf shall be contesting diligently and in good faith by proper legal proceedings, provided the action to enforce the same has not proceeded to final non-appealable judgment and adequate provision has been made for the payment thereof in accordance with GAAP and in a manner acceptable to the Buyer;
(d)Liens on property existing at the time of acquisition of such property, provided that such liens were in existence prior to such acquisition and not incurred in contemplation thereof;
(e)Permitted Licenses, including any interest or title of a licensee under a Permitted License;
(f)Liens granted pursuant to the Collateral Documents (as defined under the Note Purchase Agreement) or otherwise securing the obligations of Holdings and its Subsidiaries under the Note Purchase Agreement;
(g)pledges or deposits in the ordinary course of business in connection with workers’ compensation, unemployment insurance and other social security legislation, other than any Lien imposed by ERISA;
9
(h)undetermined or inchoate Liens arising or potentially arising under statutory provisions which have not at the time been filed or registered in accordance with applicable law or of which written notice has not been duly given in accordance with applicable law or which, although filed or registered, relate to obligations not due or delinquent;
(i)deposits to secure the performance of bids, trade contracts and leases (other than Indebtedness), statutory obligations, surety and appeal bonds, indemnity and performance bonds and other obligations of a like nature incurred in the ordinary course of business;
(j)easements, rights-of-way, restrictions and other similar encumbrances affecting real property which, in the aggregate, are not substantial in amount, and which do not materially interfere with the ordinary conduct of the business of the applicable Person;
(k)licenses, sublicenses, leases or subleases granted to others in the ordinary course of business and not interfering in any material respect with the Revenue Participation Right, the Product Rights, the Product Collateral, or the Back-Up Security Interest;
(l)any interest of title of a lessor under, and Liens arising from any protective UCC financing statements (or equivalent filings, registrations or agreements in foreign jurisdictions) relating to, leases permitted by this Agreement;
(m)normal and customary banker’s Liens and rights of setoff upon deposits of cash and securities in favor of banks or other depository institutions;
(n)Liens of a collection bank arising under Section 4-210 of the UCC on items in the course of collection;
(o)Liens on specific items of inventory or other goods (and the proceeds thereof) securing obligations in respect of bankers’ acceptances issued or created to facilitate the purchase, shipment or storage of such inventory or other goods;
(p)Liens arising out of conditional sale, title retention, consignment or similar arrangements for the sale of goods entered into in the ordinary course of business;
(q)Liens in favor of customs and revenue authorities arising as a matter of law to secure payment of custom duties in connection with the importation of goods in the ordinary course;
(r)other Liens of sellers of goods to Holdings and any of its Subsidiaries arising under Article 2 of the UCC or similar provisions of applicable law in the ordinary course of business, covering only the goods sold and securing only the unpaid purchase price for such goods and related expenses;
(s)Liens in the nature of rights of setoff in favor of counterparties to contractual agreements; and
(t)Liens securing Permitted Indebtedness (to the extent permitted to be secured pursuant to the definition thereof); provided that the Permitted Indebtedness in clauses (a) and (d) shall be secured solely by cash and cash equivalents, clause (b) shall only be secured by the assets financed by such Indebtedness, and, with respect to any such Permitted Indebtedness provided by any Senior Debt Providers, the requirements of Section 6.8 are satisfied.
“Permitted Out-License” is defined in Section 6.7(a).
“Person” means any individual, firm, corporation, company, partnership, limited liability company, trust, joint venture, association, estate, trust, Governmental Entity or other entity, enterprise, association or organization.
10
“Prime Rate” means the prime rate published by The Wall Street Journal, from time to time, as the prime rate.
“Product” means, individually and collectively, (a) Etripamil, (b) any radioisomer, stereoisomer, racemates, solvates, salt forms, bases, anhydrides, hydrates, polymorphs, metabolites, ester forms, deuterated forms or pro-drugs of such compound, and (c) any pharmaceutical product that contains any of the foregoing, in each case in any form, formulation, dosing regimen, strength, or route of administration.
“Product Collateral” means the Seller’s or any of its Affiliates’ right, title and interest in (a) all Products (including all inventory of all Products), (b) the Product Rights owned, in-licensed or otherwise held by the Seller or its Affiliates, and (c) any proceeds from either (a) or (b), including all accounts receivable and general intangibles resulting from the sale, license or other disposition of any Products by a Related Party in the Territory. Notwithstanding the foregoing, “Product Collateral” shall not include any Excluded Collateral.
“Product Rights” means any and all of the following, as they exist in the Territory (a) Intellectual Property Product Rights, (b) regulatory filings, submissions and approvals, including Marketing Approvals, with or from any Regulatory Authorities with respect to the Products, (c) In-Licenses and (d) Out-Licenses.
“PSVT” is defined in the definition of “Approved Indication.”
“Purchase Price” is defined in Section 2.2.
“Qualified Buyer” means any global pharmaceutical company with (a) a market capitalization greater than [***] for [***] trading days on a Major Stock Exchange, or (b) annual net revenue greater than [***], in each case ((a) and (b)), measured as of the date the definitive agreement for such Change of Control is executed.
“Receiving Party” is defined in Section 8.1.
“Regulatory Authority” means any national or supranational Governmental Entity, including the FDA, or such equivalent regulatory authority, or any successor agency thereto, that has responsibility in granting a Marketing Approval.
“Regulatory Exclusivity Period” shall mean, with respect to a Product in the Territory, any period of data, market or other regulatory exclusivity (other than Patent exclusivity) granted or afforded by law or by a Regulatory Authority in the Territory that confers exclusive marketing rights with respect to such Product in the Territory or prevents another party from using or otherwise relying on any data supporting the Marketing Approval for such Product.
“Regulatory Updates” means [***].
“Related Party” is defined in the definition of “Net Sales.”
“Release Event” means the earliest of the following dates: (a) the date on which the aggregate amount of all Royalty Payments received by the Buyer from the Seller equals at least [***], (b) the date of consummation of a Change of Control with a Qualified Buyer and (c) the date on which Holdings’ market capitalization is at least [***] for [***] trading days on a Major Stock Exchange. “Report” is defined in Section 6.1.
“Reporting Date” means, for each Calendar Quarter, the earlier of (a) forty-five (45) calendar days after the end of such Calendar Quarter (in the case of the first three Calendar Quarters of any calendar year) or ninety (90) calendar days after the end of such Calendar Quarter (in the case of the fourth Calendar Quarter of any calendar year), and (b) five (5) Business Days after the earlier of (X) the date that Holdings has filed with the SEC its Form 10-Q or Form 10-K, as applicable, for the immediately preceding Calendar Quarter, and (Y) the filing due date for such Form 10-Q or Form 10-K.
11
“Representative” means, with respect to any Person, (a) any direct or indirect member or partner of such Person and (b) any manager, director, trustee, officer, employee, agent, advisor or other representative (including attorneys, accountants, consultants, contractors, actual and potential lenders, investors, co-investors and assignees, bankers and financial advisers) of such Person.
“Restricted Indebtedness” mean any financing or factoring of royalties on the Products, or any Indebtedness, in each case other than the following (collectively “Permitted Indebtedness”):
(a)Indebtedness under the Note Purchase Agreement;
(b)Indebtedness existing on the date hereof and disclosed to the Buyer on Schedule A of the Disclosure Schedule;
(c)unsecured Indebtedness to trade creditors incurred in the ordinary course of business;
(d)Indebtedness incurred in the ordinary course of business consisting of capitalized lease obligations and purchase money Indebtedness, in each case incurred to finance the acquisition, repair, improvement or construction of fixed or capital assets of Holdings or any of its Subsidiaries; provided, however, that, (i) the total of all such Indebtedness for Holdings and its Subsidiaries taken together shall not exceed an aggregate principal amount of [***] at any one time outstanding. For the avoidance of doubt, leases that would have been treated as operating leases prior to the adoption of Accounting Standards Codification 842 will not be deemed to constitute as “Restricted Indebtedness”;
(e)Indebtedness consisting of a revolving line of credit (which may be first lien senior secured) in an aggregate principal amount not to exceed [***] at any time;
(f)letters of credit, bankers’ acceptances, guarantees or other similar instruments or obligations issued or relating to liabilities or obligations incurred in the ordinary course of business; provided that the aggregate outstanding amount of such letters of credit, bankers’ acceptances, guarantees or other similar instruments or other obligations issued thereunder shall not exceed [***] at any time outstanding;
(g)Indebtedness between or among Holdings and any of its Subsidiaries, including, without limitation, any intercompany obligations incurred pursuant to this Agreement;
(h)Indebtedness incurred in the ordinary course of business owed to any Person providing property, casualty, liability, or other insurance to Holdings or any of its Subsidiaries, including to finance insurance premiums, so long as the amount of such Indebtedness is not in excess of the amount of the unpaid cost of, and shall be incurred only to defer the cost of, such insurance for the policy year in which such Indebtedness is incurred and such Indebtedness is outstanding only during such policy year;
(i)to the extent constituting Indebtedness, obligations under this Agreement;
(j)other unsecured Indebtedness;
(k)Cash Management Obligations and other Indebtedness in respect of netting services, automatic clearinghouse arrangements, overdraft protections, employee credit card programs and other cash management and similar arrangements in the ordinary course of business or the honoring by a bank or other financial institution of a check, draft, or similar instrument drawn against insufficient funds in the ordinary course of business; and
(l)extensions, refinancings, modifications, amendments and restatements of Indebtedness set forth in clauses (i) through (k) above, provided that the principal amount thereof is not increased and the terms thereof are not modified to impose materially more burdensome terms upon Holdings or such Subsidiary.
12
“Revenue Participation Right” means the right to receive the Royalty Payments. “Royalty Payments” means, for each Calendar Quarter, an amount equal to the aggregate Net Sales of Products in the Territory during such Calendar Quarter multiplied by the applicable Royalty Rate.
“Royalty Rate” means the percentage based on the applicable level of aggregate Net Sales of Products in the Territory in a calendar year as set forth in the chart below:
Payment Tiers based on Annual Net Sales | Royalty Rate |
A. Annual Net Sales less than or equal to $500,000,000 | 7.0% (the “Initial Tier Royalty”) |
B. Annual Net Sales greater than $500,000,000 and less than or equal to $800,000,000 | 4.0% |
C. Annual Net Sales greater than $800,000,000 | 1.0% |
provided that, if annual Net Sales of Products do not equal at least Fifty Million Dollars ($50,000,000) for calendar year 2025, One Hundred Million Dollars ($100,000,000) for calendar year 2026, or One Hundred Fifty Million Dollars ($150,000,000) for calendar year 2027 (each such threshold, a “Sales Threshold”), the Initial Tier Royalty will increase to 9.5% beginning on January 1 of the following calendar year until a subsequent Sales Threshold is attained, at which time the Initial Tier Royalty would revert to seven percent (7.0%) beginning January 1 of the following calendar year; provided that, for any calendar year after calendar year 2027, if annual Net Sales equals or exceeds Five Hundred Million Dollars ($500,000,000), then the Initial Tier Royalty will revert to seven percent (7.0%) beginning January 1 of the following calendar year. For clarity, (a) if Net Sales of Products for calendar year 2025 are less than Fifty Million Dollars ($50,000,000), then the Initial Tier Royalty would increase to 9.5% for all Net Sales of Products on or after January 1, 2026 until a subsequent Sales Threshold is attained; (b) if the Sales Threshold for calendar year 2027 is not attained then the Initial Tier Royalty will increase to 9.5% until January 1 of the calendar year following a calendar year in which the Seller has achieved Net Sales equal to or exceeding Five Hundred Million Dollars ($500,000,000); (c) the Initial Tier Royalty will not exceed 9.5%, regardless of how many of the Sales Thresholds fail to be met; and (d) once a Sales Threshold is met, the Initial Tier Royalty will revert to seven percent (7.0%) beginning January 1 of the following calendar year.
The Royalty Rates above are incremental rates, which apply only for the respective increment of annual Net Sales described in the “Payment Tiers based on Annual Net Sales” column.
“Royalty Report” is defined in Section 6.2(b).
“Sale” means any sale, transfer, assignment or other disposition, not constituting an Out-License or Other License, of any material development, manufacturing, or Commercialization rights relating to Etripamil in the Territory, excluding for all purposes hereunder a Change of Control.
“Sales Threshold” is defined in the definition of “Net Sales”.
“SEC” means the U.S. Securities and Exchange Commission.
“Securities Act” means the Securities Act of 1933, as amended.
“Seller” is defined in the preamble.
“Seller Certificate” is defined in Section 5.1(i)
“Seller Indemnified Parties” is defined in Section 7.1(b)
“Senior Debt Provider” means, collectively, the lenders or providers (or its or their agents or representatives, as applicable) of Permitted Indebtedness secured by Lien(s) on any of the Product Collateral that enter into an
13
Acceptable Intercreditor Agreement executed and delivered by the Buyer, the Seller. Holdings and the applicable Senior Debt Provider.
“Senior Discharge Event” is defined in Section 10.15.
“Senior Obligations” is defined in Section 10.15.
“Solvent” means that (a) the fair saleable value of the Seller’s and Holdings’ consolidated assets is greater than the sum of its debts, liabilities and other obligations, including known contingent liabilities, (b) the present fair saleable value of the Seller’s and Holdings’ consolidated assets is greater than the amount that would be required to pay its liabilities on its existing debts, liabilities and other obligations, including known contingent liabilities, as they become absolute and matured, (c) each of the Seller and Holdings is able to realize upon its assets and pay its debts, liabilities and other obligations, including known contingent obligations, as they mature, (d) neither the Seller nor Holdings has any present plans or intentions to incur, debts or other obligations or liabilities beyond their abilities to pay such debts or other obligations or liabilities as they become absolute and matured, (e) neither the Seller nor Holdings has become subject to any Bankruptcy Event, (f) neither the Seller nor Holdings has been rendered insolvent within the meaning of any applicable law, and (g) no step has been taken or is intended by the Seller or Holdings or, to the Knowledge of the Seller and Holdings, any other Person to make the Seller or Holdings subject to a Bankruptcy Event. For purposes of this definition, the amount of contingent liabilities at any time shall be computed as the amount that, in light of all the facts and circumstances existing at such time, represents the amount that can reasonably be expected to become an actual or matured liability.
“Subsidiary” means any and all corporations, partnerships, limited liability companies, joint ventures, associations and other entities controlled (by contract or otherwise) by Holdings directly or indirectly through one or more intermediaries. For purposes hereof, Holdings shall be deemed to control a partnership, limited liability company, association or other business entity if Holdings, directly or indirectly through one or more intermediaries, shall be allocated a majority of partnership, limited liability company, association or other business entity gains or losses or shall be or control the managing director or general partner of such partnership, limited liability company, association or other business entity. Unless specified otherwise, as used herein, a Subsidiary shall mean a Subsidiary of Holdings, which shall include the Seller.
“Subordinated Obligations” is defined in Section 10.15.
“Tax” or “Taxes” means any federal, state, local or foreign income, gross receipts, license, payroll, employment, excise, severance, occupation, premium, windfall profits, environmental, customs duties, capital stock, franchise, profits, withholding, social security, unemployment, disability, real property, personal property, abandoned property, value added, alternative or add-on minimum, estimated or other tax of any kind whatsoever, including any interest, penalty or addition thereto, whether disputed or not.
“Territory” means the United States of America.
“Third Party” means any Person that is not the Seller or the Seller’s Affiliates.
“Transaction Documents” means this Agreement, the Note Purchase Agreement, any security documents contemplated under the Note Purchase Agreement, and any other ancillary agreement or transaction document contemplated by this Agreement or the Note Purchase Agreement.
“Transaction Expenses” means the aggregate amount of any and all documented out-of-pocket fees and expenses reasonably incurred by or on behalf of, or paid directly by, the Buyer in connection with the transactions contemplated hereby, including diligence and the negotiation, preparation, and execution of the Transaction Documents, and the consummation of the transactions contemplated hereby; [***].
“Treasury Management Arrangement” means any agreement or other arrangement governing the provision of treasury or cash management services, including Deposit Accounts, investment accounts or other accounts in which funds are held or invested to or for the credit or account of Holdings or any of its Subsidiaries, netting services,
14
overdraft, credit or debit card, funds transfer, automated clearinghouse, zero balance accounts, returned check concentration, controlled disbursement, lockbox, account reconciliation and reporting, direct debit, cash concentration, trade finance services, payment processors, and other cash management services.
“Unexpected Contraindications” means (a) any contraindications for Etripamil’s use in the combined group of patients taking calcium channel blockers or beta blockers, or (b) other contraindications or limitations that, taken as a whole, would have a similar impact on the total addressable market for Etripamil in the Territory as the contraindications described in clause (a), in each case of (a) and (b), to the extent set forth in the approved label in connection with U.S. Marketing Approval.
“U.S. Marketing Approval” is defined in Section 5.1(a)
“UCC” means the Uniform Commercial Code as in effect from time to time in the State of New York; provided that, if, with respect to any financing statement or by reason of any provisions of applicable law, the perfection or the effect of perfection or non-perfection of the Back-Up Security Interest or any portion thereof granted pursuant to Section 2.1(b) is governed by the Uniform Commercial Code as in effect in a jurisdiction of the United States other than the State of New York, then “UCC” means the Uniform Commercial Code as in effect from time to time in such other jurisdiction for purposes of the provisions of this Agreement and any financing statement relating to such perfection or effect of perfection or non-perfection.
“UCC Financing Statements” means the UCC-1 financing statements or equivalent in the applicable jurisdiction, in form and substance reasonably satisfactory to the Buyer and the Seller, that shall be filed by the Buyer, with the assistance of the Seller as reasonably requested by the Buyer, at or promptly following the Closing, as well as any additional UCC-1 financing statements or amendments thereto as reasonably requested from time to time, to perfect the Buyer’s ownership interest in the Revenue Participation Right and the Buyer’s Back-up Security Interest in the Revenue Participation Right and Product Collateral.
“Valid Claim” shall mean: (a) any claim of an issued and unexpired Patent included within the Patent Rights, that shall not have been withdrawn, lapsed, abandoned, revoked, canceled or disclaimed, or held invalid or unenforceable by a court, Governmental Entity, national or regional patent office or other appropriate body that has competent jurisdiction in a decision being final and unappealable or unappealed within the time allowed for appeal; and (b) a claim of a pending Patent application included within the Patent Rights that is filed and being prosecuted in good faith and that has not been finally abandoned or finally rejected and which has been pending for no more than seven (7) years from the date of filing of the earliest Patent application to which such pending Patent application claims priority.
“Withholding Action” means (a) a permitted assignment of this Agreement (in whole or in part) by the Buyer to an Affiliate or a Third Party resident in a different jurisdiction; but excluding, in each case, any assignment of this Agreement by the Buyer to an Affiliate of the Buyer or RTW Investments, LP that provides the documentation described in clauses (c)(i) and/or (c)(ii), as applicable; (b) a redomiciliation of Buyer to a jurisdiction other than Ireland; and (c) a failure to deliver, prior to the Closing, either (i) (x) an IRS Form W-9 or (y) an IRS Form W-8BEN-E certifying that it is exempt from U.S. federal withholding Tax in respect of all payments with respect to the Revenue Participation Right under the tax treaty between the United States and Ireland; or (ii) an IRS Form W-8IMY, accompanied by IRS Form(s) W-8BEN-E certifying that the applicable beneficial owner(s) is/are exempt from U.S. federal withholding Tax in respect of all payments with respect to the Revenue Participation Right under an applicable United States income Tax treaty or IRS Form(s) W-9, from the applicable beneficial owner(s), as applicable.
Certain Interpretations. Except where expressly stated otherwise in this Agreement, the following rules of interpretation apply to this Agreement:
(a)“either” and “or” are not exclusive and “include,” “includes” and “including” are not limiting and shall be deemed to be followed by the words “without limitation”;
(b)“extent” in the phrase “to the extent” means the degree to which a subject or other thing extends, and such phrase does not mean simply “if”;
15
(c)“hereof,” “hereto,” “herein” and “hereunder” and words of similar import when used in this Agreement refer to this Agreement as a whole and not to any particular provision of this Agreement;
(d)references to a “Person” are also to its permitted successors and assigns;
(e)definitions are applicable to the singular as well as the plural forms of such terms;
(f)references to an “Article,” “Section” or “Exhibit” refer to an Article or Section of, or an Exhibit to, this Agreement, and references to a “Schedule” refer to the corresponding part of the Disclosure Schedule;
(g)references to “$” or otherwise to dollar amounts refer to the lawful currency of the United States; and
(h)references to a law include any amendment or modification to such law and any rules and regulations issued thereunder, whether such amendment or modification is made, or issuance of such rules and regulations occurs, before or after the date of this Agreement.
Section 1.2Interpretation (Quebec).
For purposes of any assets, liabilities or entities located in the Province of Québec, Canada, including any Product Collateral charged under the security documents entered into in connection with this Agreement, and for all other purposes pursuant to which the interpretation or construction of this Agreement or any other document related thereto may be subject to the laws of the Province of Québec, Canada or a court or tribunal exercising jurisdiction in the Province of Québec, Canada (i) “personal property” shall be deemed to include “movable property”, (ii) “real property” shall be deemed to include “immovable property”, (iii) “tangible property” shall be deemed to include “corporeal property”, (iv) “intangible property” shall be deemed to include “incorporeal property”, (v) “security interest”, “mortgage” and “lien” shall be deemed to include a “hypothec”, “prior claim”, “reservation of ownership” and a “resolutory clause”, (vi) all references to filing, registering or recording under the UCC, the Personal Property Security Act shall be deemed to include publication under the Civil Code of Québec, (vii) all references to “perfection” of or “perfected” liens or security interest shall be deemed to include a reference to an “opposable” or “set up” hypothec as against third parties, (viii) any “right of offset”, “right of setoff” or similar expression shall be deemed to include a “right of compensation”, (ix) “goods” shall be deemed to include “corporeal movable property” other than chattel paper, documents of title, instruments, money and securities, (x) an “agent” shall be deemed to include a “mandatary”, (xi) “construction liens” shall be deemed to include “legal hypothecs in favour of persons having taken part in the construction or renovation of an immovable”; (xii) “joint and several” shall be deemed to include “solidary”; (xiii) “gross negligence or willful misconduct” shall be deemed to be “intentional or gross fault”; (xiv) “beneficial ownership” shall be deemed to include “ownership”; (xv) “legal title” shall be deemed to include “holding title on behalf of an owner as mandatary or prête nom”; (xvi) “easement” shall be deemed to include “servitude”; (xvii) “priority” shall be deemed to include “rank” or “prior claim”, as applicable; (xviii) “survey” shall be deemed to include “certificate of location and plan”; (xix) “fee simple title” shall be deemed to include “ownership” (including ownership under a right of superficies); (xx) “ground lease” shall be deemed to include “emphyteusis” or a “lease with a right of superficies”, as applicable; (xxi) “leasehold interest” shall be deemed to include “a valid lease”; (xxii) “lease” shall be deemed to include a “leasing contract”; (xxiii) “guarantee” and “guarantor” shall include “suretyship” and “surety”, respectively; (xxiv) “foreclosure” shall be deemed to include “the exercise of a hypothecary right”; (xxv) “merger” shall be deemed to include “amalgamation”; (xxvi) “accounts receivables” shall be deemed to include and “claims” (including monetary claims) as referenced in the Civil Code of Québec; and (xxvii) “deposit account” shall be deemed to include “financial account” (as defined in Article 2713.6 of the Civil Code of Québec). The parties hereto confirm that it is their wish that this Agreement and any other document executed in connection with the transactions contemplated herein be drawn up in the English language only (except if another language is required under any applicable law) and that all other documents contemplated thereunder or relating thereto, including notices, may also be drawn up in the English language only. Les parties aux présentes confirment que c’est leur volonté que cette convention et les autres documents de crédit soient rédigés en langue anglaise seulement et que tous les documents, y compris tous avis, envisages par cette convention et les autres documents peuvent être rédigés en langue anglaise seulement, sauf, dans chaque cas, si une autre langue est requise en vertu d’une loi applicable.
16
Article 2.
PURCHASE, SALE AND ASSIGNMENT OF THE REVENUE PARTICIPATION RIGHT
Section 2.1Purchase, Sale and Assignment.
(a)At the Closing and upon the terms and subject to the conditions of this Agreement, the Seller shall sell, transfer, assign and convey to the Buyer, without recourse (except as expressly provided herein), and the Buyer shall purchase, acquire and accept from the Seller, the Revenue Participation Right, free and clear of all Liens. Immediately upon the sale to the Buyer by the Seller of the Revenue Participation Right pursuant to this Section 2.1 all of the Seller’s right, title and interest in and to the Revenue Participation Right shall terminate, and all such right, title and interest shall vest in the Buyer, free and clear of all Liens.
(b)It is the intention of the parties hereto that the sale, transfer, assignment and conveyance contemplated by this Agreement be, and is, a true, complete, absolute and irrevocable sale, transfer, assignment and conveyance by the Seller to the Buyer of all of the Seller’s right, title and interest in and to the Revenue Participation Right. Neither the Seller nor the Buyer intends the transactions contemplated by this Agreement to be, or for any purpose (other than for accounting purposes) characterized as, a loan from the Buyer to the Seller or a pledge, a security interest, a financing transaction or a borrowing. It is the intention of the parties hereto that the beneficial interest in and title to the Revenue Participation Right and any “proceeds” (as such term is defined in the UCC) thereof shall not be part of the Seller’s estate in the event of the filing of a petition by or against the Seller under any Bankruptcy Laws. Each of the Seller and the Buyer hereby waives, to the maximum extent permitted by applicable law, any right to contest or otherwise assert that this Agreement does not constitute a true, complete, absolute and irrevocable sale, transfer, assignment and conveyance by the Seller to the Buyer of all of the Seller’s right, title and interest in and to the Revenue Participation Right under applicable law, which waiver shall, to the maximum extent permitted by applicable law, be enforceable against the Seller in any bankruptcy or insolvency proceeding relating to the Seller. Accordingly, the Seller shall treat the sale, transfer, assignment and conveyance of the Revenue Participation Right as a sale of an “account” or a “payment intangible” (as appropriate) in accordance with the UCC and the Seller hereby authorizes the Buyer to file UCC Financing Statements (and continuation statements with respect to such financing statements when applicable) naming the Seller as the debtor/Seller and the Buyer as the secured party/Buyer in respect to the Revenue Participation Right. For sake of clarification, the foregoing statements in this Section 2.1(b) shall not bind either party regarding the reporting of transactions contemplated hereby for GAAP or securities law reporting purposes. Not in derogation of the foregoing statement of the intent of the parties hereto in this regard, and for the purposes of providing additional assurance to the Buyer in the event that, despite the intent of the parties hereto, the sale, transfer, assignment and conveyance contemplated hereby is hereafter held not to be a sale, each of Holdings and the Seller does hereby grant to the Buyer, as security for the payment of amounts to the Buyer equal to the sum of the Purchase Price (including a market rate of return thereon) less all Royalty Payments received by the Buyer pursuant to this Agreement, a first priority security interest in and to all of its respective right, title and interest in, to and under the Revenue Participation Right and the Royalty Payments and a security interest in and to all right, title and interest in, to and under the Product Collateral, and Holdings and the Seller do hereby authorize the Buyer, from and after the Closing, to file UCC Financing Statements, and continuation statements with respect to such financing statements when applicable meeting the requirements of applicable law, in such manner and such jurisdictions as are necessary or appropriate to perfect such security interest (the “Back-Up Security Interest”) (or, in respect of any subsequent Permitted Out-License, in each case with respect to Commercialization of a Product in the Territory, a first priority security interest in and to all right, title and interest in, to and under the Revenue Participation Right and the Royalty Payments) and naming the Seller or Holdings, as applicable, as the debtor and the Buyer as the secured party in respect to the Revenue Participation Right.
(c)[***].
Section 2.2Purchase Price. At the Closing and upon the terms and subject to the conditions of this Agreement, the Buyer shall pay the Seller the purchase price of Seventy-Five Million Dollars ($75,000,000) in cash, as consideration to the Seller for the sale, transfer, assignment and conveyance of the Revenue Participation Right to the Buyer (the “Purchase Price”).
17
Section 2.3No Assumed Obligations, Etc. Notwithstanding any provision in this Agreement to the contrary, the Buyer is only agreeing, on the terms and conditions set forth in this Agreement, to purchase, acquire and accept the Revenue Participation Right and is not assuming any liability or obligation of the Seller of whatever nature, whether presently in existence or arising or asserted hereafter. All such liabilities and obligations shall be retained by and remain obligations and liabilities of the Seller or its Affiliates.
Article 3.
CLOSING
Section 3.1Closing. The Closing shall take place remotely via the exchange of documents and signatures on the date (the “Closing Date”) that is thirty (30) calendar days, or such period as mutually agreed upon by the parties hereto, after the date on which the condition set forth in Section 5.1(a) is satisfied, subject to the satisfaction or waiver of the other conditions set forth in ARTICLE 5
Section 3.2Payment of Purchase Price. At the Closing, the Buyer shall deliver (or cause to be delivered) payment of the Purchase Price to the Seller by electronic funds transfer or wire transfer of immediately available funds to one or more accounts specified by the Seller.
Section 3.3Bill of Sale. At the Closing, upon confirmation of the receipt of the Purchase Price, the Seller shall deliver to the Buyer a duly executed bill of sale evidencing the sale, transfer, assignment and conveyance of the Revenue Participation Right in substantially the form attached hereto as Exhibit B.
Article 4.
REPRESENTATIONS AND WARRANTIES
Section 4.1Seller’s Representations and Warranties. Except as set forth on the Disclosure Schedules attached hereto, Holdings and the Seller represent and warrant to the Buyer that as of the date hereof or as of the Amendment Date, as applicable, and, unless specified otherwise, as of the Closing Date:
(a)Existence; Good Standing. The Seller is a corporation duly incorporated, validly existing and in good standing under the laws of Delaware and Holdings is a corporation duly incorporated, validly existing and in good standing under the laws of Quebec. Each of the Seller and Holdings is duly licensed or qualified to do business and is in corporate good standing in each jurisdiction in which the nature of the business conducted by it or the character or location of the properties and assets owned, leased or operated by it makes such licensing or qualification necessary, except where the failure to be so licensed or qualified and in corporate good standing has not and would not reasonably be expected to have, either individually or in the aggregate, a Material Adverse Effect.
(b)Authorization. Each of the Seller and Holdings has all requisite corporate power and authority to execute, deliver and perform its obligations under this Agreement. The execution, delivery and performance of this Agreement, and the consummation of the transactions contemplated hereby, have been duly authorized by all necessary corporate action on the part of the Seller or Holdings, as applicable.
(c)Enforceability. This Agreement has been duly executed and delivered by an authorized officer of the Seller and Holdings and constitutes the valid and binding obligation of the Seller and Holdings, enforceable against the Seller and Holdings in accordance with its terms, except as may be limited by applicable Bankruptcy Laws or by general principles of equity (whether considered in a proceeding in equity or at law).
(d)No Conflicts. The execution, delivery and performance by the Seller and Holdings of this Agreement and the consummation of the transactions contemplated hereby and thereby do not and will not (i) contravene or conflict with the certificate of incorporation or bylaws of the Seller or Holdings, (ii) contravene or conflict with or constitute a material default under any law binding upon or applicable to the Seller, Holdings or the Revenue Participation Right or (iii) contravene or conflict with or constitute a material default under any material agreement or Judgment binding upon or applicable to the Seller, Holdings or the Revenue Participation Right.
18
(e)Consents. Except for the consents that have been obtained on or prior to the Closing, the UCC financing statements contemplated by Section 2.1(b), or any filings required by the federal securities laws or stock exchange rules, no consent, approval, license, order, authorization, registration, declaration or filing with or of any Governmental Entity or other Person is required to be done or obtained by the Seller or Holdings in connection with (i) the execution and delivery by the Seller and Holdings of this Agreement, (ii) the performance by the Seller and Holdings of its obligations under this Agreement or (iii) the consummation by the Seller and Holdings of any of the transactions contemplated by this Agreement.
(f)No Litigation. Neither Holdings nor any of its Subsidiaries is a party to, or has received any written notice of, any action, suit, investigation or proceeding pending before any Governmental Entity and, to the Knowledge of the Seller and Holdings, no such action, suit, investigation or proceeding has been threatened against the Seller or Holdings, that, individually or in the aggregate, has had or would reasonably be expected to have a Material Adverse Effect.
(g)Compliance.
(i)All applications, submissions, information and data related to a Product submitted or utilized as the basis for any request to any Regulatory Authority by or on behalf of the Seller or Holdings were true and correct in all material respects as of the date of such submission or request, and, to the Knowledge of the Seller and Holdings, any material updates, changes, corrections or modification to such applications, submissions, information or data required under applicable laws or regulations have been submitted to the necessary Regulatory Authorities.
(ii)Neither Holdings nor any of its Subsidiaries has committed any act, made any statement or failed to make any statement that would reasonably be expected to provide a basis for the FDA or other Regulatory Authority to invoke its policy with respect to “Fraud, Untrue Statements of Material Facts, Bribery, and Illegal Gratuities,” or similar policies, set forth in any applicable laws or regulations.
(iii)Neither Holdings, its Subsidiaries, or to the Knowledge of the Seller and Holdings, any Third Party acting on behalf of Seller or Holdings, has used in any capacity the services of a person debarred, excluded or disqualified (or convicted of any crime or engaged in any conduct for which debarment, exclusion or disqualification is mandated) under 21 U.S.C. § 335a.
(iv)Each of the Seller and Holdings has provided to the Buyer prior to the Amendment Date in a data room available to the Buyer true and correct copies of all material written communications sent or received by the Seller and any of its Affiliates to or from any Regulatory Authorities in the United States and Canada that relate to each Product since March 27, 2023, including but not limited to any written reports or other written communications received from a Governmental Entity that would indicate that any Regulatory Authority (A) is likely to reject, condition, or delay any application for Marketing Approval, or (B) is likely to pursue any material compliance actions against the Seller.
(v)As of the Amendment Date, Etripamil has not been the subject of a prior Marketing Approval in the Territory and as of the Closing Date, Etripamil has not been the subject of a prior Marketing Approval in the Territory, other than the U.S. Marketing Approval.
(h)Licenses.
(i)In-Licenses. Except for In-Licenses that are permitted under this Agreement, there are no In-Licenses.
(i)Out-Licenses. Except for Permitted Out-Licenses, there are no Out-Licenses.
(ii)No Liens; Title to Revenue Participation Right. None of the Product Collateral is subject to any Lien, except for Permitted Liens and Liens created hereunder as of the Closing Date. The Seller has the full right to sell, transfer, convey and assign to Buyer all of the Seller’s rights and interests in
19
and to the Revenue Participation Right being sold, transferred, conveyed and assigned to Buyer pursuant to this Agreement without any requirement to obtain the consent of any Person (including Holdings). The claims and rights of Buyer created by this Agreement in and to the Revenue Participation Right and any other Product Collateral are not subordinated in right of payment to any creditor of the Seller or any other Person (except to the extent specified in any Acceptable Intercreditor Agreement). Upon the Closing, the Buyer will have acquired good and valid title to the Revenue Participation Right, free and clear of all Liens, except for Liens for Taxes, assessments or governmental charges or levies not yet due.
(j)Manufacturing; Supply. All Products have, since June 30, 2019, been manufactured, transported, stored and handled in all material respects in accordance with applicable law and with good manufacturing practices. Since June 30, 2019, neither the Seller nor any Affiliate of the Seller has experienced any failures in the manufacturing or supply of any Product that, individually or in the aggregate, have had or would reasonably be expected to result in a Material Adverse Effect. The Seller has on hand or has made adequate provisions to secure sufficient clinical quantities of Etripamil to complete all clinical trials and all activities required for Marketing Approvals, in each case, that are ongoing or planned as of the Amendment Date. As of the Closing Date, the Seller has on hand or has made adequate provisions to secure sufficient quantities of Etripamil to support the commercial launch of Products in the Territory, and Seller and Holdings will each take such additional actions as may be reasonably necessary during the Term to ensure that Seller has reasonably sufficient quantities of Product to permit Seller to meet demand for Products in the Territory during the Term. Holdings may not, and shall not, privilege its own needs with respect to Product supply over the needs of Seller.
(k)Intellectual Property.
(l)As of the Amendment Date, Schedule 4.1(k)(i) of the Disclosure Schedule lists all of the currently existing Patents included within the Patent Rights in the Territory (the “Existing Patent Rights”). As of the Closing Date, the updated Schedule 4.1(k)(i) of the Disclosure Schedule delivered as of the Closing Date lists all of the then existing Patents included within the Patent Rights as of the Closing Date. The Seller or Holdings is the sole and exclusive owner of all of the Existing Patent Rights. Schedule 4.1(k)(i) of the Disclosure Schedule specifies as to each listed patent or patent application the jurisdictions by or in which each such patent has issued as a patent or such patent application has been filed, including the respective patent or application numbers.
(i)Neither Holdings nor any of its Subsidiaries is a party to any pending and, to the Knowledge of the Seller and Holdings, there is no threatened litigation, interference, reexamination, opposition or like procedure involving any of the Existing Patent Rights in the Territory or any currently existing Patent included within the Patent Rights outside of the Territory.
(ii)All of the issued patents within the Existing Patent Rights are (A) to the Knowledge of the Seller and Holdings, valid and enforceable, and (B) in full force and effect. None of the issued patents within the Existing Patent Rights have lapsed, expired or otherwise terminated. Neither Holdings nor any of its Subsidiaries has received any written notice relating to the lapse, expiration or other termination of any of the issued patents within the Existing Patent Rights in the Territory, and neither Holdings nor its Subsidiaries has received any written legal opinion that alleges that, an issued patent within any of the Existing Patent Rights is invalid or unenforceable.
(iii)Neither Holdings nor any of its Subsidiaries has received any written notice that there is any, and, to the Knowledge of the Seller and Holdings, there is no Person who is or claims to be an inventor under any of the Existing Patent Rights who is not a named inventor thereof.
(iv)Neither Seller nor its Affiliates has received any written notice of any claim by any Person challenging the inventorship or ownership of, the rights of the Seller or Holdings in and to, or the patentability, validity or enforceability of, any of the Existing Patent Rights, or asserting that the development, manufacture, importation, sale, offer for sale or use of a Product infringes, misappropriates or otherwise violates or will infringe, misappropriate or otherwise violate such Person’s Patents or other intellectual property rights.
20
(v)To the Knowledge of the Seller and Holdings, the discovery, development manufacture, importation, sale, offer for sale or use of each Product, in each case in the form such Product exists as of the Amendment Date and as such activity is currently contemplated by the Seller, has not and will not, infringe, misappropriate or otherwise violate any Patents or other intellectual property rights owned by any Third Party.
(vi)To the Knowledge of the Seller and Holdings, no Person has infringed, misappropriated or otherwise violated, or is infringing, misappropriating or otherwise violating, any of the Intellectual Property Rights.
(vii)The Seller or Holdings has paid all maintenance fees, annuities and like payments required as of the Amendment Date with respect to each of the Existing Patent Rights.
(viii)Indebtedness; No Undisclosed Indebtedness. Other than any Indebtedness owed under the Note Purchase Agreement upon signing and closing the transaction contemplated by the Note Purchase Agreement and Permitted Indebtedness, (i) neither Holdings or any of its Subsidiaries has any outstanding Indebtedness, and (ii) there are no liabilities (excluding Indebtedness or obligations under the Intercompany License) of Holdings or its Subsidiaries related to a Product not incurred in the ordinary course.
(m)Solvency. Each of Holdings and the Seller has determined that, and by virtue of its entering into the transactions contemplated by the respective Transaction Documents to which each is party and the respective authorization, execution and delivery of the Transaction Documents to which each is party, each of Holdings’ and the Seller’s incurrence of any liability hereunder or thereunder or contemplated hereby or thereby is in its own best interests. Upon consummation of the transactions contemplated by the Transaction Documents on the Amendment Date and the application of the proceeds therefrom, each of Holdings and the Seller will be Solvent.
(n)Lien Related Representation and Warranties. The Seller’s exact legal name is, and for the immediately preceding five (5) years has been, “Milestone Pharmaceuticals USA, Inc.” The Seller is, and for the prior five (5) years has been, incorporated in the state of Delaware. Holdings’ exact legal name is, and for the immediately preceding ten (10) years has been, “Milestone Pharmaceuticals Inc.” The Seller is, and for the prior ten (10) years has been, incorporated in a jurisdiction of Canada.
(o)Brokers’ Fees. Except for Cowen Inc., the fees of which will be paid by the Seller or Holdings, there is no investment banker, broker, finder, financial advisor or other intermediary who has been retained by or is authorized to act on behalf of the Seller or Holdings who might be entitled to any fee or commission in connection with the transactions contemplated by this Agreement.
(p)Public Company Reporting Obligations. Holdings has filed or furnished (as applicable) with or to the SEC all registration statements, forms, reports, certifications and other documents required to be filed or furnished by Holdings with or to the SEC since January 1, 2022 (all such registration statements, forms, reports, certifications and other documents (including those that Holdings may file or furnish after the Amendment Date until the Closing and Holdings’ Annual Report on Form 10-K for the year ended December 31, 2022) are referred to herein as the “Holdings SEC Documents”). Holdings SEC Documents (i) were filed or furnished on a timely basis or has received a valid extension of such time of filing and has filed any such Holdings SEC Documents prior to the expiration of any such extension, (ii) at the time filed or furnished, were prepared in compliance as to form in all material respects with the applicable requirements of the Securities Act and the Securities Exchange Act of 1934, as amended, as the case may be, and the rules and regulations of the SEC thereunder applicable to such Holdings SEC Documents, except to the extent that information contained in any Holdings SEC Document has been revised or superseded by a later filed Holdings SEC Document filed and made publicly available prior to the date of this Agreement, and (iii) did not at the time they were filed or furnished contain any untrue statement of a material fact or omit to state a material fact required to be stated in such Holdings SEC Documents or necessary in order to make the statements in such Holdings SEC Documents, in the light of the circumstances under which they were made, not materially misleading. Holdings’ financial statements included within Holdings SEC Documents have been prepared in accordance with accounting principles generally accepted in the United States and such financial statements do not contain an untrue statement of a material fact or omit to state a material fact required to be stated therein or necessary to make the statements therein not materially misleading at the time made.
21
Section 4.2Buyer’s Representations and Warranties. The Buyer hereby represents and warrants to the Seller as of the date hereof or as of the Amendment Date, as applicable, that:
(a)Existence; Good Standing. The Buyer is a designated activity company duly incorporated, validly existing and in good standing under the laws of Ireland.
(b)Authorization. The Buyer has the requisite corporate power and authority to execute, deliver and perform its obligations under this Agreement. The execution, delivery and performance of this Agreement, and the consummation of the transactions contemplated hereby, have been duly authorized by all necessary action on the part of the Buyer.
(c)Enforceability. This Agreement has been duly executed and delivered by an authorized person of the Buyer and constitutes the valid and binding obligation of the Buyer, enforceable against the Buyer in accordance with its terms, except as may be limited by applicable Bankruptcy Laws or by general principles of equity (whether considered in a proceeding in equity or at law).
(d)No Conflicts. The execution, delivery and performance by the Buyer of this Agreement do not and will not (i) contravene or conflict with the organizational documents of the Buyer, (ii) contravene or conflict with or constitute a default under any material provision of any law binding upon or applicable to the Buyer or (iii) contravene or conflict with or constitute a default under any material contract or other material agreement or Judgment binding upon or applicable to the Buyer.
(e)Consents. Except for any filings required by the federal securities laws or stock exchange rules, no consent, approval, license, order, authorization, registration, declaration or filing with or of any Governmental Entity or other Person is required to be done or obtained by the Buyer in connection with (i) the execution and delivery by the Buyer of this Agreement, (ii) the performance by the Buyer of its obligations under this Agreement or (iii) the consummation by the Buyer of any of the transactions contemplated by this Agreement.
(f)No Litigation. There is no action, suit, investigation or proceeding pending or, to the knowledge of the Buyer, threatened before any Governmental Entity to which the Buyer is a party that would, if determined adversely, reasonably be expected to prevent or materially and adversely affect the ability of the Buyer to perform its obligations under this Agreement.
(g)Financing. The Buyer will have sufficient cash to pay the Purchase Price at the Closing. The Buyer acknowledges that its obligations under this Agreement are not contingent on obtaining financing.
(h)Brokers’ Fees. There is no investment banker, broker, finder, financial advisor or other intermediary who has been retained by or is authorized to act on behalf of the Buyer who might be entitled to any fee or commission in connection with the transactions contemplated by this Agreement.
Section 4.3No Implied Representations and Warranties. The Buyer acknowledges and agrees that, other than the express representations and warranties of the Seller specifically contained in ARTICLE 4 (a) there are no representations or warranties of the Seller either expressed or implied with respect to the Patent Rights or Royalty Payment and that the Buyer does not rely on, and shall have no remedies in respect of, any representation or warranty not specifically set forth in ARTICLE 4 and all other representations and warranties are hereby expressly disclaimed, and (b) nothing contained herein guarantees that sales of the Products or the aggregate Royalty Payments due to the Buyer will achieve any specific amounts (it being understood and agreed that nothing in this Section 4.3 shall limit in any way the Seller’s obligations under ARTICLE 8). Notwithstanding the foregoing, claims for fraud, gross negligence, or willful misconduct shall not be waived or limited in any way by this Section 4.3. Except for the Revenue Participation Right, Back-Up Security Interest and the Buyer’s rights under Section 6.5(e) the Buyer further acknowledges and agrees that no licenses or assignments under any assets (including the Patent Rights or any other intellectual property) of the Seller and its Affiliates are granted pursuant to this Agreement, including by implication, estoppel, exhaustion or otherwise.
22
Article 5.
CONDITIONS TO CLOSING
Section 5.1Conditions to the Buyer’s Obligations. The obligations of the Buyer to consummate the transactions contemplated hereunder on the Closing Date are subject to the satisfaction or waiver, at or prior to the Closing Date, of each of the following conditions precedent:
(a)The Seller or Holdings shall have received Marketing Approval from the FDA, based on the active ingredient Etripamil, for the Commercialization of Etripamil in the United States (“U.S. Marketing Approval”) for the Approved Indication on or prior to the Marketing Approval Deadline.
(b)The Seller and Holdings shall have performed and complied in all material respects with all agreements, covenants, obligations and conditions required to be performed and complied with by it under this Agreement at or prior to the Closing Date, and the Buyer shall have received a certificate executed by a duly authorized officer of the Seller on the Closing Date certifying on behalf of the Seller to the effect of the foregoing.
(c)The Seller shall have delivered to Buyer an updated Schedule 4.1(k)(i);
(d)The representations and warranties of the Seller and Holdings contained in Section 4.1 shall have been true and correct in all material respects as of the date hereof and shall be true and correct in all material respects as of the Closing Date as though made at and as of the date hereof and as of the Closing Date, respectively, except to the extent any such representation or warranty expressly speaks as of a particular date, in which case it shall be true and correct in all material respects as of such date; provided that (A), to the extent that any such representation or warranty is qualified by the term “material” or “Material Adverse Effect” such representation or warranty (as so written, including the term “material” or “Material Adverse Effect”) shall have been true and correct in all respects as of the date hereof and shall be true and correct in all respects as of the Closing Date or such other date, as applicable, and (B) notwithstanding the foregoing, the representations and warranties of Holdings and the Seller contained in Sections Section 4.1(g) and 4.1(k)(ii) through (viii) shall be true and correct as of the Closing Date, except as would not reasonably be expected to have a Material Adverse Effect. The Buyer shall have received a certificate executed by an authorized officer of the Seller and Holdings on the Closing Date certifying on behalf of the Seller and Holdings to the effect of the foregoing.
(e)No event or events shall have occurred, or be reasonably likely to occur, that, individually or in the aggregate, have had or would reasonably be expected to result in (or, with the giving of notice, the passage of time or otherwise, would result in) a material adverse effect [***]. The Buyer shall have received a certificate executed by a duly authorized officer of the Seller on the Closing Date certifying on behalf of the Seller to the effect of the foregoing.
(f)There shall not have been issued and be in effect any Judgment of any Governmental Entity enjoining, preventing or restricting the consummation of the transactions contemplated by this Agreement.
(g)There shall not have been instituted or be pending any action or proceeding by any Governmental Entity or any other Person (i) challenging or seeking to make illegal, to delay materially or otherwise directly or indirectly to restrain or prohibit the consummation of the transactions contemplated hereby, (ii) seeking to obtain material damages in connection with the transactions contemplated hereby or (iii) seeking to restrain or prohibit the Buyer’s purchase of the Revenue Participation Right.
(h)The Buyer shall have received a fully executed copy of the Note Purchase Agreement and no Event of Default (as defined in the Note Purchase Agreement) shall have occurred and be continuing. Upon consummation of the transactions contemplated by the Transaction Documents on the Closing Date and the application of the proceeds therefrom, each of the Seller and Holdings will be Solvent.
(i)The Buyer shall have received a certificate of an officer of the Seller and Holdings, dated the Closing Date, certifying as to (i) the incumbency of each officer of the Seller and Holdings executing this
23
Agreement and (ii) the attached thereto copies of (A) each of Holdings’ and the Seller’s formation document, (B) bylaws or other operating agreement, and (C) resolutions adopted by the applicable governing bodies of Holdings and the Seller authorizing the respective execution and delivery by Holdings and the Seller of this Agreement and the consummation by Holdings and the Seller of the transactions contemplated hereby (the “Seller Certificate”).
(j)The Seller shall have confirmed it has scheduled delivery to Buyer of a CD or USB containing copies of all documents uploaded to the data room related to the transactions contemplated by this Agreement, as of the date hereof, maintained by the Seller and made available to the Buyer, including all documents referred to in Section 4.1(g)(iv)
(k)At or prior to (i) the date hereof, the Seller shall have paid the Transaction Expenses incurred prior to or on the date hereof, and (ii) the Closing Date, the Seller shall have paid the aggregate amount of, if applicable, any and all other Transaction Expenses incurred prior to or on the Closing Date; provided that the condition set forth in this Section 5.1(k)(ii) will be satisfied by the transfer by the Buyer of an amount equal to the Purchase Price minus the Transaction Expenses owed by the Seller under this Section 5.1(k)(ii).
Section 5.2Conditions to the Seller’s Obligations. The obligations of the Seller to consummate the transactions contemplated hereunder on the Closing Date are subject to the satisfaction or waiver, at or prior to the Closing Date, of each of the following conditions precedent:
(a)The Buyer shall have performed and complied in all material respects with all agreements, covenants, obligations and conditions required to be performed and complied with by it under this Agreement at or prior to the Closing Date, and the Seller shall have received a certificate executed by a duly authorized person of the Buyer, on the Closing Date certifying on behalf of the Buyer to the effect of the foregoing.
(b)The representations and warranties of the Buyer contained in Section 4.2 shall have been true and correct in all material respects as of the date hereof and shall be true and correct in all material respects as of the Closing Date as though made at and as of the date hereof and Closing Date, respectively, except to the extent any such representation or warranty expressly speaks as of a particular date, in which case it shall be true and correct in all material respects as of such date; provided, that to the extent that any such representation or warranty is qualified by the term “material,” or “Material Adverse Effect” such representation or warranty (as so written, including the term “material” or “Material Adverse Effect”) shall have been true and correct in all respects as of the date hereof and shall be true and correct in all respects as of the Closing Date or such other date, as applicable. The Seller shall have received a certificate executed by a duly authorized person of Buyer, on the Closing Date certifying on behalf of the Buyer to the effect of the foregoing.
(c)There shall not have been issued and be in effect any Judgment of any Governmental Entity enjoining, preventing or restricting the consummation of the transactions contemplated by this Agreement.
(d)There shall not have been instituted or be pending any action or proceeding by any Governmental Entity or any other Person (i) challenging or seeking to make illegal, to delay materially or otherwise directly or indirectly to restrain or prohibit the consummation of the transactions contemplated hereby, (ii) seeking to obtain material damages in connection with the transactions contemplated hereby or (iii) seeking to restrain or prohibit the Buyer’s purchase of the Revenue Participation Right.
(e)The Seller shall have received a fully executed copy of the Note Purchase Agreement.
(f)The Seller shall have received a certificate of an authorized person of the Buyer, dated the Closing Date, certifying as to the incumbency of the officers executing this Agreement on behalf of the Buyer.
Article 6.
COVENANTS
24
Section 6.1Reports; Meetings. From and after the date hereof, the Seller (or Holdings on behalf of the Seller) shall provide the Buyer:
(a)Following the end of each Calendar Quarter, but in any event, in each case, no later than the Reporting Date for such Calendar Quarter, a reasonably detailed quarterly report setting forth, with respect to such same period, (i) the Clinical Updates, (ii) the Commercial Updates, (iii) the Regulatory Updates, and (iv) the Intellectual Property Updates (the “Reports”).
(b)The Seller shall include in the Commercial Updates, any (i) material [***] updates and (ii) details as to the achievement of any development, sales, regulatory or other milestone event set forth in any Permitted Out-License and any material Other Licenses.
(c)The Seller shall promptly notify Buyer (and in no event more than [***] of Seller’s or Holdings’ Knowledge of the following events) of (i) any material action, demand, suit, claim, cause of action, proceeding or investigation pending or, to the Knowledge of the Seller and Holdings, threatened (in writing) by or against Holdings or any of its Subsidiaries, or (ii) any material proceeding or inquiry of any Governmental Entity pending or, to the Knowledge of the Seller and Holdings, threatened (in writing) against Holdings or any of its Subsidiaries, in each case, related to any Product, the Product Collateral or any Transaction Document.
(d)During the term of this Agreement, in the event that the Seller or any of its Affiliates enters into any Permitted Out-License, commercialization, co-promotion, collaboration, distribution, marketing or partnering program with respect to the Product, in each case that grants a license with respect to the Intellectual Property Product Rights to any Subsidiary of Holdings, at least [***] prior to the consummation of any such transaction, the Seller shall give the Buyer written notice thereof and will prior to such consummation cause any such Subsidiary to execute and deliver to the Buyer a joinder agreement and other documents reasonably requested and satisfactory to the Buyer in order to cause such Subsidiary to become a party to the applicable Transaction Documents as if such Subsidiary was a party thereto as of the date hereof.
(e)The Seller shall also provide the Buyer with such additional information regarding the updates included in each Report or Royalty Report as the Buyer may reasonably request from time to time, provided if there is more than one Buyer, such inspection and/or audit shall only be exercised by one Buyer Representative. At the Buyer Representative’s election, the Buyer Representative shall be entitled to a [***] update meeting, or as the Buyer Representative may otherwise reasonably request, to discuss the Reports and Royalty Reports delivered by the Seller pursuant to Section 6.1 and Section 6.2(b), respectively. The Seller shall prepare and maintain and shall cause Holdings and Holdings’ Subsidiaries and use Commercially Reasonable Efforts to cause its Affiliates and Licensees to prepare and maintain reasonably complete and accurate records of the information to be disclosed in each Report and each Royalty Report. All Reports and Royalty Reports, and the Confidential Information contained therein, shall be the Confidential Information of the Seller and Holdings and subject to the obligations of confidentiality set forth in ARTICLE 8
Section 6.2Royalty Payments; Royalty Reports; Buy-Back Option.
(a)From and after the First Commercial Sale of a Product in the Territory, the Seller shall pay to the Buyer, without any setoff or offset (subject, in each case, to Section 6.12 and Section 6.13), the Royalty Payment for each Calendar Quarter on or prior to the Reporting Date for such Calendar Quarter; provided that for any Net Sales made by a Related Party (except Holdings) for which payment is received by the Seller fewer than ten (10) calendar days prior to the applicable Reporting Date, such payment to the Buyer will be paid with the following Calendar Quarter’s Royalty Payment, and further provided that any adjustments to the Royalty Payment for a Calendar Quarter based on or arising out of any discrepancies with Holdings’ Form 10-Q or Form 10-K filed with the SEC will be paid with or credited against, as applicable, the following Calendar Quarter’s Royalty Payment. The Royalty Payment payable for the Calendar Quarter during which the Closing occurs shall be based on the entire period of such Calendar Quarter, and not prorated from the Closing Date through the end of such Calendar Quarter. A late fee of [***] over the Prime Rate (calculated on a per annum basis) will accrue on all unpaid amounts with respect to any Royalty Payment from the date such obligation was due. The imposition and payment of a late fee shall not constitute a waiver of the Buyer’s rights with respect to such payment default.
25
(b)From and after the First Commercial Sale of a Product in the Territory, for each Calendar Quarter, no later than the Reporting Date for such Calendar Quarter, the Seller shall provide to the Buyer a report (a “Royalty Report”), in substantially the form attached to this Agreement as Exhibit C, setting forth in reasonable detail with respect to all Products, (A) Gross Sales and Net Sales (1) for the applicable Calendar Quarter, (2) the most recent period of four (4) consecutive Calendar Quarters ended as of the last day of such Calendar Quarter, and (3) the calendar year to date, in each case, on a Product-by-Product basis (including a reasonably detailed break-down of all permitted deductions from Gross Sales used to determine Net Sales and any Net Sales described in Section 6.5(e)); (B) the calculation of the Royalty Payment payable to the Buyer for the applicable Calendar Quarter, identifying, on a Product-by-Product basis, the number of units of each Product sold by the Seller, its Affiliates, and each Licensee; and (C) if applicable, foreign currency exchange rates used (which shall be rates of exchange determined in a manner consistent with Holdings’ method for calculating rates of exchange in the preparation of Holdings’ annual financial statements in accordance with GAAP); provided that for any reports received by the Seller with respect to Net Sales by Licensees or Distributors fewer than [***] prior to the Reporting Date, the Seller shall deliver to the Buyer the relevant information from such reports in the following Calendar Quarter’s report and, provided, further, that any adjustments to the Royalty Payment for a Calendar Quarter based on or arising out of any discrepancies with Holdings’ Form 10-Q or Form 10-K filed with the SEC will be paid with or credited against, as applicable, the following Calendar Quarter’s Royalty Payment.
(c)Any payments required to be made by either party under this Agreement shall be made in United States Dollars via electronic funds transfer or wire transfer of immediately available funds to such bank account as the other party shall designate in writing prior to the date of such payment.
(d)If Holdings enters into a definitive agreement with a Person that is not an Affiliate of Seller to consummate a Change of Control (a “CoC Agreement”), Holdings or such Person shall have the option to prepay (the “Buy-Back Option”), or the Buyer Representative (if more than one Buyer, on behalf of all Buyers) may, other than in a Change of Control involving a Qualified Buyer, require the Seller to prepay or cause a prepayment (the “Buy-Back Requirement”) of, a prespecified amount (“the CoC Payment”) on the date of consummation of such Change of Control (the “CoC Date”) to the Buyer and terminate this Agreement and all obligations hereunder and in respect of the Revenue Participation Right, with such pre-specified payment amount to be based on the CoC Date, as follows: (a) [***] if the CoC Date is after [***] but prior to or on [***], (b) [***], [***] if the CoC Date is after [***] but prior to or on [***], (c) [***], [***] if the CoC Date is after [***] but prior to or on [***], and (d) an amount equal to [***] if the CoC Date is after [***]. Upon entering into a CoC Agreement by Holdings, the Seller shall promptly but no later than [***] thereafter deliver notice of entering into such CoC Agreement to the Buyer, including whether the Seller is exercising a Buy-Back Option and whether the acquiring party is a Qualified Buyer. If the Seller elects to exercise the Buy-Back Option, or if the Buyer Representative (if more than one Buyer, on behalf of all Buyers) exercises the Buy-Back Requirement by delivering notice to the Seller within [***] after receiving such notice from the Seller, the Seller shall or shall cause to, promptly but no later than [***] following the consummation of such Change of Control, pay the CoC Payment to the Buyer. The Seller’s obligation to pay the CoC Payment following the Seller’s exercise of the Buy-Back Option or the Buyer Representative’s exercise of the Buy-Back Requirement shall be contingent upon the consummation of such Change of Control; if such Change of Control is not consummated, the exercise of such Buy-Back Option or such Buy-Back Requirement shall be void. The payment of the CoC Payment shall be made by wire transfer of immediately available funds to one or more accounts specified by the Buyer or, if not timely designated by Buyer, to the account to which payments under the Royalty Payments were transmitted or are to be transmitted pursuant to Section 6.2 Upon Buyer’s receipt of the CoC Payment, except as set forth in Section 9.5 this Agreement shall terminate and all rights and obligations of the parties hereunder and in respect of the Revenue Participation Right shall automatically without any further action of the parties be deemed to be released and irrevocably terminated. Notwithstanding any of the foregoing to the contrary, in the event that the Buy-Back Option or Buy-Back Requirement is exercised during any Calendar Quarter prior to or on the second anniversary of the Closing Date, during which the Seller has earned Net Sales and would otherwise be obligated to make a Royalty Payment to the Buyer, the Seller shall be obligated to make all Royalty Payments otherwise due in accordance with Section 6.2 for all such earned Net Sales up to the date that is [***] prior to the date such CoC Payment is due. The Seller shall remit the CoC Payment to the Buyer on the CoC Payment due date, and such CoC Payment shall include the foregoing amount of such final Royalty Payment.
Section 6.3Disclosures. Except for a press release and Holdings’ Form 10-K or Form 8-K describing the material terms of this Agreement and the transactions contemplated by this Agreement, in each case, previously
26
approved in form and substance by the Seller and the Buyer or any other public announcement using substantially the same text or disclosing substantially the same substance as such press release, Form 10-K, or Form 8-K, neither the Buyer nor the Seller shall, and each party hereto shall cause its respective Representatives, Affiliates and Affiliates’ Representatives not to, issue a press release or other public announcement or otherwise make any public disclosure with respect to this Agreement or the subject matter hereof without the prior written consent of the other party hereto (which consent shall not be unreasonably withheld or delayed), except as may be required by applicable law or stock exchange rule (in which case the party hereto required to make the press release or other public announcement or disclosure shall allow the other party hereto reasonable time to comment on such press release or other public announcement or disclosure in advance of such issuance, and such party hereto required to make the press release or other public announcement or disclosure shall reasonably consider such other party hereto’s reasonable comments), provided that no review or consent shall be required with respect to disclosures by either party hereto otherwise previously approved pursuant to this Section 6.3 and (b) notwithstanding anything herein to the contrary, each party hereto may, without the review or consent of the other party hereto, disclose (and nothing herein shall be construed to restrict either party hereto from disclosing) the Purchase Price and the amount and nature of the Revenue Participation Right and Royalty Payments (and relating accounting disclosures of the transactions contemplated hereby) in such party’s periodic reports and financial statements (other than such party’s initial disclosure regarding the transactions contemplated by this Agreement); provided that with respect to disclosing the nature of the Revenue Participation Right, such disclosure is consistent with prior disclosures.
Section 6.4Inspections and Audits of the Seller. Following the Closing, upon at least [***] written notice and during normal business hours, no more frequently than [***] per [***], the Buyer Representative may cause an inspection and/or audit by an independent public accounting firm reasonably acceptable to the Seller to be made of the Seller’s (or Holdings’) books of account for the [***] prior to the audit for the purpose of determining the correctness of Royalty Payments made under this Agreement, provided if there is more than one Buyer, such inspection and/or audit shall only be exercised by one Buyer Representative. Upon the Buyer Representative’s reasonable request, no more frequently than [***] per [***] while any Out-License remains in effect, the Seller or Holdings, as applicable, shall use Commercially Reasonable Efforts to exercise any rights it may have under any Out-License relating to a Product to cause an inspection and/or audit by an independent public accounting firm to be made of the books of account of any counterparty thereto for the purpose of determining the correctness of Royalty Payments made under this Agreement. All of the out-of-pocket expenses of any inspection or audit requested by the Buyer Representative hereunder (including any audits of any Related Parties and the fees and expenses of such independent public accounting firm designated for such purpose) shall be borne solely by the Buyer, unless the independent public accounting firm determines that Royalty Payments previously paid during the period of the audit were underpaid by an amount greater than [***] of the Royalty Payments actually paid during such period, in which case such expenses shall be borne by the Seller. Any such accounting firm shall not disclose the confidential information of the Seller, Holdings or any such Licensee relating to a Product to any Buyer, except to the extent such disclosure is necessary to determine the correctness of Royalty Payments or otherwise would be included in a Report or a Royalty Report. All information obtained by the Buyer as a result of any such inspection or audit shall be Confidential Information subject to ARTICLE 8. If any audit discloses any underpayments by the Seller to the Buyer, then such underpayment shall be paid by the Seller to the Buyer within [***] of it being so disclosed. If any audit discloses any overpayments by the Seller to the Buyer, then (i) the Seller shall have the right to credit the amount of the overpayment against each subsequent quarterly Royalty Payment due to the Buyer until the overpayment has been fully applied, or (ii) if requested by the Seller in writing, the Buyer shall refund an amount equal to any such remaining overpayment within [***] of receipt of the Seller’s written request.
Section 6.5Intellectual Property Matters.
(a)The Seller and Holdings shall, at their sole expense, either directly or by causing any Licensee to do so, use Commercially Reasonable Efforts to take such actions (including taking legal action to specifically enforce the applicable terms of any In-License or Out-License), and prepare, execute, deliver and file any and all agreements, documents or instruments which are necessary to diligently prosecute and maintain, and avoid disclaimer or abandonment of, the Patent Rights in the Territory. The Seller and Holdings shall use Commercially Reasonable Efforts to ensure that all patent applications corresponding to the Patent Rights are diligently prosecuted with the intent to protect all Products in the Territory. In the exercise of its reasonable business discretion, the Seller and Holdings shall use Commercially Reasonable Efforts to diligently defend or assert the Patent Rights against infringement or interference by any other Persons, and against any claims of invalidity (including any reexamination,
27
inter partes review, opposition, or like proceeding) or unenforceability, including, without limitation, by bringing any legal action for infringement or defending any counterclaim of invalidity or action of a Third Party for declaratory judgment of non-infringement or non-interference, in each case in the Territory. Notwithstanding any other obligation of Seller or Holdings under this Section 6.5(a) Seller or Holdings shall (i) timely bring and prosecute infringement actions in response to certifications made under paragraph IV of 21 U.S.C. §355(j)(2)(A)(vii) or §355(b)(2)(A) with respect to any Orange Book Patent, (ii) defend the Orange Book Patents against any claim of invalidity or unenforceability, and (iii) not disclaim or abandon any Orange Book Patent.
(b)The Seller shall provide to the Buyer a copy of any written notice received by the Seller or Holdings from a Third Party alleging or claiming that the making, having made, using, importing, offering for sale or selling of a Product infringes or misappropriates any Patents or other intellectual property rights of such Third Party, together with copies of material correspondence sent or received by the Seller or Holdings related thereto, as soon as practicable and in any event not more than [***] following such delivery or receipt.
(c)The Seller shall promptly inform the Buyer of any infringement by a Third Party of any Patent Right in the Territory of which any of the individuals named in the definition of “Knowledge of the Seller and Holdings” (or the successors of such Person at Holdings) becomes aware. Without limiting the foregoing, the Seller shall provide to the Buyer a copy of any written notice of any suspected infringement of any Patent Rights in the Territory delivered or received by the Seller or Holdings, as well as copies of material correspondence related thereto, as soon as practicable and in any event not more than [***] following such delivery or receipt.
(d)Within [***] of initiating, or permitting a Licensee to initiate, an enforcement action regarding any suspected infringement by a Third Party of any Patent Right in the Territory, the Seller shall provide the Buyer with written notice of such enforcement action.
(e)If the Seller or Holdings recovers monetary damages from a Third Party in an action brought for such Third Party’s infringement of any Patent Rights in the Territory relating to a Product, where such damages, whether in the form of judgment or settlement, are awarded for such infringement of such Patent Rights, (i) such recovery will be allocated first to the reimbursement of any expenses incurred by the Seller or Holdings (or any party to an In-License or Permitted Out-License of such Patent Rights entitled to such reimbursement under any such In-License or Permitted Out-License ) in bringing such action (including all reasonable attorney’s fees), (ii) any remaining amounts will be reduced, if applicable, to comply with allocation of recovered damages with licensors of such Patent Rights required under any In-Licenses or Licensees of such Patent Rights under any Permitted Out-Licenses, if any, and (iii) any residual amount of such damages after application of (i) and (ii) will be treated as Net Sales to the extent set forth therein with respect to the applicable Product for the purposes of this Agreement.
Section 6.6In-Licenses.
(a)The Seller shall promptly (and in any event within [***]) provide the Buyer with (i) executed copies of any In-License entered into by the Seller or its Affiliates, and (ii) executed copies of each amendment, supplement, modification or written waiver of any provision of any In-License.
(b)Each of the Seller and Holdings shall use Commercially Reasonable Efforts to comply in all material respects with its obligations under any In-Licenses it enters into and shall not take any action or forego any action that would reasonably be expected to result in a material breach thereof. Promptly, and in any event within ten (10) Business Days, after receipt by the Seller or Holdings of any (written or oral) notice from a counterparty to any In-License or its Affiliates of an alleged material breach under any In-License, the Seller shall provide the Buyer a copy thereof (or if restricted by applicable confidentiality obligations under such In-License, notification thereof and a reasonably detailed summary of such material breach). Each of the Seller and Holdings shall use its Commercially Reasonable Efforts to cure any such material breaches by it under any In-License and shall give written notice to the Buyer upon curing any such breach. The Seller shall provide the Buyer with written notice following becoming aware of a counterparty’s material breach of its obligations under any In-License.
Neither the Seller nor Holdings shall terminate any In-License without providing the Buyer prior written notice. Promptly, and in any event within [***] following the Seller’s or Holdings’ notice to a counterparty to any In-
28
License of an alleged breach by such counterparty under any such In-License, the Seller shall provide the Buyer a copy thereof.
Section 6.7Permitted Licenses; Sales. [***]:
(a)Subject to compliance with this Section 6.7, the Seller or Holdings may enter into (i) an Out-License (A) solely with the Buyer Representative’s prior written consent (such consent not to be unreasonably withheld, conditioned or delayed) or (B) in the case of a Change of Control, (1) with a Qualified Buyer (if the Qualified Buyer assumes the obligations hereunder in accordance with Section 10.3) or (2) if the CoC Payment is made on the applicable CoC Date (each of (A) and (B)(1), a “Permitted Out-License”); (ii) distribution, marketing, contract manufacturing, logistics, clinical trial or sales force agreements, and similar vendor or fee-for-service arrangements, and non-exclusive research, non-exclusive development and other non-exclusive licenses, in each case without the Buyer Representative’s prior written consent (“Other License”), provided that (x) such Other License does not convey any Commercialization rights (other than by Distributors or Commercialization services performed by service providers for the benefit of the Seller, Holdings, or any of its Affiliates), and (y) such Permitted Out-License or Other License, as may be applicable, shall not assign, or otherwise convey title to or impose any Lien (other than the grant of the license, sublicense or other right) on any Product Collateral in favor of any Third Party; and (iii) the Intercompany License (any such Permitted Out-License, Other License or the Intercompany License, a “Permitted License”). Neither Seller nor Holdings shall enter into definitive agreement with respect to any Sale without the Buyer Representative’s prior written consent, which may be exercised in its sole and absolute discretion.
(b)The Seller shall promptly (and in any event within [***]) provide the Buyer with (i) executed copies of each Permitted License, as may be applicable, and (ii) executed copies of each amendment, supplement, modification or written waiver of any material provision of a Permitted License, as may be applicable.
(c)The Seller and Holdings shall include in all Permitted Out-Licenses as may be applicable, provisions (i) requiring the Licensee (as applicable) to provide to Seller all information that Seller is required to provide in the Royalty Reports within the same time frame as required under Section 6.2(b), including but not limited to Gross Sales and Net Sales for the applicable Calendar Quarter and four (4) consecutive Calendar Quarters ended as of the last day of such Calendar Quarter and calendar year to date, on a Product-by-Product basis (including a reasonably detailed break-down of all permitted deductions from Gross Sales used to determine Net Sales and any Net Sales described in Section 6.5(e)) (collectively, the “Licensee Reports”), (ii) allowing the Seller to provide such Licensee Reports to Buyer (and the Seller hereby covenants to provide such Licensee Reports (as applicable) promptly to the Buyer but in no event later than delivery of the respective Royalty Report under Section 6.2(b)), and (iii) for inspection and audit rights in favor of the Seller or Holdings, as applicable, substantially similar in nature and scope as provided to the Buyer pursuant to Section 6.4. Holdings agrees to comply with the foregoing requirements with respect to the Intercompany License and subject to such compliance, Buyer acknowledges and agrees that the Intercompany License in effect as of the Amendment Date complies with this Section 6.7(c).
(d)Each of the Seller and Holdings shall provide the Buyer prompt (and in any event within [***]) written notice of a material breach by a Licensee or of any of its obligations under any Permitted Out-License, of which any of the individuals named in the definition of “Knowledge of the Seller and Holdings” (or the successors of such Person at Holdings) becomes aware.
(e)The Seller shall provide the Buyer with written notice promptly (and in any event within [***]) following the termination of any Permitted Out-License.
(f)The Buyer acknowledges and agrees that in the event that Seller or Holdings enters into (or plans to enter into) any Permitted License or other license (e.g. outside of the Territory), the Buyer shall, at the reasonable request of the Seller, enter into non-disturbance and similar agreements in a form reasonably acceptable to the Buyer in connection with such Permitted License to the extent reasonably requested by the counterparty (or prospective counter-party) to such Permitted License.
Section 6.8Restricted Indebtedness. Prior to the occurrence of a Release Event, Holdings shall not, and shall not permit any of its Subsidiaries to, create, incur, assume or suffer to exist any Restricted Indebtedness without the Buyer Representative’s prior consent. As a condition to the incurrence at any time of any secured
29
Indebtedness for borrowed money with one or more Senior Debt Providers, the Buyer shall enter, and the Seller and Holdings shall enter and cause such Senior Debt Provider to enter, into an Acceptable Intercreditor Agreement to be mutually agreed by the parties thereto. The Buyer and the Seller and Holdings agree to negotiate such Acceptable Intercreditor Agreement in good faith.
Section 6.9Diligence.
(a)The Seller and Holdings shall use Commercially Reasonable Efforts to (i) complete clinical development of Etripamil in the Territory, (ii) obtain and maintain Marketing Approvals for all Products for the Approved Indication in the Territory, (iii) conduct and complete any post-marketing Clinical Trials required by the FDA as a condition for maintaining U.S. Marketing Approval, and (iv) Commercialize all Products in the Territory. In furtherance of the foregoing, the Seller and Holdings shall use Commercially Reasonable Efforts to prepare, execute, deliver and file any and all agreements, documents or instruments that are necessary or desirable to secure and maintain such Marketing Approval required to Commercialize all Products in the Territory and the Seller and Holdings shall use Commercially Reasonable Efforts to not withdraw or abandon, or fail to take any action necessary to prevent the withdrawal or abandonment of, any such Marketing Approval in the Territory.
(b)On a Product-by-Product basis, if a Loss of Market Exclusivity has occurred in the Territory for a Product, the Seller’s and Holdings’ obligations under Section 6.9(a) shall no longer apply in the Territory for such Product.
Section 6.10Efforts to Consummate Transactions. Subject to the terms and conditions of this Agreement, the Seller will use its Commercially Reasonable Efforts and the Buyer will use its commercially reasonable efforts prior to the Closing to take, or cause to be taken, all actions and to do, or cause to be done, all things reasonably necessary under applicable law to consummate the transactions contemplated by this Agreement and the Transaction Documents. Each of the Buyer and the Seller agrees to execute and deliver such other documents, certificates, agreements and other writings and to take such other actions as may be reasonably necessary in order to consummate or implement expeditiously the transactions contemplated by this Agreement and to vest in Buyer good and valid right, title, and interest in and to the Revenue Participation Right, which is, as of the Closing, free and clear of all Liens, except for (a) Liens created in favor of Buyer on or after the Closing pursuant to the Note Purchase Agreement, and (b) liens for taxes or other government charges arising by operation of law in the ordinary course of business for sums which are not yet due and payable.
Section 6.11Continuing Efforts; Further Assurances.
(a)After the Amendment Date, the Seller and the Buyer agree to execute and deliver such other documents, certificates, agreements and other writings and to take such other actions as may be reasonably necessary in order to give effect to the transactions contemplated by this Agreement. After the Amendment Date, the Seller shall use its Commercially Reasonable Efforts to obtain and maintain any required consents, acknowledgements, certificates or waivers so that the transactions contemplated by this Agreement or any other Transaction Document may be consummated and shall not result in any default or breach or termination of any material contract in respect of the Revenue Participation Right or the Product Collateral.
(b)The Buyer, the Seller and Holdings shall cooperate and provide assistance as reasonably requested by the other party, at the expense of such other party (other than expenses that are Losses subject to indemnification in accordance with Article 7), in connection with any Third Party litigation, arbitration or other Third Party proceeding with respect to the Revenue Participation Right, or the Product Collateral (whether threatened, existing, initiated, or contemplated prior to, on or after the date hereof) to which any party hereto or any of its officers, directors, shareholders, agents or employees is or may become a party or is or may become otherwise directly or indirectly affected or as to which any such Persons have a direct or indirect interests, in each case relating to this Agreement, any other Transaction Document, the Revenue Participation Right or any other Product Collateral, or the transactions described herein or therein.
Section 6.12Non-Impairment; Back-Up Security Interest. Holdings shall not, and shall not permit any Subsidiary to, as may be applicable, (a) enter into any contracts or arrangement or otherwise knowingly take any action or knowingly fail to act in a manner that would, individually or in the aggregate, reasonably be expected to
30
materially and adversely affect the Buyer’s rights to the Revenue Participation Right or the Back-Up Security Interest, or (b) take any action or engage in any transaction (or series of actions or transactions), whether by reorganization, transfer of assets, merger, dissolution, amendment of organizational documents or otherwise, the primary purpose of which is to evade, avoid or seek to avoid the performance or observance of the covenants, agreements or obligations of the Seller or Holdings under the Transaction Documents; provided, however, that such clauses (a) and (b) shall not restrict the incurrence of Indebtedness or Liens or the entry into Out-Licenses, in each case to the extent not otherwise prohibited under this Agreement or otherwise be deemed to expand or modify any affirmative or negative covenants in this ARTICLE 6. At the Closing Date, each of the Seller and Holdings shall grant in favor of Buyer, and take such additional actions as reasonably requested by Buyer to ensure that thereafter, prior to the occurrence of a Release Event, Buyer has a valid, continuing, first priority security interest in and to all right, title and interest in, to and under the Revenue Participation Right and the Royalty Payments and a security interest in and to all right, title and interest in, to and under the Product Collateral in accordance with the terms set forth in Section 2.1
Section 6.13Certain Tax Matters.
(a)The Seller and the Buyer agree that for U.S. federal Tax purposes, the Seller and the Buyer shall treat the transactions contemplated by this Agreement as a sale of the Revenue Participation Right. The Seller shall not, by reason of its duties and functions hereunder, be deemed to be acting as a partner of or to be engaged in a joint venture, association or syndication with, the Buyer for tax purposes. The parties hereto agree not to take any position that is inconsistent with the provisions of this Section 6.13(a) on any U.S. tax return or in any audit or other tax-related administrative or judicial proceeding in the U.S. unless the other party hereto has consented in writing (such consent not to be unreasonably withheld, conditioned or delayed) to such actions. If there is an inquiry by any Governmental Entity of the Buyer or the Seller related to the treatment described in this Section 6.13(a) the parties hereto shall cooperate with each other in responding to such inquiry in a reasonable manner which is consistent with this Section 6.13(a)
(b)The parties hereto acknowledge and agree that, as of the date hereof, all payments by the Seller to the Buyer and by the Buyer to the Seller shall be made without deduction or withholding for any Taxes. If any change in applicable law requires the deduction or withholding of any Tax from any payment by the Seller, then the Seller shall be entitled to withhold and deduct (or cause to be withheld and deducted) from any amount payable under this Agreement to the Buyer any Tax that it is required to withhold and deduct under applicable law. The Seller shall timely pay the full amount deducted or withheld to the relevant governmental authority in accordance with applicable law. For the avoidance of doubt and notwithstanding anything to the contrary in this Agreement, any amount payable by the Seller to the Buyer shall be increased as necessary so that after deduction or withholding of any Tax has been made (including such deductions and withholdings applicable to additional sums payable under this Section 6.13(b)) (whether as a result of a change in applicable law or otherwise), the Buyer receives an amount equal to the sum it would have received had no such deduction or withholding been made; provided, however, that if, as a result of a Withholding Action by the Buyer, the amount of any tax withheld or deducted exceeds the amount of such withholding or deduction that would have been required in the absence of such Withholding Action, the Seller shall be required to pay an additional amount only to the extent that the Seller would be required to pay any additional amount to the Buyer pursuant to this Section 6.13(b) if the Buyer had not committed such Withholding Action (except to the extent such excess additional amounts results from a change in applicable law that occurs after the date of such Withholding Action).
Section 6.14Use of Proceeds. The Seller shall use proceeds received from Buyer pursuant to this Agreement in support of Commercialization of the Products.
Section 6.15Intercompany License. The Seller and Holdings shall not (i) enter into any related party agreement that materially adversely affects Buyer’s rights under this Agreement (other than the Intercompany License) or (ii) amend, modify or supplement the Intercompany License in a way that materially adversely affects the Buyer’s interests under this Agreement (and any other ancillary agreement or transaction document contemplated by this Agreement), without prior written consent of the Buyer Representative. The Seller and Holdings shall provide the Buyer Representative with a copy of any amendments, modifications or supplements to the Intercompany License, whether or not the Buyer Representative’s consent is required pursuant to the foregoing. Without limiting the foregoing, neither Holdings nor Seller may terminate the Intercompany License without the prior written consent of
31
the Buyer Representative. All obligations of Holdings and Seller under the Intercompany License, including indemnification obligations, are subordinated to Buyer’s rights under this Agreement pursuant to Section 10.15.
Section 6.16Ownership of Seller. Holdings does and shall continue to hold directly or indirectly one hundred percent (100%) of the outstanding voting securities of Seller.
Article 7.
INDEMNIFICATION
Section 7.1General Indemnity. From and after the Closing:
(a)the Seller hereby agrees to indemnify, defend and hold harmless the Buyer and its Affiliates and its and their directors, managers, trustees, officers, agents and employees (the “Buyer Indemnified Parties”) from, against and in respect of all Losses suffered or incurred by the Buyer Indemnified Parties to the extent arising out of or resulting from (i) any breach of any of the representations or warranties of the Seller in this Agreement, (ii) any breach of any of the covenants or agreements of the Seller in this Agreement and (iii) any Indemnified Taxes; and
(b)the Buyer hereby agrees to indemnify, defend and hold harmless the Seller and its Affiliates and its and their directors, officers, agents and employees (the “Seller Indemnified Parties”) from, against and in respect of all Losses suffered or incurred by the Seller Indemnified Parties to the extent arising out of or resulting from (i) any breach of any of the representations or warranties of the Buyer in this Agreement and (ii) any breach of any of the covenants or agreements of the Buyer in this Agreement.
Section 7.2Notice of Claims. If either a Buyer Indemnified Party, on the one hand, or a Seller Indemnified Party, on the other hand (such Buyer Indemnified Party on the one hand and such Seller Indemnified Party on the other hand being hereinafter referred to as an “Indemnified Party”), has suffered or incurred any Losses for which indemnification may be sought under this ARTICLE 7, the Indemnified Party shall so notify the other party from whom indemnification is sought under this ARTICLE 7 (the “Indemnifying Party”) promptly in writing describing such Loss, the amount or estimated amount thereof, if known or reasonably capable of estimation, and the method of computation of such Loss, all with reasonable particularity and containing a reference to the provisions of this Agreement in respect of which such Loss shall have occurred. If any claim, action, suit or proceeding is asserted or instituted by or against a Third Party with respect to which an Indemnified Party intends to claim any Loss under this ARTICLE 7 such Indemnified Party shall promptly notify the Indemnifying Party of such claim, action, suit or proceeding and tender to the Indemnifying Party the defense of such claim, action, suit or proceeding. A failure by an Indemnified Party to give notice and to tender the defense of such claim, action, suit or proceeding in a timely manner pursuant to this Section 7.2 shall not limit the obligation of the Indemnifying Party under this ARTICLE 7 except to the extent such Indemnifying Party is actually prejudiced thereby.
Section 7.3Claim Procedures. In case any such action is brought against an Indemnified Party and it notifies the Indemnifying Party of the commencement thereof, the Indemnifying Party will be entitled to participate therein and, to the extent that it may wish, jointly with any other Indemnifying Party similarly notified, to assume the defense thereof, with counsel reasonably satisfactory to such Indemnified Party, and after notice from the Indemnifying Party to such Indemnified Party of its election so to assume the defense thereof, the Indemnifying Party will not be liable to such Indemnified Party under this Section 7.3 for any legal or other expenses subsequently incurred by such Indemnified Party in connection with the defense thereof other than reasonable costs of investigation. In any such proceeding, an Indemnified Party shall have the right to retain its own counsel reasonably satisfactory to the Indemnifying Party, but the reasonable fees and expenses of such counsel shall be at the expense of such Indemnified Party unless (a) the Indemnifying Party has assumed the defense of such proceeding and has failed within a reasonable time to retain counsel reasonably satisfactory to such Indemnified Party or (b) the named parties to any such proceeding (including any impleaded parties) include both the Indemnifying Party and the Indemnified Party and representation of both parties by the same counsel would be inappropriate due to actual or potential conflicts of interests between them based on the advice of such counsel. It is agreed that the Indemnifying Party shall not, in connection with any proceeding or related proceedings in the same jurisdiction, be liable for the reasonable fees and expenses of more than one separate law firm (in addition to local counsel where necessary) for all such Indemnified Parties. The Indemnifying Party shall not be liable for any settlement of any proceeding effected without its written
32
consent. No Indemnifying Party shall, without the prior written consent of the Indemnified Party, which shall not be unreasonably withheld, conditioned, or delayed, effect any settlement of any pending or threatened proceeding in respect of which any Indemnified Party is or could have been a party and indemnity could have been sought hereunder by such Indemnified Party, unless such settlement includes an unconditional release of such Indemnified Party from all liability on claims that are the subject matter of such proceeding.
Section 7.4Limitations on Liability.
(a)Except for claims arising from a breach of confidentiality obligations under ARTICLE 8 or in cases of fraud, gross negligence, or willful misconduct, no party hereto shall be liable for any consequential, punitive, special or incidental damages under this ARTICLE 7 (and no claim for indemnification hereunder shall be asserted) as a result of any breach or violation of any covenant or agreement of such party (including under this ARTICLE 7) in or pursuant to this Agreement. In connection with the foregoing, the parties hereto acknowledge and agree that (a) the Buyer’s damages, if any, for any such action or claim will include Losses for Royalty Payments that the Buyer was entitled to receive in respect of its ownership of the Royalty Payments but did not receive timely or at all due to such indemnifiable event and (b) the Buyer shall be entitled to make claims for all such missing or delayed Royalty Payments as Losses hereunder, and such missing or Royalty Payments shall not be deemed consequential, punitive, special, indirect or incidental damages.
Section 7.5Exclusive Remedy. Except as set forth in Section 10.10, from and after Closing, the rights of the parties hereto pursuant to (and subject to the conditions of) this ARTICLE 7 shall be the sole and exclusive remedy of the parties hereto and their respective Affiliates with respect to any Losses (whether based in contract, tort or otherwise) resulting from or relating to any breach of the representations, warranties covenants and agreements made under this Agreement or any certificate, document or instrument delivered hereunder, and each party hereto hereby waives, to the fullest extent permitted under applicable law, and agrees not to assert after Closing, any other claim or action in respect of any such breach. Notwithstanding the foregoing, claims for fraud, gross negligence, or willful misconduct shall not be waived or limited in any way by this ARTICLE 7
Section 7.6Tax Treatment of Indemnification Payments. For all purposes hereunder, any indemnification payments made pursuant to this ARTICLE 7 will be treated as an adjustment to the Purchase Price for Tax purposes, including U.S. federal income tax to the fullest extent permitted by applicable law.
Section 7.7Survival. All representations and warranties made herein and in any other Transaction Document, any certificates or in any other writing delivered pursuant hereto or thereto shall survive the execution and delivery of this Agreement and shall continue to survive until the expiration or termination of this Agreement in accordance with ARTICLE 9
Article 8.
CONFIDENTIALITY
Section 8.1Confidentiality. Except as provided in this ARTICLE 8 Section 10.4 or otherwise agreed in writing by the parties, the parties hereto agree that, during the term of this Agreement and for [***] thereafter, each party (the “Receiving Party”) shall keep confidential and shall not publish or otherwise disclose and shall not use for any purpose other than as provided for in this Agreement (which includes the exercise of any rights or the performance of any obligations hereunder) any information furnished to it by or on behalf of the other party (the “Disclosing Party”) pursuant to this Agreement (such information, “Confidential Information” of the Disclosing Party), except for that portion of such information that:
(a)was already known to the Receiving Party, other than under an obligation of confidentiality, at the time of disclosure by the Disclosing Party;
(b)was generally available to the public or otherwise part of the public domain at the time of its disclosure to the Receiving Party;
33
(c)became generally available to the public or otherwise part of the public domain after its disclosure and other than through any act or omission of the Receiving Party in breach of this Agreement or any other agreement;
(d)is independently developed by the Receiving Party or any of its Affiliates, as evidenced by written records, without the use of or reference of the Confidential Information; or
(e)is subsequently disclosed to the Receiving Party on a non-confidential basis by a Third Party without obligations of confidentiality with respect thereto.
Section 8.2Authorized Disclosure.
(a)Either party hereto may disclose Confidential Information to the extent such disclosure is reasonably necessary in the following situations:
(i)prosecuting or defending litigation;
(ii)complying with applicable laws and regulations, including regulations promulgated by securities exchanges;
(iii)complying with a valid order of a court of competent jurisdiction or other Governmental Entity;
(iv)for regulatory, Tax or customs purposes;
(v)for audit purposes, provided that each recipient of Confidential Information must be bound by customary and reasonable obligations of confidentiality and non-use prior to any such disclosure;
(vi)disclosure to its Affiliates and Representatives on a need-to-know basis, provided that each such recipient of Confidential Information must be bound by contractual or professional obligations of confidentiality and non-use at least as stringent as those imposed upon the parties hereunder prior to any such disclosure;
(vii)upon the prior written consent of the Disclosing Party;
(viii)disclosure to its potential investors, and other sources of funding, including debt financing, or potential partners, collaborators or acquirers, and their respective accountants, financial advisors and other professional representatives, provided that such disclosure shall be made only to the extent customarily required to consummate such investment, financing transaction partnership, collaboration or acquisition and that each recipient of Confidential Information must be bound by customary obligations of confidentiality and non-use prior to any such disclosure;
(ix)as is necessary in connection with a permitted assignment pursuant to Section 10.3.
(b)Notwithstanding the foregoing, in the event the Receiving Party is required to make a disclosure of the Disclosing Party’s Confidential Information pursuant to Section 8.2(a)(i) (ii) (iii) or (iv) it will, except where impracticable, give reasonable advance notice to the Disclosing Party of such disclosure and use reasonable efforts to secure confidential treatment of such information. In any event, the Buyer shall not file any patent application based upon or using the Confidential Information of the Seller provided hereunder.
(c)Notwithstanding any provision in this Agreement to the contrary, materials and documentation relating to the Seller’s Intellectual Property Rights may be only disclosed to or accessed by the Buyer, its attorneys and auditors, and prospective assignees without further disclosure to any other Representative of the Buyer.
34
Article 9.
TERMINATION
Section 9.1Mutual Termination. This Agreement may be terminated by mutual written agreement of the Buyer and the Seller.
Section 9.2Termination upon Failure to Achieve U.S. Marketing Approval.
This Agreement shall automatically terminate upon Seller’s or Holdings’ failure to achieve U.S. Marketing Approval by the Marketing Approval Deadline unless such date is extended by the mutual written agreement of the Seller and the Buyer.
Section 9.3Buyer Termination for Unexpected Contraindications. If the Seller or Holdings receives U.S. Marketing Approval with one or more Unexpected Contraindications, then the Buyer Representative (if more than one Buyer, on behalf of all Buyers) may terminate this Agreement by delivering written notice to the Seller of such election.
Section 9.4Automatic Termination. Unless earlier terminated as provided in this ARTICLE 9 this Agreement shall continue in full force and effect until sixty (60) days after such time as the Seller is no longer obligated to make any Royalty Payments under this Agreement, at which point this Agreement shall automatically terminate, except with respect to any rights that shall have accrued prior to such termination.
Section 9.5Survival. Notwithstanding anything to the contrary in this ARTICLE 9 the following provisions shall survive termination of this Agreement: Section 6.3 (Disclosures), ARTICLE 7 (Indemnification), ARTICLE 8 (Confidentiality), this Section 9.5 (Survival) and ARTICLE 10 (Miscellaneous) Termination of the Agreement shall not relieve any party of liability in respect of breaches under this Agreement by any party on or prior to termination.
Article 10.
MISCELLANEOUS
Section 10.1Headings. The table of contents and the descriptive headings of the several Articles and Sections of this Agreement and the Exhibits and Schedules are for convenience only, do not constitute a part of this Agreement and shall not control or affect, in any way, the meaning or interpretation of this Agreement.
Section 10.2Notices. All notices and other communications under this Agreement shall be in writing and shall be by email with PDF attachment, facsimile, courier service or personal delivery to the following addresses, or to such other addresses as shall be designated from time to time by a party hereto in accordance with this Section 10.2:
If to the Seller:
Milestone Pharmaceuticals Inc.
1111 Dr.-Frederik-Philips Blvd., Ste. 420
Montreal, Quebec
H4M 2X6 Canada
Attention: Chief Financial Officer
Email: [***]
with a copy to:
Cooley LLP
3 Embarcadero Center, 20th Floor
35
San Francisco, CA 94111-4004
Attention: Gian-Michele a Marca, Jason Savich
Email: gmamarca@cooley.com, jsavich@cooley.com
If to the Buyer:
RTW ROYALTY I DAC
2nd Floor
Palmserston House
Denzille Lane
Dublin 2
D02 WD37
Ireland
and
RTW Investments, LP, as Buyer Representative for the Buyer
40 10th Avenue, Floor 7
New York, NY 10014
Attn: [***] and [***]
Email: [***]; [***] and legalops@rtwfunds.com
with a copy to:
Gibson, Dunn & Crutcher LLP
555 Mission Street
San Francisco, CA 94105
Attention: Ryan Murr
E-mail: rmurr@gibsondunn.com
All notices and communications under this Agreement shall be deemed to have been duly given (a) when delivered by hand, if personally delivered, (b) when sent, if by email with PDF attachment, with an acknowledgement of receipt being produced by the recipient’s email account, or (c) one (1) Business Day following sending within the United States by overnight delivery via commercial one-day overnight courier service.
Section 10.3Assignment. The Seller may not assign in whole or in part this Agreement or any of its rights or obligations hereunder without the Buyer Representative’s prior written consent, except in connection with a Change of Control or a Permitted Out-License, and only if upon closing any such transaction, the Seller causes such acquirer or Licensee, as applicable, to deliver a writing to the Buyer in which it assumes all of the obligations of the Seller to the Buyer under this Agreement, and such acquirer or Licensee as applicable, shall be deemed an assignee of the Seller under this Agreement; provided that, for the avoidance of doubt, nothing in this Section 10.3 shall restrict a Change of Control or the Seller licensing any Product Rights pursuant to a Permitted License. Following the Closing, Buyer may assign, without the consent of Seller, its obligations and rights under this Agreement to any Person, including to any Third Party or to one or more of its Affiliates, [***]. This Agreement shall be binding upon, inure to the benefit of and be enforceable by, the parties hereto and their respective permitted successors and assigns. Any purported assignment in violation of this Section 10.3 shall be null and void.
Section 10.4Amendment and Waiver.
(a)This Agreement may be amended, modified or supplemented only in a writing signed by each of the parties hereto. Any provision of this Agreement may be waived only in a writing signed by the party hereto granting such waiver.
(b)No failure or delay on the part of any party hereto in exercising any right, power or remedy hereunder shall operate as a waiver thereof, nor shall any single or partial exercise of any such right, power or remedy
36
preclude any other or further exercise thereof or the exercise of any other right, power or remedy. No course of dealing between the parties hereto shall be effective to amend, modify, supplement or waive any provision of this Agreement.
Section 10.5Entire Agreement. This Agreement, the Exhibits annexed hereto and the Disclosure Schedule constitute the entire understanding between the parties hereto with respect to the subject matter hereof and supersede all other understandings and negotiations with respect thereto.
Section 10.6No Third-Party Beneficiaries.
This Agreement is for the sole benefit of the Seller and the Buyer and their permitted successors and assigns and nothing herein expressed or implied shall give or be construed to give to any Person, other than the parties hereto and such successors and assigns, any legal or equitable rights hereunder, except that the Indemnified Parties shall be third-party beneficiaries of the benefits provided for in Section 7.1.
Section 10.7Governing Law. This Agreement shall be governed by, and construed in accordance with, the laws of the State of New York without giving effect to any choice or conflict of law provision or rule that would cause the application of the laws of any other jurisdiction.
Section 10.8Jurisdiction; Venue.
(a)EACH OF THE PARTIES HERETO HEREBY IRREVOCABLY AND UNCONDITIONALLY SUBMITS, FOR ITSELF AND ITS RESPECTIVE PROPERTY AND ASSETS, TO THE EXCLUSIVE JURISDICTION OF ANY NEW YORK STATE COURT OR FEDERAL COURT OF THE UNITED STATES OF AMERICA SITTING IN NEW YORK COUNTY, NEW YORK, AND ANY APPELLATE COURT THEREOF, IN ANY ACTION OR PROCEEDING ARISING OUT OF OR RELATING TO THIS AGREEMENT, OR FOR RECOGNITION OR ENFORCEMENT OF ANY JUDGMENT IN RESPECT THEREOF, AND THE BUYER AND THE SELLER HEREBY IRREVOCABLY AND UNCONDITIONALLY AGREE THAT ALL CLAIMS IN RESPECT OF ANY SUCH ACTION OR PROCEEDING MAY BE HEARD AND DETERMINED IN ANY SUCH NEW YORK STATE COURT OR, TO THE FULLEST EXTENT PERMITTED BY APPLICABLE LAW, IN SUCH FEDERAL COURT. THE BUYER AND THE SELLER HEREBY AGREE THAT A FINAL JUDGMENT IN ANY SUCH ACTION OR PROCEEDING SHALL BE CONCLUSIVE AND MAY BE ENFORCED IN OTHER JURISDICTIONS BY SUIT ON THE JUDGMENT OR IN ANY OTHER MANNER PROVIDED BY APPLICABLE LAW. EACH OF THE BUYER AND THE SELLER HEREBY SUBMITS TO THE EXCLUSIVE PERSONAL JURISDICTION AND VENUE OF SUCH NEW YORK STATE AND FEDERAL COURTS. THE BUYER AND THE SELLER AGREE, TO THE FULLEST EXTENT PERMITTED BY APPLICABLE LAW, THAT PROCESS MAY BE SERVED ON THE BUYER OR THE SELLER IN THE SAME MANNER THAT NOTICES MAY BE GIVEN PURSUANT TO SECTION 10.2.
(b)EACH OF THE PARTIES HERETO HEREBY IRREVOCABLY AND UNCONDITIONALLY WAIVES, TO THE FULLEST EXTENT IT MAY LEGALLY AND EFFECTIVELY DO SO, ANY OBJECTION THAT IT MAY NOW OR HEREAFTER HAVE TO THE LAYING OF VENUE OF ANY ACTION OR PROCEEDING ARISING OUT OF OR RELATING TO THIS AGREEMENT IN ANY NEW YORK STATE OR FEDERAL COURT. EACH OF THE BUYER AND THE SELLER HEREBY IRREVOCABLY WAIVES, TO THE FULLEST EXTENT PERMITTED BY APPLICABLE LAW, THE DEFENSE OF AN INCONVENIENT FORUM TO THE MAINTENANCE OF SUCH ACTION OR PROCEEDING IN ANY SUCH COURT.
(c)EACH PARTY HEREBY JOINTLY AND SEVERALLY WAIVES ANY AND ALL RIGHT TO TRIAL BY JURY IN ANY ACTION OR PROCEEDING RELATING TO THIS AGREEMENT OR ANY OTHER DOCUMENT DELIVERED HEREUNDER OR IN CONNECTION HEREWITH, OR ANY TRANSACTION ARISING FROM OR CONNECTED TO ANY OF THE FOREGOING. EACH OF THE PARTIES REPRESENTS THAT THIS WAIVER IS KNOWINGLY, WILLINGLY, AND VOLUNTARILY GIVEN.
Section 10.9Severability. If any term or provision of this Agreement shall for any reason be held to be invalid, illegal or unenforceable in any situation in any jurisdiction, then, to the extent that the economic and legal
37
substance of the transactions contemplated hereby is not affected in a manner that is materially adverse to either party hereto, all other terms and provisions of this Agreement shall nevertheless remain in full force and effect and the enforceability and validity of the offending term or provision shall not be affected in any other situation or jurisdiction.
Section 10.10Specific Performance. Each party hereto acknowledges and agrees that the other party would be damaged irreparably in the event any of the provisions of this Agreement are not performed in accordance with their specific terms or otherwise are breached or violated. Accordingly, each of the parties agrees that, without posting bond or other undertaking, the other party will be entitled to seek an injunction or injunctions to prevent breaches or violations of the provisions of this Agreement and to seek to enforce specifically this Agreement and the terms and provisions hereof in any action, suit or other proceeding instituted in any court of the United States or any state thereof having jurisdiction over the parties and the matter in addition to any other remedy to which it may be entitled, at law or in equity. Each of the parties further agrees that, in the event of any action for specific performance in respect of such breach of violation, it will not assert the defense that a remedy at law would be adequate.
Section 10.11Counterparts. This Agreement may be executed in any number of counterparts and by the parties hereto in separate counterparts, each of which when so executed shall be deemed to be an original and all of which taken together shall constitute one and the same agreement. Copies of executed counterparts transmitted by telecopy, facsimile or other similar means of electronic transmission, including “PDF,” shall be considered original executed counterparts, provided receipt of such counterparts is confirmed.
Section 10.12Relationship of the Parties. The relationship between the Buyer and the Seller is solely that of purchaser and seller, and neither the Buyer nor the Seller has any fiduciary or other special relationship with the other party or any of its Affiliates. This Agreement is not a partnership or similar agreement, and nothing contained herein shall be deemed to constitute the Buyer and the Seller as a partnership, an association, a joint venture or any other kind of entity or legal form for any purposes, including any Tax purposes. The Buyer and the Seller agree that they shall not take any inconsistent position with respect to such treatment in a filing with any Governmental Entity.
Section 10.13Expenses. On the date hereof, the Seller shall reimburse the Buyer for the Transaction Expenses incurred prior to or on the date hereof. Upon the earliest of (a) the Closing, (b) termination of this Agreement, and (c) termination of any of the transactions contemplated hereby, the Seller shall reimburse the Buyer for, if applicable, any and all other Transaction Expenses incurred prior to or on the Closing Date, which shall not, in the aggregate, exceed amounts permitted under the definition of “Transaction Expenses”.
Section 10.14Limited Recourse and Non-Petition.
(a)Notwithstanding any of the provisions of this Agreement, each of the parties hereto hereby agrees that if the net proceeds from a liquidation of the unsecured assets of the Buyer are less than the aggregate amount payable by the Buyer to the Seller in respect of its obligations under this Agreement (such negative amount being referred to herein as a shortfall), the amount payable by the Buyer to that party in respect of the Buyer’s obligations under this Agreement will be reduced to such amount of the net proceeds which are available to satisfy such payment obligation. In such circumstances the other assets of the Buyer will not be available for payment of such shortfall, and the Seller’s right to receive any further amounts in respect of such obligations shall be extinguished and that party may not take any further action to recover such amounts. [***].
(b)No party shall be entitled at any time to institute against the Buyer, or join in any institution against the Buyer of, any bankruptcy, examinership, reorganization, arrangement, insolvency or liquidation proceedings or other proceedings under any applicable bankruptcy or similar law in connection with any obligation of the Buyer under this Agreement, save for lodging a claim in the liquidation of the Buyer which is initiated by another non-affiliated party or taking proceedings to obtain a declaration or judgment as to the obligations of the Buyer in relation thereto.
(c)Each of the Buyer and the Seller hereby agrees that no recourse under any obligation, covenant, or agreement of either party contained in this Agreement may be sought against any shareholder, officer, agent, employee or director of the Buyer, by the enforcement of any assessment or by any proceeding, by virtue of any statute or otherwise, it being expressly agreed and understood that this Agreement contains corporate obligations of the Buyer. Each of the parties hereto agrees that no personal liability shall attach to or be incurred by the
38
shareholders, officers, agents, employees or directors of the Buyer, or any of them, under or by reason of any of the obligations, covenants or agreements of the Buyer contained in this Agreement, or implied therefrom, and any and all personal liability of every such shareholder, officer, agent, employee or director for breaches by the Buyer of any such obligations, covenants or agreements, either at law or by statute or constitution is hereby deemed expressly waived by the parties hereto.
(d)The provisions of this Section 10.14 shall survive the termination of this Agreement.
Section 10.15Subordination of Obligations to Holdings.
(a)Holdings hereby agrees that all liabilities or other obligations, including any costs, charges, expenses, reimbursement obligations, indemnification obligations and any other amounts, in each case, whether now existing or hereafter arising, of the Seller to Holdings in respect of the Intercompany License, whether now existing or hereafter arising, and whether due or to become due, absolute or contingent, liquidated or unliquidated, determined or undetermined (the “Subordinated Obligations”) shall be subject, subordinate, and junior, in right of payment and exercise of remedies, to the payment in full in cash in immediately available funds of all liabilities or other obligations (except for and any contingent indemnification Obligations for which a claim or demand for payment has been not made at such time), including any costs, charges, expenses, reimbursement obligations, indemnification obligations and any other amounts, in each case, of the Seller to the Buyer hereunder, whether now existing or hereafter arising, and whether due or to become due, absolute or contingent, liquidated or unliquidated, determined or undetermined and whether or not allowed in any Bankruptcy Event (the “Senior Obligations” and such payment in full in cash of such Senior Obligations, a “Senior Discharge Event”).
(b)In the event of any payment or distribution of assets of the Seller of any kind or character, whether in cash, property, or securities, upon the dissolution, winding up, or total or partial liquidation or reorganization, readjustment, arrangement, or similar proceeding relating to the Seller or its property, or in bankruptcy, insolvency, receivership, arrangement, or similar proceedings or upon an assignment for the benefit of creditors, or upon any other marshaling or composition of the assets and liabilities of Seller, or upon the occurrence of a Bankruptcy Event or otherwise upon any bankruptcy, insolvency, receivership, examinership or other similar proceeding relating to Seller or its property (such events, collectively, the “Insolvency Events”): (i) the Senior Discharge Event must have occurred before any payments on account of Subordinated Obligations is made; and (ii) any payment on account of Subordinated Obligations to which Holdings would be entitled except for the provisions hereof, shall be paid or delivered by the trustee in bankruptcy, receiver, assignee for the benefit of creditors, or other liquidating agent making such payment or distribution directly to Buyer until the Senior Discharge Event has occurred.
(c)So long as no breach of this Agreement by Seller or Holdings has occurred, Seller may make, and Holdings shall be entitled to accept and receive, payments and repayments on account of the Subordinated Obligations. No payment or other amount due or payable by the Seller under the Intercompany License shall be made or permitted or required to be made by Seller or shall be accepted by Holdings if, at the time of the making of such payment, the Seller or Holdings is in breach of this Agreement.
(d)During the term of this Agreement, Holdings shall not, without the prior written consent of Buyer Representative, take any liens or security interests in any of the assets of Seller to secure amounts owing pursuant to the Intercompany License and any liens and security interests of Holdings which may exist in breach of the foregoing shall be and hereby are subordinated for all purposes and in all respects to the liens and security interests of Buyer granted under this Agreement.
(e)This Section 10.15 shall continue to be effective or shall be reinstated (and the amount of Senior Obligations shall be reinstated), as the case may be, if, for any reason, any payment of the Senior Obligations shall be rescinded or must otherwise be restored by Buyer, whether as a result of any bankruptcy, insolvency, receivership, examinership or other similar proceeding or Bankruptcy Event, or otherwise, all as though such payment had not been made. The provisions of this Section 10.15 are intended to be and shall be enforceable as a subordination agreement within the meaning of Section 510 of the Bankruptcy Code.
39
Article 11.
GUARANTEE
Section 11.1Guarantee. Holdings unconditionally guarantees, as a primary obligor and not merely as a surety, the due and punctual payment and performance of the Seller’s obligations hereunder (the “Obligations”). Holdings further agrees that the Obligations may be extended or renewed, in whole or in part, without notice to or further assent from it, and that it will remain bound upon its guarantee notwithstanding any extension or renewal of any Obligation. Holdings waives presentment to, demand of payment from and protest to the Seller, and also waives notice of acceptance of its guarantee and notice of protest for nonpayment.
Section 11.2Guarantee of Payment. Holdings further agrees that its guarantee hereunder constitutes a guarantee of payment when due and not of collection, and waives any right to require that any resort be had by the Buyer to any security held for the payment of the Obligations or credit on the books of the Buyer in favor of the Seller or any other person.
Section 11.3No Limitations, Etc.
(a)Except for (i) termination of this Agreement pursuant to Article 9 or Section 11.8 or (ii) the occurrence of a Release Event, the obligations of Holdings hereunder shall not be subject to any reduction, limitation, impairment or termination for any reason, including any claim of waiver, release, surrender, alteration or compromise, and shall not be subject to any defense or setoff, counterclaim, recoupment or termination whatsoever by reason of the invalidity, illegality or unenforceability of the Obligations or otherwise. Without limiting the generality of the foregoing, the obligations of Holdings hereunder shall not be discharged or impaired or otherwise affected by (i) the failure of the Buyer to assert any claim or demand or to enforce any right or remedy under the provisions of this Agreement or otherwise, (ii) any rescission, waiver, amendment or modification of, or any release from any of the terms or provisions of, this Agreement or any other agreement, (iii) the release of, or any impairment of or failure to perfect any Lien on or security interest in, any security held by the Buyer for the Obligations or any of them, (iv) any default, failure or delay, willful or otherwise, in the performance of the Obligations, or (v) any other act or omission that may or might in any manner or to any extent vary the risk of Holdings or otherwise operate as a discharge of Holdings as guarantor as a matter of law or equity. Holdings expressly authorizes the Buyer to take and hold security for the payment and performance of the Obligations, to exchange, waive or release any or all such security (with or without consideration), to enforce or apply such security and direct the order and manner of any sale thereof in its sole discretion or to release or substitute any one or more other guarantors or obligors upon or in respect of the Obligations, all without affecting the obligations of Holdings as a guarantor hereunder.
(b)To the fullest extent permitted by applicable law, Holdings waives any defense based on or arising out of any defense of the Seller or the unenforceability of the Obligations or any part thereof from any cause, or the cessation from any cause of the liability of the Seller, other than the fact that Seller is no longer obligated to make any Royalty Payments under this Agreement.
Section 11.4Reinstatement. Holdings agrees that its guarantee hereunder shall continue to be effective or be reinstated, as the case may be, if at any time payment, or any part thereof, of any Obligation is rescinded or must otherwise be restored by the Buyer upon the bankruptcy or reorganization of the Seller or otherwise.
Section 11.5Agreement To Pay; Subrogation. In furtherance of the foregoing and not in limitation of any other right that the Buyer has at law or in equity against Holdings by virtue hereof, upon the failure of the Seller to pay any Obligation when and as the same shall become due, Holdings hereby promises to and will forthwith pay, or cause to be paid, to Buyer in cash the amount of such unpaid Obligation. Upon payment by Holdings of any sums to the Buyer as provided above, all rights of Holdings against the Seller arising as a result thereof by way of right of subrogation, contribution, reimbursement, indemnity or otherwise shall in all respects be subject to Section 11.7.
Section 11.6Information. Holdings assumes all responsibility for being and keeping itself informed of the Seller’s financial condition and assets and of all other circumstances bearing upon the risk of nonpayment of the Obligations and the nature, scope and extent of the risks that Holdings assumes and incurs hereunder, and agrees that
40
the Buyer will not have any duty to advise Holdings of information known to it or any of them regarding such circumstances or risks.
Section 11.7Subordination.
(a)Notwithstanding any provision of this Agreement to the contrary, all of Holdings’ rights of indemnity, contribution or subrogation under applicable law or otherwise shall be fully subordinated to the payment in full the Obligations (except for and any contingent indemnification Obligations for which a claim or demand for payment has been not made at such time).
(b)Holdings hereby agrees that all Indebtedness and other monetary obligations owed by it to Seller shall be fully subordinated to the payment in full of the Obligations (except for and any contingent indemnification Obligations for which a claim or demand for payment has been not made at such time).
Section 11.8Termination. Holdings’ obligations as a guarantor under this Article 11 shall automatically terminate, without any action by any Person, upon the occurrence of a Release Event.
[Signature Pages Follow]
41
IN WITNESS WHEREOF, the parties hereto have caused this Agreement to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| MILESTONE PHARMACEUTICALS USA, INC. | |
| | |
| By: | /s/Amit Hasija |
| Name: Amit Hasija | |
| Title: Chief Financial Officer | |
| Milestone Pharmaceuticals Inc. | |
| | |
| By: | /s/ Amit Hasija |
| Name: Amit Hasija | |
| Title: Chief Financial Officer | |
[Signature Page to Purchase and Sale Agreement]
IN WITNESS WHEREOF, the parties hereto have caused this Agreement to be executed and delivered by their respective representatives thereunto duly authorized as of the date first above written.
| RTW ROYALTY I DAC | |
| | |
| By: | /s/Roderick Wang, M.D. |
| Name: Roderick Wang, M.D. | |
|---|---|---|
| Title: Authorized Attorney | |
[Signature Page to Purchase and Sale Agreement]
Exhibit A
Description of Etripamil
[***]
44
Exhibit B
Form of Bill of Sale
[***]
[Signature Page to Bill of Sale]
Exhibit C
Form of Royalty Report
[***]
46
Exhibit D
Assignment Agreement
[***]
Schedule 4.1(k)(i)
Patents
[***]
Exhibit 23.1
CONSENT OF INDEPENDENT REGISTERED PUBLIC ACCOUNTING FIRM
We hereby consent to the incorporation by reference in the Registration Statements on Form S-8 (Nos. 333-231347, 333-236971, 333-254838, 333-263807, 333-271522, 333-283160, 333-285765 and 333-289509) and on Form S-3 (Nos. 333-283162 and 333-271949) of Milestone Pharmaceuticals Inc. of our report dated March 20, 2026, relating to the consolidated financial statements, which appears in this Form 10-K.
/s/PricewaterhouseCoopers LLP
Montréal, Canada
March 20, 2026
Exhibit 31.1
CERTIFICATION PURSUANT TO
RULES 13a-14(a) AND 15d-14(a) UNDER THE SECURITIES EXCHANGE ACT OF 1934,
AS ADOPTED PURSUANT TO SECTION 302 OF THE SARBANES-OXLEY ACT OF 2002
I, Joseph Oliveto, certify that:
1. I have reviewed this Annual Report on Form 10-K of Milestone Pharmaceuticals Inc.;
2. Based on my knowledge, this report does not contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements made, in light of the circumstances under which such statements were made, not misleading with respect to the period covered by this report;
3. Based on my knowledge, the financial statements, and other financial information included in this report, fairly present in all material respects the financial condition, results of operations and cash flows of the registrant as of, and for, the periods presented in this report;
4. The registrant’s other certifying officer(s) and I are responsible for establishing and maintaining disclosure controls and procedures (as defined in Exchange Act Rules 13a-15(e) and 15d-15(e)) and internal control over financial reporting (as defined in Exchange Act Rules 13a-15(f) and 15d-15(f)) for the registrant and have:
(a) Designed such disclosure controls and procedures, or caused such disclosure controls and procedures to be designed under our supervision, to ensure that material information relating to the registrant, including its consolidated subsidiaries, is made known to us by others within those entities, particularly during the period in which this report is being prepared;
(b) Designed such internal control over financial reporting, or caused such internal control over financial reporting to be designed under our supervision, to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with generally accepted accounting principles;
(c) Evaluated the effectiveness of the registrant’s disclosure controls and procedures and presented in this report our conclusions about the effectiveness of the disclosure controls and procedures, as of the end of the period covered by this report based on such evaluation; and
(d) Disclosed in this report any change in the registrant’s internal control over financial reporting that occurred during the registrant’s most recent fiscal quarter (the registrant’s fourth fiscal quarter in the case of an annual report) that has materially affected, or is reasonably likely to materially affect, the registrant’s internal control over financial reporting; and
5. The registrant’s other certifying officer(s) and I have disclosed, based on our most recent evaluation of internal control over financial reporting, to the registrant’s auditors and the audit committee of the registrant’s board of directors (or persons performing the equivalent functions):
(a) All significant deficiencies and material weaknesses in the design or operation of internal control over financial reporting which are reasonably likely to adversely affect the registrant’s ability to record, process, summarize and report financial information; and
(b) Any fraud, whether or not material, that involves management or other employees who have a significant role in the registrant’s internal control over financial reporting.
Date: March 20, 2026 | |
| /s/ Joseph Oliveto |
| Joseph Oliveto |
| President and Chief Executive Officer |
| (Principal Executive Officer) |
Exhibit 31.2
CERTIFICATION PURSUANT TO
RULES 13a-14(a) AND 15d-14(a) UNDER THE SECURITIES EXCHANGE ACT OF 1934,
AS ADOPTED PURSUANT TO SECTION 302 OF THE SARBANES-OXLEY ACT OF 2002
I, Amit Hasija, certify that:
1. I have reviewed this Annual Report on Form 10-K of Milestone Pharmaceuticals Inc.;
2. Based on my knowledge, this report does not contain any untrue statement of a material fact or omit to state a material fact necessary to make the statements made, in light of the circumstances under which such statements were made, not misleading with respect to the period covered by this report;
3. Based on my knowledge, the financial statements, and other financial information included in this report, fairly present in all material respects the financial condition, results of operations and cash flows of the registrant as of, and for, the periods presented in this report;
4. The registrant’s other certifying officer(s) and I are responsible for establishing and maintaining disclosure controls and procedures (as defined in Exchange Act Rules 13a-15(e) and 15d-15(e)) and internal control over financial reporting (as defined in Exchange Act Rules 13a-15(f) and 15d-15(f)) for the registrant and have:
(a) Designed such disclosure controls and procedures, or caused such disclosure controls and procedures to be designed under our supervision, to ensure that material information relating to the registrant, including its consolidated subsidiaries, is made known to us by others within those entities, particularly during the period in which this report is being prepared;
(b) Designed such internal control over financial reporting, or caused such internal control over financial reporting to be designed under our supervision, to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with generally accepted accounting principles;
(c) Evaluated the effectiveness of the registrant’s disclosure controls and procedures and presented in this report our conclusions about the effectiveness of the disclosure controls and procedures, as of the end of the period covered by this report based on such evaluation; and
(d) Disclosed in this report any change in the registrant’s internal control over financial reporting that occurred during the registrant’s most recent fiscal quarter (the registrant’s fourth fiscal quarter in the case of an annual report) that has materially affected, or is reasonably likely to materially affect, the registrant’s internal control over financial reporting; and
5. The registrant’s other certifying officer(s) and I have disclosed, based on our most recent evaluation of internal control over financial reporting, to the registrant’s auditors and the audit committee of the registrant’s board of directors (or persons performing the equivalent functions):
(a) All significant deficiencies and material weaknesses in the design or operation of internal control over financial reporting which are reasonably likely to adversely affect the registrant’s ability to record, process, summarize and report financial information; and
(b) Any fraud, whether or not material, that involves management or other employees who have a significant role in the registrant’s internal control over financial reporting.
Date: March 20, 2026 | |
| /s/ Amit Hasija |
| Amit Hasija |
| Chief Financial Officer |
| (Principal Financial and Accounting Officer) |
Exhibit 32.1
CERTIFICATION PURSUANT TO
18 U.S.C. SECTION 1350, AS ADOPTED PURSUANT TO
SECTION 906 OF THE SARBANES-OXLEY ACT OF 2002
Pursuant to the requirement set forth in Rule 13a-14(b) of the Securities Exchange Act of 1934, as amended, (the “Exchange Act”) and Section 1350 of Chapter 63 of Title 18 of the United States Code (18 U.S.C. §1350), Joseph Oliveto, Chief Executive Officer of Milestone Pharmaceuticals Inc. (the “Company”), and Amit Hasija, Chief Financial Officer of the Company, each hereby certifies that, to the best of his or her knowledge:
1. The Company’s Annual Report on Form 10-K for the period ended December 31, 2025, to which this Certification is attached as Exhibit 32.1 (the “Periodic Report”), fully complies with the requirements of Section 13(a) or Section 15(d) of the Exchange Act; and
2. The information contained in the Periodic Report fairly presents, in all material respects, the financial condition and results of operations of the Company.
Dated: March 20, 2025 | | |
| | |
| | |
/s/ Joseph Oliveto | | /s/ Amit Hasija |
Joseph Oliveto | | Amit Hasija |
Chief Executive Officer | | Chief Financial Officer |
(Principal Executive Officer) | | (Principal Financial and Accounting Officer) |
“This certification accompanies the Form 10-K to which it relates, is not deemed filed with the Securities and Exchange Commission and is not to be incorporated by reference into any filing of Milestone Pharmaceuticals Inc under the Securities Act of 1933, as amended, or the Securities Exchange Act of 1934, as amended (whether made before or after the date of the Form 10-K), irrespective of any general incorporation language contained in such filing.”